Related Experiment Video
Updated: Jan 5, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Pseudodiagonalization-based wavefunction optimization with contracted planewave basis functions
Duncan W Stuart1, Nicholas J Mosey1
1Department of Chemistry, 90 Bader Lane, Queen's University, Kingston, Ontario, Canada, K7L 3N6.

Abstract:
Electronic structure calculations representing the molecular orbitals (MOs) with contracted planewave basis functions (CPWBFs) have been reported recently. CPWBFs are Fourier-series representations of atom-centered basis functions. The mathematical features of CPWBFs permit the construction of matrix-vector products, FC o , involving the application of the Fock matrix, F, to the set of occupied MOs, C o , without the explicit evaluation of F. This approach offers a theoretical speed-up of M/n over F-based methods, where M and n are the number of basis functions and occupied MOs, respectively. The present study reports methodological advances that permit FC o -based optimization of wavefunction formed from CPWBFs. In particular, a technique is reported for optimizing wavefunctions by combining pseudodiagonalization techniques based on an exact representation of FC o , approximate information regarding the virtual orbital energies, and direct inversion of the iterative subspace optimization schemes to guide the wavefunction to a converged solution. This method is found to speed-up wavefunction optimizations by factors of up to ~6 - 8 over F-based optimization methods while providing identical results. Further, the computational cost of this technique is relatively insensitive to basis set size, thus providing further benefits in calculations using large CPWBF basis sets. The results of density functional theory calculations show that this method permits the use of hybrid exchange-correlation (XC) functionals with a small increase in effort over analogous calculations using generalized gradient approximation XC functionals. © 2019 Wiley Periodicals, Inc.
More Related Videos


Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 01:13
Graphing the Wave Function
 01:10
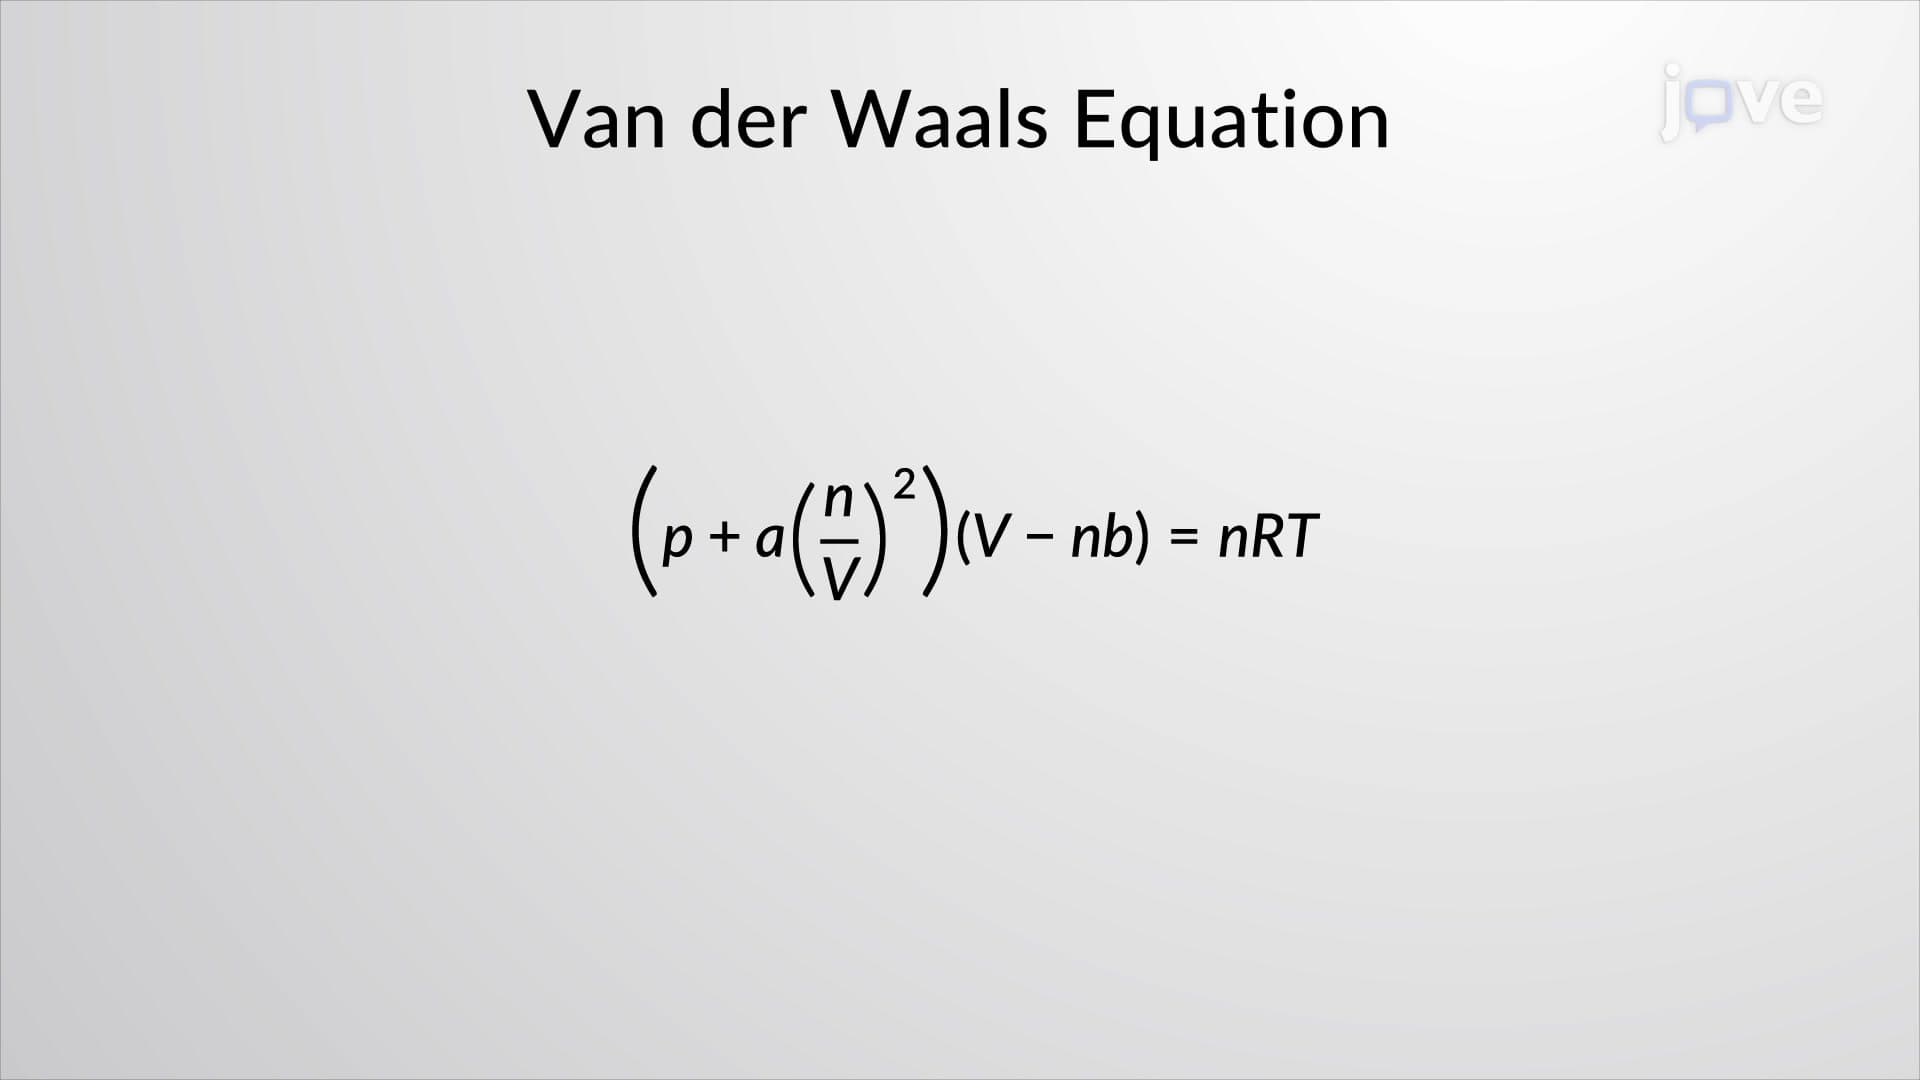
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...