Well-Balanced Force Field ff03CMAP for Folded and Disordered Proteins
Yangpeng Zhang1, Hao Liu1, Sheng Yang1
1State Key Laboratory of Microbial metabolism, Department of Bioinformatics and Biostatistics, National Experimental Teaching Center for Life Sciences and Biotechnology, School of Life Sciences and Biotechnology , Shanghai Jiao Tong University , Shanghai 200240 , China.

Abstract:
Molecular dynamics simulation as an important complement of experiment is widely used to study protein structures and functions. However, previous studies indicate that the current force fields cannot, simultaneously, provide accurate descriptions of folded proteins and intrinsically disordered proteins (IDPs). Therefore, a correction maps (CMAP)-optimized force field based on the Amber ff03 force field (termed ff03CMAP herein) was developed for a balanced sampling of folded proteins and IDPs. Extensive validations of short peptides, folded proteins, disordered proteins, and fast-folding proteins show that simulated chemical shifts, J-coupling constants, order parameters, and residual dipolar couplings (RDCs) with the ff03CMAP force field are in very good agreement with nuclear magnetic resonance measurements and are more accurate than other ff03-series force fields. The influence of solvent models was also investigated. It was found that the combination of ff03CMAP/TIP4P-Ew is suitable for folded proteins, and that of ff03CMAP/TIP4PD is better for disordered proteins. These findings confirm that the newly developed force field ff03CMAP can improve the balance of conformer sampling between folded proteins and IDPs.
Related Concept Videos
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:30
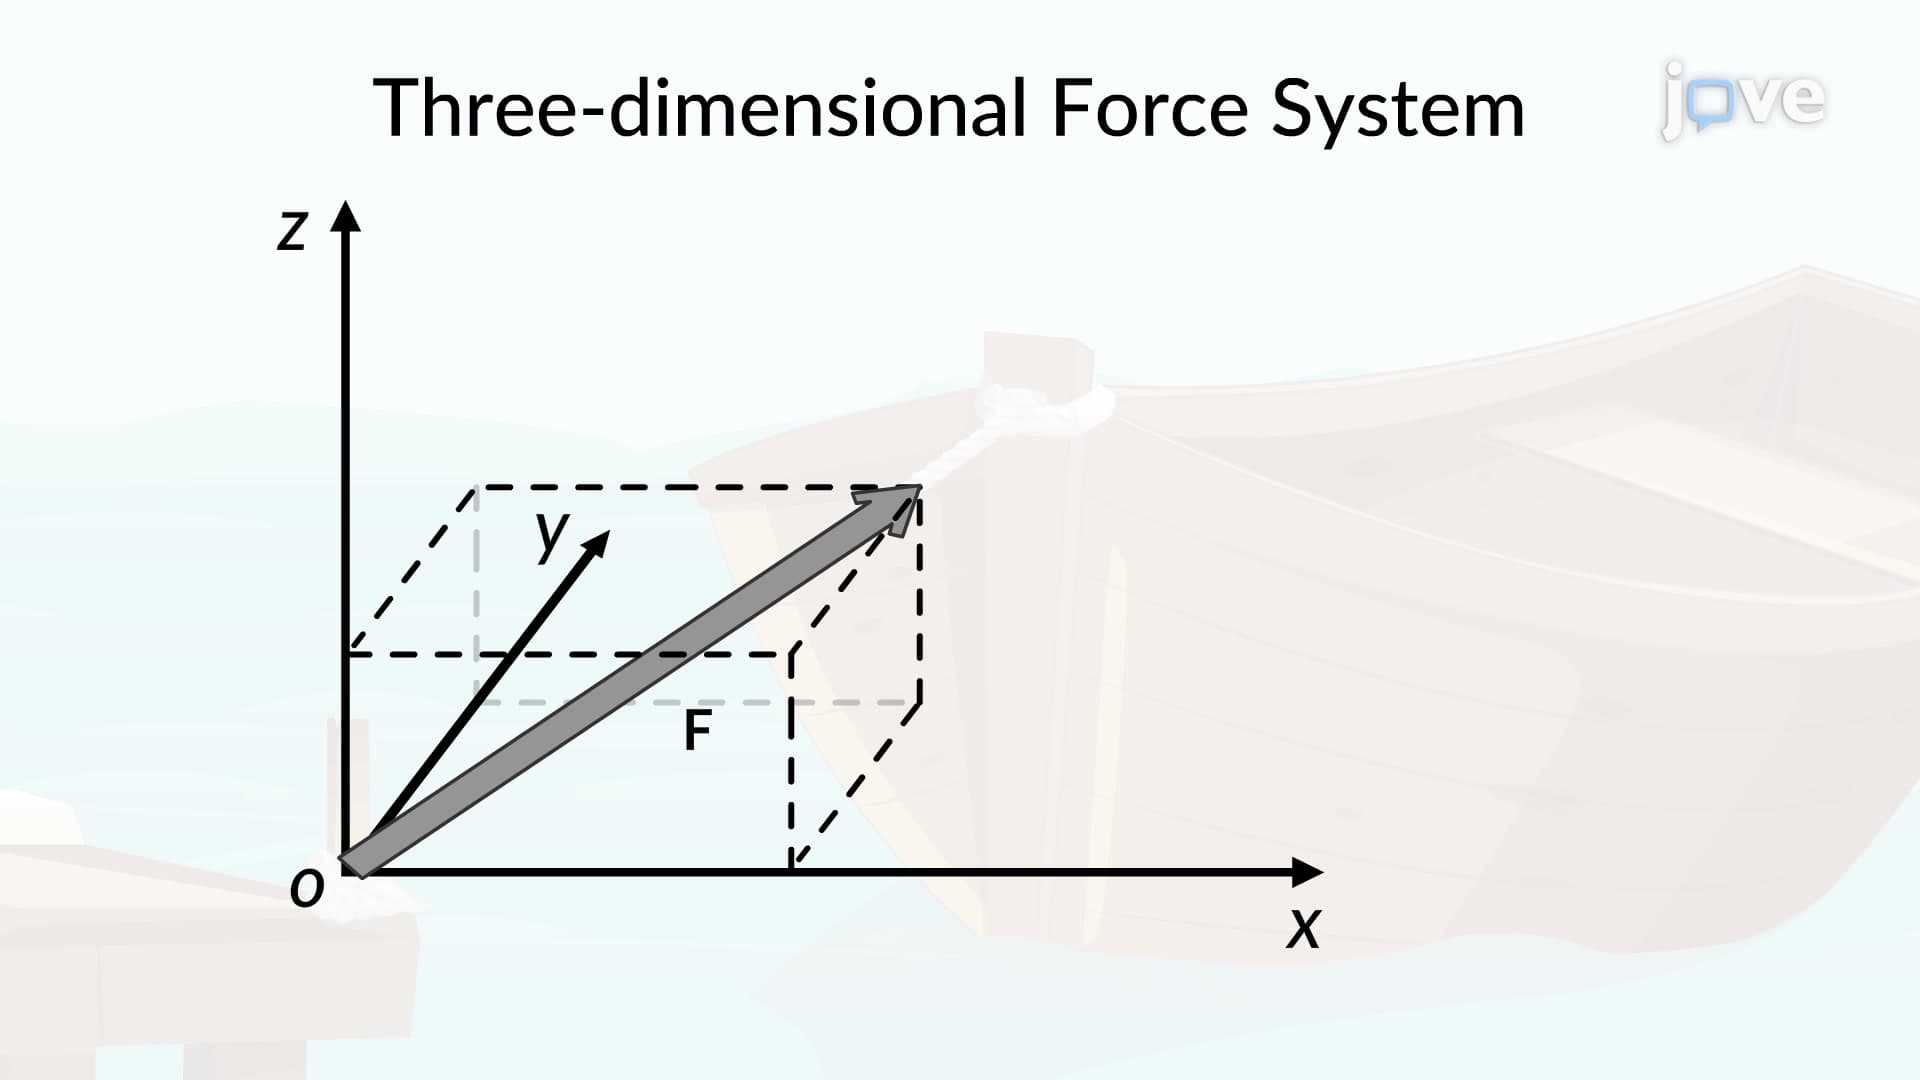
Three-Dimensional Force System