Related Experiment Video
Updated: Jan 3, 2026
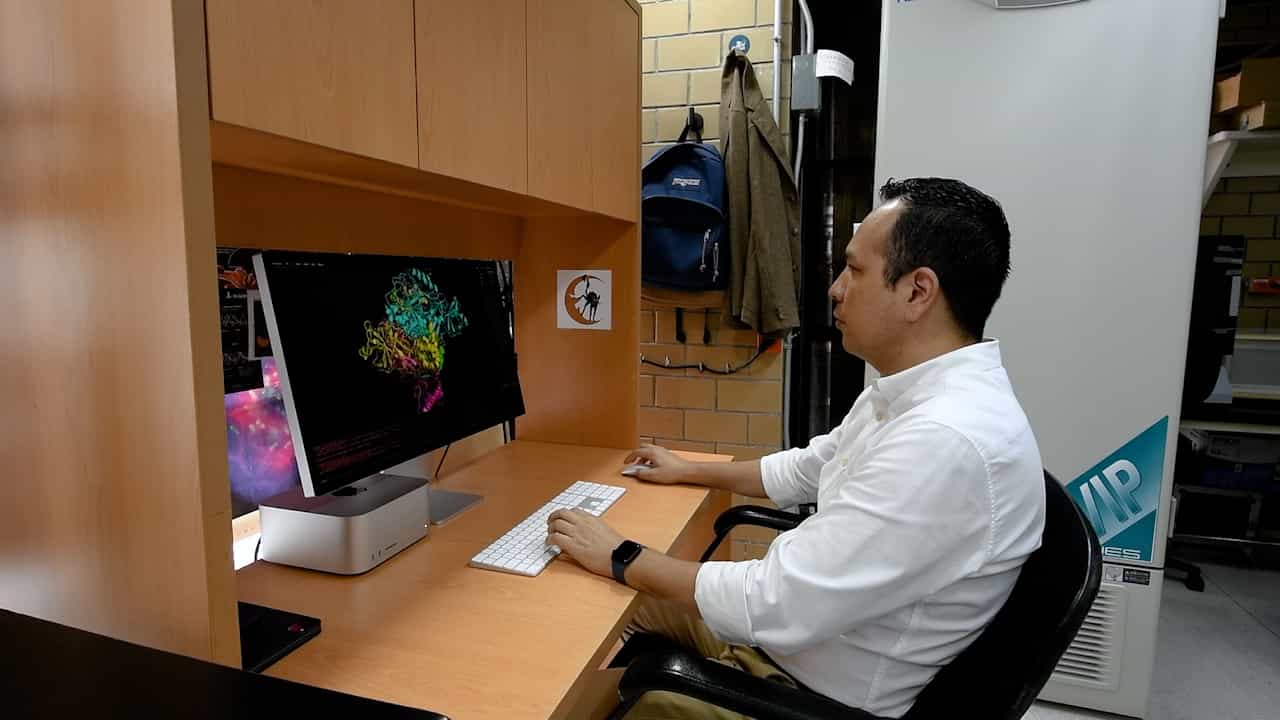

Pharmacophore Modeling for Targets with Extensive Ligand Libraries: A Case Study on SARS-CoV-2 Mpro
Published on: September 26, 2025
Ligand-Based Pharmacophore Modeling and Virtual Screening to Discover Novel CYP1A1 Inhibitors
Rana Adnan Tahir1,2, Farwa Hassan1, Abdul Kareem3
1Department of Biosciences, COMSATS University Islamabad, Sahiwal Campus, Sahiwal, Pakistan.

Abstract:
Backgound: Cytochrome P450, family 1, subfamily A, polypeptide 1 (CYP1A1) is an imperative enzyme due to its immersion in the biotransformation of a wide range of drugs and other xenobiotics. The involvement of enzymes in drug metabolism indicates an effective drug target for the development of novel therapeutics. The discovery of CYP1A1 specific inhibitors would be of particular relevance for the clinical pharmacology.
Methods:
In the current work, in silico approaches were utilized to identify the novel potential compounds through a diverse set of reported inhibitors against CYP1A1. A dataset of reported compounds against CYP1 belongs to 10 different classes (alkaloids, coumarins, flavonoids, natural compounds, synthetic inhibitors, drugs, MBI's, PAHs, naphthoquinone and stilbenoids) was retrieved and utilized for the comparative molecular docking analyses followed by pharmacophore modeling. The total eleven novel compounds were scrutinized on the basis of the highest binding affinities and least binding energy values.
Results:
ZINC08792486 compound attained the highest gold fitness score of 90.11 against CYP1A1 among all the scrutinized molecules.
Conclusion:
It has been elucidated that the residues Phe-224, Gly-316 and Ala-317 were conserved in all ligand-receptor interactions and critical for the development of effective therapies. The ADMET property analyses also predict better absorption and distribution of the selected hits that may be used in the future for in vitro validations and drug development.
More Related Videos
 10:25
Screening Traditional Chinese Medicine Compounds for Inhibiting UCHL3 Activity Based on Molecular Docking and Deubiquitinating Enzyme Probe Technology
Published on: November 22, 2024
 08:49
Identification of Mediators of T-cell Receptor Signaling via the Screening of Chemical Inhibitor Libraries
Published on: January 22, 2019
Related Concept Videos
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...