Related Experiment Video
Updated: Jan 3, 2026


Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
A shared-weight neural network architecture for predicting molecular properties
Trevor A Profitt1, Jason K Pearson
1Department of Chemistry, University of Prince Edward Island, Charlottetown, PE, Canada. jpearson@upei.ca.

Abstract:
Quantum chemical methods scale poorly with increasing molecular size and machine learning models have emerged as a promising, computationally-efficient alternative. We present a shared-weight neural network architecture based on modified atom-centered symmetry functions (ACSFs) and show that it performs similarly to the more computationally expensive per-element neural networks of previous work with ACSFs. The model achieves chemically accurate predictions, with a mean absolute error as low as 0.63 kcal mol-1 on energy predictions in the QM9 data set. Additionally, we show that it can reliably predict atomic forces.

Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:24
Predicting Reaction Outcomes
 02:00
Molecular Models
 01:01
Polymers: Defining Molecular Weight
The number average molecular weight (Mn) is the summation of the number...
 01:08
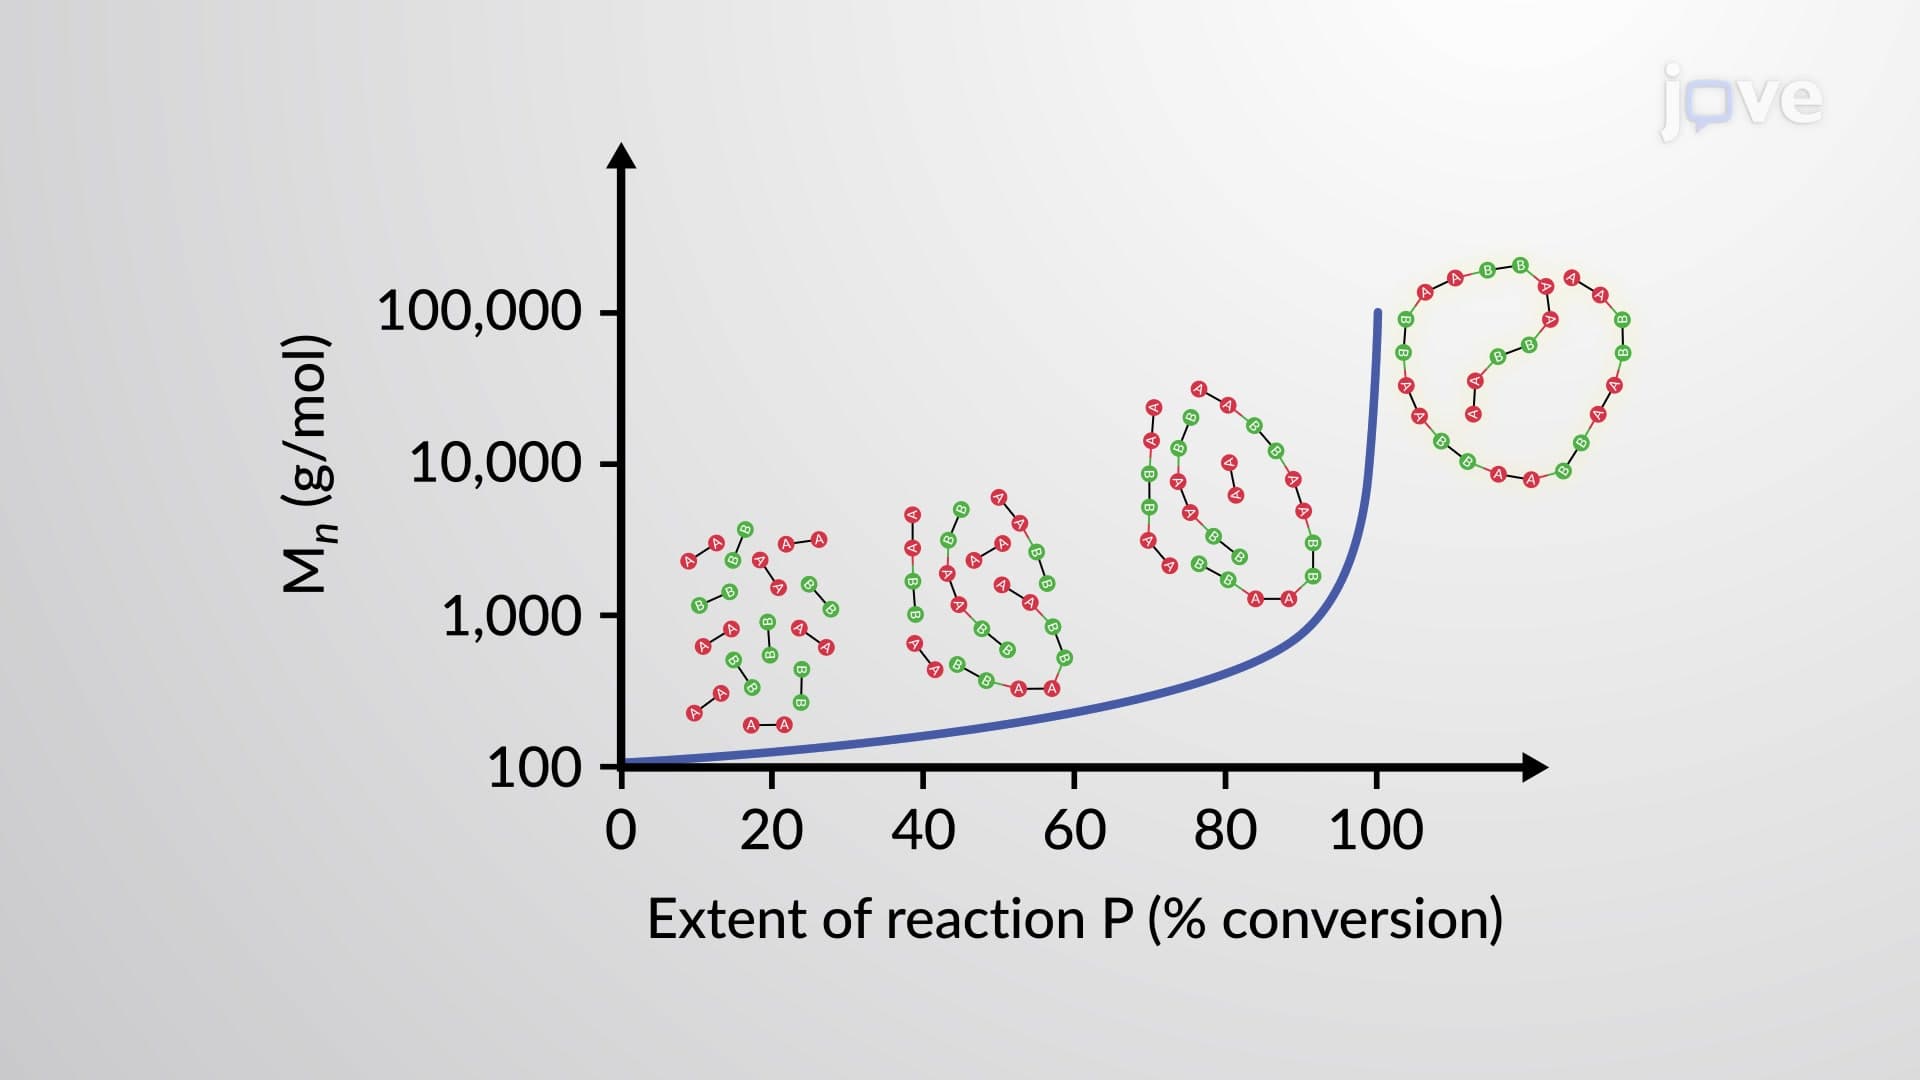
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 01:30
Classification of Neurotransmitters