Related Experiment Video
Updated: Jan 2, 2026
Paramagnetic Relaxation Enhancement for Detecting and Characterizing Self-Associations of Intrinsically Disordered Proteins
Published on: September 23, 2021
Disordered Function Conjunction: On the in-silico function annotation of intrinsically disordered regions
Sina Ghadermarzi1, Akila Katuwawala, Christopher J Oldfield
1Department of Computer Science, Virginia Commonwealth University, 401 West Main Street, Richmond, VA 23284, U.S.A.

Abstract:
Intrinsically disorder regions (IDRs) lack a stable structure, yet perform biological functions. The functions of IDRs include mediating interactions with other molecules, including proteins, DNA, or RNA and entropic functions, including domain linkers. Computational predictors provide residue-level indications of function for disordered proteins, which contrasts with the need to functionally annotate the thousands of experimentally and computationally discovered IDRs. In this work, we investigate the feasibility of using residue-level prediction methods for region-level function predictions. For an initial examination of the multiple function region-level prediction problem, we constructed a dataset of (likely) single function IDRs in proteins that are dissimilar to the training datasets of the residue-level function predictors. We find that available residue-level prediction methods are only modestly useful in predicting multiple region-level functions. Classification is enhanced by simultaneous use of multiple residue-level function predictions and is further improved by inclusion of amino acids content extracted from the protein sequence. We conclude that multifunction prediction for IDRs is feasible and benefits from the results produced by current residue-level function predictors, however, it has to accommodate inaccuracy in functional annotations.
More Related Videos
Related Concept Videos
 02:18
Intrinsically Disordered Proteins
02:18Intrinsically Disordered Proteins
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:04
Protein-protein Interfaces
 03:36
Genome Annotation and Assembly
 01:24
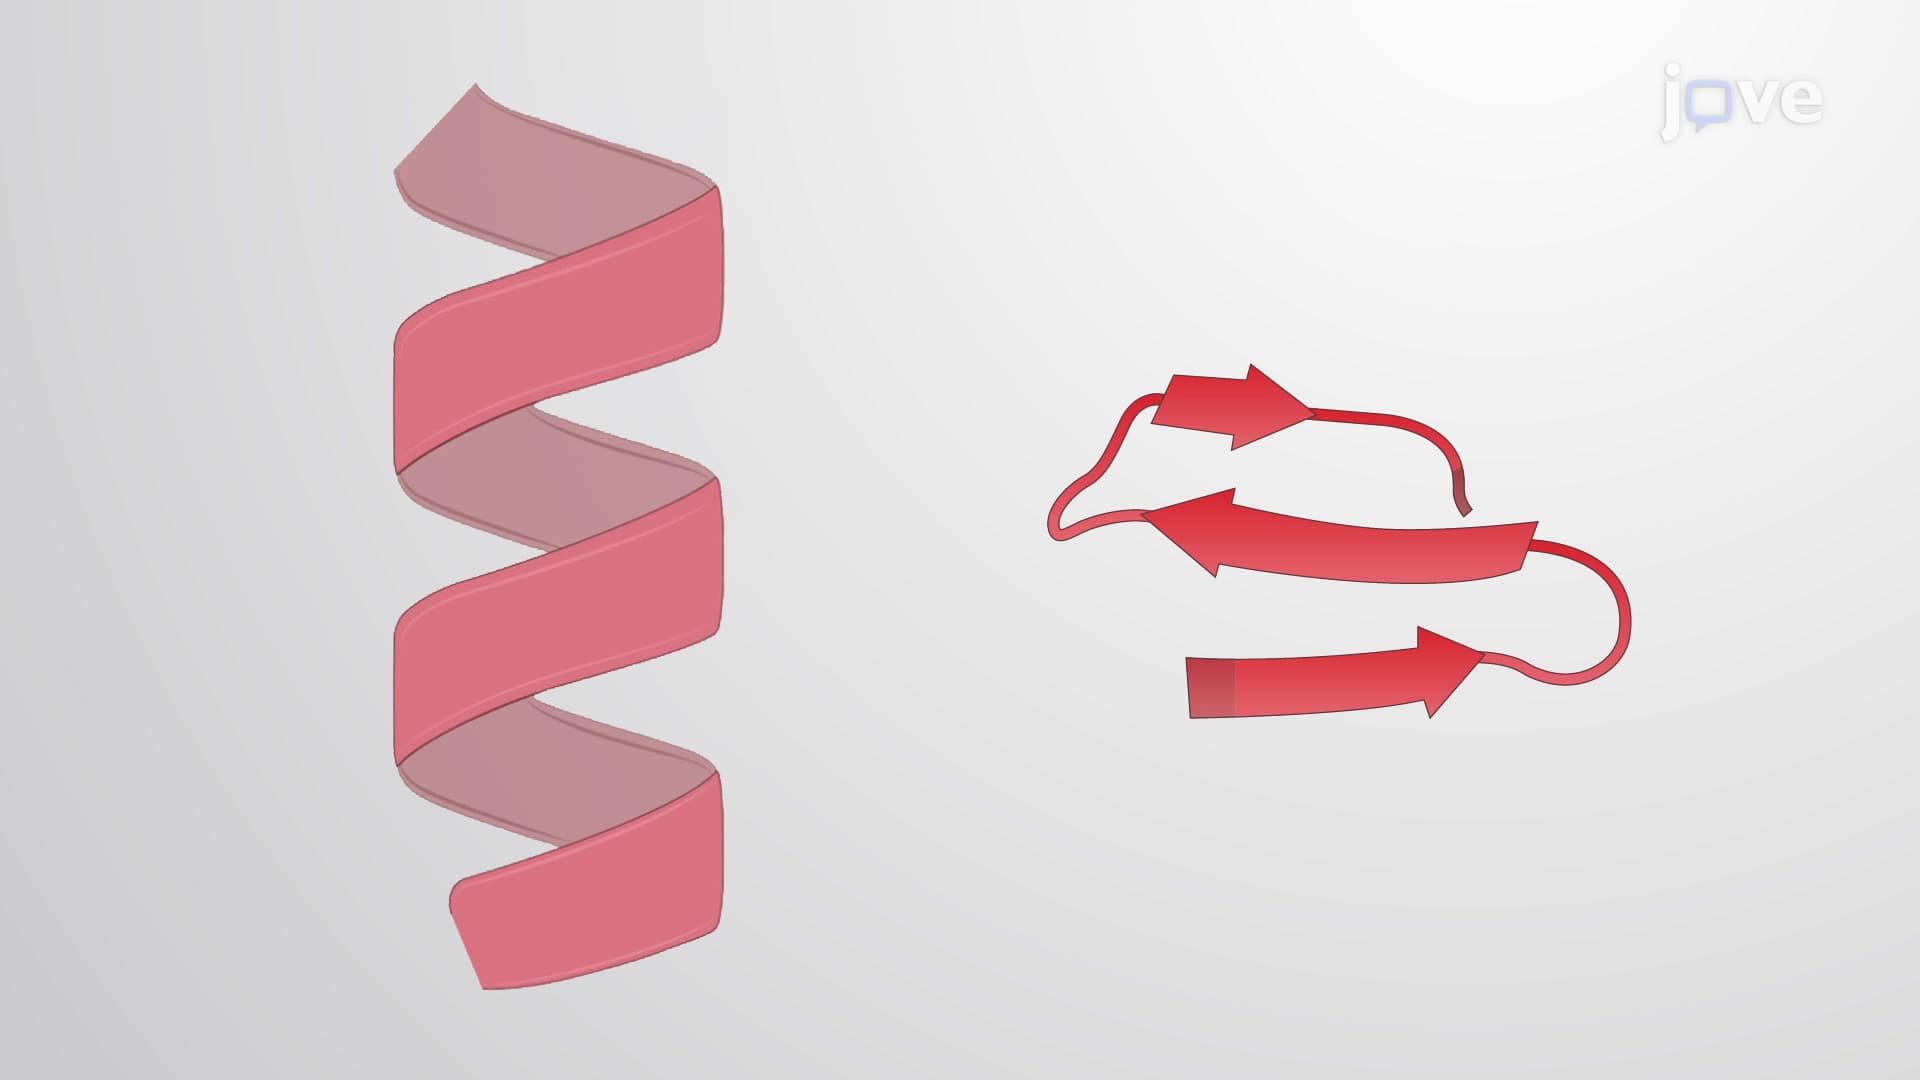
Protein Organization
The primary structure of a protein is its amino acid sequence....