Related Experiment Video
Updated: Jan 2, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Predicting HSE band gaps from PBE charge densities via neural network functionals
Levi C Lentz1, Alexie M Kolpak1
1Massachusetts Institute of Technology, Mechanical Engineering, Cambridge, MA 02139, United States of America.

Abstract:
Density functional theory (DFT) has become a standard method for ab initio calculations of material properties. However, it has a number of shortcomings, particularly in predicting key properties, such as band gap and optical spectra, which are dependent on excited states. To treat such properties, more accurate approaches such as GW or DFT with hybrid functionals (including HSE, PBE0, and B3LYP, to name a few) can be employed; however, these approaches are unfeasible for many large and/or complex systems due to their high computational cost and large memory requirements. In this work, we investigate the ability to train neural networks of the traditional DFT charge density computed with a standard PBE functional to accurately predict HSE band gaps. We show that a single network PBE charge density functional can predict the HSE band gap of seven different materials-silicon, gallium arsenide, molybdenum disulfide, germanium, tin phosphate, titanium phosphate, and zirconium phosphate-under a wide variety of conditions with an RSME of 172.6 meV, which is 34% better accuracy than standard regression between the PBE and HSE band gaps. This approach, which, in principle, can be used to map PBE charge densities to band gaps or other properties computed with any higher accuracy method, has the potential to decrease computational costs, increase prediction accuracy, and enable accurate high-throughput screening for a wide variety of complex materials systems.
More Related Videos


Related Concept Videos
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states...
 02:35
Band Theory
The energy difference between these bands is known as the band gap.
Conductor, Semiconductor,...
 01:18
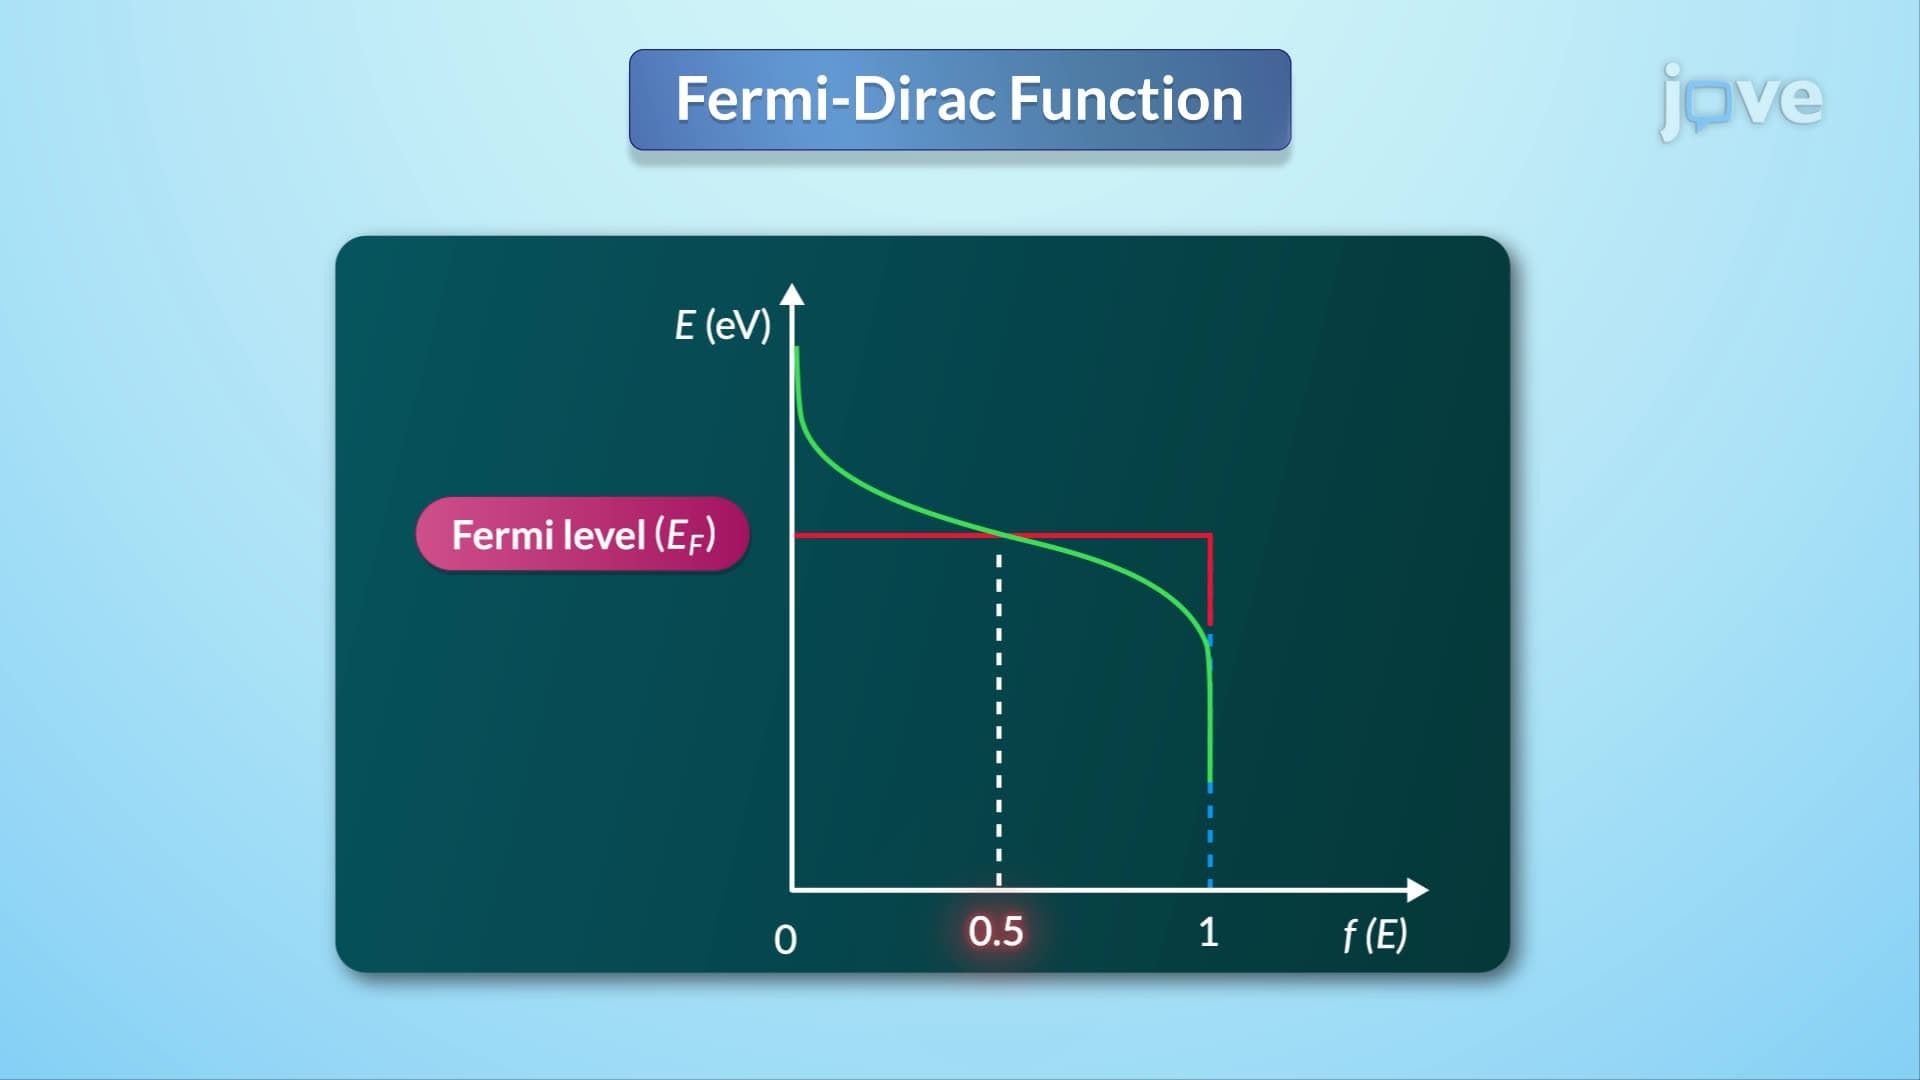
Fermi Level
At absolute zero temperature, electrons fill all energy states up to the Fermi level, leaving upper states empty. As the temperature rises,...
 01:12
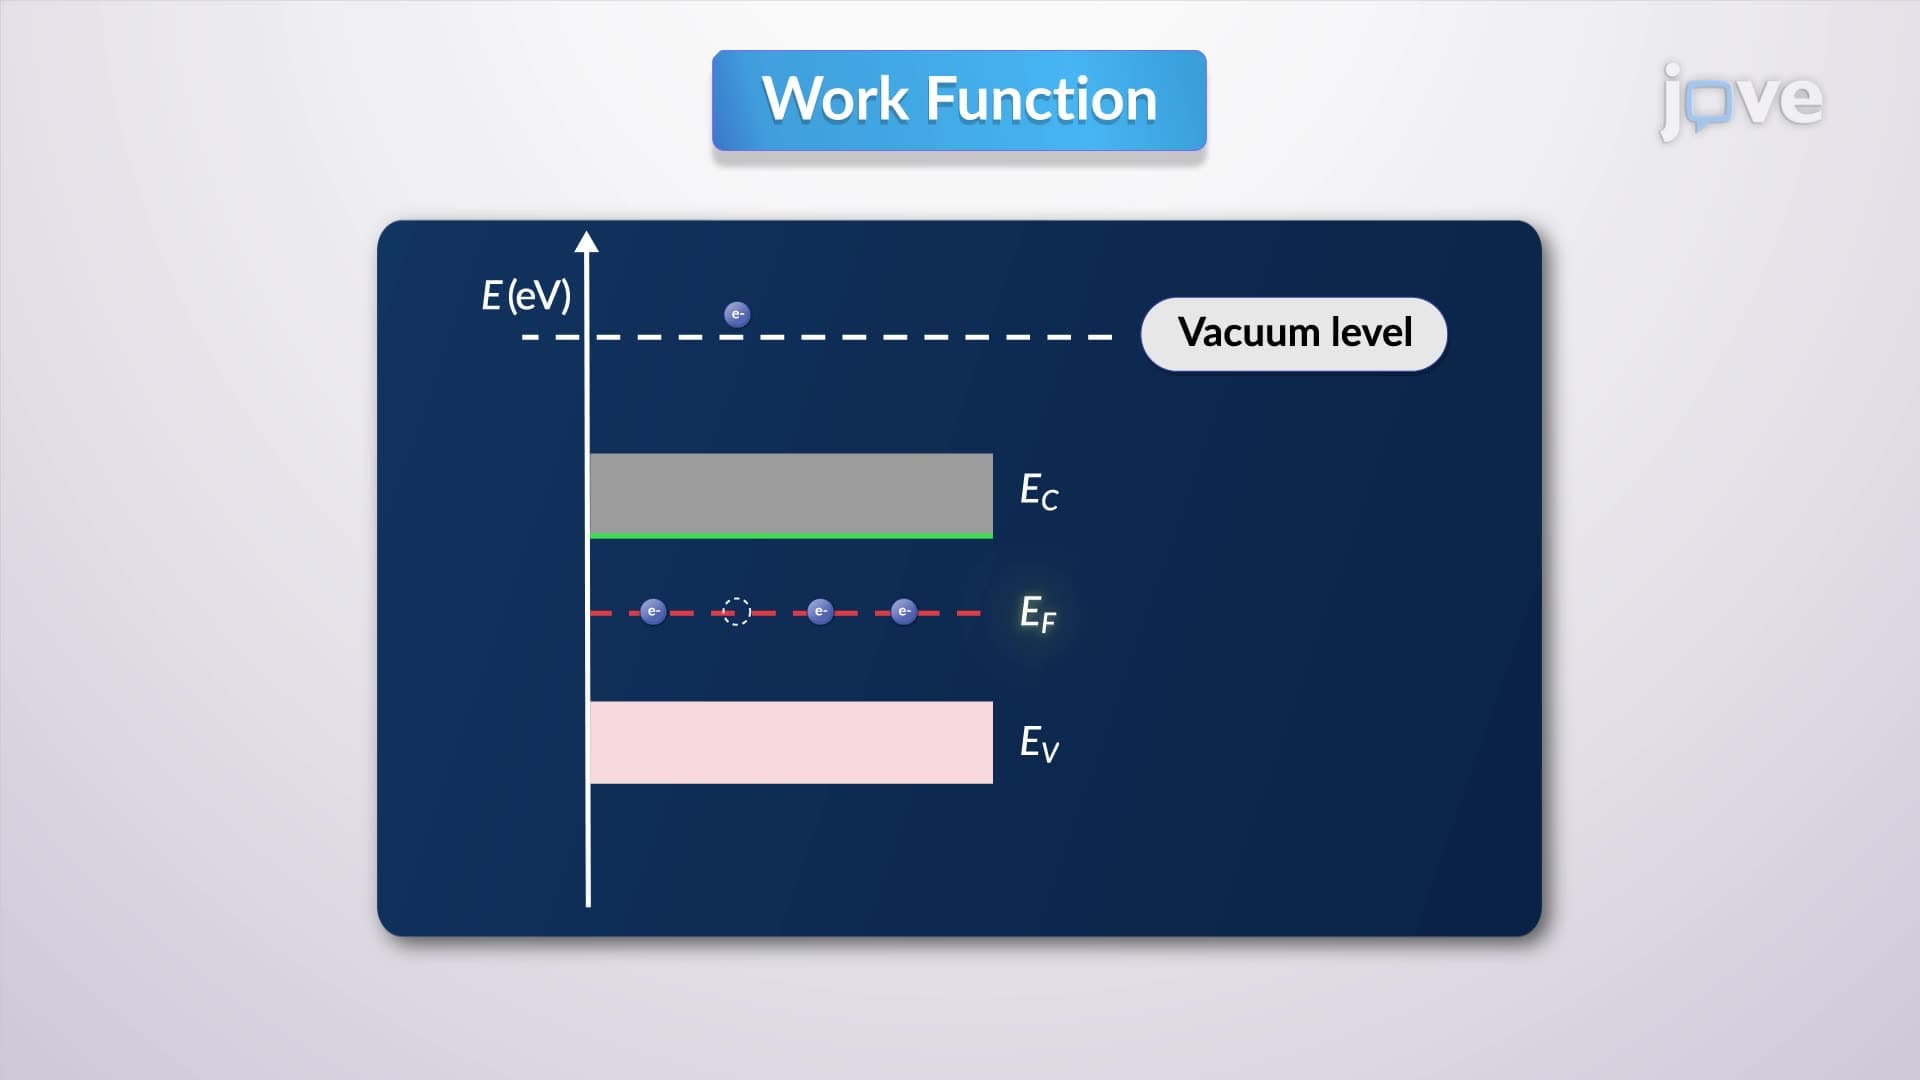
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:22
Semiconductors
Metals such as copper (Cu), zinc (Zn), or lead (Pb) have low resistivity and feature conduction bands that are either not fully occupied or overlap with the valence band, making a bandgap non-existent. This allows electrons in the highest energy levels of the valence band to easily transition to the conduction band upon gaining...
 02:27
Predicting Molecular Geometry