Related Experiment Video
Updated: Jan 2, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Self-Consistent Range-Separated Density-Functional Theory with Second-Order Perturbative Correction via the
Szymon Śmiga1, Ireneusz Grabowski1, Mateusz Witkowski1
1Institute of Physics, Faculty of Physics, Astronomy and Informatics , Nicolaus Copernicus University , 87-100 Toruń , Poland.

Abstract:
We extend the range-separated double-hybrid RSH+MP2 method (Ángyán, J. G.; et al. Phys. Rev. A 2005, 72, 012510), combining long-range HF exchange and MP2 correlation with a short-range density functional to a fully self-consistent version using the optimized-effective-potential technique in which the orbitals are obtained from a local potential including the long-range HF and MP2 contributions. We test this approach, that we name RS-OEP2, on a set of small closed-shell atoms and molecules. For the commonly used value of the range-separation parameter μ = 0.5 bohr-1, we find that self-consistency does not seem to bring any improvement for total energies, ionization potentials, and electronic affinities. However, contrary to the non-self-consistent RSH+MP2 method, the present RS-OEP2 method gives a LUMO energy which physically corresponds to a neutral excitation energy and gives local exchange-correlation potentials which are reasonably good approximations to the corresponding Kohn-Sham quantities. At a finer scale, we find that RS-OEP2 gives largely inaccurate correlation potentials and correlated densities, which points to the need of further improvement of this type of range-separated double hybrids.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:51
Molecular Orbital Theory II
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 01:10
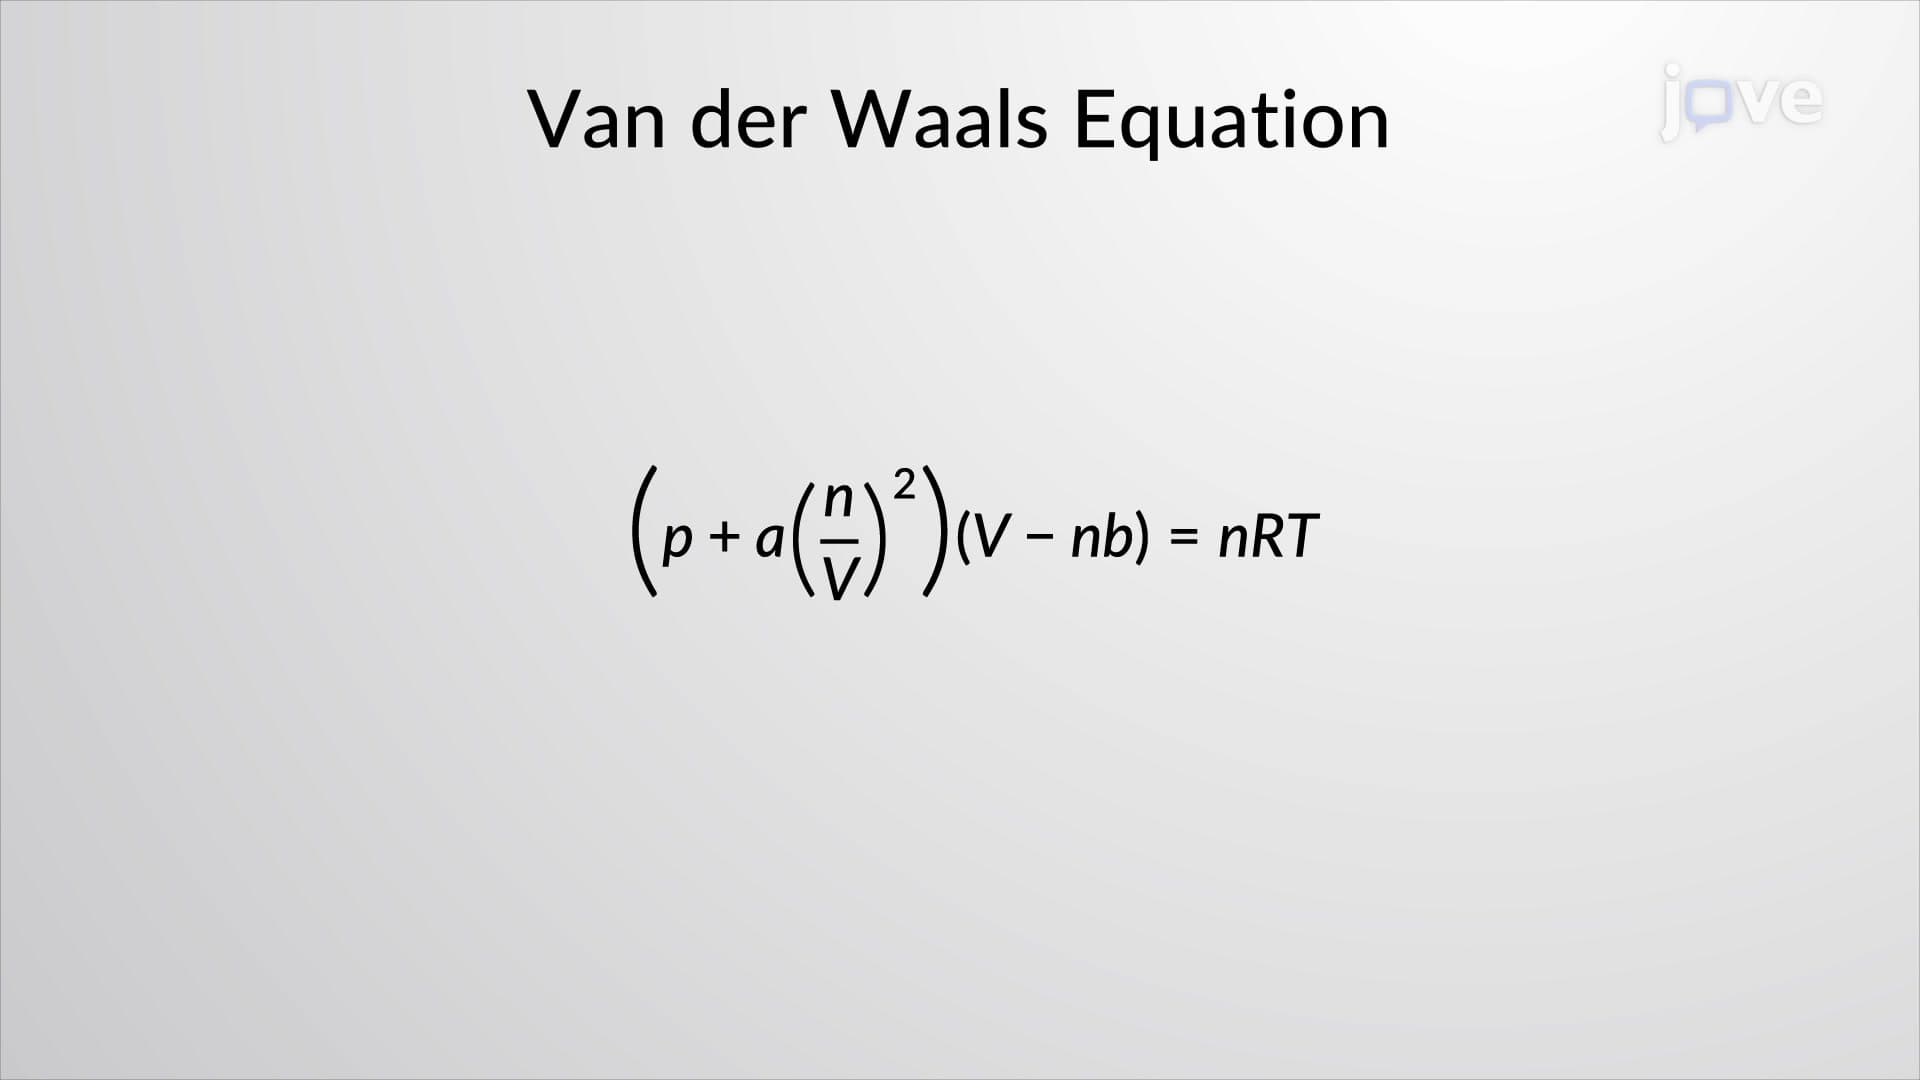
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...