Related Experiment Video
Updated: Jan 1, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Unbiasing the initiator approximation in full configuration interaction quantum Monte Carlo
Khaldoon Ghanem1, Alexander Y Lozovoi1, Ali Alavi1
1Max Planck Institute for Solid State Research, Heisenbergstr. 1, 70569 Stuttgart, Germany.

Abstract:
We identify and rectify a crucial source of bias in the initiator full configuration interaction quantum Monte Carlo algorithm. Noninitiator determinants (i.e., determinants whose population is below the initiator threshold) are subject to a systematic undersampling bias, which in large systems leads to a bias in the energy when an insufficient number of walkers are used. We show that the acceptance probability (pacc), that a noninitiator determinant has its spawns accepted, can be used to unbias the initiator bias, in a simple and accurate manner, by reducing the applied shift to the noninitiator proportionately to pacc. This modification preserves the property that in the large walker limit, when pacc → 1, the unbiasing procedure disappears, and the initiator approximation becomes exact. We demonstrate that this algorithm shows rapid convergence to the FCI limit with respect to the walker number and, furthermore, largely removes the dependence of the algorithm on the initiator threshold, enabling highly accurate results to be obtained even with large values of the threshold. This is exemplified in the case of butadiene/ANO-L-pVDZ and benzene/cc-pVDZ, correlating 22 and 30 electrons in 82 and 108 orbitals, respectively. In butadiene 5 × 10⁷ and in benzene 108 walkers suffice to obtain an energy within a millihartree of the coupled cluster singles doubles triples and perturbative quadruples [CCSDT(Q)] result in Hilbert spaces of 1026 and 1035, respectively. Essentially converged results require ∼108 walkers for butadiene and ∼109 walkers for benzene and lie slightly lower than CCSDT(Q). Owing to large-scale parallelizability, these calculations can be executed in a matter of hours on a few hundred processors. The present method largely solves the initiator-bias problems that the initiator method suffered from when applied to medium-sized molecules.
Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 01:27
Biasing of Metal-Semiconductor Junctions
In Schottky junctions, where the semiconductor is n-type, applying a positive voltage to the metal relative to the semiconductor reduces its Fermi...
 03:35
Hybridization of Atomic Orbitals II
 01:16
Biasing of P-N Junction
In equilibrium, no external voltage is applied across the p-n junction. The depletion region is formed at the junction interface due to the diffusion of carriers, which leaves behind charged dopants, acceptors on the p-side, and donors on the n-side. These immobile charges create an electric field that prevents further diffusion of carriers. The related energy band...
 04:08
The Uncertainty Principle
 01:22
Biasing of FET
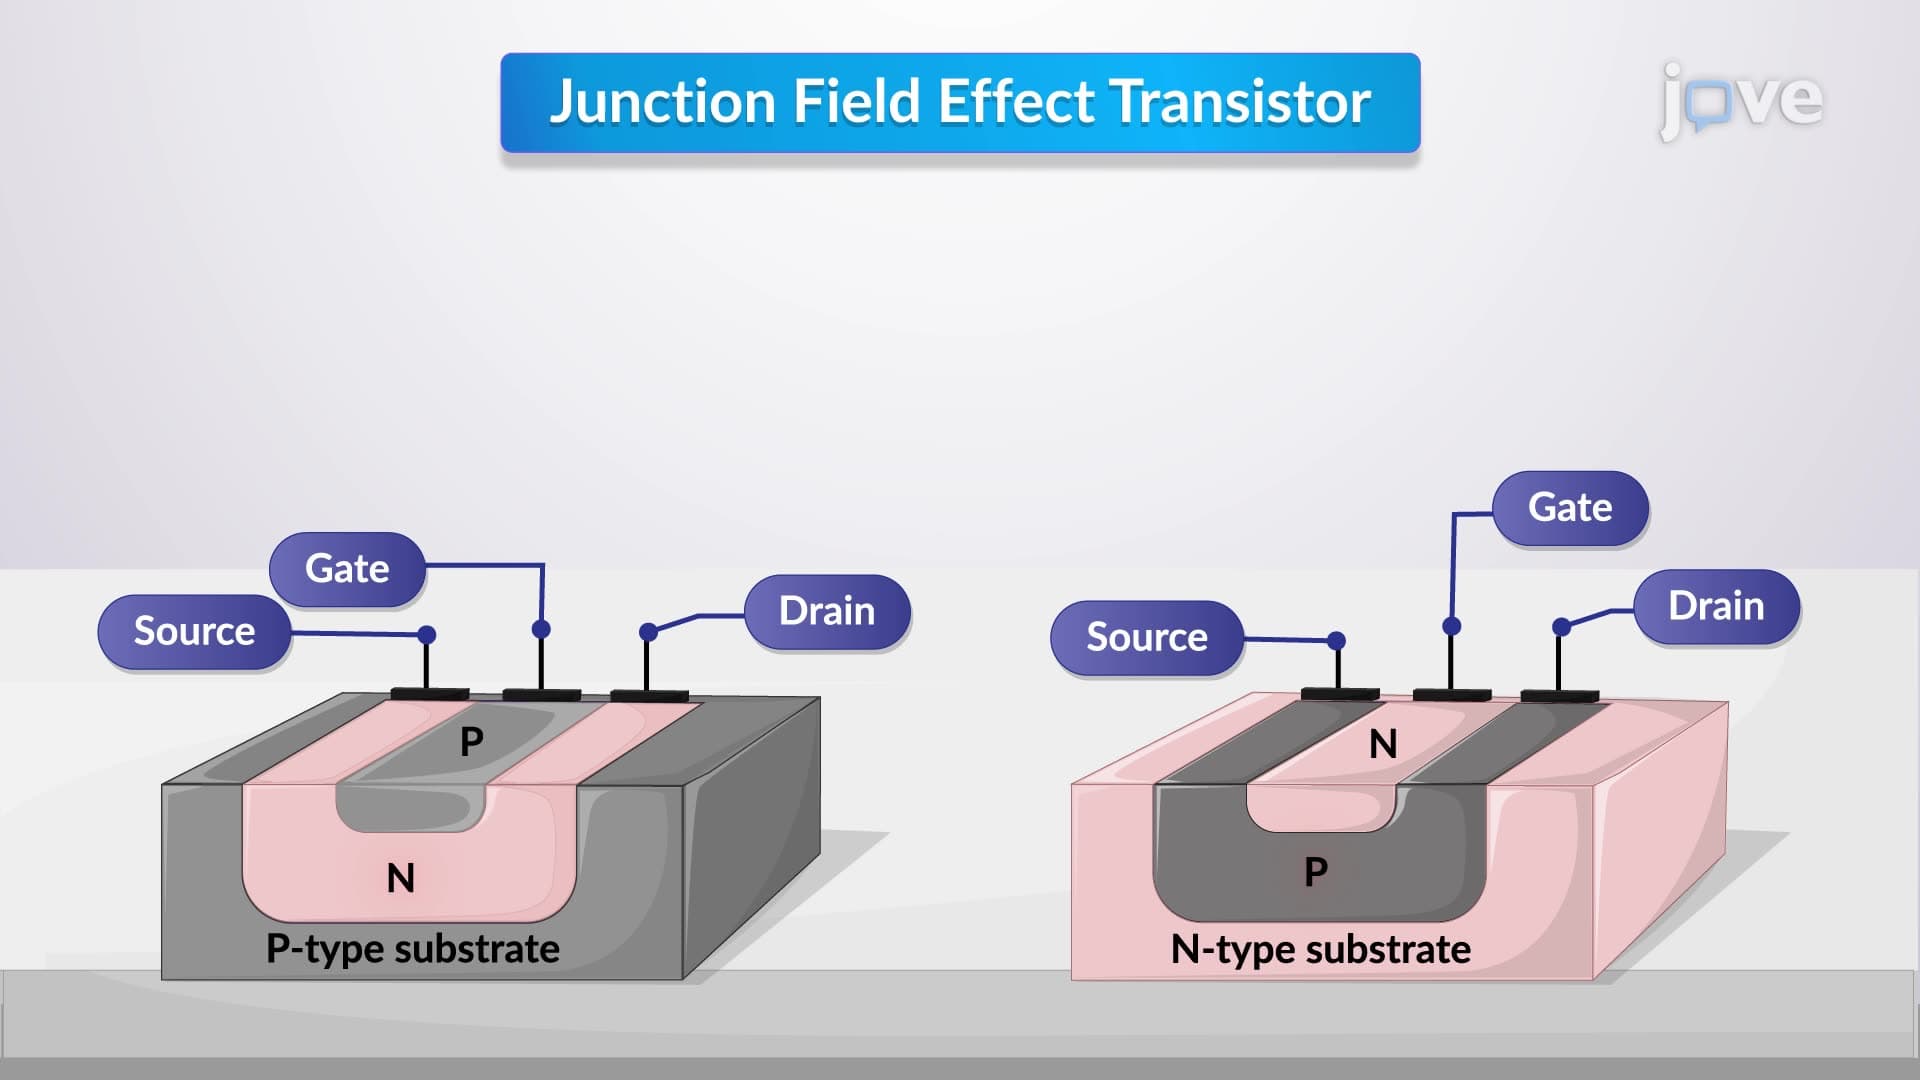
In an N-channel JFET, the structure consists of N-type material forming the channel on a P-type substrate, with the...