Related Experiment Video
Updated: Jan 1, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Dirac-Fock calculations on molecules in an adaptive multiwavelet basis
Joel Anderson1, Bryan Sundahl1, Robert Harrison1
1Institute for Advanced Computational Science, Stony Brook University, Stony Brook, New York 11794-5250, USA.

Abstract:
We report the first fully numerical approach for relativistic quantum chemical calculations applicable to molecules. The approach uses an adaptive basis of multiwavelet functions to solve the full four-component Dirac-Coulomb equation to a user-specified accuracy. The accuracy of the code is demonstrated by comparison with ground state energy calculations of atoms performed in GRASP, and the applicability to molecules is shown via ground state calculations of some simple molecules, including water analogs up to H2Po. In the case of molecules, comparison is made with Gaussian basis set calculations in DIRAC.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:29
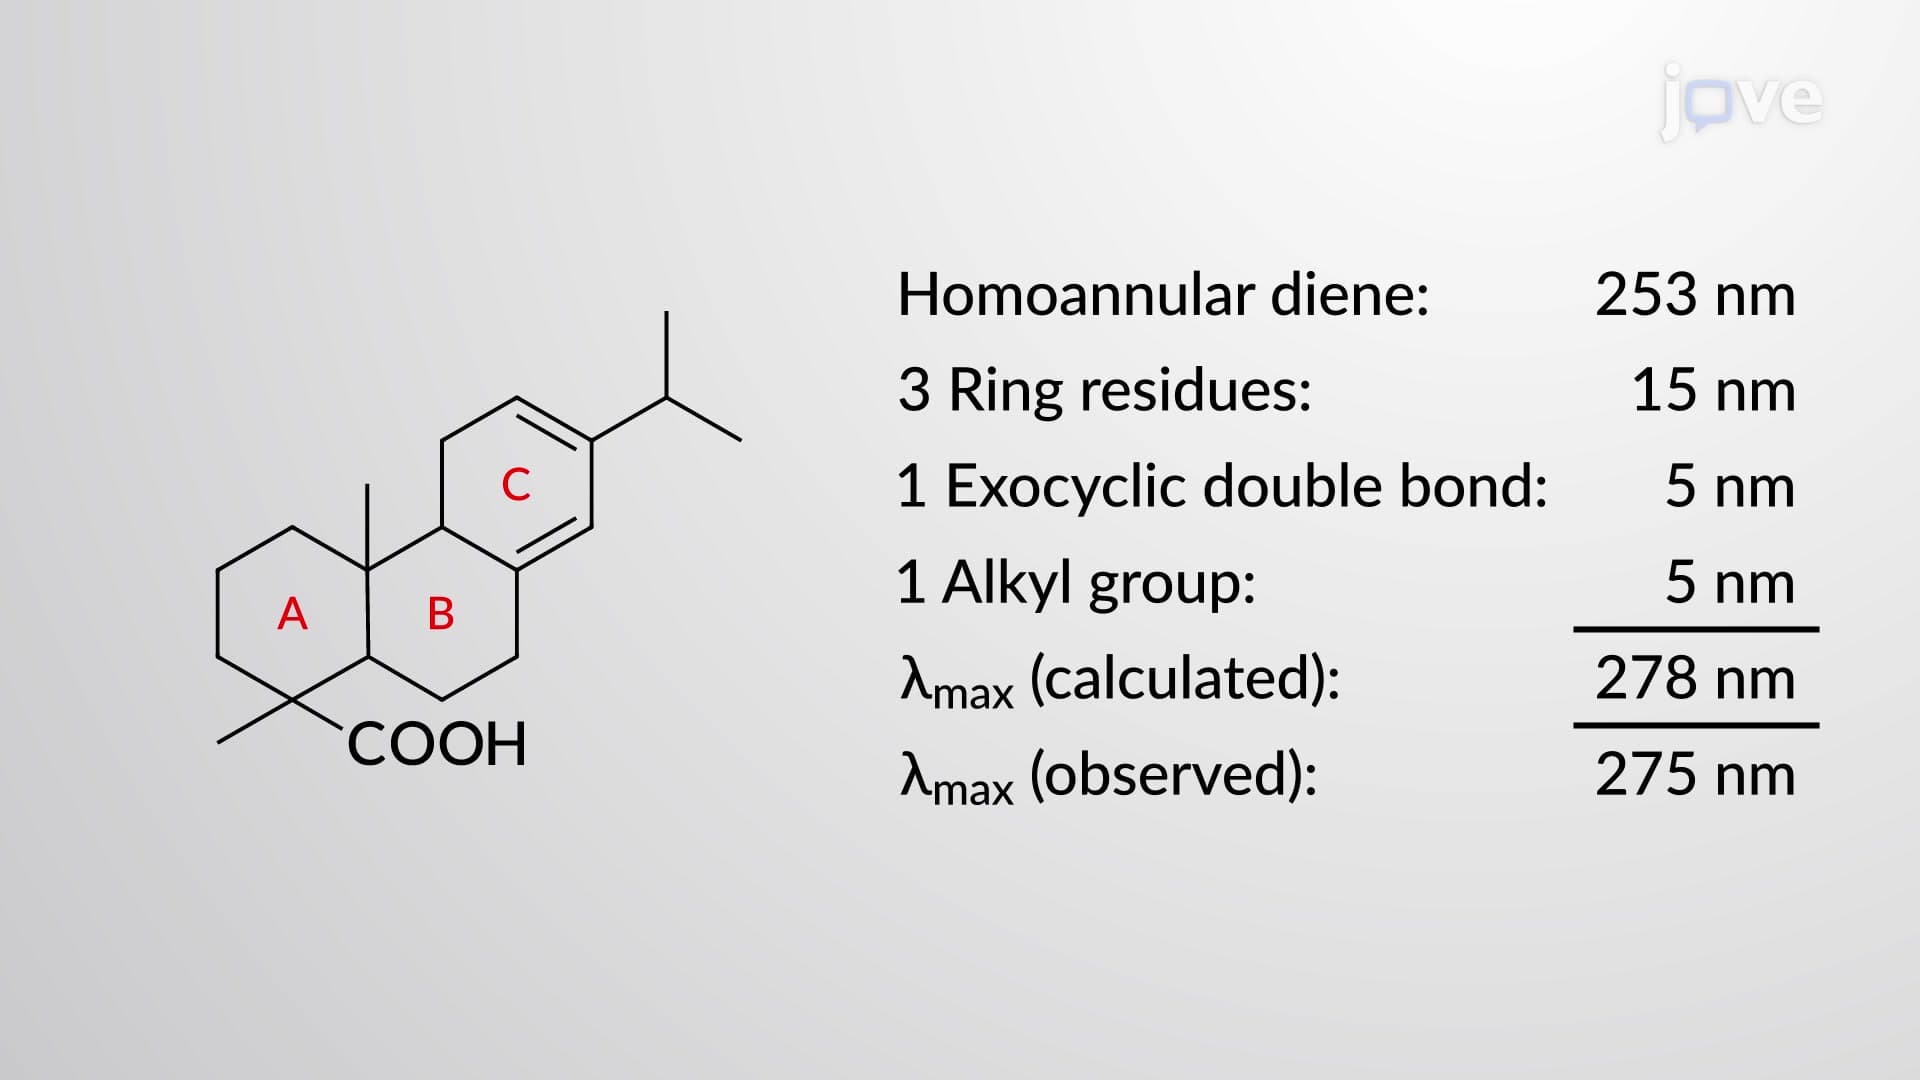
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 02:32
The de Broglie Wavelength