Related Experiment Video
Updated: Jan 1, 2026
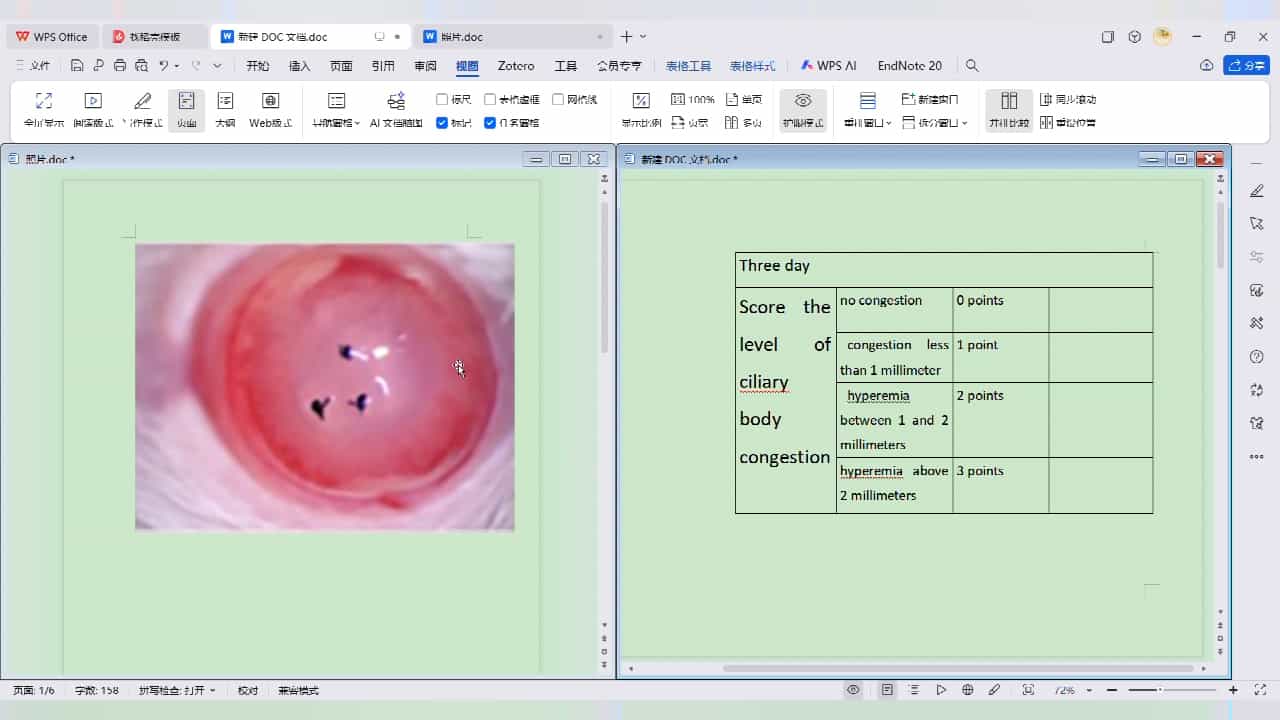

Establishing a Severe Corneal Inflammation Model in Rats Based on Corneal Epithelium Curettage Combined with Corneal Sutures
Published on: November 22, 2024
Autoinflammatory disease with corneal and mucosal dyskeratosis caused by a novel NLRP1 variant
Troels Herlin1, Sofie E Jørgensen2,3, Christian Høst1
1Department of Pediatrics.

Objectives:
Here we investigated a patient with inflammatory corneal intraepithelial dyskeratosis, mucosal inflammation, tooth abnormalities and, eczema to uncover the genetic and immunological basis of the disease.
Methods:
On suspicion of an autoinflammatory condition, Sanger sequencing of nucleotide-binding oligomerization domain-like, leucine-rich repeat pyrin domain containing 1 (NLRP1) was performed and combined with an in vitro inflammasome reconstitution assay to measure caspase-1-mediated IL-1β cleavage, stimulation of patient peripheral blood mononuclear cells (PBMCs) and whole blood to measure IL-1β, IL-18 production and quantification of apoptosis-associated speck-like protein containing CARD (ASC) speck formation as a measure of inflammasome activation by flow cytometry.
Results:
Sanger sequencing revealed a novel mutation (c.175G>C, p.A59P; NM_33004.4) in the inflammasome molecule NLRP1 segregating with disease, although with incomplete penetrance, in three generations. We found that patient PBMCs produced increased IL-1β in response to inflammatory stimuli, as well as increased constitutive levels of IL-18. Moreover, we demonstrate that expression of the identified NLRP1 A59P variant caused spontaneous IL-1β cleavage to mature IL-1β. In addition, patient PBMCs responded to NLRP1 stimulation with increased ASC speck formation as a reflection of elevated inflammasome activity.
Conclusion:
We demonstrate that this novel NLRP1 A59P variant caused increased activation of the NLRP1 inflammasome, resulting in constitutively and inducibly elevated IL-1β and IL-18 synthesis. We suggest the NLRP1 mutation underlies the pathogenesis of this rare autoinflammatory dyskeratotic disease inherited in an autosomal dominant manner with incomplete penetrance in the patient and within the family for several generations.