

Performance of the LRESC Model on top of DFT Functionals for Relativistic NMR Shielding Calculations
Juan I Melo1, Alejandro F Maldonado2, Gustavo A Aucar2
1Departamento de Física, Facultad de Ciencias Exactas y Naturales , Universidad de Buenos Aires and IFIBA CONICET , Buenos Aires 1428 , Argentina.

Abstract:
The linear response within the elimination of the small component model (LRESC) is an insightful and computationally efficient method for including relativistic effects on molecular properties like the nuclear magnetic shielding constants, spin-rotation constant, g-tensor, and electric field gradient of heavy atom containing molecules with atoms belonging up to the sixth row of the periodic table. One of its main advantages is its capacity to analyze the electronic origin of the different relativistic correcting terms. Until now, it was always applied on top of Hartree-Fock ground-state wave functions (LRESC/HF) to calculate and analyze NMR shieldings. In this work, we show the performance of the LRESC formalism on top of some density functional theory (DFT) functionals to compute tin shielding constants in SnX4 (X = H, F, Cl, Br, I) molecular systems. We analyze the performance of each LRESC/DFT scheme on reproducing the electronic mechanisms of the shieldings, taking as a benchmark the results of relativistic calculations at the RPA level of approach (4c/RPA). As in previous works, we divide the LRESC relativistic correcting terms into two groups: core-dependent and ligand-dependent contributions. It is shown here that core-dependent corrections are well-reproduced for the selected DFT functionals, but some differences arise in the ligand-dependent ones. We focus on the performance of different functionals, including the same electron correlation part but containing different amounts of HF exchange. The best results are obtained for the BHandHLYP functional (50% of HF exchange) and the worst for BLYP (0%). When the percentage of HF exchange increases, ligand-dependent contributions are better described, and the final LRESC/DFT results are closer to those obtained with LRESC/HF and 4c/RPA methods. The spin-orbit correction to the shielding constant is one of the main ligand-dependent contributions (there are two more) with total value depending on the amount of HF exchange included in the functional. When the amount of HF exchange decreases, the spin-orbit contribution becomes larger, overestimating the shielding constant even when nonrelativisitc DFT values are much smaller than the nonrelativistic HF ones, as it happens for the heaviest molecular system studied here (SnI4).
More Related Videos
 10:52
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
 11:19
Site Directed Spin Labeling and EPR Spectroscopic Studies of Pentameric Ligand-Gated Ion Channels
Published on: July 4, 2016
Related Concept Videos
 01:14
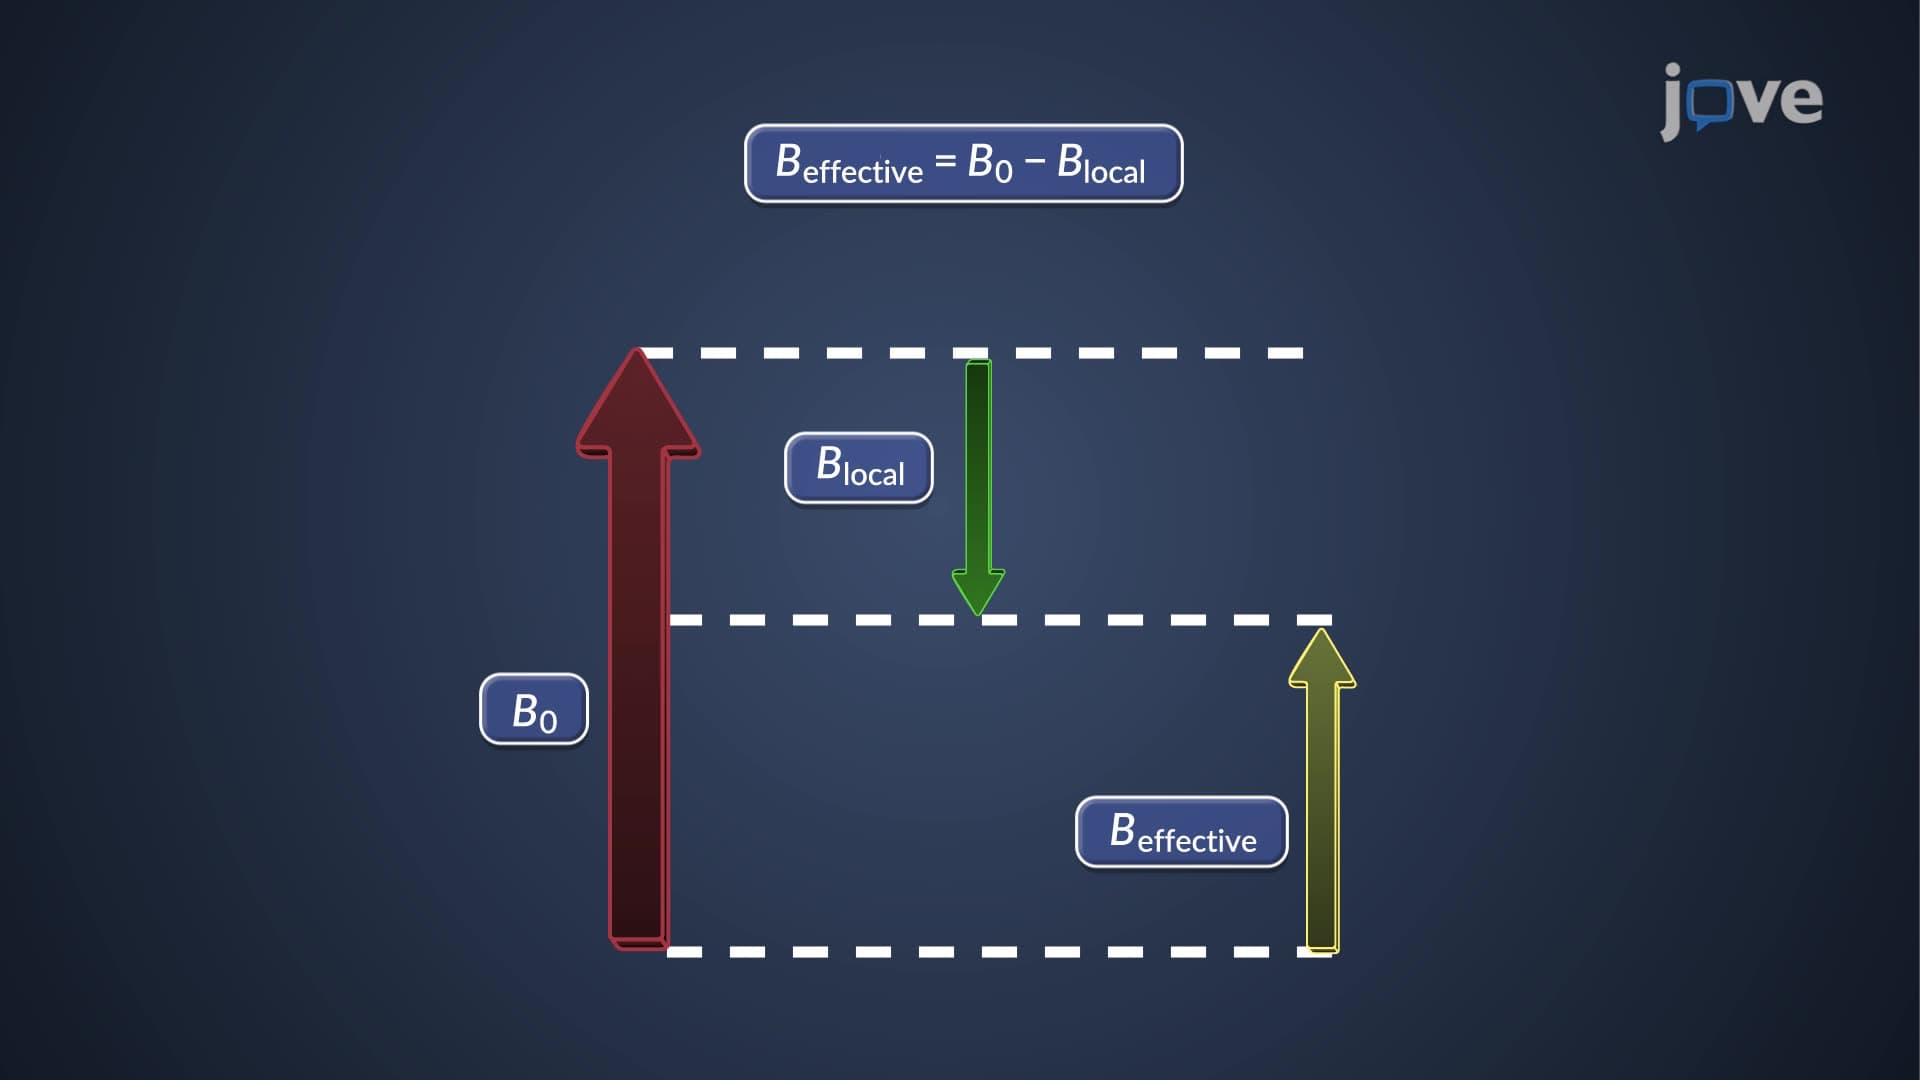
Diamagnetic Shielding of Nuclei: Local Diamagnetic Current
 03:21
The Energies of Atomic Orbitals
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:14
NMR Spectrometers: Resolution and Error Correction
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:27
π Electron Effects on Chemical Shift: Overview