Related Experiment Video
Updated: Dec 31, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A Novel Method for Calculation of Molecular Energies and Charge Distributions by Thermodynamic Formalization
TongIl Kim1, ChungIl Ri2, HakSung Yun2
1Institute of Chemistry and Biology, University of Science, Pyongyang, 950003, Democratic People's Republic of Korea. kimtil@163.com.

Abstract:
The paper describes a new approach to the thermodynamic formalization for calculation of molecular energy and charge distribution in ground state by means of the variational equation of DFT. In order to thermodynamically formalize the molecular calculation, the pseudo chemical potential (PCP) is conceptualized, where a molecule is broken into multi-phase(atom) one-component(electron) systems and the energy of system is represented as PCP. Calculation of the molecular energy and atomic charge by PCP is put forward, thereafter the approach is proved to be valid and its efficiency (accuracy and calculation speed) is verified.
More Related Videos


Related Concept Videos
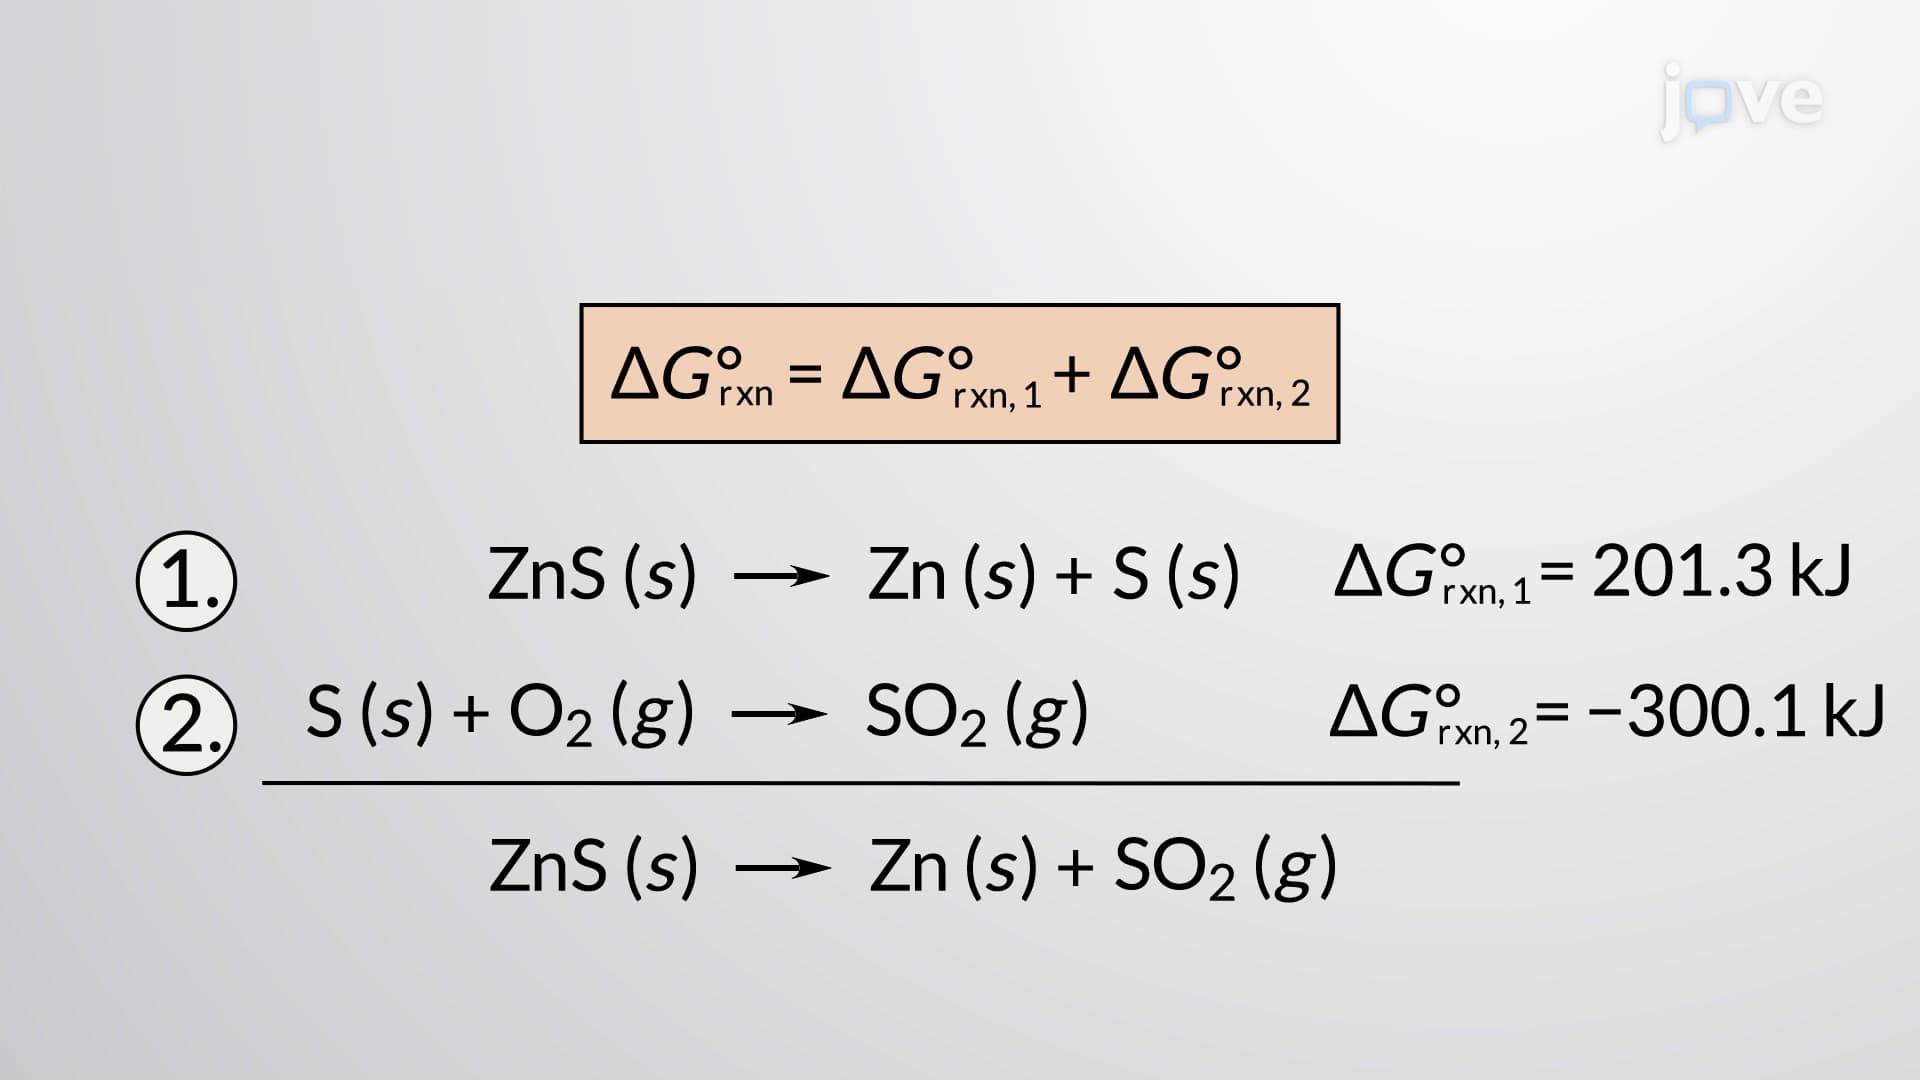 02:49
Calculating Standard Free Energy Changes
 01:26
Thermodynamic Potentials
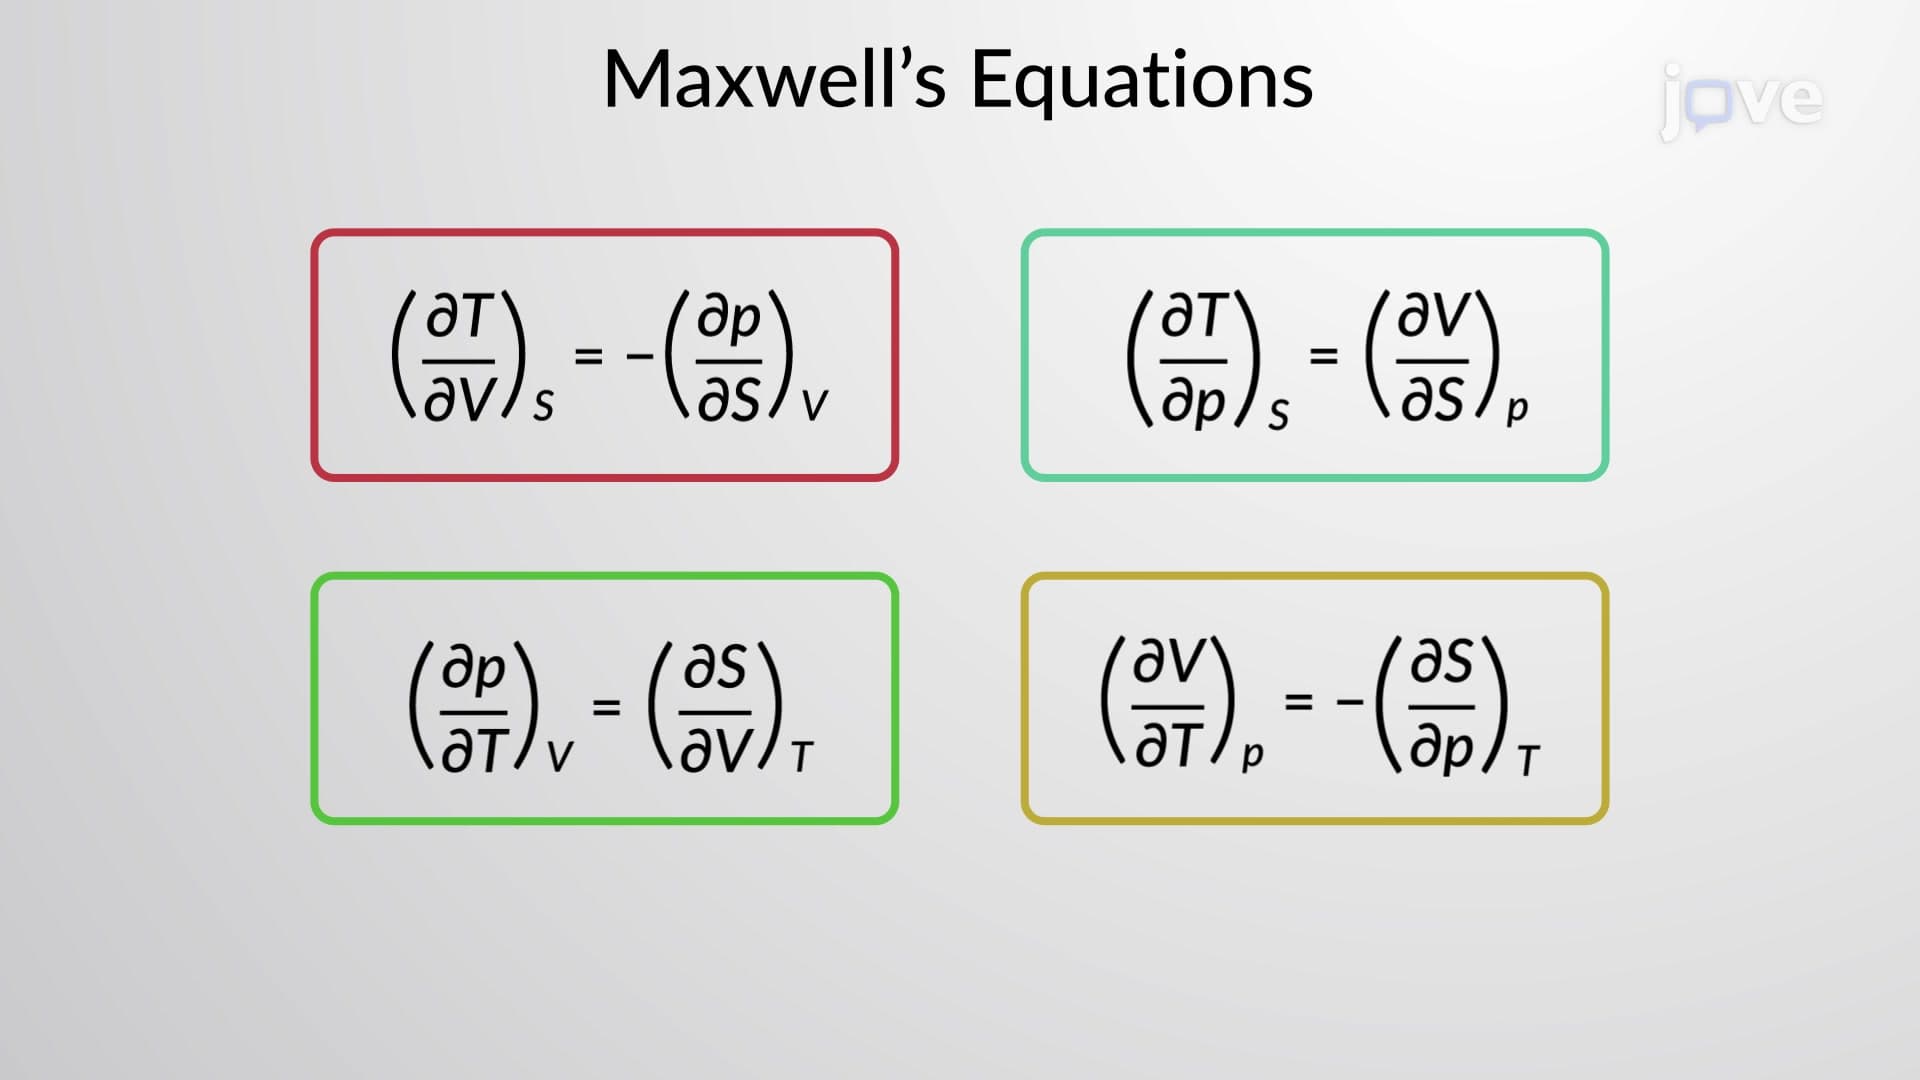 01:23
Maxwell's Thermodynamic Relations
All thermodynamic potentials are exact differentials. Therefore, their second-order...
 01:10
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 02:55
Thermochemical Equations
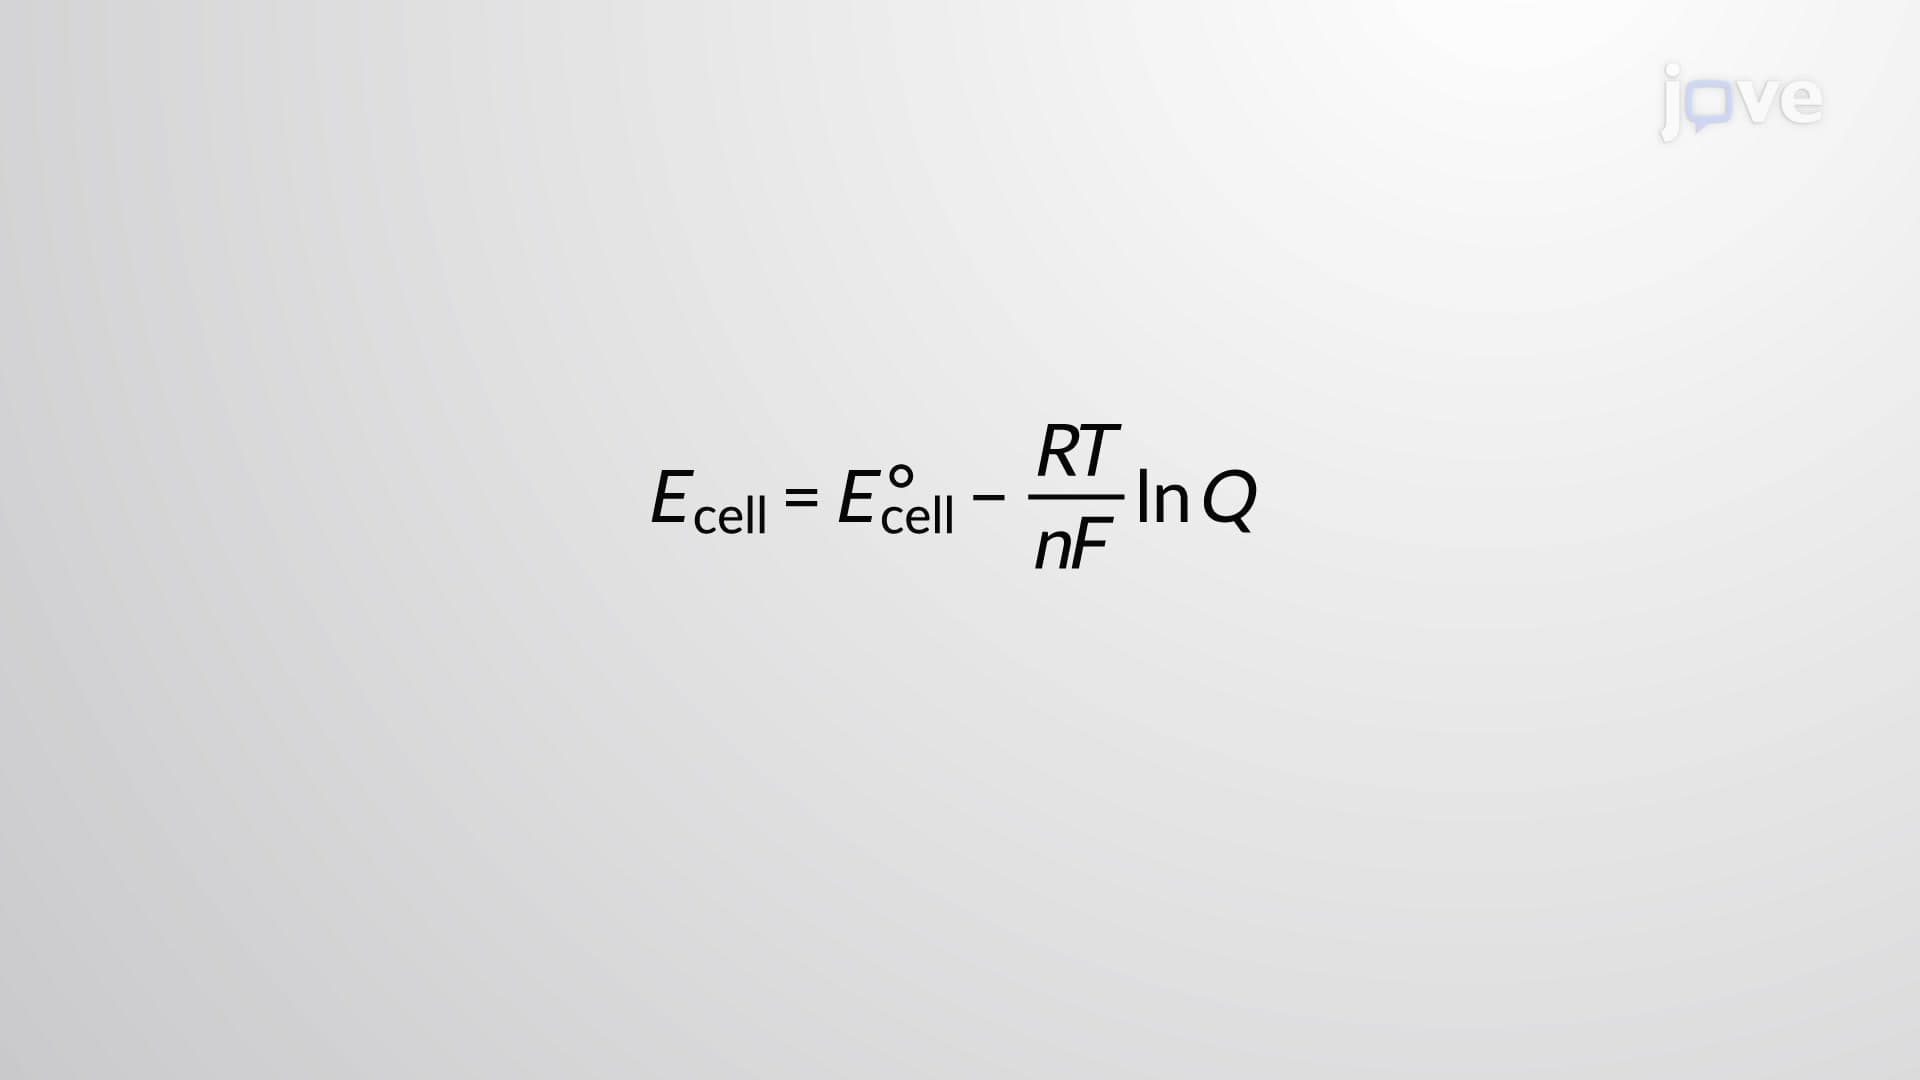 02:59
The Nernst Equation
The interconnection between standard cell potentials and various thermodynamic parameters such as the standard free energy change ΔG° and equilibrium constant K has been previously explored. For example, a redox reaction involving zinc(II) and tin(II) ions at 1 M concentration with Eºcell = +0.291 V and ΔG° = −56.2 kJ is spontaneous.