
Isothiourea-Catalyzed Atropselective Acylation of Biaryl Phenols via Sequential Desymmetrization/Kinetic Resolution
Elizabeth S Munday1, Markas A Grove2, Taisiia Feoktistova2
1EaStCHEM School of Chemistry, University of St Andrews, North Haugh, St Andrews, KY16 9ST, UK.

Abstract:
Axially chiral phenols are attractive targets in organic synthesis. This motif is central to many natural products and widely used as precursors to, or directly, as chiral ligands and catalysts. Despite their utility few simple catalytic methods are available for their synthesis in high enantiopurity. Herein the atropselective acylation of a range of symmetric biaryl diols is investigated using isothiourea catalysis. Studies on a model biaryl diol substrate shows that the high product er observed in the process is a result of two successive enantioselective reactions consisting of an initial enantioselective desymmetrization coupled with a second chiroablative kinetic resolution. Extension of this process to a range of substrates, including a challenging tetraorthosubstituted biaryl diol, led to highly enantioenriched products (14 examples, up to 98:2 er), with either HyperBTM or BTM identified as the optimal catalyst depending upon the substitution pattern within the substrate. Computation has been used to understand the factors that lead to high enantiocontrol in this process, with maintenance of planarity to maximize a 1,5-S⋅⋅⋅O interaction within the key acyl ammonium intermediate identified as the major feature that determines atropselective acylation and thus product enantioselectivity.
More Related Videos
Related Concept Videos
 02:27
Preparation of Alkynes: Alkylation Reaction
Alkylation of terminal alkynes with primary alkyl halides in the presence of a strong base like sodium amide is one of the common methods for the synthesis of longer carbon-chain alkynes. For example, treatment of 1-propyne with sodium amide followed by reaction with ethyl bromide yields 2-pentyne.
 01:30
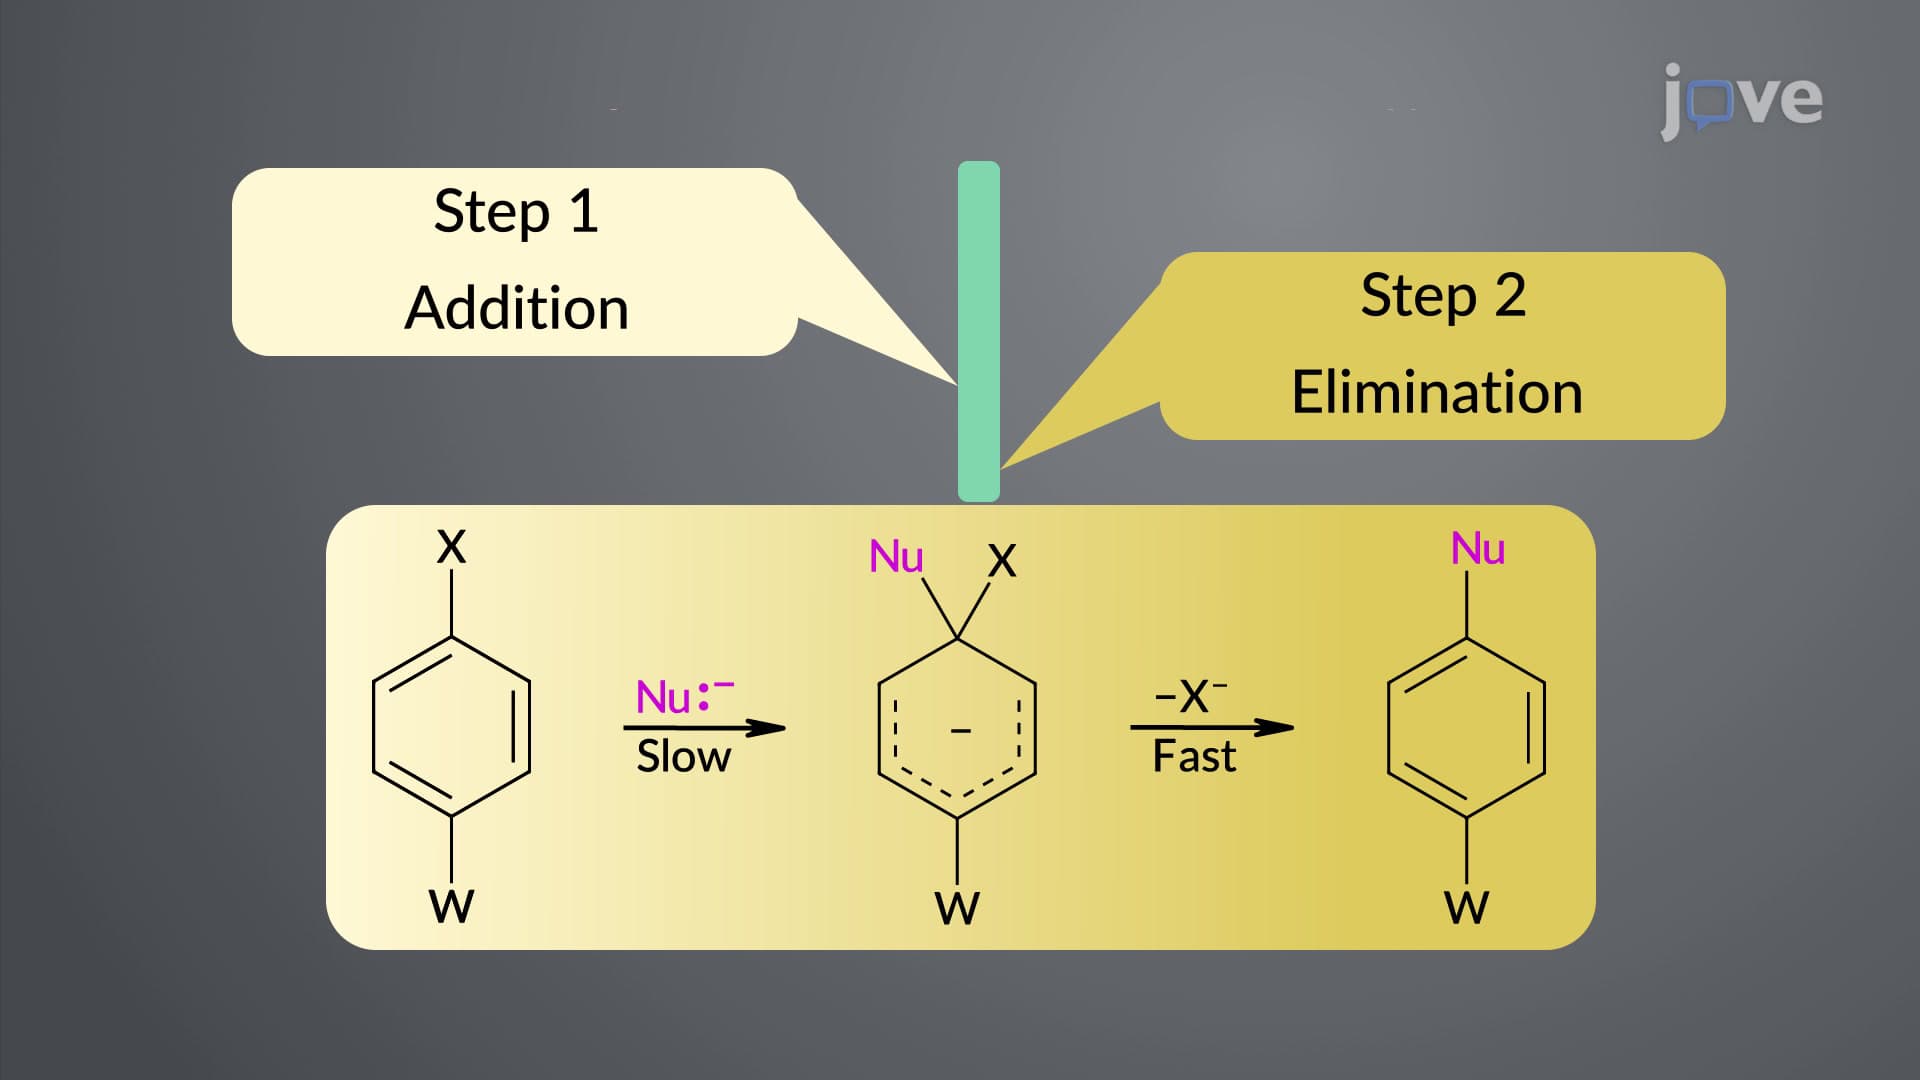
Nucleophilic Aromatic Substitution: Addition–Elimination (SNAr)
The reaction begins with an attack of the nucleophile on the carbon that holds the leaving group. This results in the delocalization of the π electrons over the ring carbons. The resonance interaction between...
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 01:11
Nucleophilic Aromatic Substitution: Elimination–Addition
 02:47
Alkynes to Aldehydes and Ketones: Hydroboration-Oxidation
One of the convenient methods for the preparation of aldehydes and ketones is via hydration of alkynes. Hydroboration-oxidation of alkynes is an indirect hydration reaction in which an alkyne is treated with borane followed by oxidation with alkaline peroxide to form an enol that rapidly converts into an aldehyde or a ketone. Terminal alkynes form aldehydes, whereas internal alkynes give ketones as the final product.
 01:13
Electrophilic Aromatic Substitution: Fluorination and Iodination of Benzene