Related Experiment Video
Updated: Dec 29, 2025

Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Multiple Free Energy Calculations from Single State Point Continuous Fractional Component Monte Carlo Simulation
Ahmadreza Rahbari1, Remco Hens1, Othonas A Moultos1
1Engineering Thermodynamics, Process & Energy Department, Faculty of Mechanical, Maritime and Materials Engineering, Delft University of Technology, Leeghwaterstraat 39, 2628 CB Delft, The Netherlands.

Abstract:
We introduce an alternative method to perform free energy calculations for mixtures at multiple temperatures and pressures from a single simulation, by combining umbrella sampling and the continuous fractional component Monte Carlo method. One can perform a simulation of a mixture at a certain pressure and temperature and accurately compute the chemical potential at other pressures and temperatures close to the simulation conditions. This method has the following advantages: (1) Accurate estimates of the chemical potential as a function of pressure and temperature are obtained from a single state simulation without additional postprocessing. This can potentially reduce the number of simulations of a system for free energy calculations for a specific temperature and/or pressure range. (2) Partial molar volumes and enthalpies are obtained directly from the estimated chemical potentials. We tested our method for a Lennard-Jones system, aqueous mixtures of methanol at T = 298 K and P = 1 bar, and a mixture of ammonia, nitrogen, and hydrogen at T = 573 K and P = 800 bar. For pure methanol (N = 410 molecules), we observed that the estimated chemical potentials from umbrella sampling are in excellent agreement with the reference values obtained from independent simulations, for ΔT = ±15 K and ΔP = 100 bar (with respect to the simulated system). For larger systems, this range becomes smaller because the relative fluctuations of energy and volume become smaller. Without sufficient overlap, the performance of the method will become poor especially for nonlinear variations of the chemical potential.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
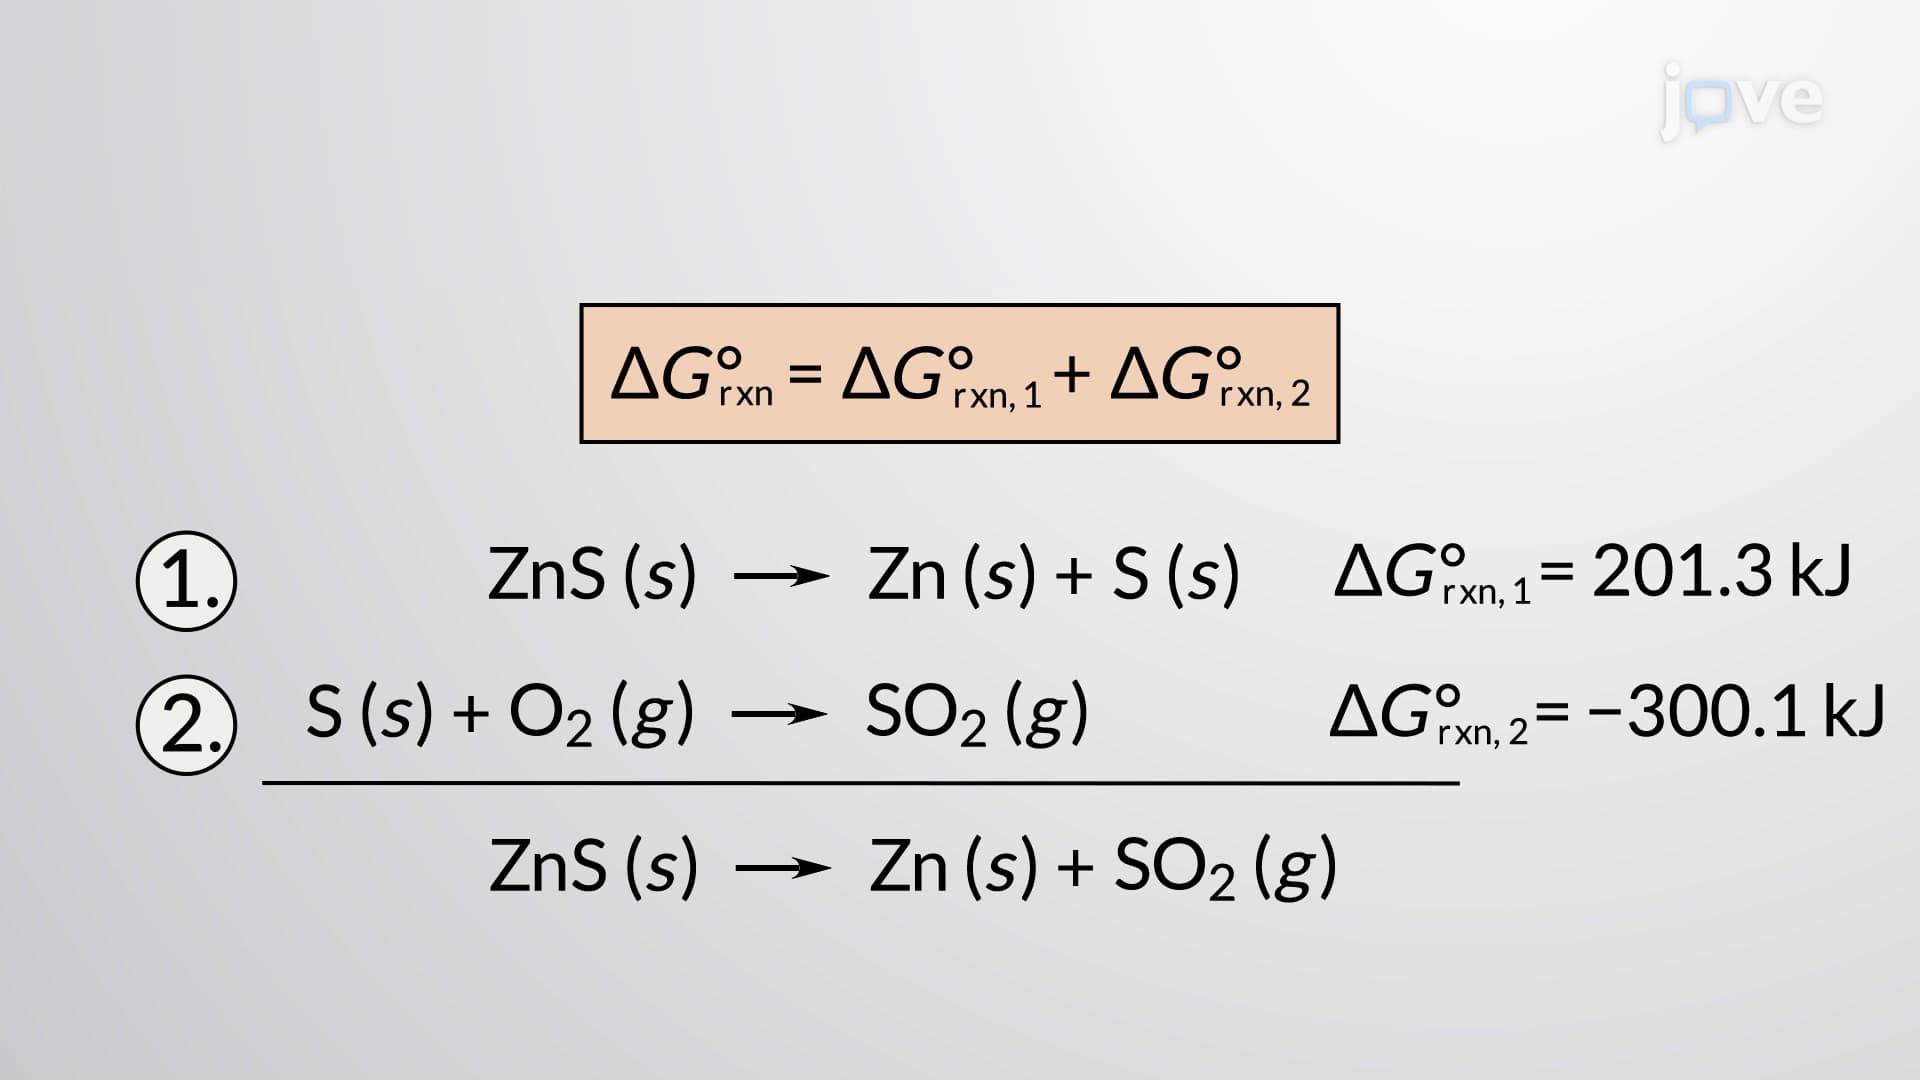 02:49
Calculating Standard Free Energy Changes
 01:20
Maxwell-Boltzmann Distribution: Problem Solving
This distribution function f(v) is defined by saying that the expected number N (v1,v2) of particles with speeds between v1 and v2 is given by
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
 01:16
Energy Conservation and Bernoulli's Equation
All the terms in the equation have the dimension of energy per unit volume. The kinetic energy per unit volume is called the kinetic energy density, and the potential energy per unit volume is...
 03:25
Free Energy Changes for Nonstandard States
 01:10
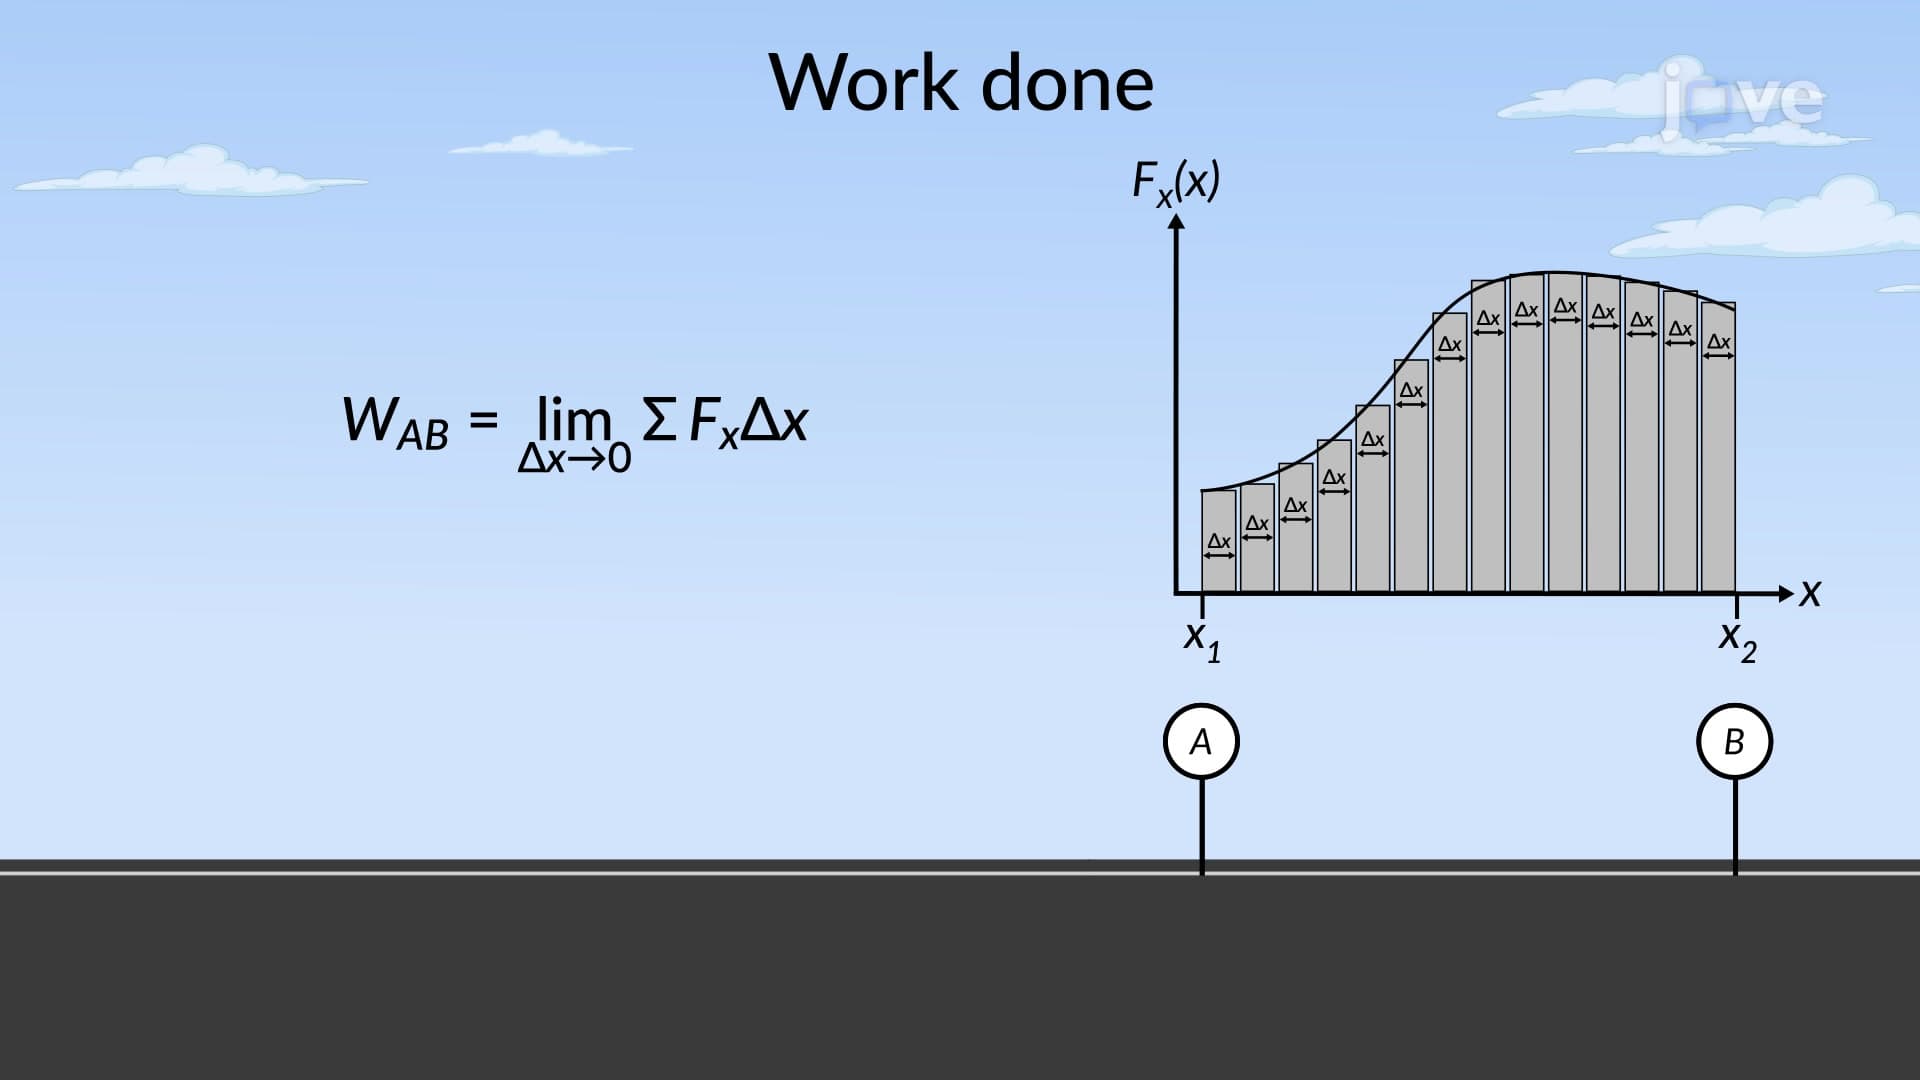
Work and Energy for Variable Forces