Related Experiment Video
Updated: Dec 29, 2025

Visualization of SARS-CoV-2 using Immuno RNA-Fluorescence In Situ Hybridization
Published on: December 23, 2020
Genomic characterisation and epidemiology of 2019 novel coronavirus: implications for virus origins and receptor
Roujian Lu1, Xiang Zhao1, Juan Li2
1NHC Key Laboratory of Biosafety, National Institute for Viral Disease Control and Prevention, Chinese Center for Disease Control and Prevention, Beijing, China.

Insights
A novel coronavirus (2019-nCoV) causing viral pneumonia in Wuhan, China, was identified. Genetic analysis reveals it
Area of Science:
- Virology
- Genomics
- Epidemiology
Background:
- First identified in Wuhan, China, in late December 2019, a novel coronavirus (2019-nCoV) caused viral pneumonia of unknown origin.
- Human-to-human transmission was confirmed, with over 2000 cases reported by January 26, 2020, primarily affecting individuals in Wuhan.
Purpose of the Study:
- To sequence and analyze the genomes of the novel coronavirus (2019-nCoV) from infected patients.
- To determine the evolutionary origins and receptor-binding properties of 2019-nCoV.
Main Methods:
- Next-generation sequencing and Sanger sequencing were used to obtain complete and partial 2019-nCoV genomes from nine patients.
- Phylogenetic analysis and homology modeling were employed to understand the virus's evolutionary history and potential receptor interactions.
Main Results:
- Ten highly similar 2019-nCoV genome sequences (>99.98% identity) were obtained.
- 2019-nCoV showed 88% identity to bat-derived SARS-like coronaviruses, distinct from SARS-CoV and MERS-CoV.
- Homology modeling indicated a similar receptor-binding domain structure to SARS-CoV, suggesting potential binding to human ACE2 receptors.
Conclusions:
- 2019-nCoV is a distinct human-infecting betacoronavirus, divergent from SARS-CoV.
- Bats are the likely original host, with a potential intermediate animal host in the Wuhan seafood market.
- Urgent investigation into the virus's future evolution, adaptation, and spread is warranted.
Background:
In late December, 2019, patients presenting with viral pneumonia due to an unidentified microbial agent were reported in Wuhan, China. A novel coronavirus was subsequently identified as the causative pathogen, provisionally named 2019 novel coronavirus (2019-nCoV). As of Jan 26, 2020, more than 2000 cases of 2019-nCoV infection have been confirmed, most of which involved people living in or visiting Wuhan, and human-to-human transmission has been confirmed.
Methods:
We did next-generation sequencing of samples from bronchoalveolar lavage fluid and cultured isolates from nine inpatients, eight of whom had visited the Huanan seafood market in Wuhan. Complete and partial 2019-nCoV genome sequences were obtained from these individuals. Viral contigs were connected using Sanger sequencing to obtain the full-length genomes, with the terminal regions determined by rapid amplification of cDNA ends. Phylogenetic analysis of these 2019-nCoV genomes and those of other coronaviruses was used to determine the evolutionary history of the virus and help infer its likely origin. Homology modelling was done to explore the likely receptor-binding properties of the virus.
Findings:
The ten genome sequences of 2019-nCoV obtained from the nine patients were extremely similar, exhibiting more than 99·98% sequence identity. Notably, 2019-nCoV was closely related (with 88% identity) to two bat-derived severe acute respiratory syndrome (SARS)-like coronaviruses, bat-SL-CoVZC45 and bat-SL-CoVZXC21, collected in 2018 in Zhoushan, eastern China, but were more distant from SARS-CoV (about 79%) and MERS-CoV (about 50%). Phylogenetic analysis revealed that 2019-nCoV fell within the subgenus Sarbecovirus of the genus Betacoronavirus, with a relatively long branch length to its closest relatives bat-SL-CoVZC45 and bat-SL-CoVZXC21, and was genetically distinct from SARS-CoV. Notably, homology modelling revealed that 2019-nCoV had a similar receptor-binding domain structure to that of SARS-CoV, despite amino acid variation at some key residues.
Interpretation:
2019-nCoV is sufficiently divergent from SARS-CoV to be considered a new human-infecting betacoronavirus. Although our phylogenetic analysis suggests that bats might be the original host of this virus, an animal sold at the seafood market in Wuhan might represent an intermediate host facilitating the emergence of the virus in humans. Importantly, structural analysis suggests that 2019-nCoV might be able to bind to the angiotensin-converting enzyme 2 receptor in humans. The future evolution, adaptation, and spread of this virus warrant urgent investigation.
Funding:
National Key Research and Development Program of China, National Major Project for Control and Prevention of Infectious Disease in China, Chinese Academy of Sciences, Shandong First Medical University.
Related Concept Videos
 01:05
Single Nucleotide Polymorphisms-SNPs
 02:50
Conjugated Proteins
Nucleoproteins are protein complexes that contain nucleic acids, categorized as deoxyribonucleoproteins (DNPs) or ribonucleoproteins (RNPs) respectively. The nucleosome is a typical example of a DNP where nuclear DNA is associated with histone proteins. The major antigen for the Covid-19 virus SARS-CoV is an RNP that is critical...
 00:36
Viral Mutations
 02:28
Leaky Scanning
 01:15
G Protein-coupled Receptors
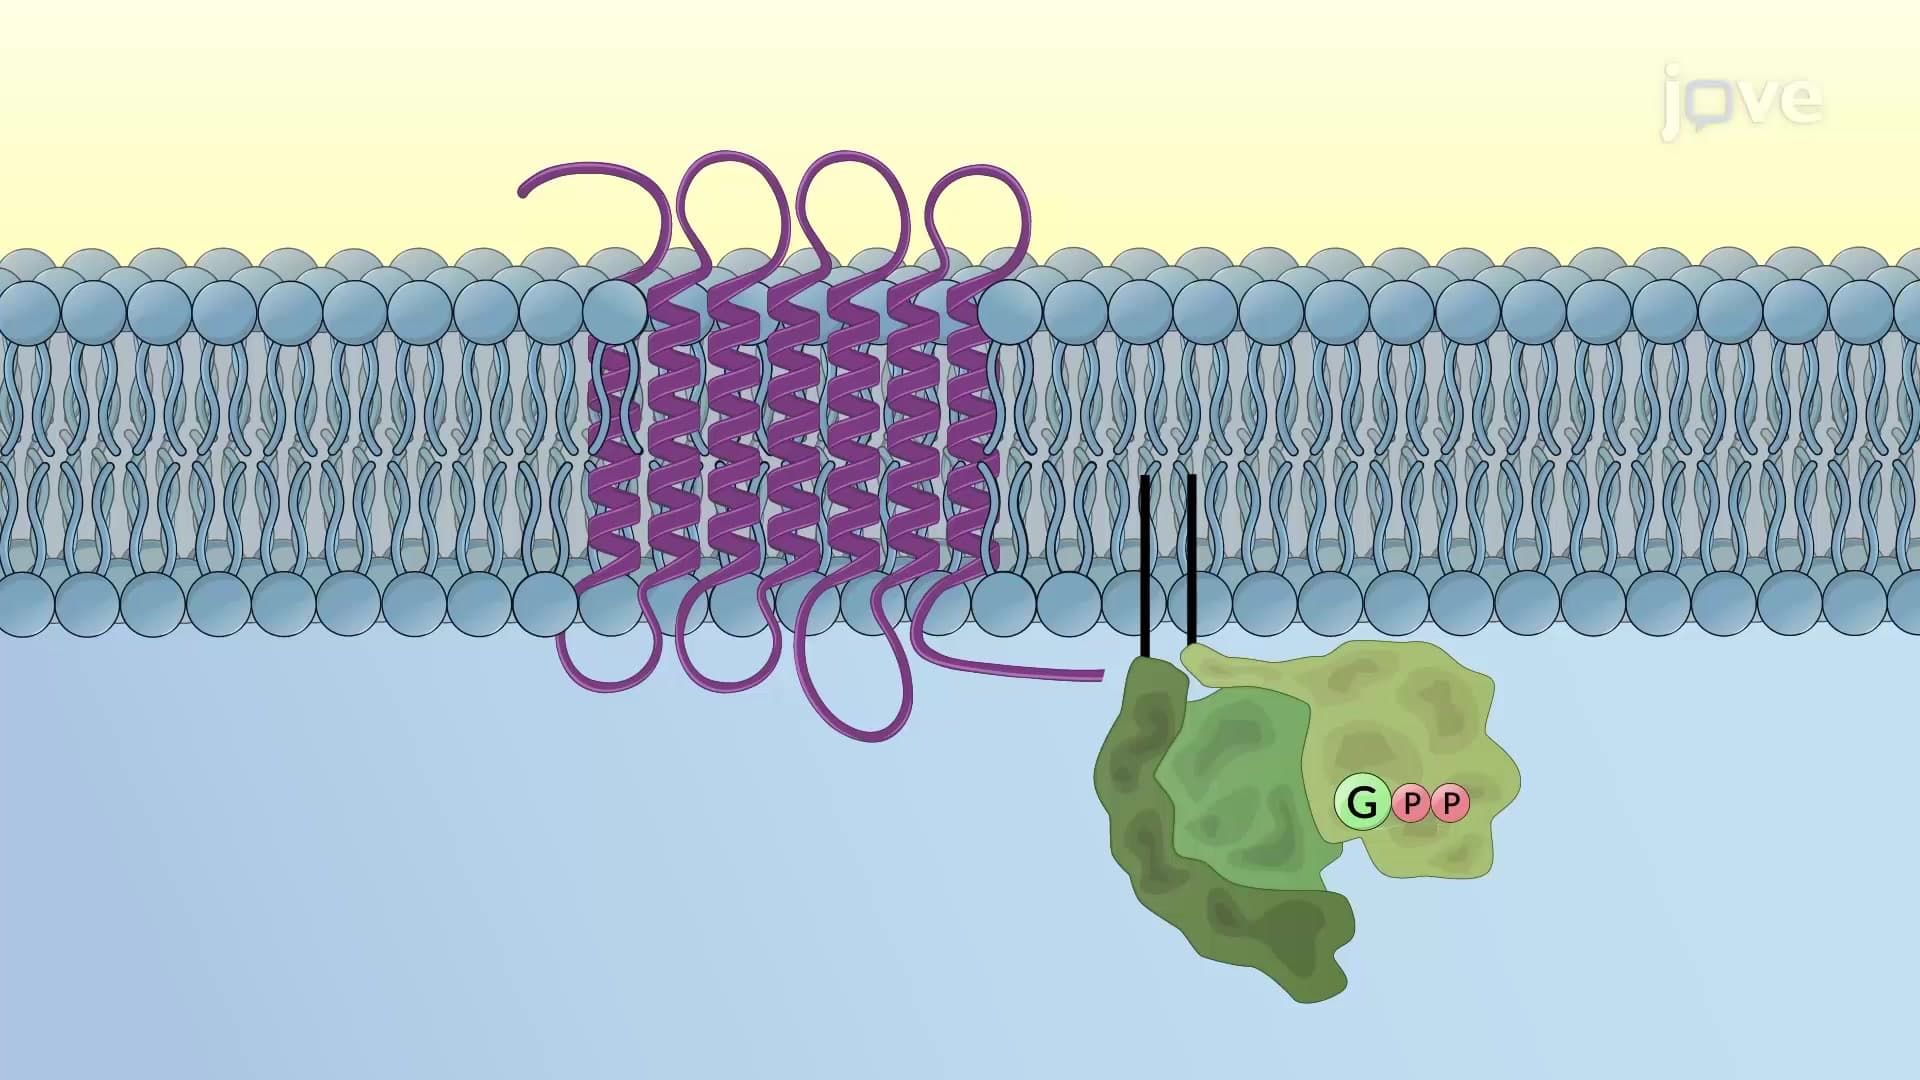
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:29
Viruses with RNA Genomes