Related Experiment Video
Updated: Dec 29, 2025


Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Ab Initio Molecular Dynamics Investigation of Beryllium Complexes
Onyekachi Raymond1,2,3, Michael Bühl3, Joseph R Lane1
1Chemistry, School of Science , University of Waikato , Private Bag 3105 , Hamilton 3240 , New Zealand.

Abstract:
Structures of aqueous [Be(H2O)4]2+, its outer-sphere and inner-sphere complexes with F-, Cl-, and SO42-, and dinuclear complexes with a [Be2(κ-OH)(κ-SO4)]+ core have been studied through Car-Parrinello molecular dynamics (CPMD) simulations with the BLYP functional. According to constrained CPMD/BLYP simulations and pointwise thermodynamic integration, the free energy of deprotonation of [Be(H2O)4]2+ and its binding free energy with F- are 9.6 and -6.2 kcal/mol, respectively, in good accord with available experimental data. The computed activation barriers for replacing a water ligand in [Be(H2O)4]2+ with F- and SO42-, 10.9 and 13.6 kcal/mol, respectively, are also in good qualitative agreement with available experimental data. These ligand-substitution reactions are indicated to follow associative interchange mechanisms with backside (SN2-like) attack of the anion relative to the aquo ligand it is displacing. Outperforming static density functional theory computations of the salient kinetic and thermodynamic quantities involving simple polarizable continuum solvent models, CPMD simulations are validated as a promising tool for studying the structures and speciation of beryllium complexes in aqueous solution.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 03:51
Molecular Orbital Theory II
 02:42
Valence Bond Theory
 01:18
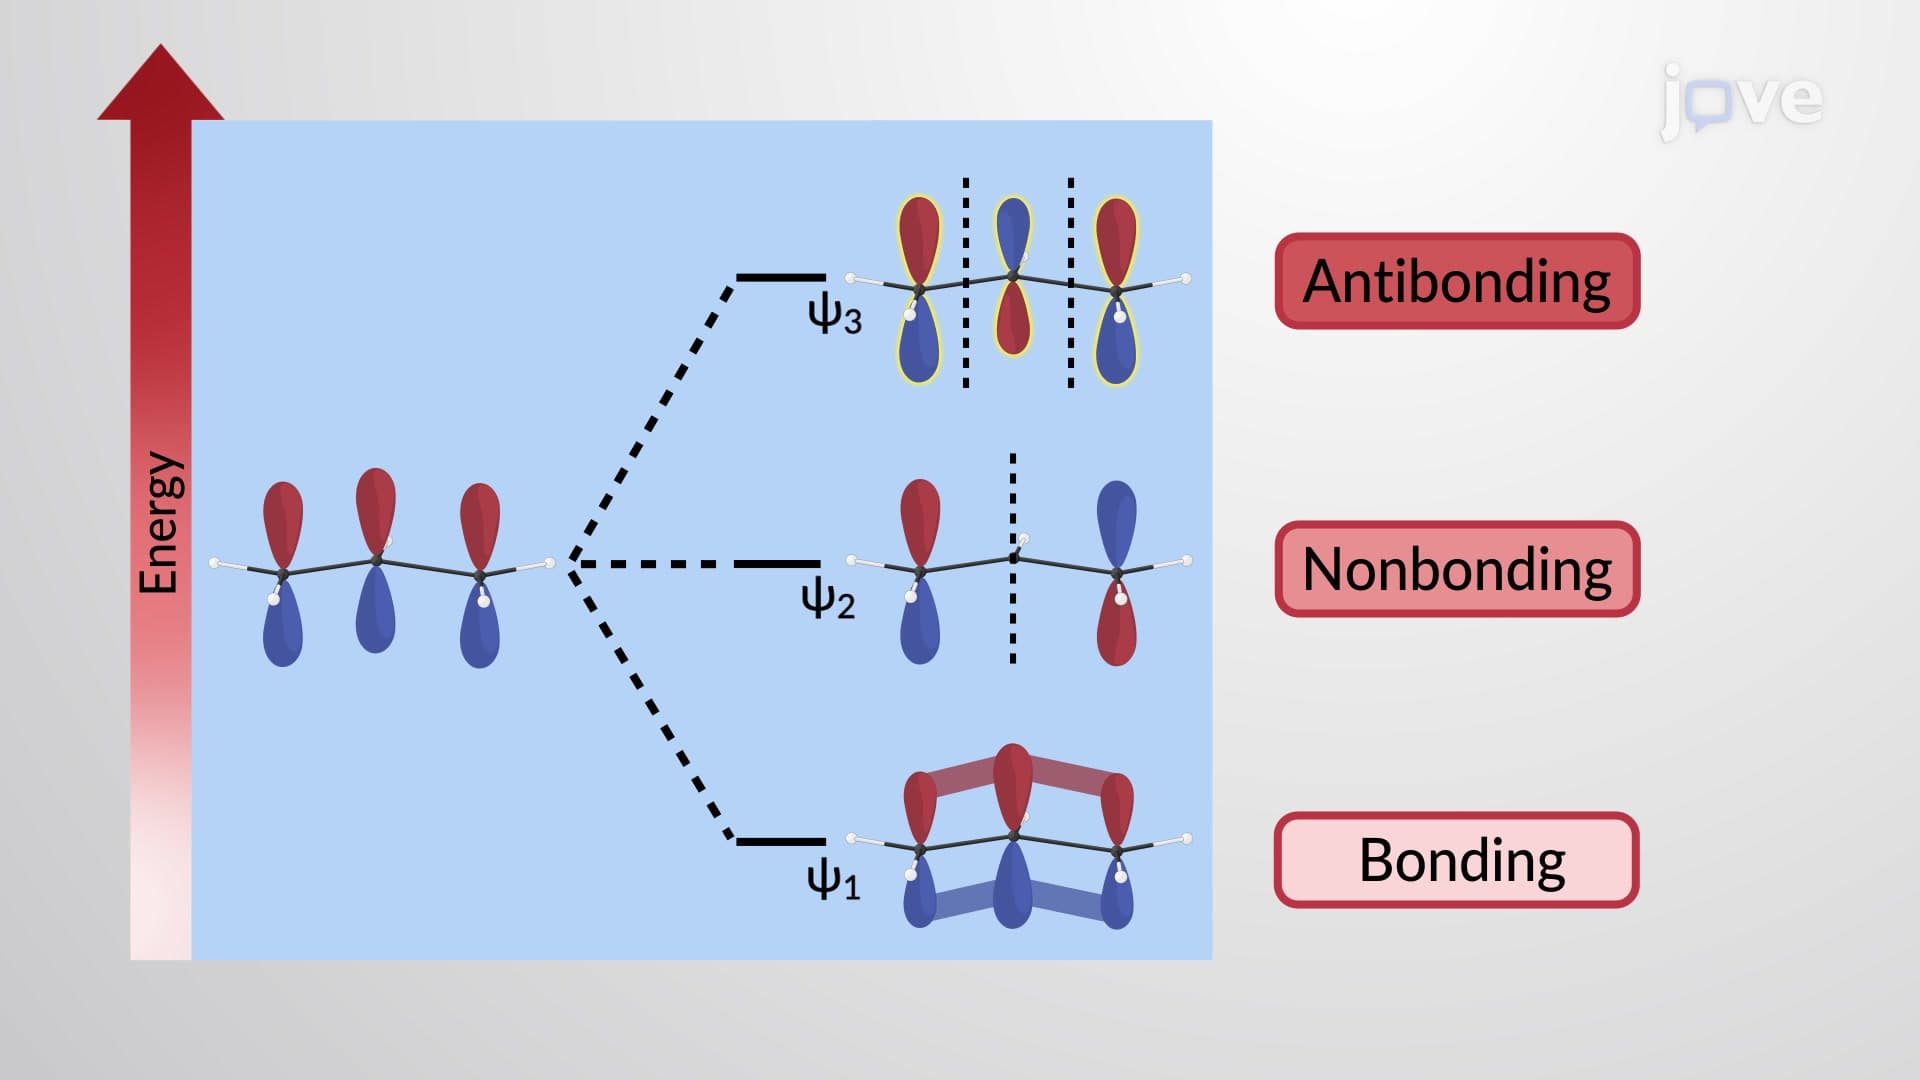
π Molecular Orbitals of the Allyl Cation and Anion
 02:40
MO Theory and Covalent Bonding