Related Experiment Video
Updated: Dec 29, 2025
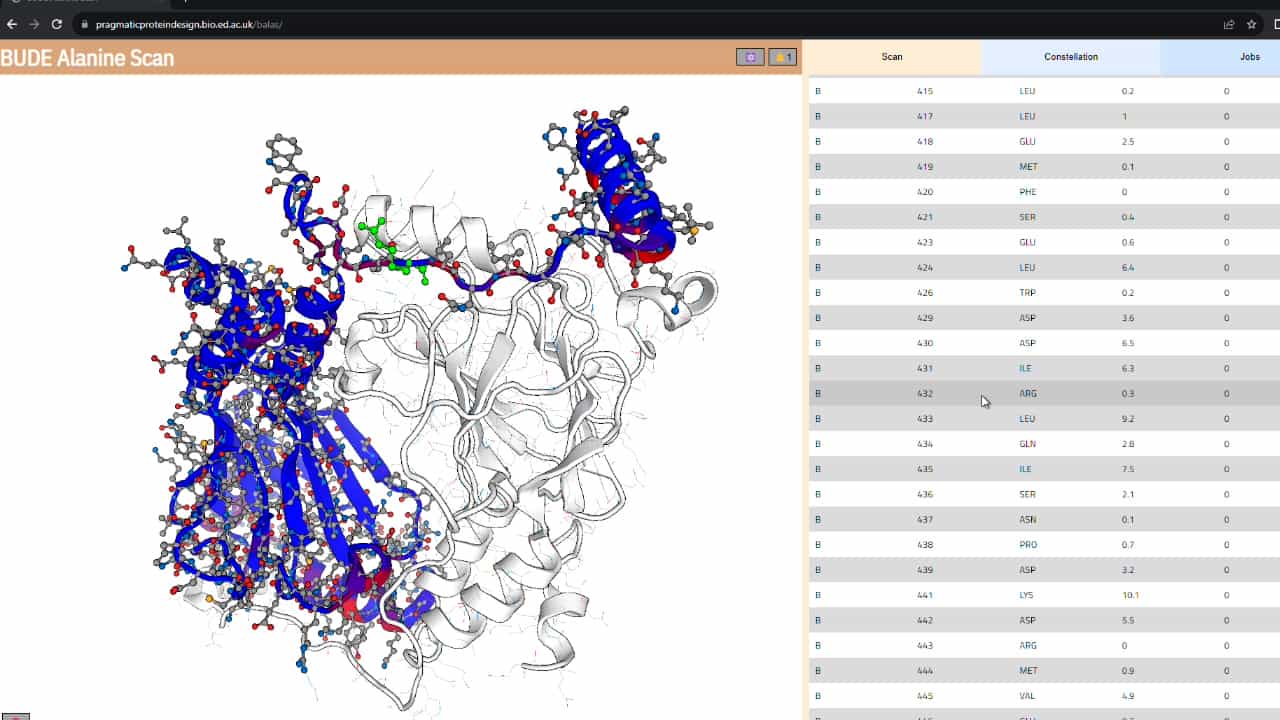
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Computational prediction of active sites and ligands in different AHL quorum quenching lactonases and acylases
Zulkar Nain1, Utpal Kumar Adhikari, Faruq Abdulla
1Department of Biotechnology and Genetic Engineering, Faculty of Biological Sciences, Islamic University, Kushtia 7003, Bangladesh.

Abstract:
With the emergence of multidrug-resistant 'superbug', conventional treatments become obsolete. Quorum quenching (QQ), enzyme-dependent alteration of quorum sensing (QS), is now considered as a promising antimicrobial therapy because of its potentiality to impede virulence gene expression without resulting in growth inhibition and antibiotic resistance. In our study, we intended to compare between two major QQ enzyme groups (i.e., AHL lactonases and AHL acylases) in terms of their structural and functional aspects. The amino acid composition-based principal component analysis (PCA) suggested that probably there is no structural and functional overlapping between the two groups of enzymes as well as within the lactonase enzymes but the acylases may functionally be affected by one another. In subcellular localization analysis, we also found that most lactonases are cytoplasmic while acylases are periplasmic. Investigation on the secondary structural features showed random coil dominates over alpha-helix and beta-sheet in all evaluated enzymes. For structural comparison, the tertiary structures of the selected proteins were modelled and submitted to the PMDB database (Accession ID: PM0081007 to PM0081018). Interestingly, sequence alignment revealed the presence of several conserved domains important for functions in both protein groups. In addition, three amino acid residues, namely aspartic acid, histidine, and isoleucine, were common in the active sites of all protein models while most frequent ligands were found to be 3C7, FEO, and PAC. Importantly, binding interactions of predicted ligands were similar to that of native QS signal molecules. Furthermore, hydrogen bonds analysis suggested six proteins are more stable than others. We believe that the knowledge of this comparative study could be useful for further research in the development of QSbased universal antibacterial strategies.
Related Concept Videos
 00:49
Ligand Binding and Linkage
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
 01:58
Cooperative Allosteric Transitions