Related Experiment Video
Updated: Dec 28, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Toward a less costly but accurate calculation of the CCSD(T)/CBS noncovalent interaction energy
Jiu-Li Chen1, Tao Sun1, Yi-Bo Wang1
1Department of Chemistry, and Key Laboratory of Guizhou High Performance Computational Chemistry, Guizhou University, Guiyang, China.

Abstract:
The popular method of calculating the noncovalent interaction energies at the coupled-cluster single-, double-, and perturbative triple-excitations [CCSD(T)] theory level in the complete basis set (CBS) limit was to add a CCSD(T) correction term to the CBS second-order Møller-Plesset perturbation theory (MP2). The CCSD(T) correction term is the difference between the CCSD(T) and MP2 interaction energies evaluated in a medium basis set. However, the CCSD(T) calculations with the medium basis sets are still very expensive for systems with more than 30 atoms. Comparatively, the domain-based local pair natural orbital coupled-cluster method [DLPNO-CCSD(T)] can be applied to large systems with over 1,000 atoms. Considering both the computational accuracy and efficiency, in this work, we propose a new scheme to calculate the CCSD(T)/CBS interaction energies. In this scheme, the MP2/CBS term keeps intact and the CCSD(T) correction term is replaced by a DLPNO-CCSD(T) correction term which is the difference between the DLPNO-CCSD(T) and DLPNO-MP2 interaction energies evaluated in a medium basis set. The interaction energies of the noncovalent systems in the S22, HSG, HBC6, NBC10, and S66 databases were recalculated employing this new scheme. The consistent and tight settings of the truncation parameters for DLPNO-CCSD(T) and DLPNO-MP2 in this noncanonical CCSD(T)/CBS calculations lead to the maximum absolute deviation and root-mean-square deviation from the canonical CCSD(T)/CBS interaction energies of less than or equal to 0.28 kcal/mol and 0.09 kcal/mol, respectively. The high accuracy and low cost of this new computational scheme make it an excellent candidate for the study of large noncovalent systems.

Related Concept Videos
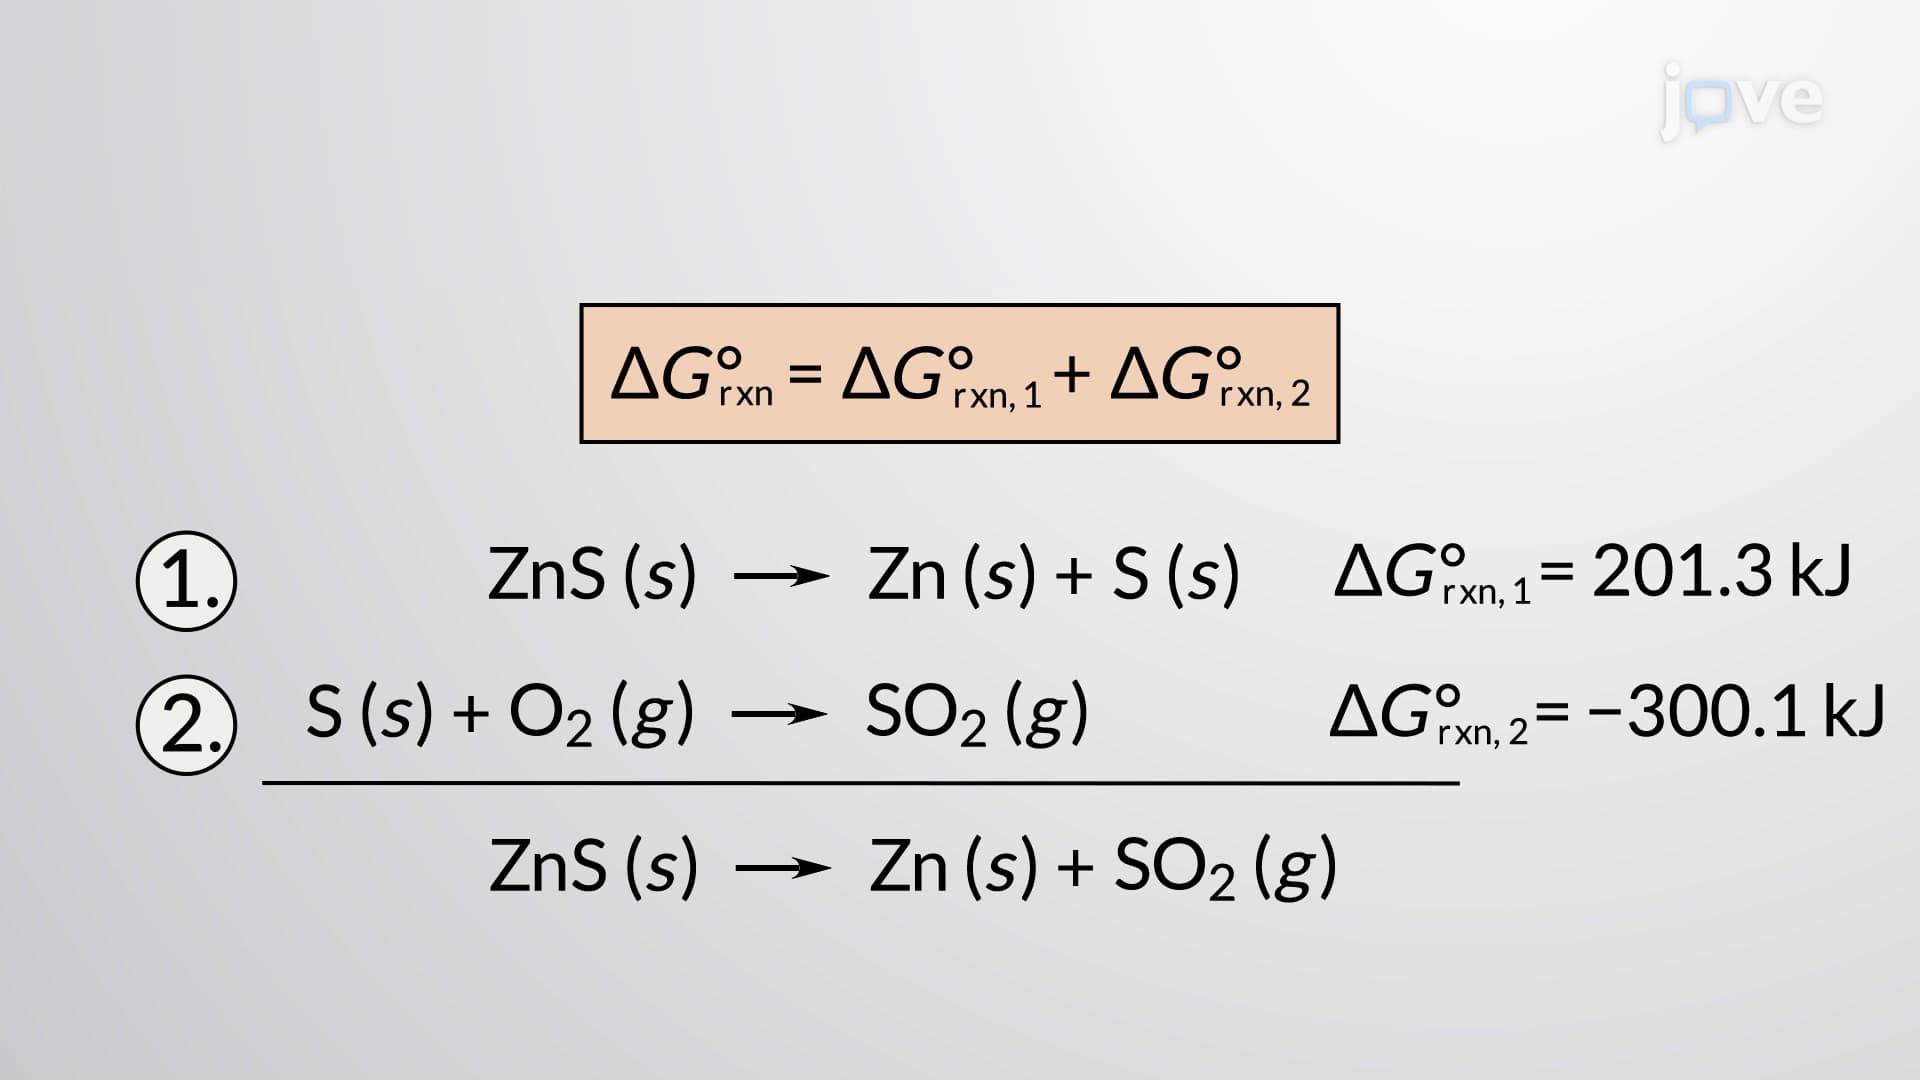 02:49
Calculating Standard Free Energy Changes
 01:24
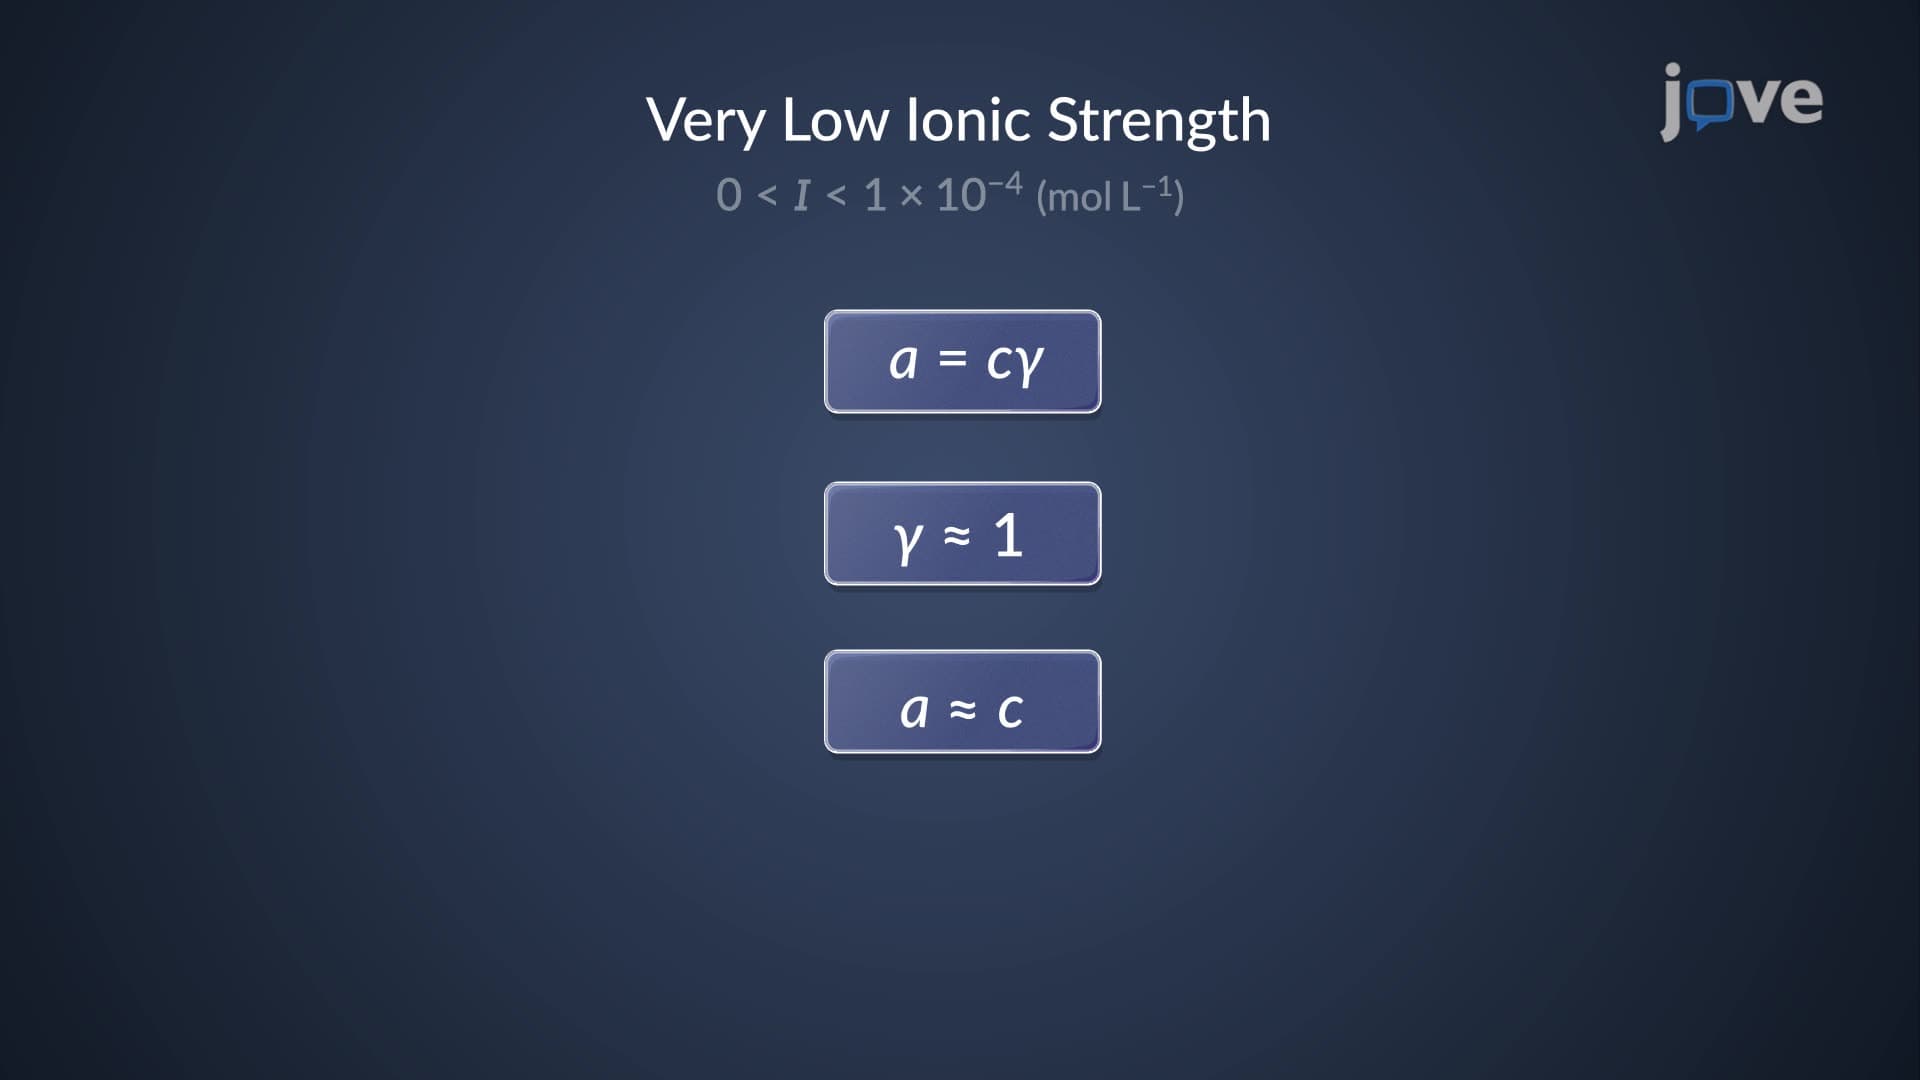
Thermodynamics: Activity Coefficient
The activity coefficient is a measure of the deviation from ideal behavior. When the ionic strength of the solution is minimal, the activity coefficient of an ionic species is close to unity, making...
 02:35
Noncovalent Attractions in Biomolecules
02:35Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
 01:24
Van der Waals Interactions
 02:46
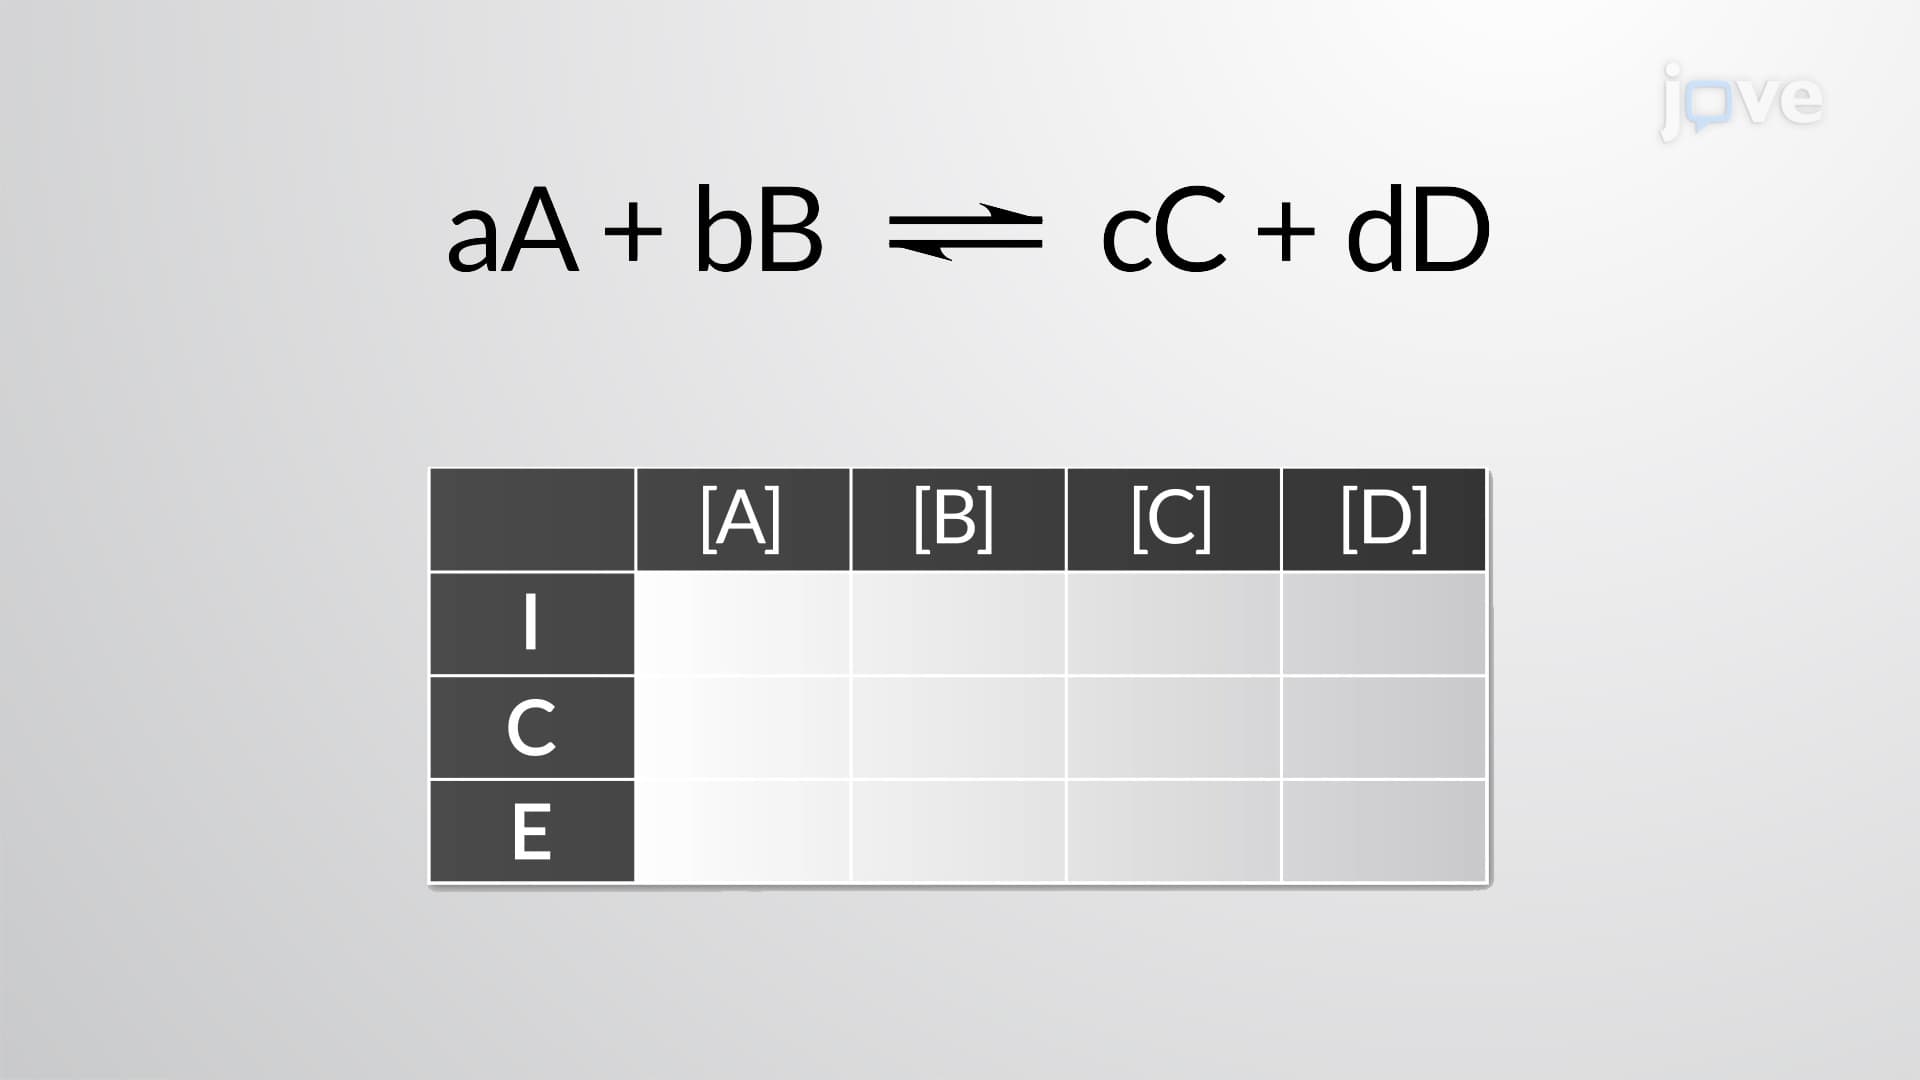
Calculating the Equilibrium Constant
For example, gaseous nitrogen dioxide forms dinitrogen tetroxide according to this equation: