Related Experiment Video
Updated: Dec 28, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
The MRCC program system: Accurate quantum chemistry from water to proteins
Mihály Kállay1, Péter R Nagy1, Dávid Mester1
1Department of Physical Chemistry and Materials Science, Budapest University of Technology and Economics, P.O. Box 91, H-1521 Budapest, Hungary.

Abstract:
MRCC is a package of ab initio and density functional quantum chemistry programs for accurate electronic structure calculations. The suite has efficient implementations of both low- and high-level correlation methods, such as second-order Møller-Plesset (MP2), random-phase approximation (RPA), second-order algebraic-diagrammatic construction [ADC(2)], coupled-cluster (CC), configuration interaction (CI), and related techniques. It has a state-of-the-art CC singles and doubles with perturbative triples [CCSD(T)] code, and its specialties, the arbitrary-order iterative and perturbative CC methods developed by automated programming tools, enable achieving convergence with regard to the level of correlation. The package also offers a collection of multi-reference CC and CI approaches. Efficient implementations of density functional theory (DFT) and more advanced combined DFT-wave function approaches are also available. Its other special features, the highly competitive linear-scaling local correlation schemes, allow for MP2, RPA, ADC(2), CCSD(T), and higher-order CC calculations for extended systems. Local correlation calculations can be considerably accelerated by multi-level approximations and DFT-embedding techniques, and an interface to molecular dynamics software is provided for quantum mechanics/molecular mechanics calculations. All components of MRCC support shared-memory parallelism, and multi-node parallelization is also available for various methods. For academic purposes, the package is available free of charge.


Related Concept Videos
 01:10
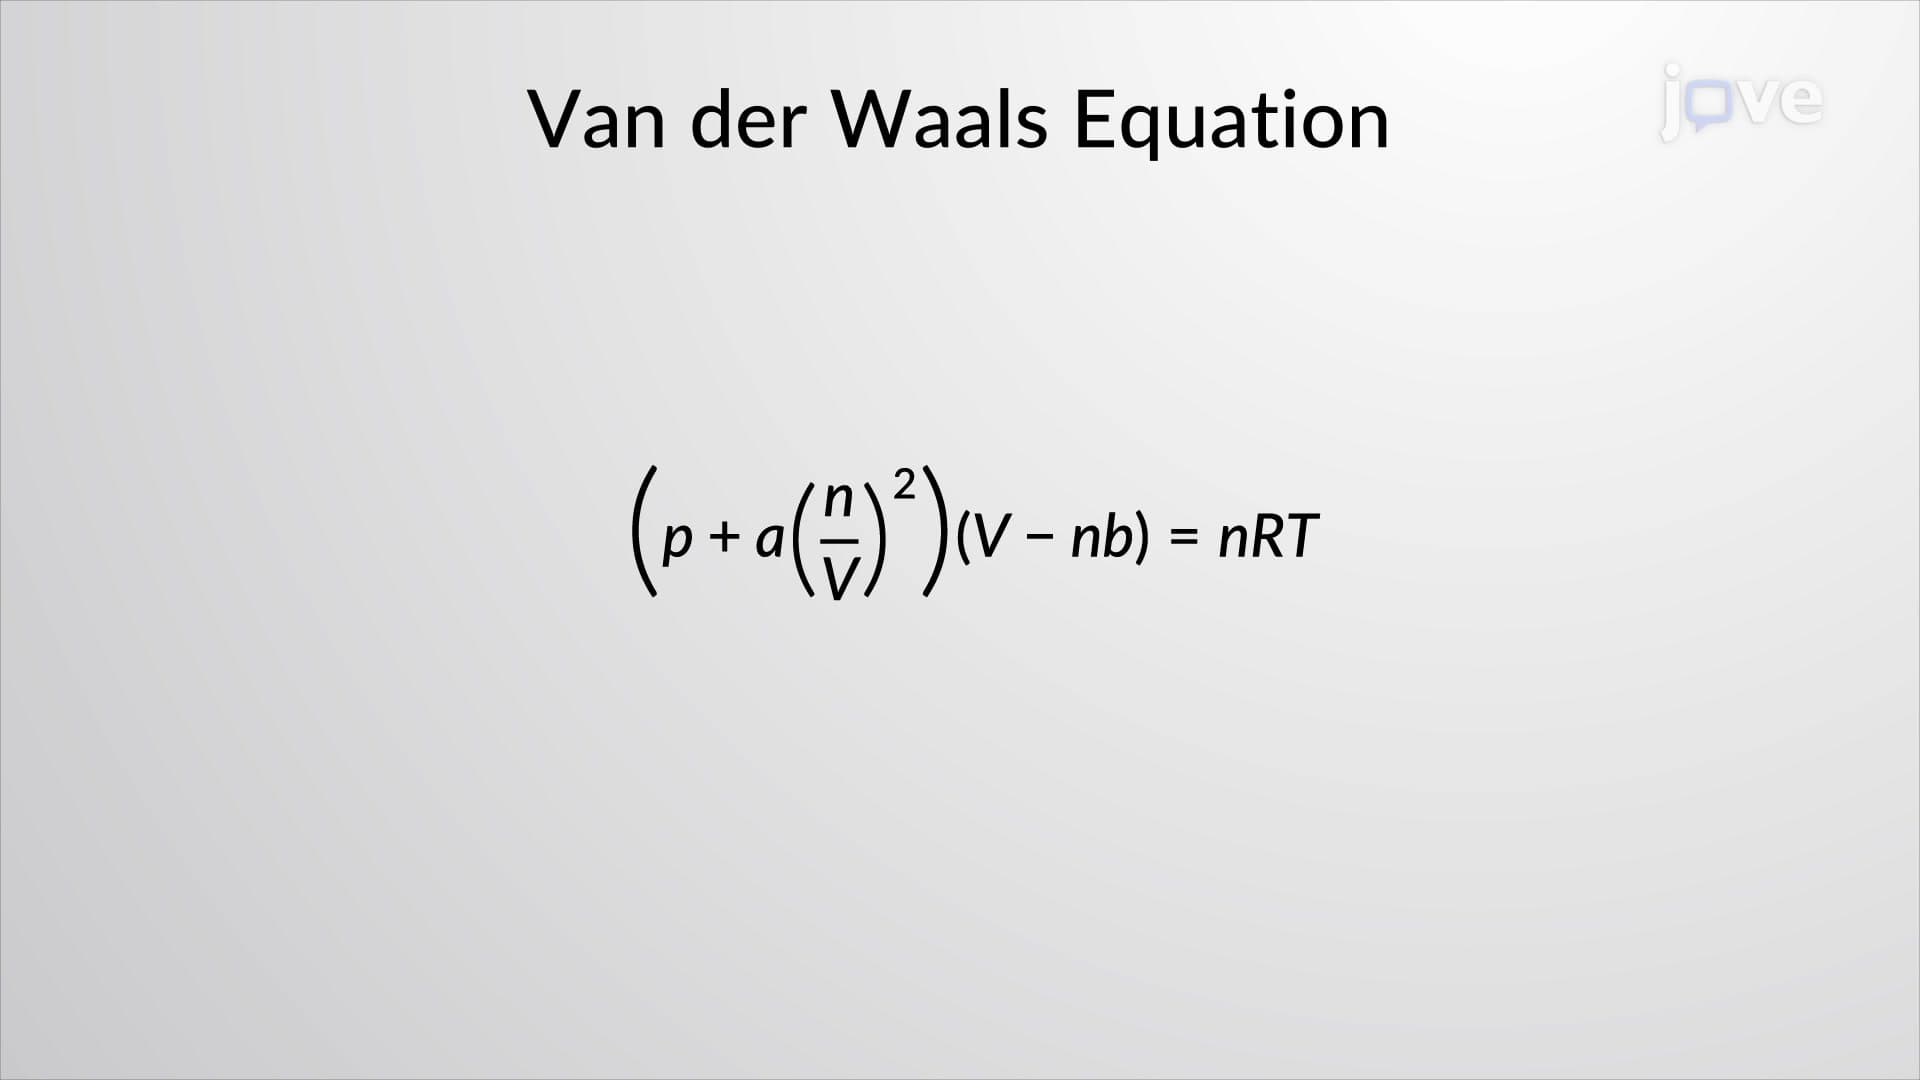
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...