The Methyl Torsion in Unsaturated Compounds
Related Experiment Video
Updated: Dec 27, 2025
Protocol for the Synthesis of Ortho-trifluoromethoxylated Aniline Derivatives
Published on: January 19, 2016
Andrea Zachariou1,2, Alexander P Hawkins1,2, Paul Collier3
1School of Chemistry, University of Glasgow, Joseph Black Building, Glasgow G12 8QQ, U.K.

Abstract:
How the methyl torsion transition energy in unsaturated systems is affected by its environment is investigated. It is strongly influenced by both its immediate neighborhood, (the number of methyl groups present in the molecule) and the intermolecular interactions. It is clear that the intermolecular interactions have a major influence on the torsion transition energy, as demonstrated unambiguously previously for mesitylene and also seen here for other systems. In part, this may be caused by the fact that the methyl torsion is rarely a pure mode (unless enforced by symmetry). Where the crystal structure is available, the assignments have been supported by CASTEP calculations of the unit cell. The agreement between the observed and calculated spectra is generally good, although not perfect, toluene being a case in point, and highlights just how demanding it is to obtain accurate transition energies for low energy modes. The disagreement between observed and calculated inelastic neutron scattering spectra for meta-xylene and 9,10 dimethylanthracene is so severe that it would suggest that there are additional phases to those presently known. Comparison between the full periodic calculations and those for the isolated molecule shows that intermolecular interactions raise the methyl torsion transition energy by at least 8% and in some cases by more than 50%. The presence of more than one methyl group in the molecule generally raises the average torsion energy from the <100 cm-1 seen for single methyl groups to 150-200 cm-1.
Related Concept Videos
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 02:29
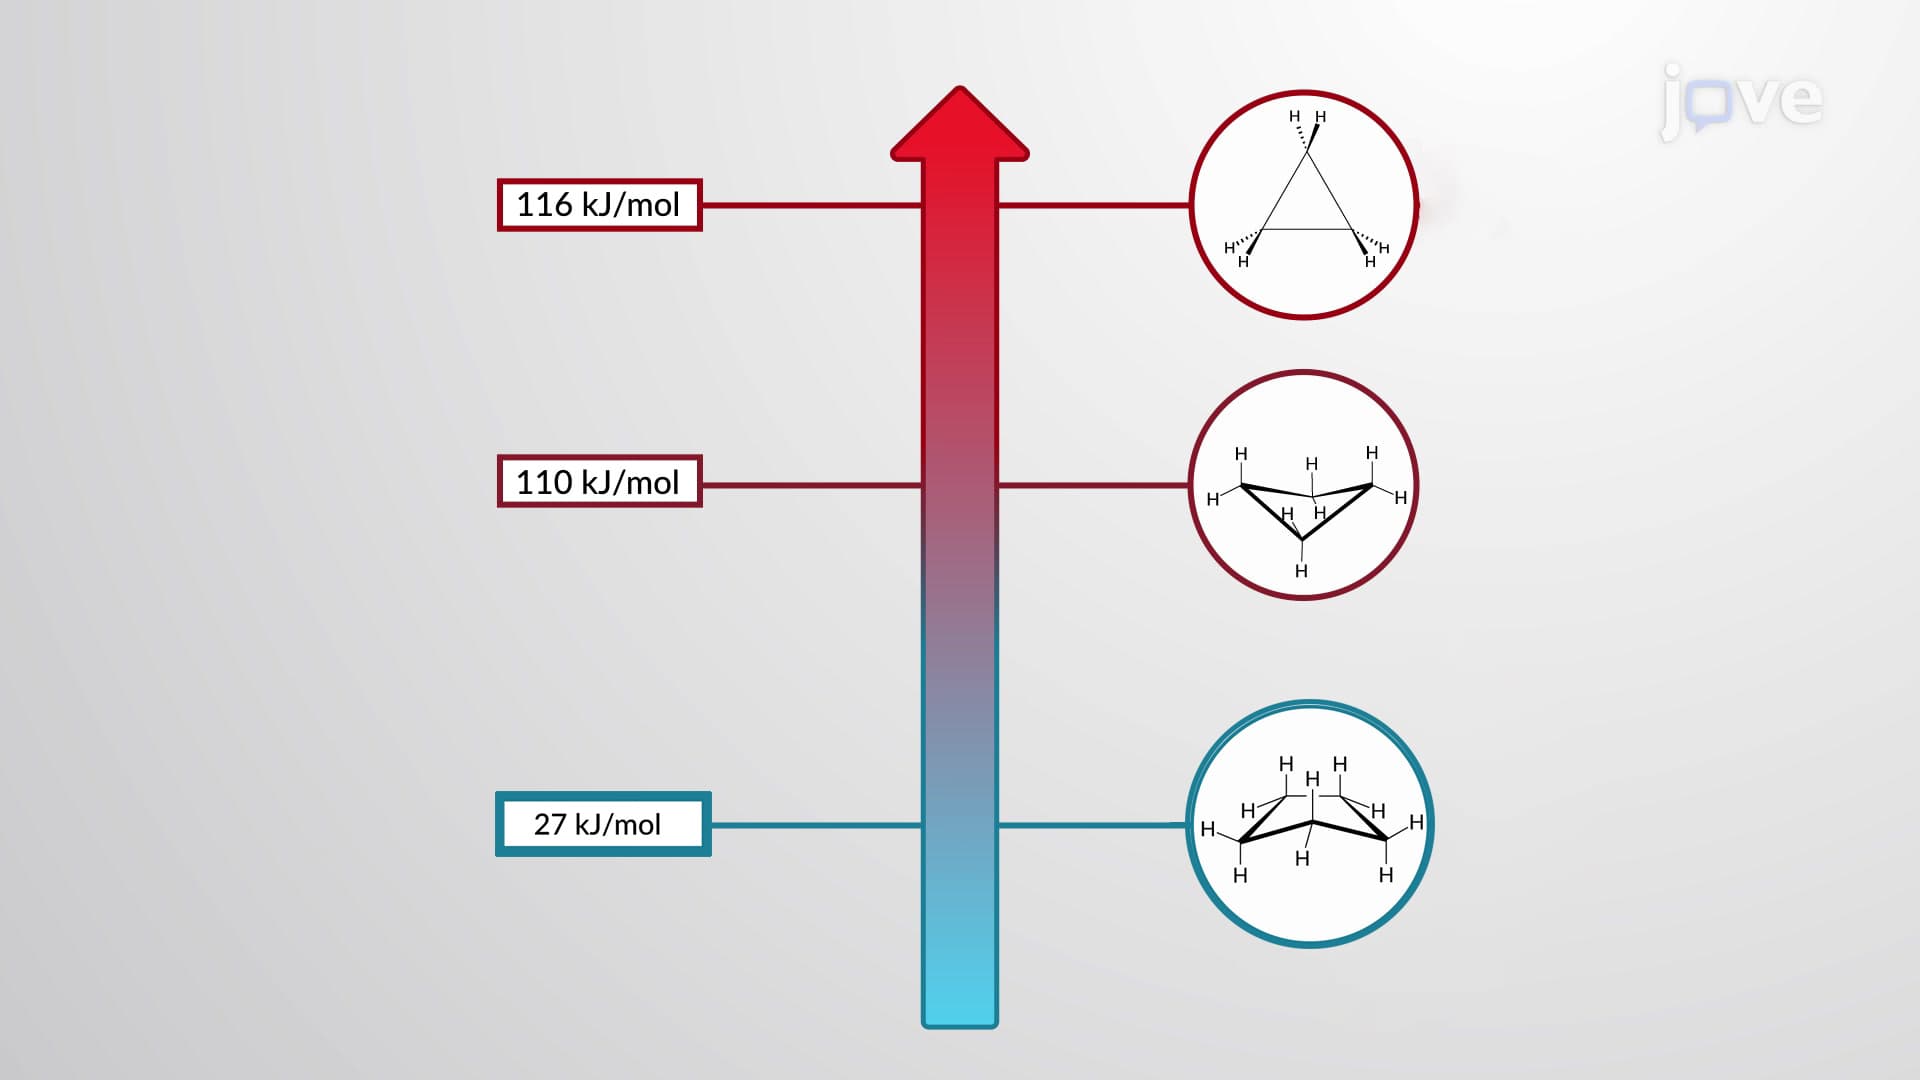
Conformations of Cycloalkanes
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
 03:24
Hybridization of Atomic Orbitals I
 02:18
Conformations of Ethane and Propane
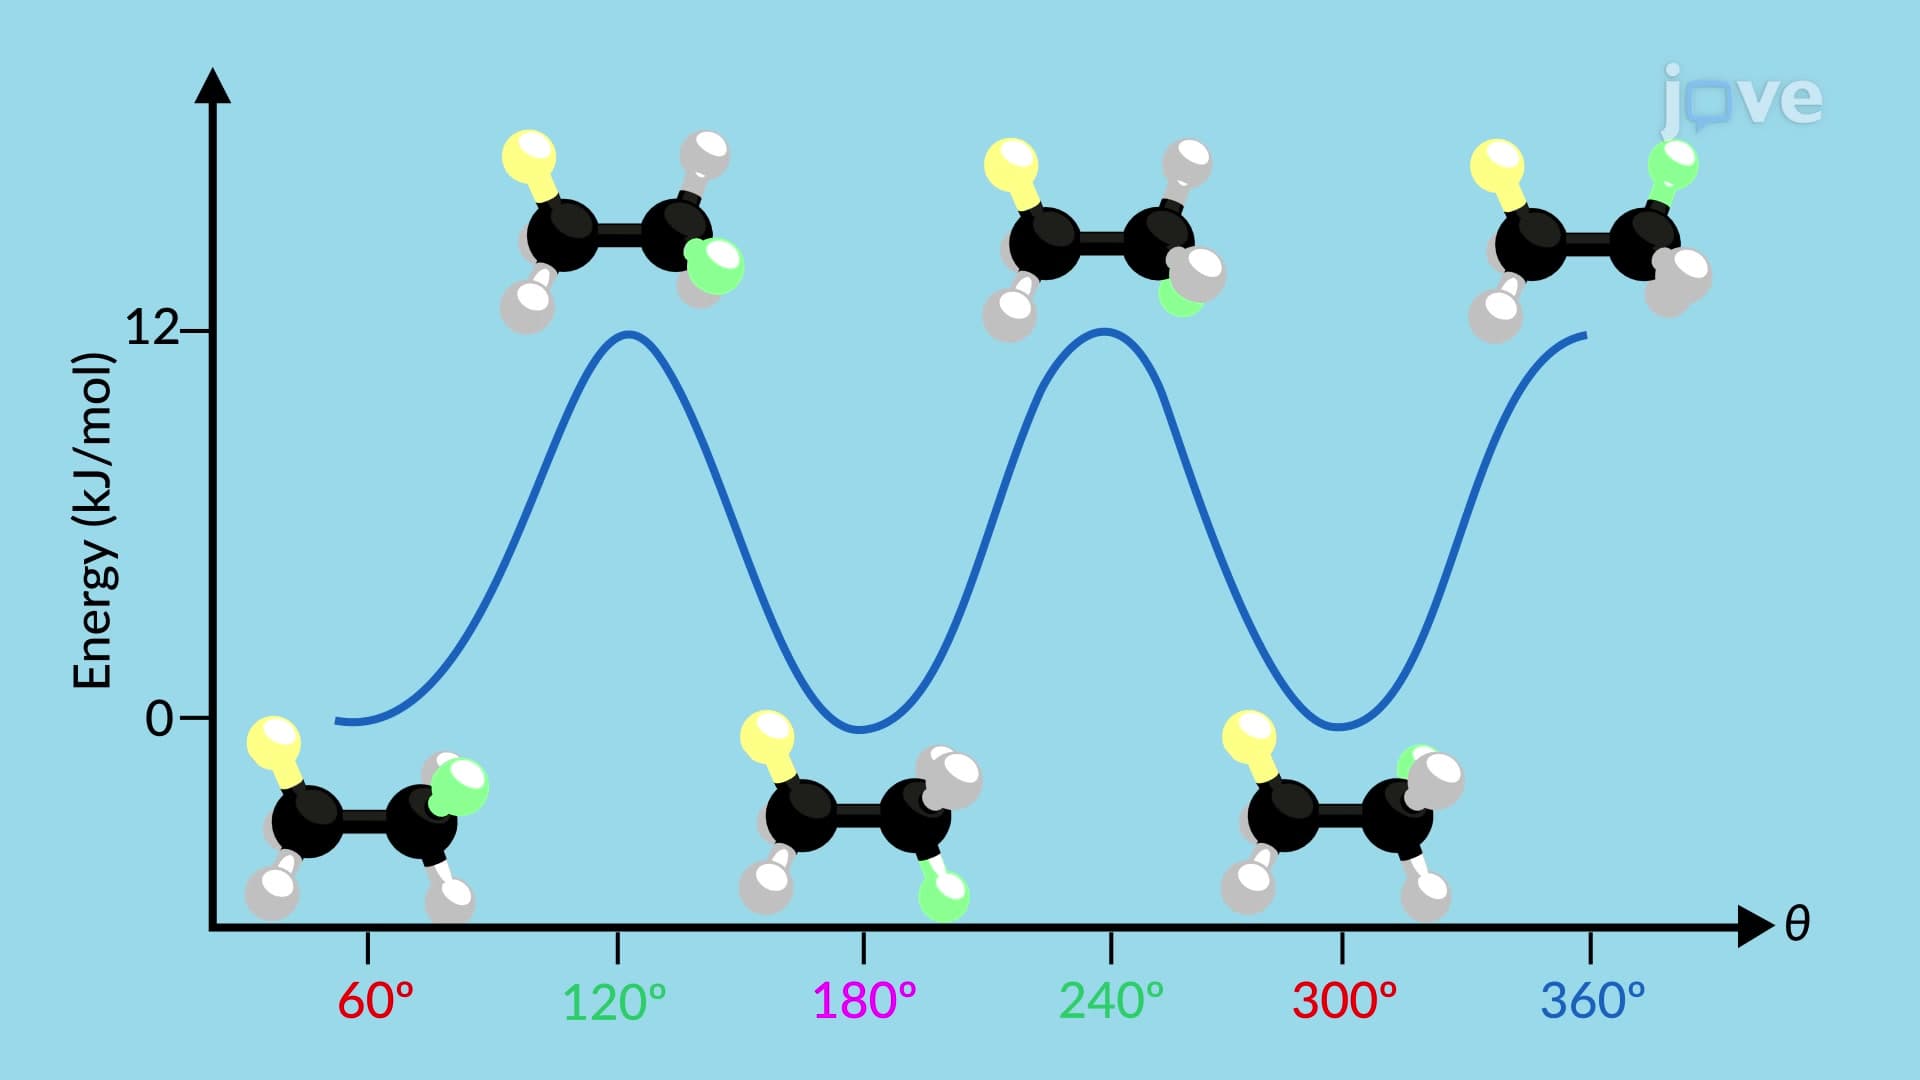
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered...