
Vibrational Dynamics of Crystalline 4-Phenylbenzaldehyde from INS Spectra and Periodic DFT Calculations
Mariela M Nolasco1, Catarina F Araujo1, Pedro D Vaz2
1CICECO, Departamento de Química, Universidade de Aveiro, P-3810-193 Aveiro, Portugal.

Abstract:
The present work emphasizes the value of periodic density functional theory (DFT) calculations in the assessment of the vibrational spectra of molecular crystals. Periodic calculations provide a nearly one-to-one match between the calculated and observed bands in the inelastic neutron scattering (INS) spectrum of crystalline 4-phenylbenzaldehyde, thus validating their assignment and correcting previous reports based on single molecule calculations. The calculations allow the unambiguous assignment of the phenyl torsional mode at ca. 118-128 cm-1, from which a phenyl torsional barrier of ca. 4000 cm-1 is derived, and the identification of the collective mode involving the antitranslational motion of CH···O bonded pairs, a hallmark vibrational mode of systems where C-H···O contacts are an important feature.
Related Concept Videos
 01:34
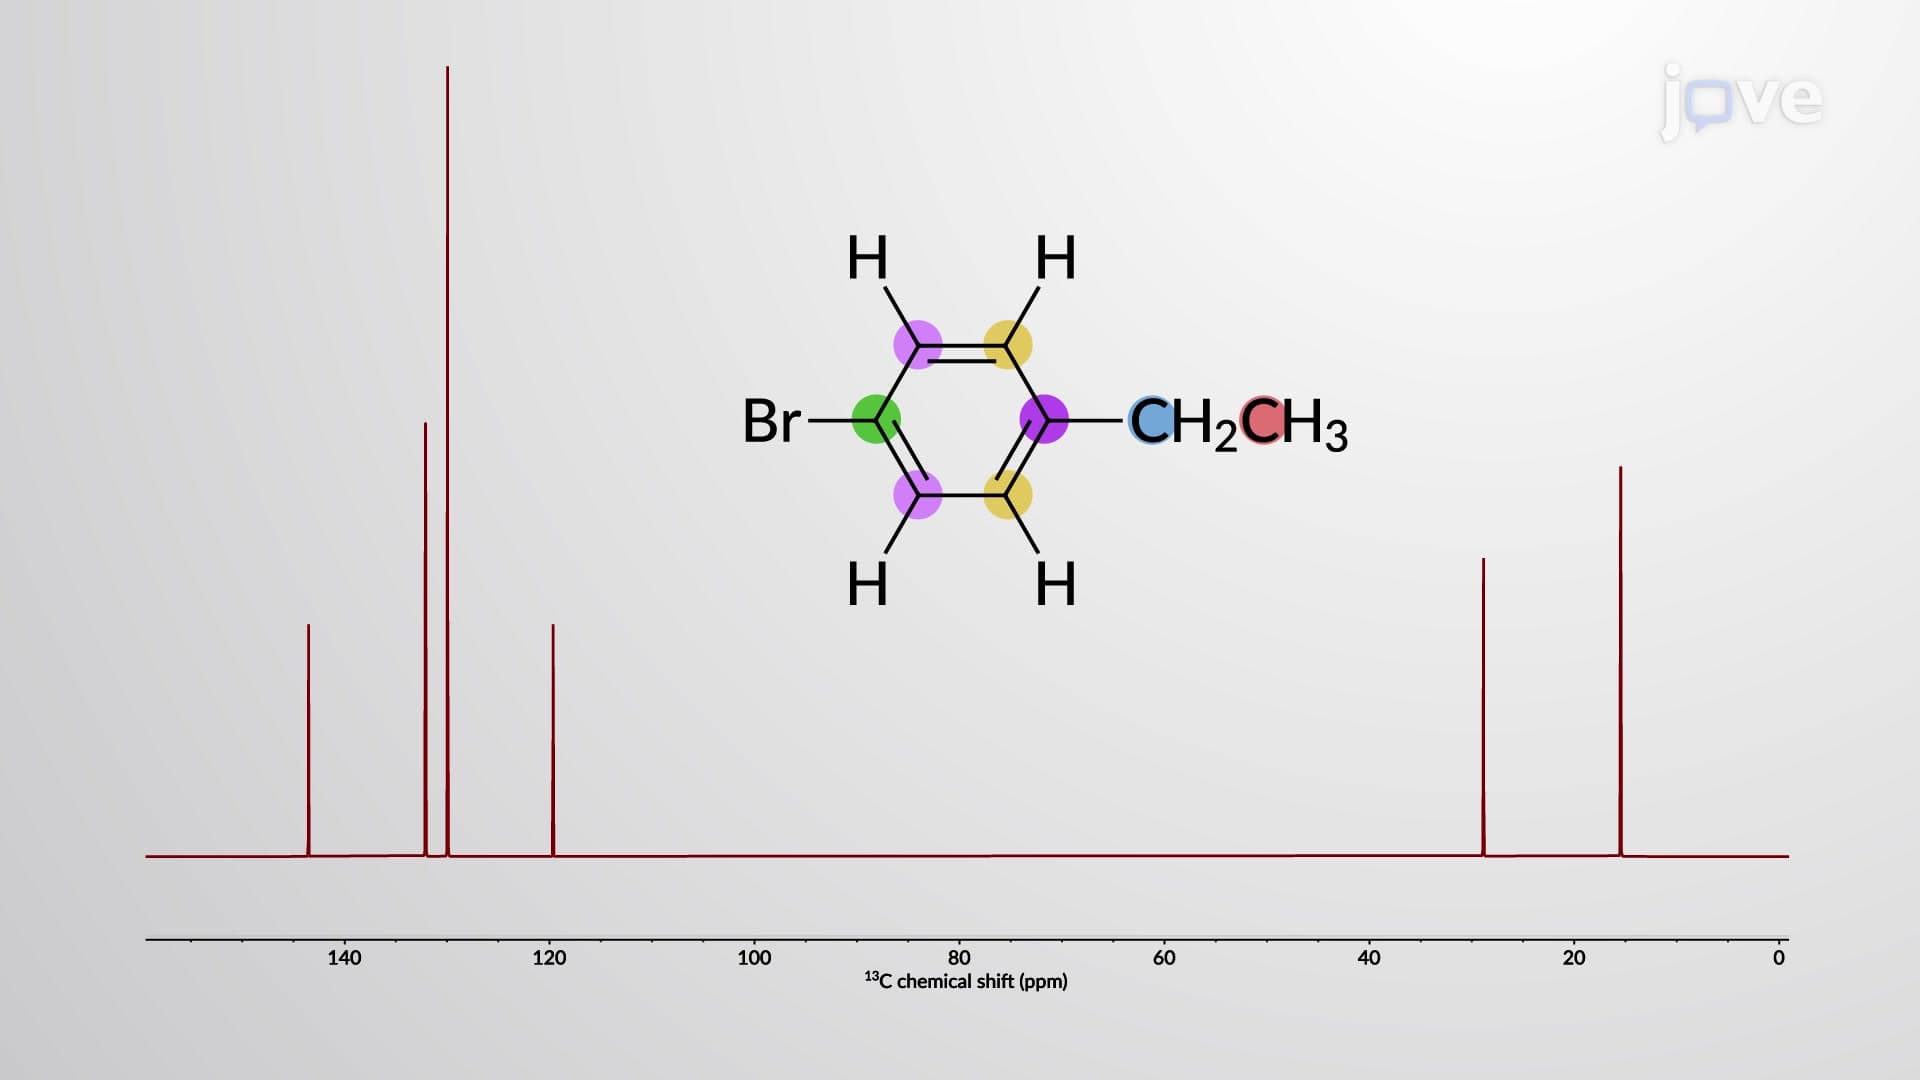
NMR Spectroscopy of Benzene Derivatives
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:29
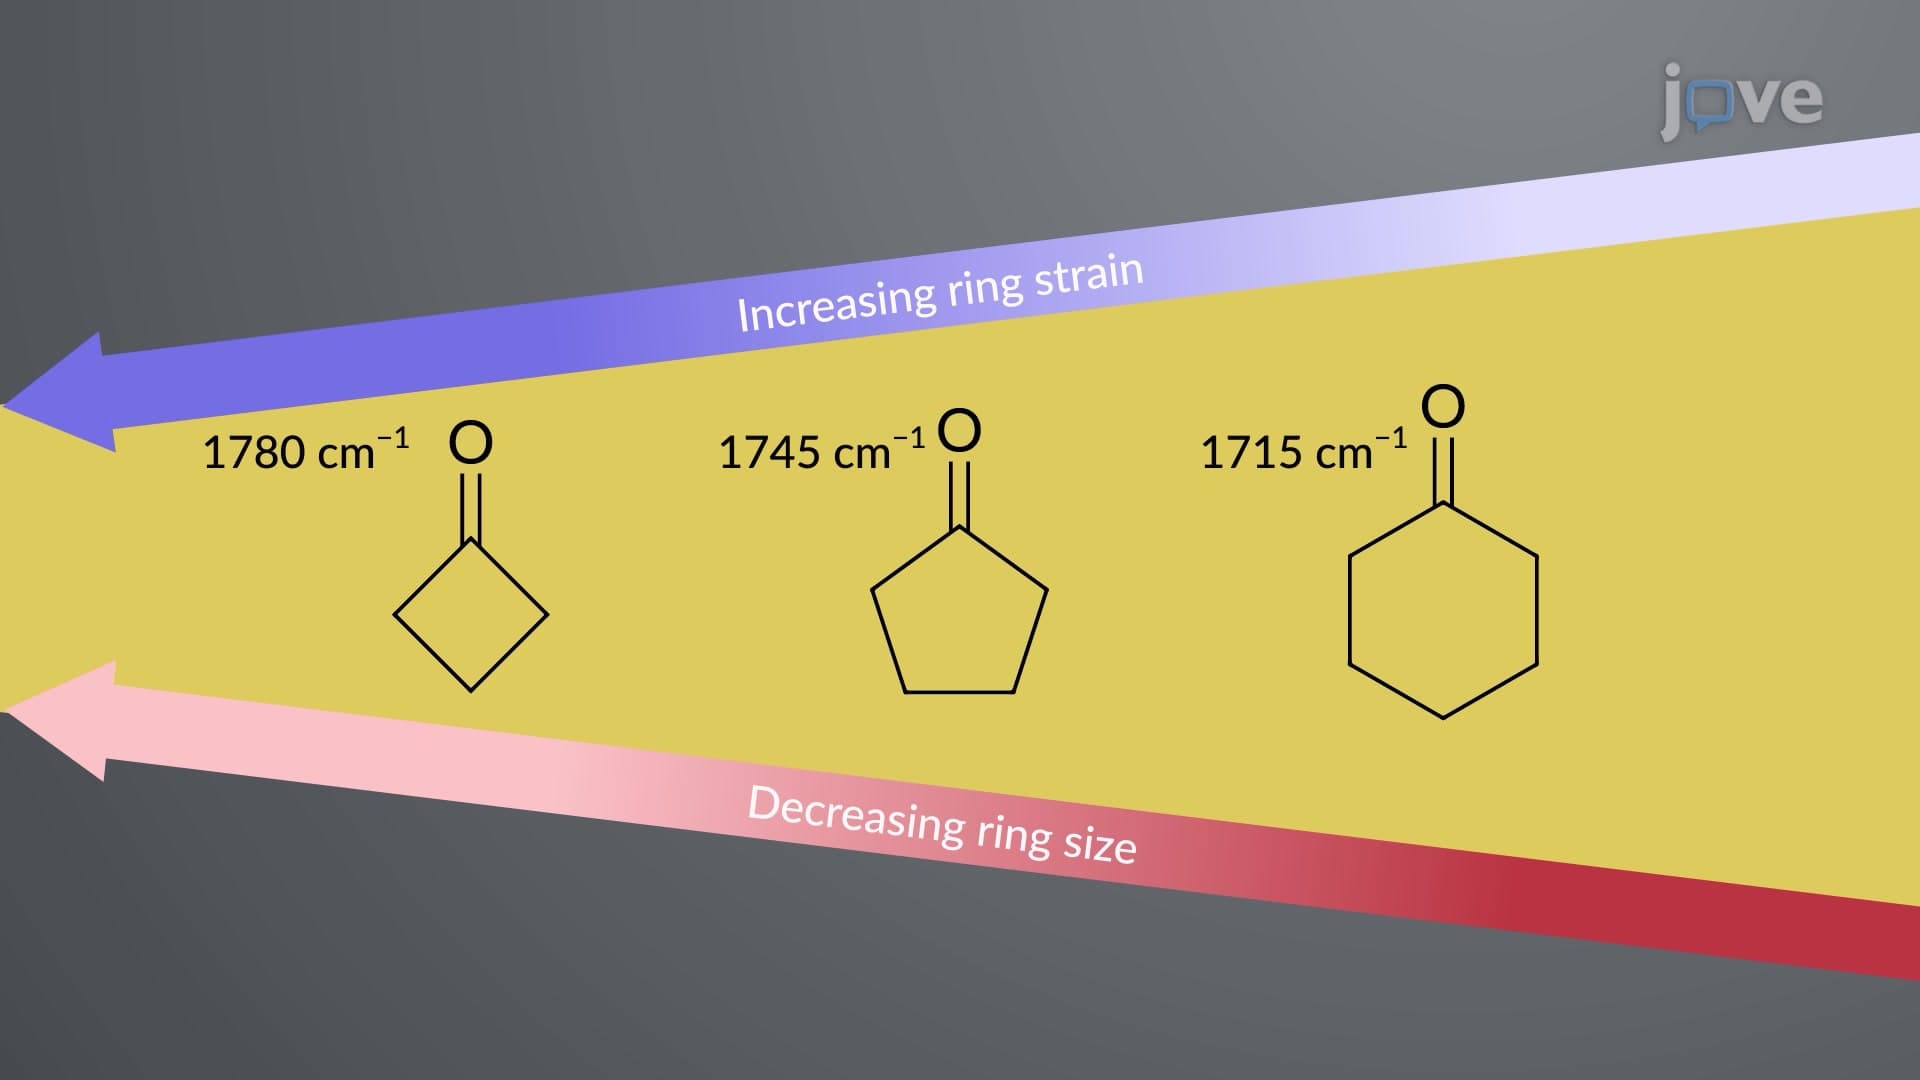
IR and UV–Vis Spectroscopy of Aldehydes and Ketones
 01:18
Structure of Benzene: Molecular Orbital Model
 01:23
IR Spectrum Peak Broadening: Hydrogen Bonding
However, the extent of hydrogen bonding influences the observed stretching frequency and band broadening. Intermolecular or intramolecular...