Related Experiment Video
Updated: Dec 24, 2025

15N CPMG Relaxation Dispersion for the Investigation of Protein Conformational Dynamics on the µs-ms Timescale
Published on: April 19, 2021
13C NMR Relaxation Analysis of Protein GB3 for the Assessment of Side Chain Dynamics Predictions by Current AMBER and
Janet S Anderson1, Griselda Hernández2, David M LeMaster2
1Department of Chemistry, Union College, Schenectady, New York 12308, United States.

Abstract:
Molecular simulations with seven current AMBER- and CHARMM-based force fields yield markedly differing internal bond vector autocorrelation function predictions for many of the 223 methine and methylene H-C bonds of the 56-residue protein GB3. To enable quantification of accuracy, 13C R1, R2, and heteronuclear NOE relaxation rates have been determined for the methine and stereochemically assigned methylene Cα and Cβ positions. With only three experimental relaxation values for each bond vector, central to this analysis is the accuracy with which MD-derived autocorrelation curves can be represented by a 3-parameter equation which, in turn, maps onto the NMR relaxation values. In contrast to the more widely used extended Lipari-Szabo order parameter representation, 95% of these MD-derived internal autocorrelation curves for GB3 can be fitted to within 1.0% rmsd over the time frame from 30 ps to 4 ns by a biexponential Larmor frequency-selective representation (LF-S2). Applying the LF-S2 representation to the experimental relaxation rates and uncertainties serves to determine the boundary range for the autocorrelation function of each bond vector consistent with the experimental data. Not surprisingly, all seven force fields predict the autocorrelation functions for the more motionally restricted 1Hα-13Cα and 1Hβ-13Cβ bond vectors with reasonable accuracy. However, for the 1Hβ-13Cβ bond vectors exhibiting aggregate order parameter S2 values less than 0.85, only 1% of the MD-derived predictions lie with 1 σ of the experimentally determined autocorrelation functions and only 7% within 2 σ. On the other hand, substantial residue type-specific improvements in predictive performance were observed among the recent AMBER force fields. This analysis indicates considerable potential for the use of 13C relaxation measurements in guiding the optimization of the side chain dynamics characteristics of protein molecular simulations.
Related Concept Videos
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:19
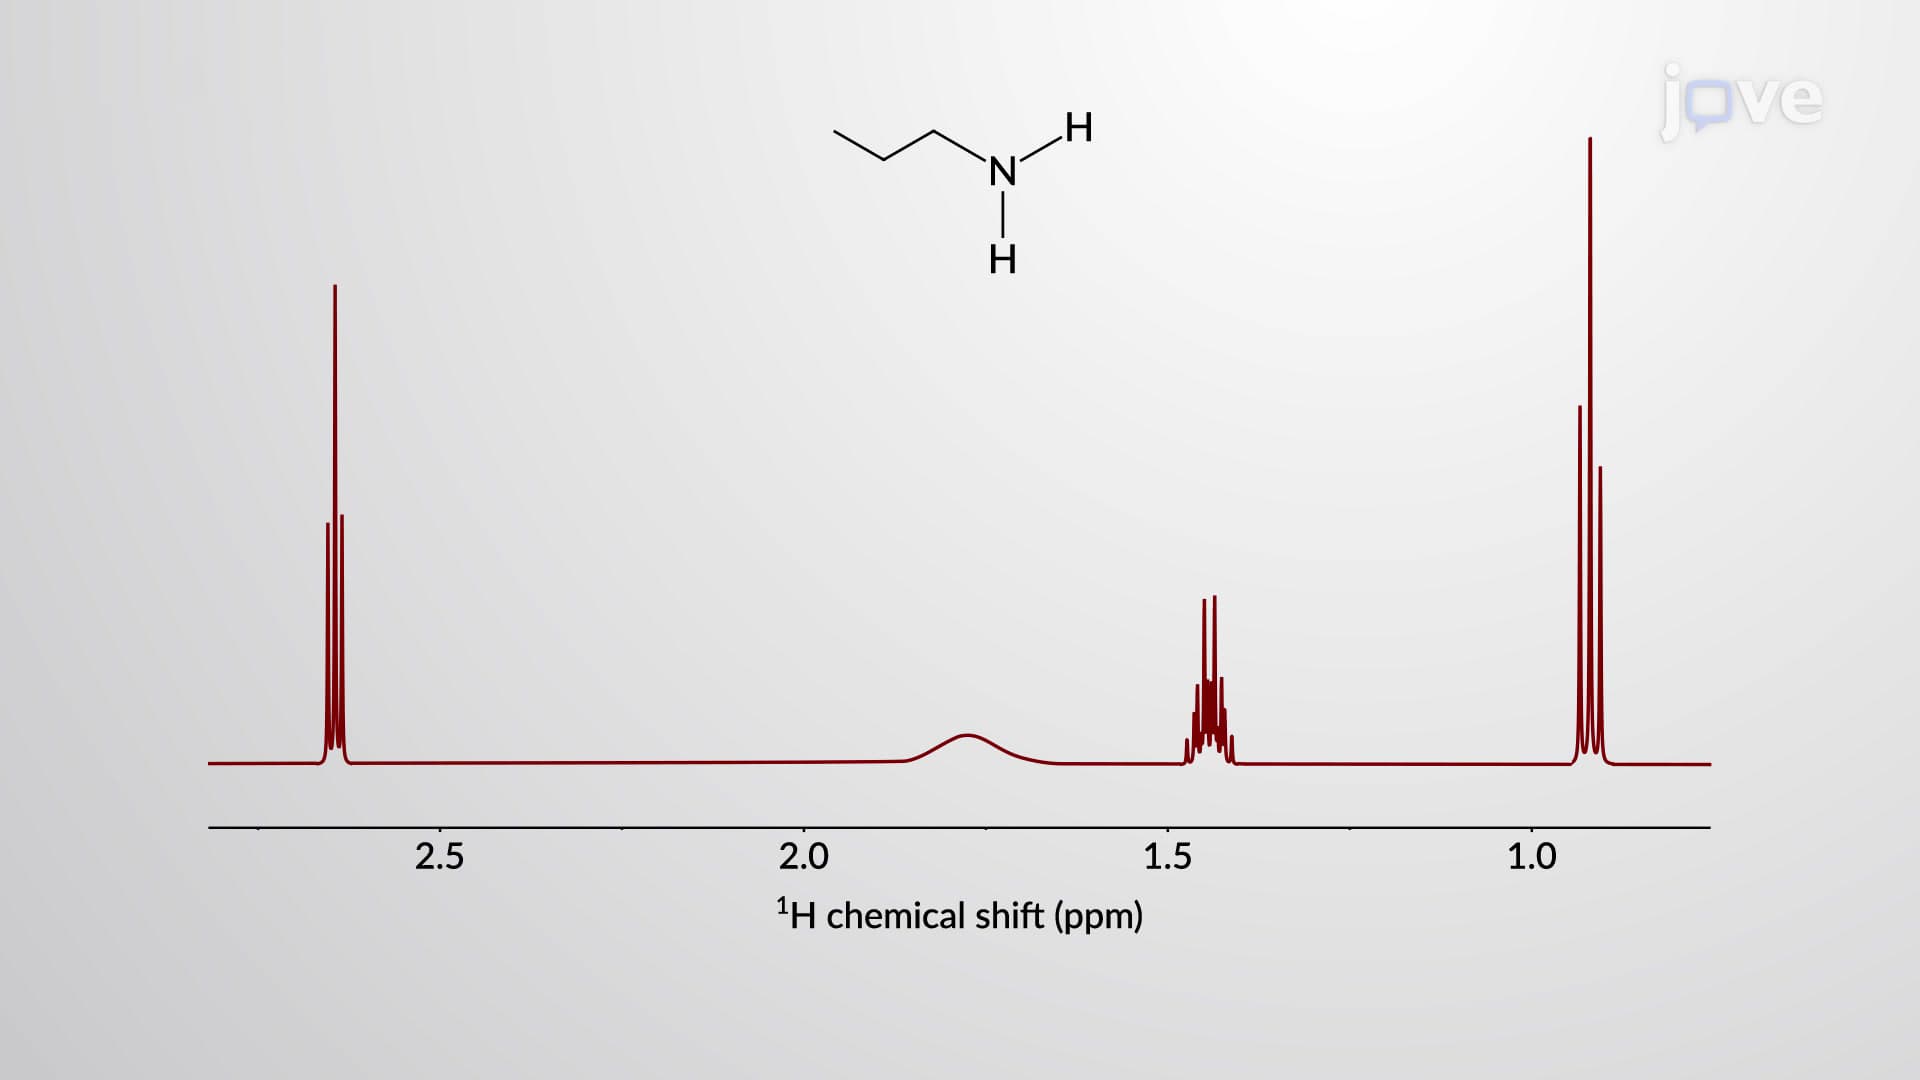
NMR Spectroscopy Of Amines
 01:10
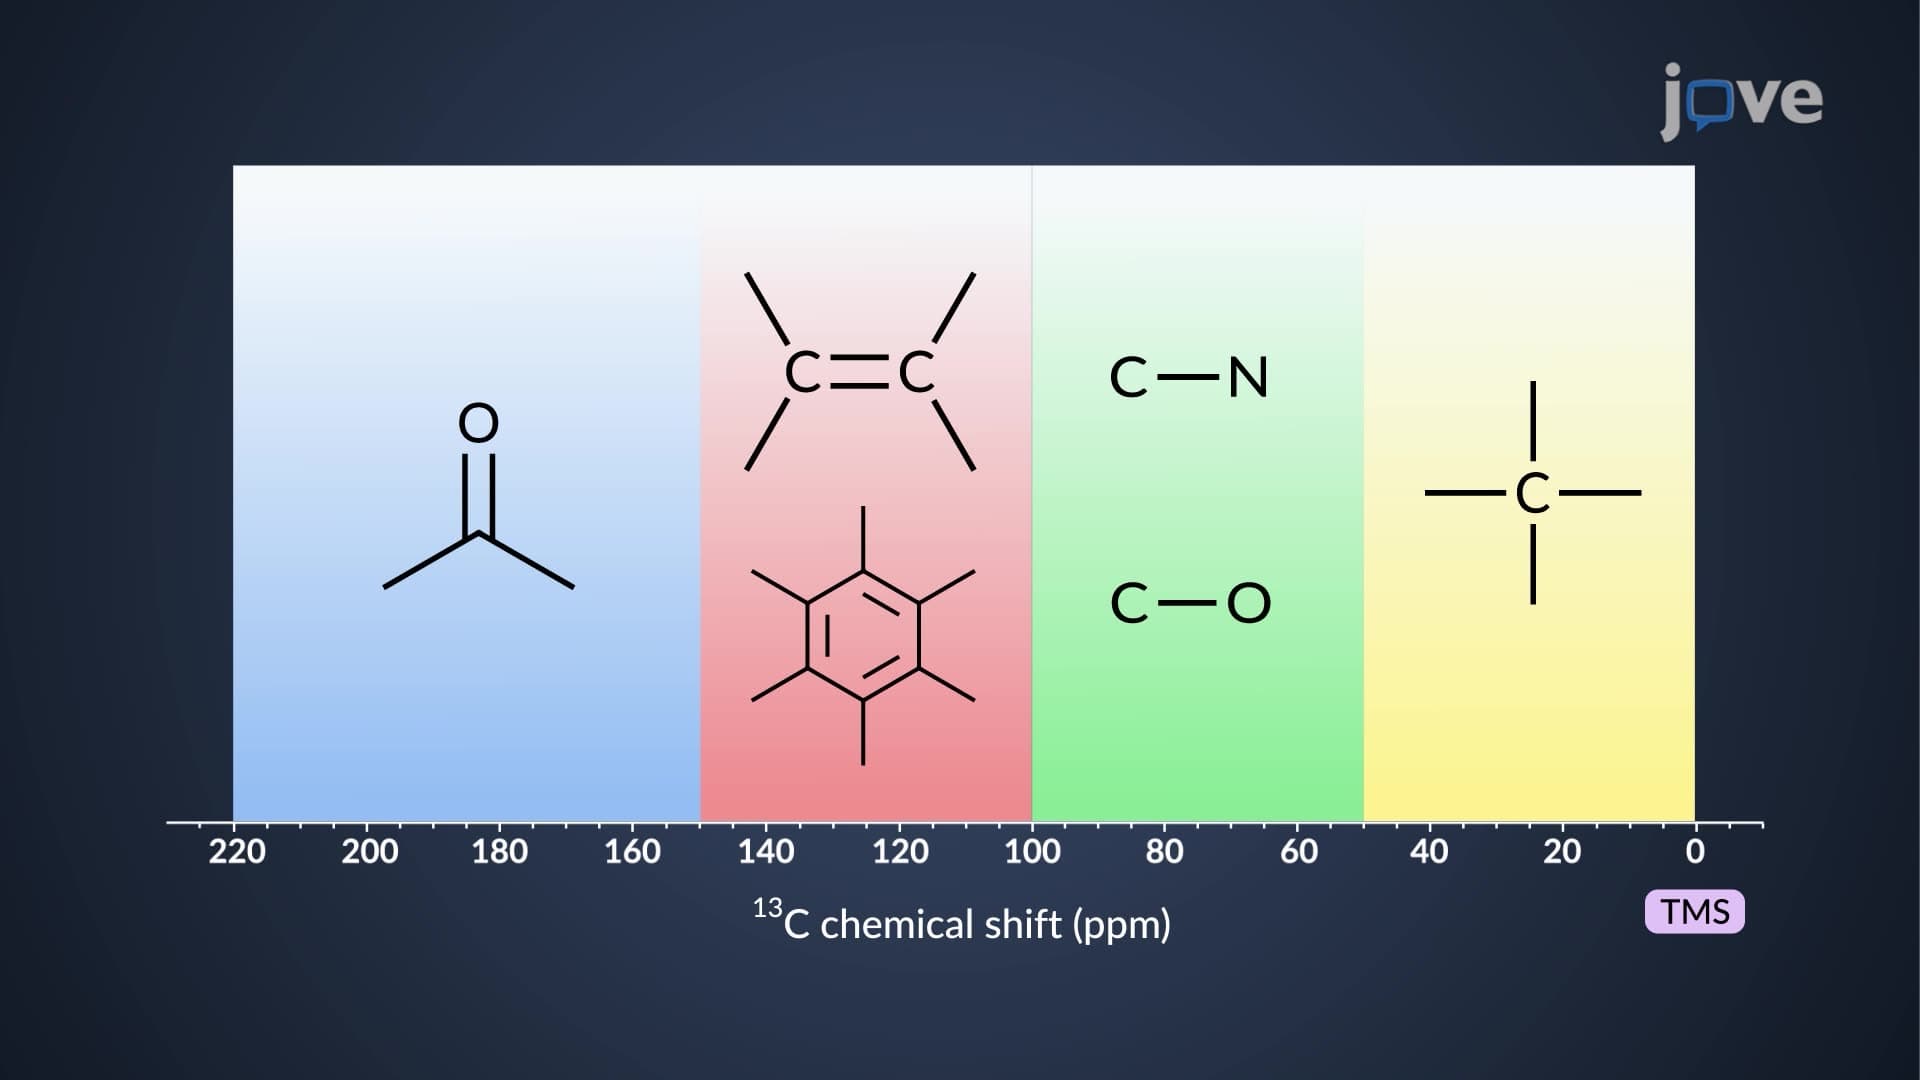
Carbon-13 (¹³C) NMR: Overview