Multicomponent Quantum Chemistry: Integrating Electronic and Nuclear Quantum Effects via the Nuclear-Electronic Orbital Method
Related Experiment Video
Updated: Dec 24, 2025
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Fabijan Pavošević1, Tanner Culpitt1, Sharon Hammes-Schiffer1
1Department of Chemistry, Yale University, 225 Prospect Street, New Haven, Connecticut 06520, United States.

Abstract:
In multicomponent quantum chemistry, more than one type of particle is treated quantum mechanically with either density functional theory or wave function based methods. In particular, the nuclear-electronic orbital (NEO) approach treats specified nuclei, typically hydrogen nuclei, on the same level as the electrons. This approach enables the incorporation of nuclear quantum effects, such as nuclear delocalization, anharmonicity, zero-point energy, and tunneling, as well as non-Born-Oppenheimer effects directly into quantum chemistry calculations. Such effects impact optimized geometries, molecular vibrational frequencies, reaction paths, isotope effects, and dynamical simulations. Multicomponent density functional theory (NEO-DFT) and time-dependent DFT (NEO-TDDFT) achieve an optimal balance between computational efficiency and accuracy for computing ground and excited state properties, respectively. Multicomponent wave function based methods, such as the coupled cluster singles and doubles (NEO-CCSD) method for ground states and the equation-of-motion counterpart (NEO-EOM-CCSD) for excited states, attain similar accuracy without requiring any parametrization and can be systematically improved but are more computationally expensive. Variants of the orbital-optimized perturbation theory (NEO-OOMP2) method achieve nearly the accuracy of NEO-CCSD for ground states with significantly lower computational cost. Additional approaches for computing excited electronic, vibrational, and vibronic states include the delta self-consistent field (NEO-ΔSCF), complete active space SCF (NEO-CASSCF), and nonorthogonal configuration interaction methods. Multireference methods are particularly important for describing hydrogen tunneling processes. Other types of multicomponent systems, such as those containing electrons and positrons, have also been studied within the NEO framework. The NEO approach allows the incorporation of nuclear quantum effects and non-Born-Oppenheimer effects for specified nuclei into quantum chemistry calculations in an accessible and computationally efficient manner.
More Related Videos

Related Concept Videos
 02:45
The Quantum-Mechanical Model of an Atom
 03:35
Hybridization of Atomic Orbitals II
 02:35
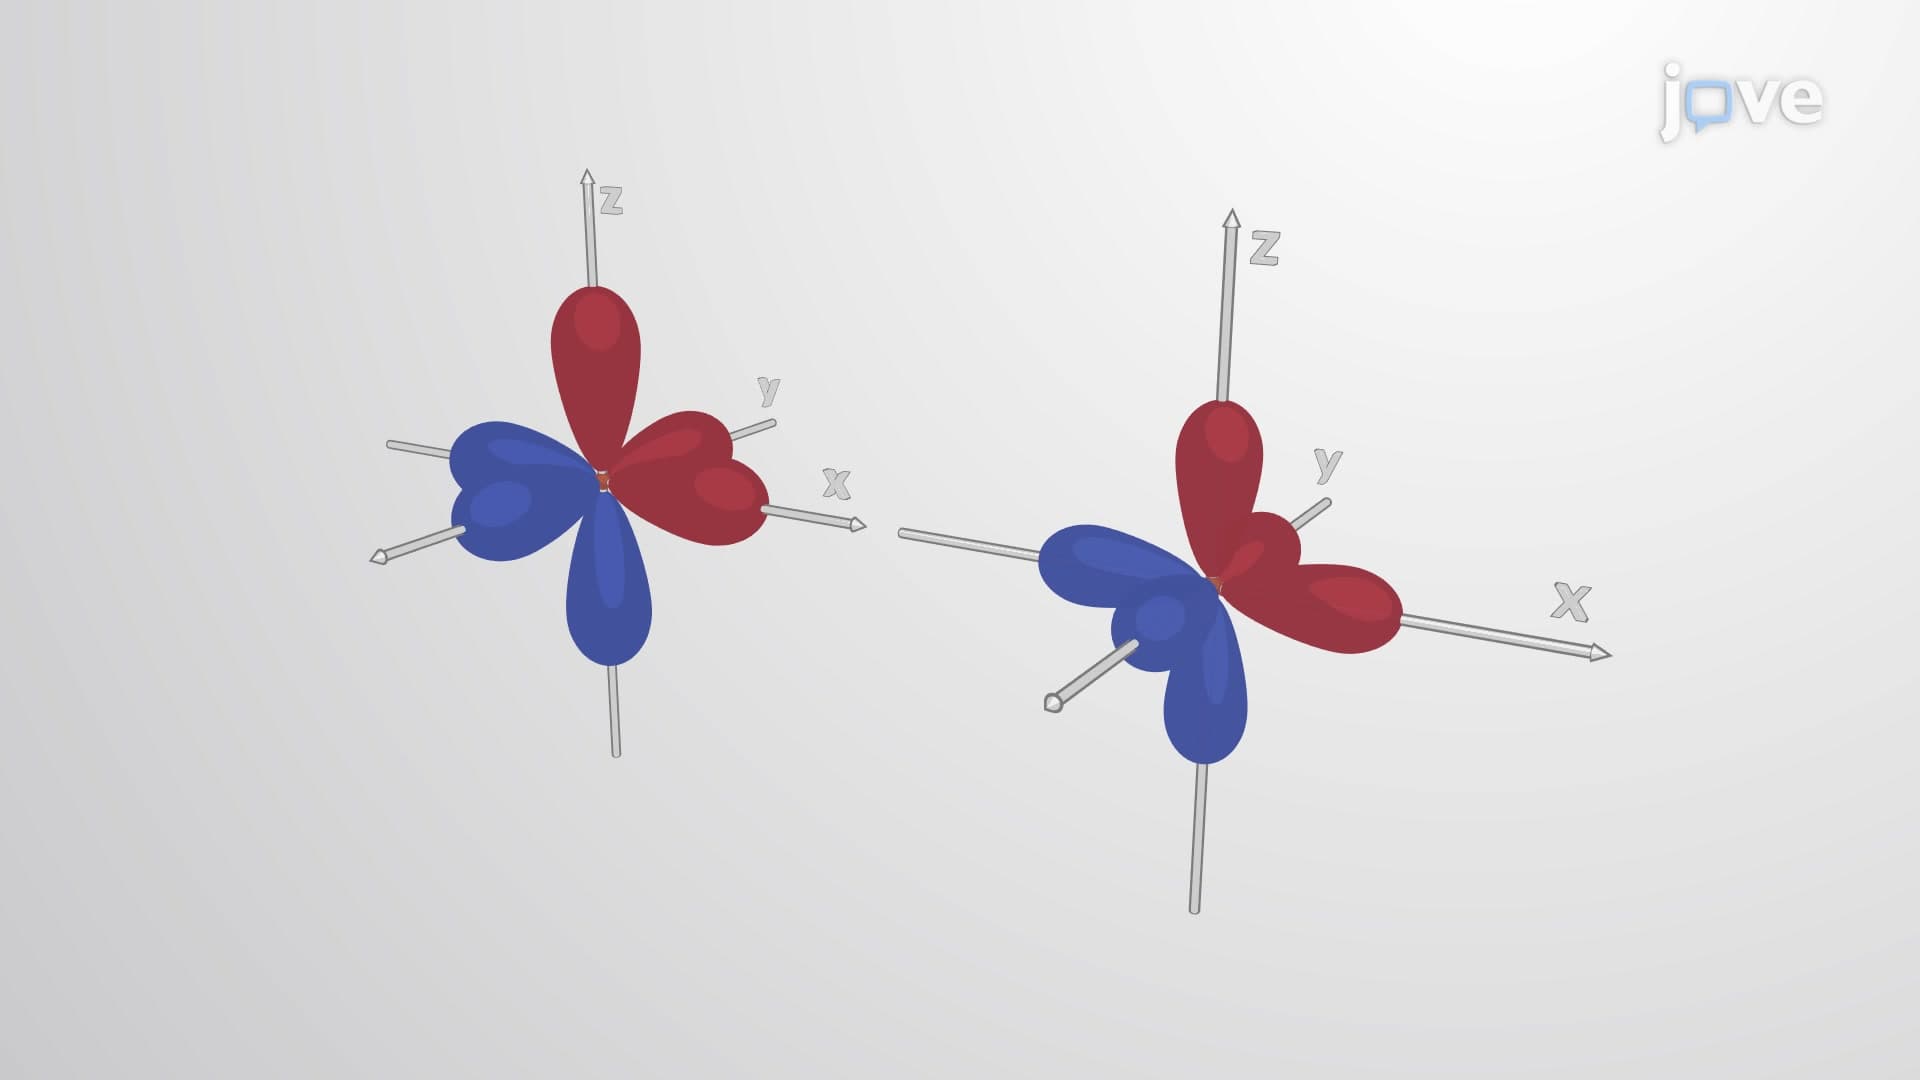
Molecular Orbital Theory I
 03:24
Hybridization of Atomic Orbitals I
 03:51
Molecular Orbital Theory II
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...