Related Experiment Video
Updated: Dec 22, 2025

Versatile CO2 Transformations into Complex Products: A One-pot Two-step Strategy
Published on: November 9, 2019
Insights on Absolute and Relative Stereocontrol in Stereodivergent Cooperative Catalysis
Avtar Changotra1, Bangaru Bhaskararao1, Christopher M Hadad2
1Department of Chemistry, Indian Institute of Technology Bombay, Powai, Mumbai 400076, India.

Abstract:
An increasing number of examples demonstrate that the use of two mutually compatible chiral catalysts in one-pot conditions can help realize the long-cherished goal of simultaneous control of absolute and relative configurations in asymmetric catalysis. Engaging two transition metal catalysts for this goal presents a considerable degree of mechanistic challenge to control the mode of substrate activation as well as origin of enantio- and diastereoselectivities, both of which are central to the burgeoning domain of stereodivergent catalysis. We have employed density functional theory (B3LYP-D3) computations to investigate an important stereodivergent reaction between azaaryl acetamide and cinnamyl methyl carbonate. These compounds participate in the stereocontrolling C-C bond formation in the form of activated substrates, respectively, when bound to chiral Cu-Walphos and Ir-phosphoramidite catalysts. Herein, we provide the molecular origin of how all four stereoisomers of the product bearing two contiguous stereogenic centers could be accessed by changing the combinations of chiral catalysts (C1(R,Rp) or C2(S,Sp) of Cu-Walphos in conjunction with P1(R,R,R) or P2(S,S,S) of Ir-phosphoramidite catalysts). The origin of stereodivergence is identified to depend on the differences in the number and nature of noncovalent interactions (NCIs) in the stereocontrolling transition states. In particular, NCIs between the chiral catalysts (C-H···π in C1-P1 catalyst dyad and C-H···π, C-H···F, and π···π in C2-P1) in stereocontrolling transition states are found to be the differentiating factors rendering one of the four stereochemically distinct transition states to be the lowest energy one for a given catalyst combination. These molecular insights suggest that subtle modifications to the catalyst framework could be further exploited in stereodivergent catalysis.
Related Concept Videos
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 01:33
Radical Halogenation: Stereochemistry
Halogenation to form a new chiral center:
 01:18
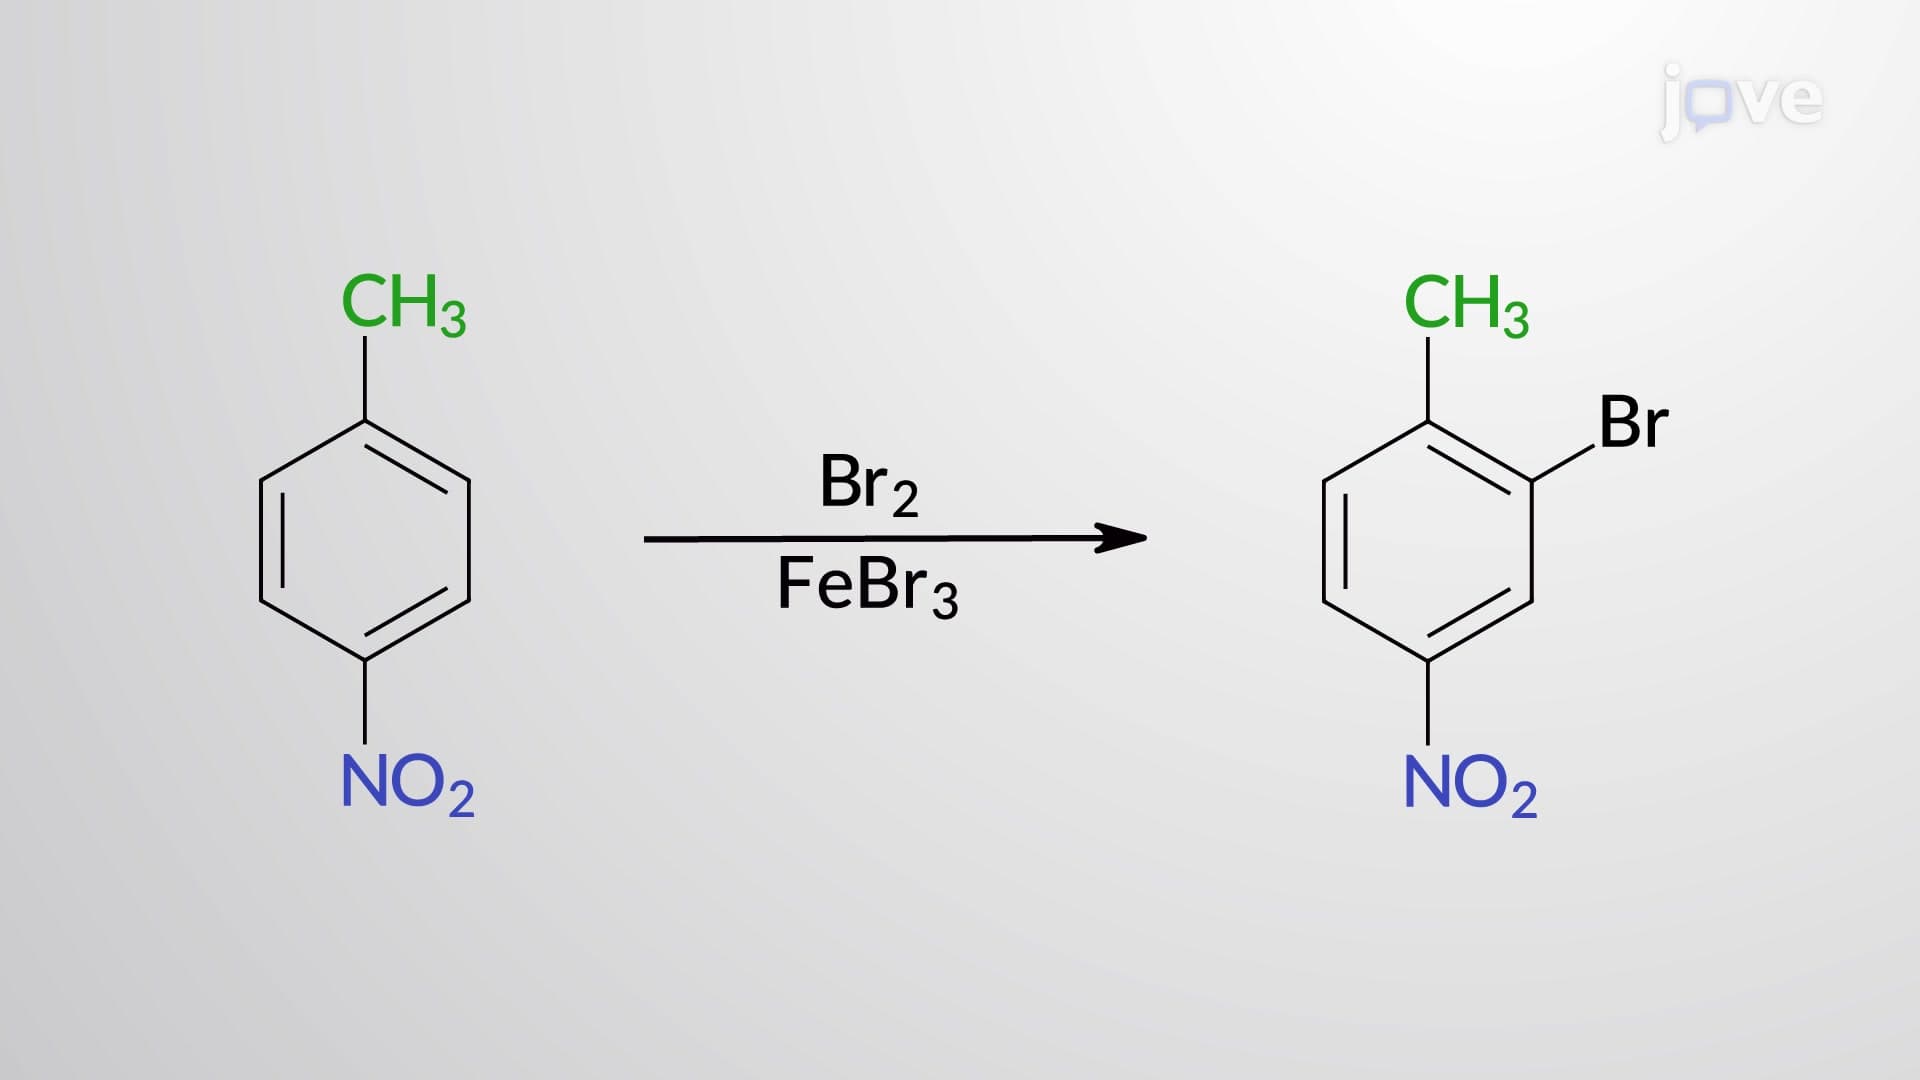
Directing and Steric Effects in Disubstituted Benzene Derivatives
 02:43
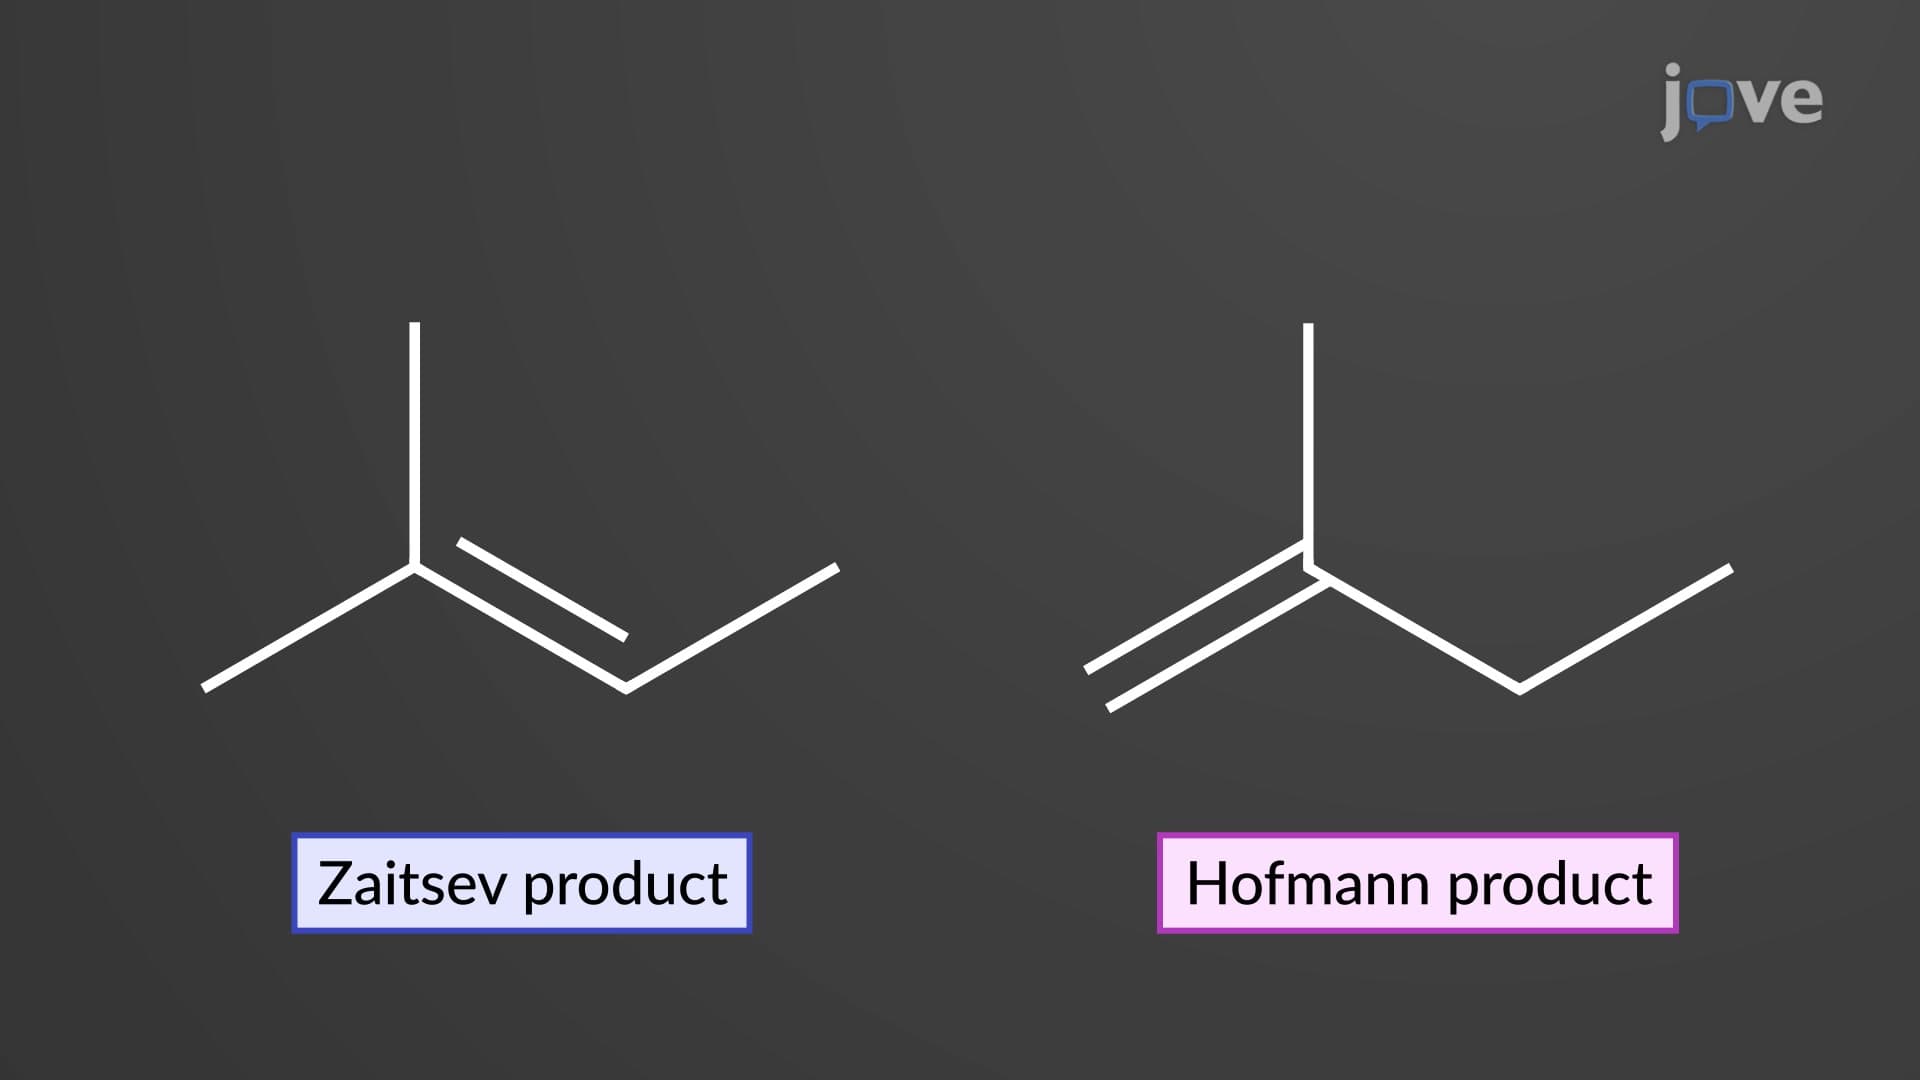
E2 Reaction: Stereochemistry and Regiochemistry
When a substrate with two different β hydrogens undergoes an E2 elimination, the presence of a strong base can yield two regioisomeric alkenes. The more-substituted alkene is the major...
 02:43
E1 Reaction: Stereochemistry and Regiochemistry