Related Experiment Video
Updated: Dec 22, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A basis-set error correction based on density-functional theory for strongly correlated molecular systems
Emmanuel Giner1, Anthony Scemama2, Pierre-François Loos2
1Laboratoire de Chimie Théorique (UMR 7616), Sorbonne Université, CNRS, Paris, France.

Abstract:
We extend to strongly correlated molecular systems the recently introduced basis-set incompleteness correction based on density-functional theory (DFT) [E. Giner et al., J. Chem. Phys. 149, 194301 (2018)]. This basis-set correction relies on a mapping between wave-function calculations in a finite basis set and range-separated DFT (RSDFT) through the definition of an effective non-divergent interaction corresponding to the electron-electron Coulomb interaction projected in the finite basis set. This enables the use of RSDFT-type complementary density functionals to recover the dominant part of the short-range correlation effects missing in this finite basis set. To study both weak and strong correlation regimes, we consider the potential energy curves of the H10, N2, O2, and F2 molecules up to the dissociation limit, and we explore various approximations of complementary functionals fulfilling two key properties: spin-multiplet degeneracy (i.e., independence of the energy with respect to the spin projection Sz) and size consistency. Specifically, we investigate the dependence of the functional on different types of on-top pair densities and spin polarizations. The key result of this study is that the explicit dependence on the on-top pair density allows one to completely remove the dependence on any form of spin polarization without any significant loss of accuracy. Quantitatively, we show that the basis-set correction reaches chemical accuracy on atomization energies with triple-ζ quality basis sets for most of the systems studied here. In addition, the present basis-set incompleteness correction provides smooth potential energy curves along the whole range of internuclear distances.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 03:51
Molecular Orbital Theory II
 01:10
Propagation of Uncertainty from Systematic Error
 01:10
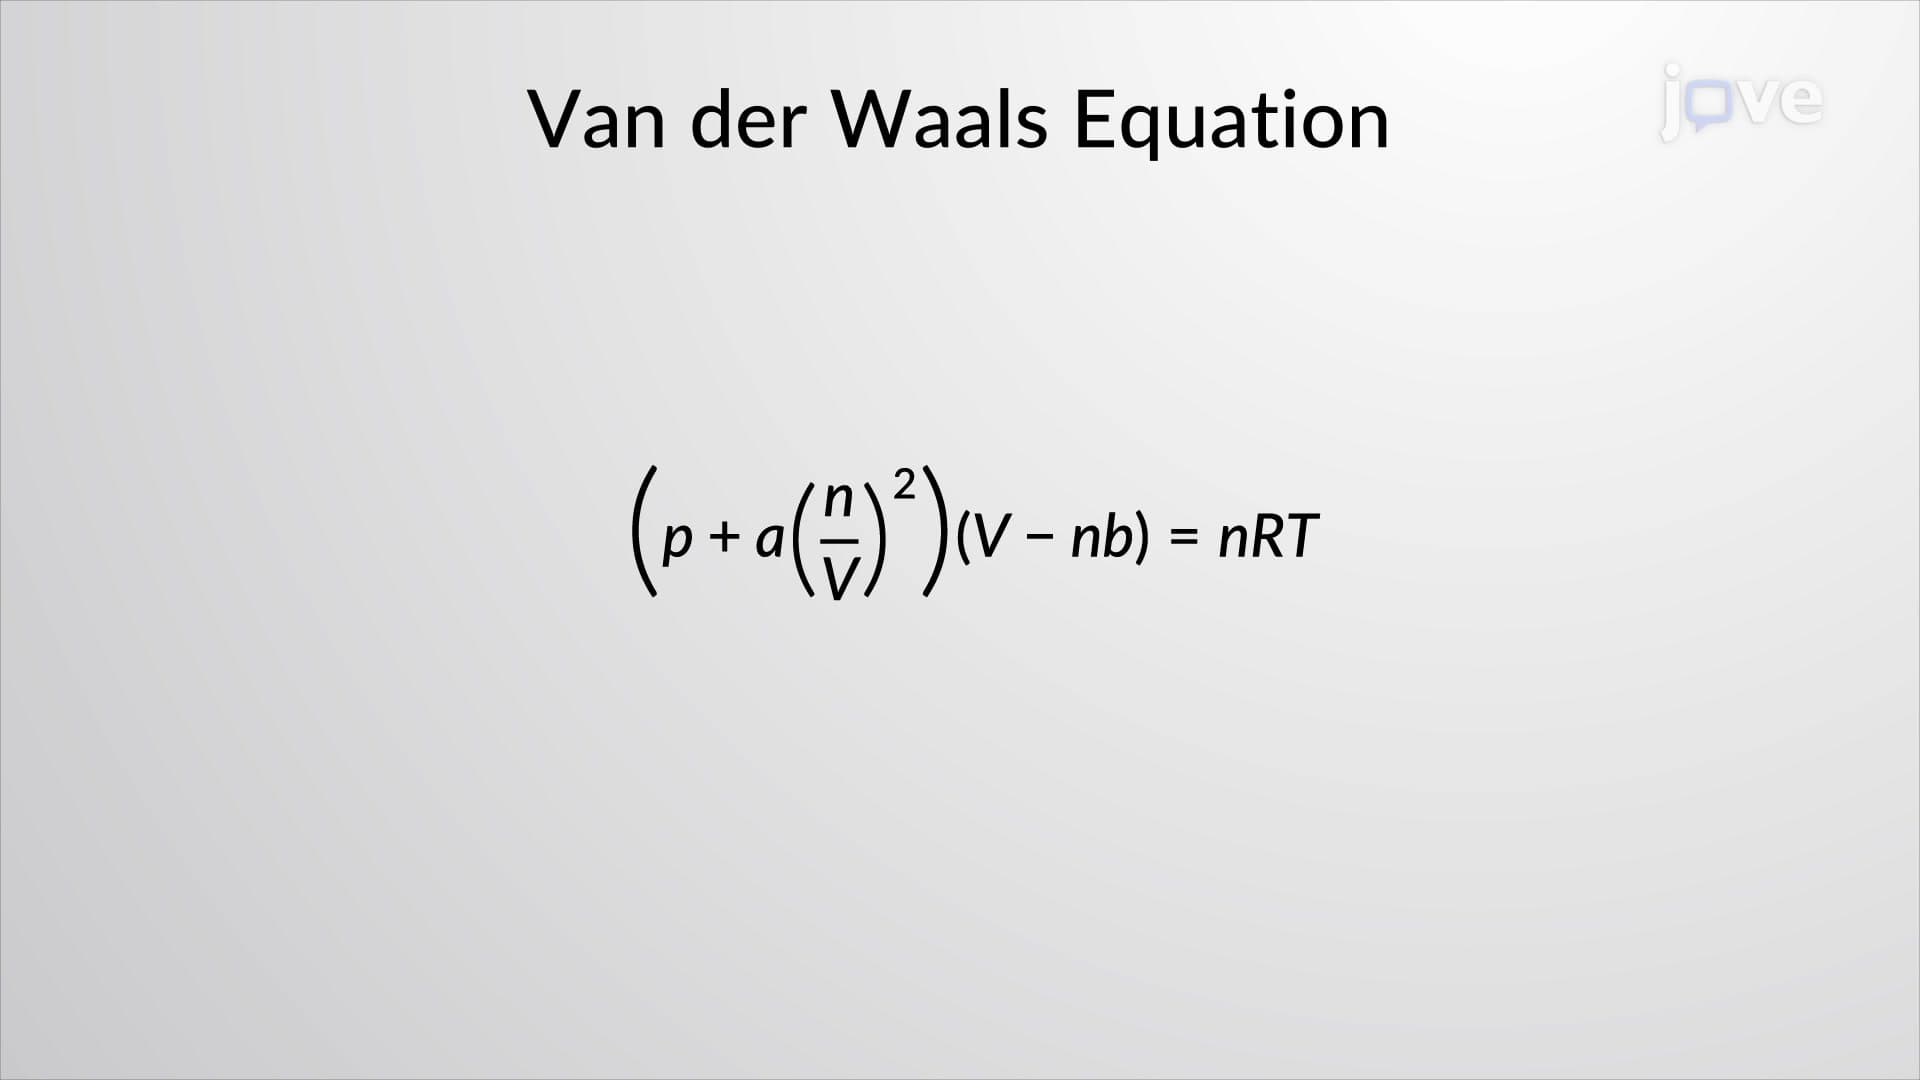
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:14
NMR Spectrometers: Resolution and Error Correction
 02:35
Molecular Orbital Theory I