Related Experiment Video
Updated: Dec 21, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
An efficient hybrid scheme for time dependent density functional theory
Marco Medves1, Luca Sementa2, Daniele Toffoli1
1Dipartimento di Scienze Chimiche e Farmaceutiche, Università di Trieste, Via Giorgieri 1, 34127 Trieste, Italy.

Abstract:
A hybrid approach able to perform Time Dependent Density Functional Theory (TDDFT) simulations with the same accuracy as that of hybrid exchange-correlation (xc-) functionals but at a fraction of the computational cost is developed, implemented, and validated. The scheme, which we name Hybrid Diagonal Approximation (HDA), consists in employing in the response function a hybrid xc-functional (containing a fraction of the non-local Hartree-Fock exchange) only for the diagonal elements of the omega matrix, while the adiabatic local density approximation is employed for the off-diagonal terms. HDA is especially (but not exclusively) advantageous when using Slater type orbital basis sets and allows one to employ them in a uniquely efficient way, as we demonstrate here by implementing HDA in a local version of the Amsterdam Density Functional code. The new protocol is tested on NH3, C6H6, and the [Au25(SCH3)18]- cluster as prototypical cases ranging from small molecules to ligand-protected metal clusters, finding excellent agreement with respect to both full kernel TDDFT simulations and experimental data. Additionally, a specific comparison test between full kernel and HDA is considered at the Casida level on seven other molecular species, which further confirm the accuracy of the approach for all investigated systems. For the [Au25(SCH3)18]- cluster, a speedup by a factor of seven is obtained with respect to the full kernel. The HDA, therefore, promises to provide a quantitative description of the optical properties of medium-sized systems (nanoclusters) at an affordable cost, thanks to its computational efficiency, especially in combination with a complex polarization algorithm version of TDDFT.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:12
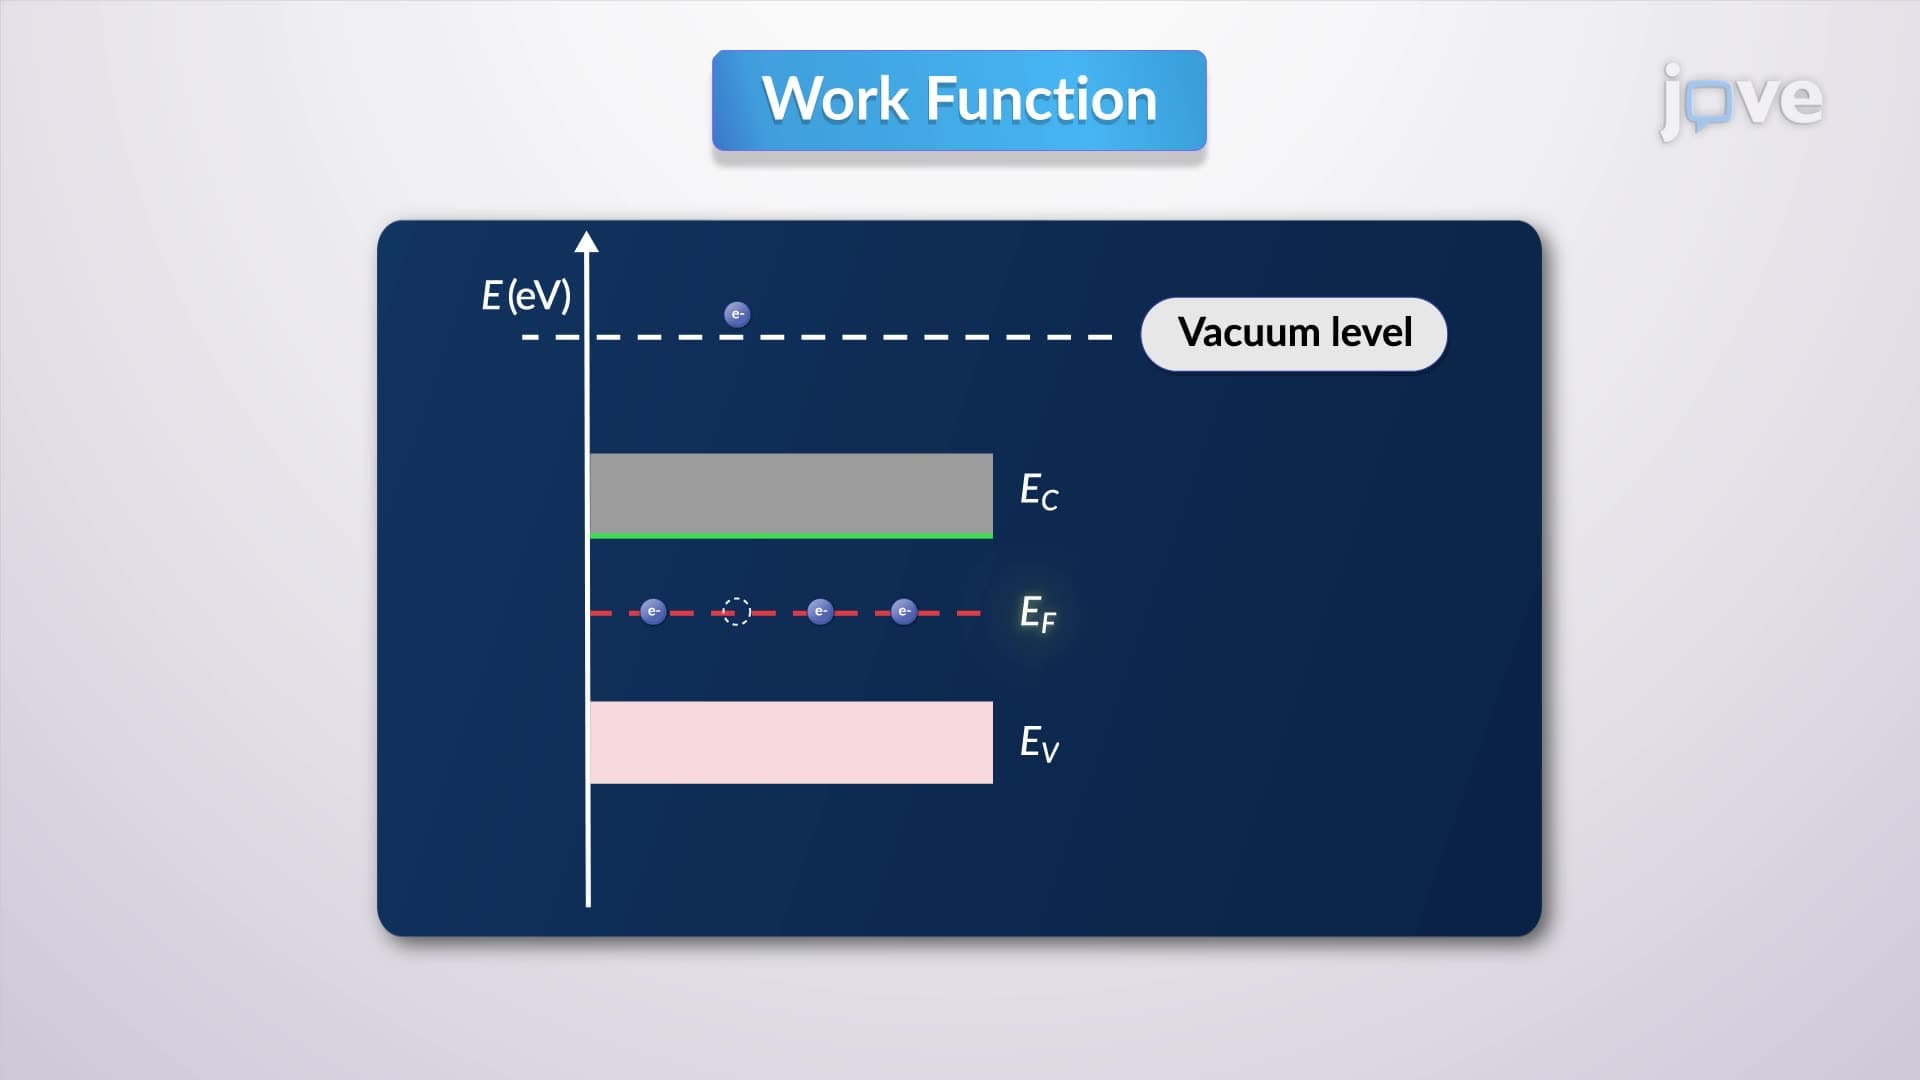
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:23
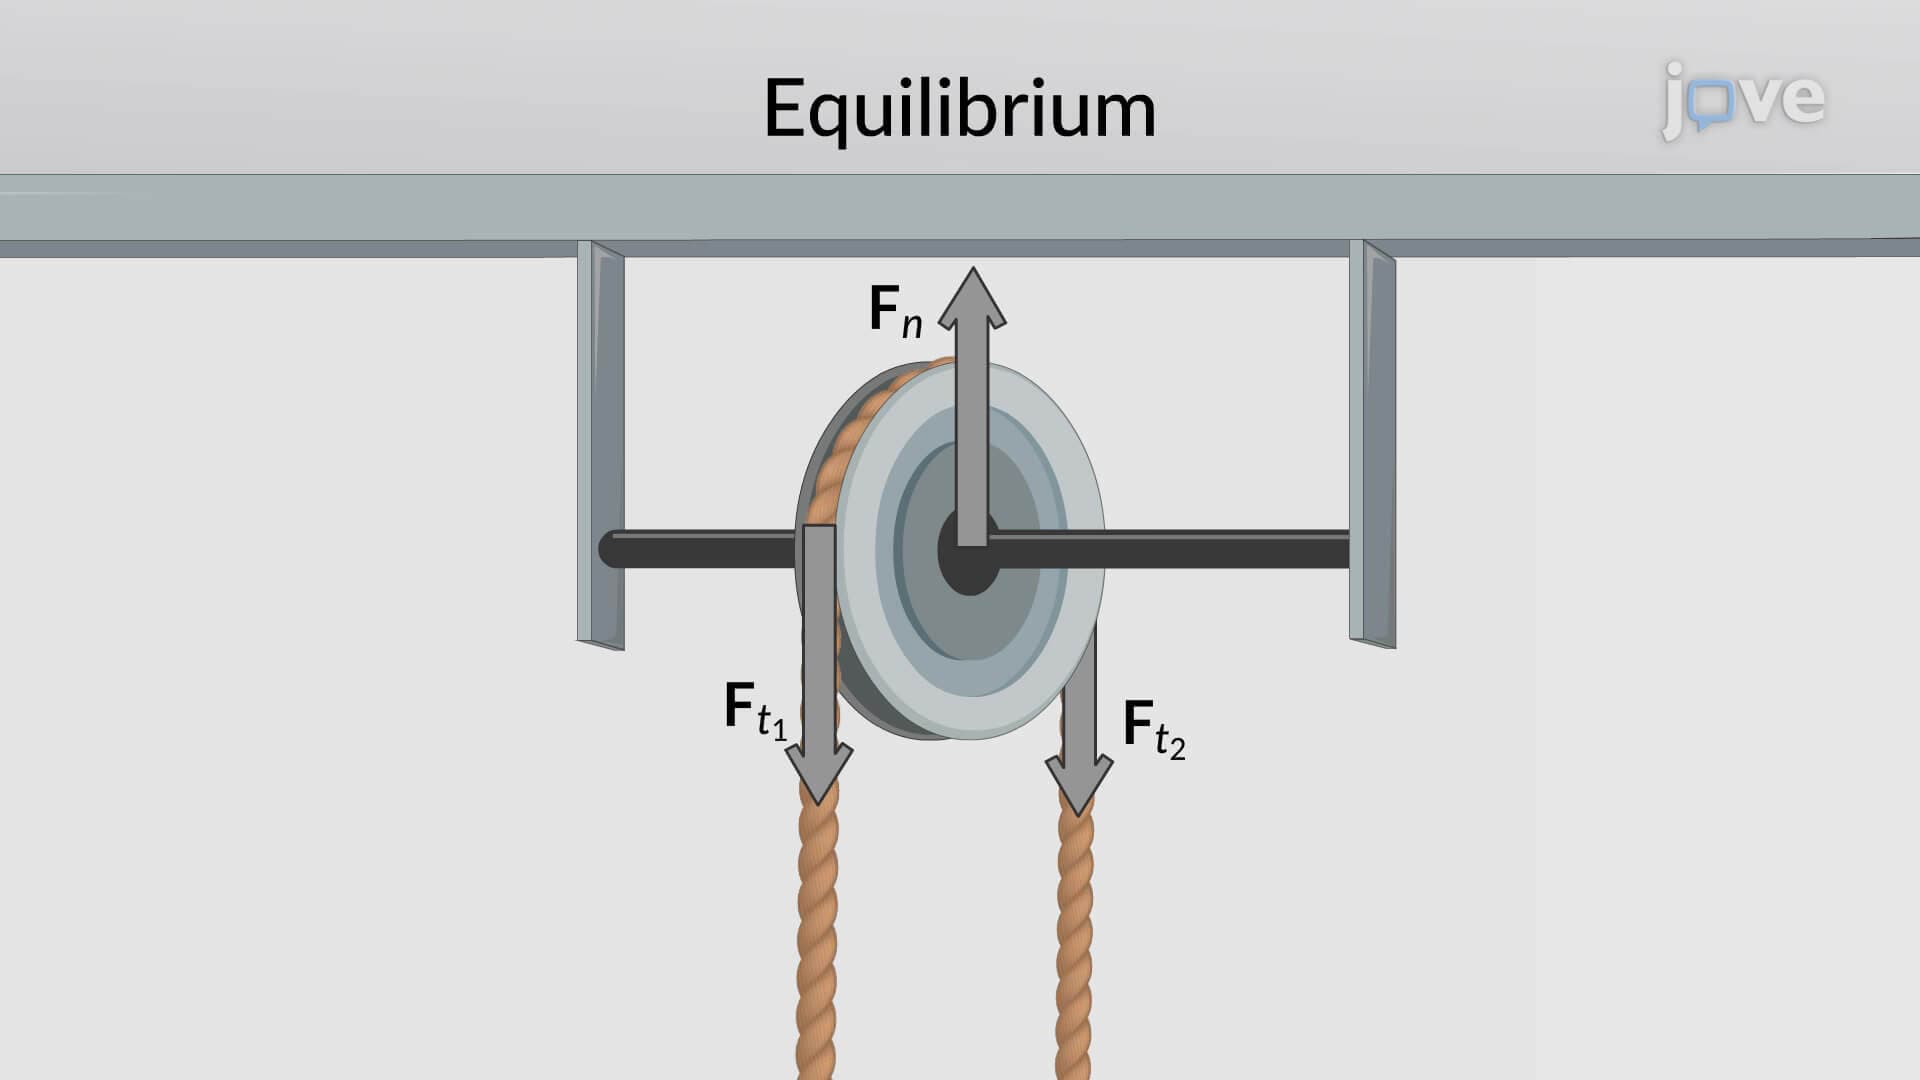
Equilibrium Conditions for a Particle
To understand the concept of equilibrium, let us first consider the forces acting on an object. When different forces act on an object, they can...
 02:45
The Quantum-Mechanical Model of an Atom