Related Experiment Video
Updated: Dec 20, 2025

Analyzing Protein Architectures and Protein-Ligand Complexes by Integrative Structural Mass Spectrometry
Published on: October 15, 2018
Multidimensional Global Optimization and Robustness Analysis in the Context of Protein-Ligand Binding
Negin Forouzesh1, Abhishek Mukhopadhyay2, Layne T Watson1,3,4,5
1Department of Computer Science, Virginia Polytechnic Institute & State University, Blacksburg, Virginia 24061, United States.

Abstract:
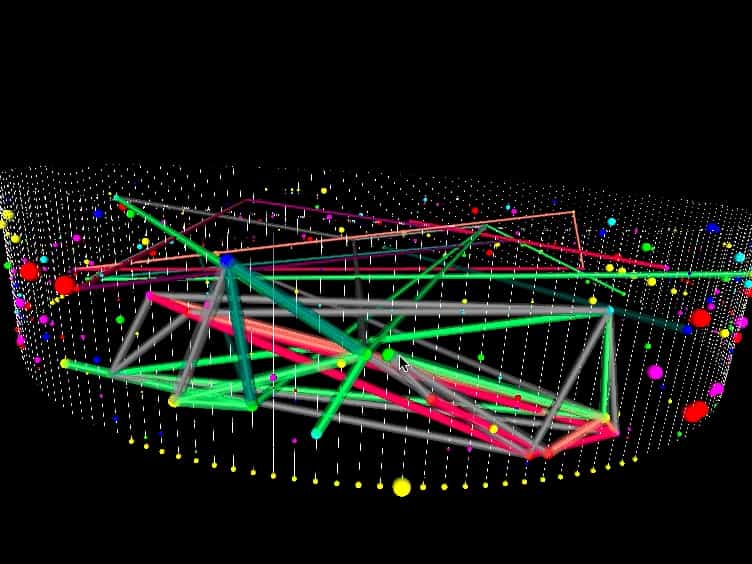
Accuracy of protein-ligand binding free energy calculations utilizing implicit solvent models is critically affected by parameters of the underlying dielectric boundary, specifically, the atomic and water probe radii. Here, a global multidimensional optimization pipeline is developed to find optimal atomic radii specifically for protein-ligand binding calculations in implicit solvent. The computational pipeline has these three key components: (1) a massively parallel implementation of a deterministic global optimization algorithm (VTDIRECT95), (2) an accurate yet reasonably fast generalized Born implicit solvent model (GBNSR6), and (3) a novel robustness metric that helps distinguish between nearly degenerate local minima via a postprocessing step of the optimization. A graph-based "kT-connectivity" approach to explore and visualize the multidimensional energy landscape is proposed: local minima that can be reached from the global minimum without exceeding a given energy threshold (kT) are considered to be connected. As an illustration of the capabilities of the optimization pipeline, we apply it to find a global optimum in the space of just five radii: four atomic (O, H, N, and C) radii and water probe radius. The optimized radii, ρW = 1.37 Å, ρC = 1.40 Å, ρH = 1.55 Å, ρN = 2.35 Å, and ρO = 1.28 Å, lead to a closer agreement of electrostatic binding free energies with the explicit solvent reference than two commonly used sets of radii previously optimized for small molecules. At the same time, the ability of the optimizer to find the global optimum reveals fundamental limits of the common two-dielectric implicit solvation model: the computed electrostatic binding free energies are still almost 4 kcal/mol away from the explicit solvent reference. The proposed computational approach opens the possibility to further improve the accuracy of practical computational protocols for binding free energy calculations.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 07:08
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
01:49Conserved Binding Sites
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
 00:49
Ligand Binding and Linkage
00:49Ligand Binding and Linkage