Related Experiment Video
Updated: Dec 20, 2025

Advanced Imaging of Lung Homing Human Lymphocytes in an Experimental In Vivo Model of Allergic Inflammation Based on Light-sheet Microscopy
Published on: April 16, 2019
ILC2 Lung-Homing in Cystic Fibrosis Patients: Functional Involvement of CCR6 and Impact on Respiratory Failure
Anja Schulz-Kuhnt1, Vicky Greif1, Kai Hildner1
1Department of Medicine 1, University Hospital of Erlangen, Erlangen, Germany.

Abstract:
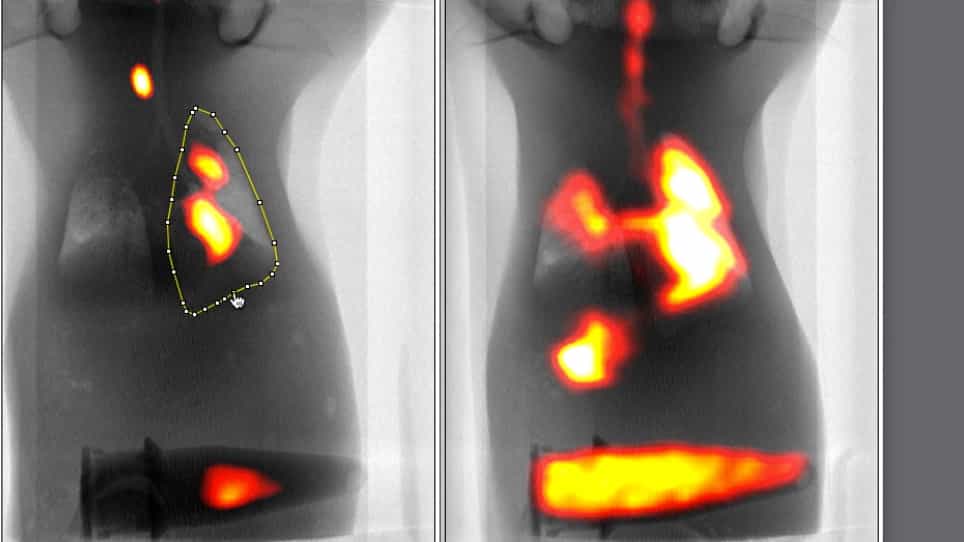
Cystic fibrosis patients suffer from a progressive, often fatal lung disease, which is based on a complex interplay between chronic infections, locally accumulating immune cells and pulmonary tissue remodeling. Although group-2 innate lymphoid cells (ILC2s) act as crucial initiators of lung inflammation, our understanding of their involvement in the pathogenesis of cystic fibrosis remains incomplete. Here we report a marked decrease of circulating CCR6+ ILC2s in the blood of cystic fibrosis patients, which significantly correlated with high disease severity and advanced pulmonary failure, strongly implicating increased ILC2 homing from the peripheral blood to the chronically inflamed lung tissue in cystic fibrosis patients. On a functional level, the CCR6 ligand CCL20 was identified as potent promoter of lung-directed ILC2 migration upon inflammatory conditions in vitro and in vivo using a new humanized mouse model with light-sheet fluorescence microscopic visualization of lung-accumulated human ILC2s. In the lung, blood-derived human ILC2s were able to augment local eosinophil and neutrophil accumulation and induced a marked upregulation of pulmonary type-VI collagen expression. Studies in primary human lung fibroblasts additionally revealed ILC2-derived IL-4 and IL-13 as important mediators of this type-VI collagen-inducing effect. Taken together, the here acquired results suggest that pathologically increased CCL20 levels in cystic fibrosis airways induce CCR6-mediated lung homing of circulating human ILC2s. Subsequent ILC2 activation then triggers local production of type-VI collagen and might thereby drive extracellular matrix remodeling potentially influencing pulmonary tissue destruction in cystic fibrosis patients. Thus, modulating the lung homing capacity of circulating ILC2s and their local effector functions opens new therapeutic avenues for cystic fibrosis treatment.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation