Related Experiment Video
Updated: Dec 20, 2025
Drug-induced Sensitization of Adenylyl Cyclase: Assay Streamlining and Miniaturization for Small Molecule and siRNA Screening Applications
Published on: January 27, 2014
In Silico Drug Design for Purinergic GPCRs: Overview on Molecular Dynamics Applied to Adenosine and P2Y Receptors
Veronica Salmaso1, Kenneth A Jacobson1
1Molecular Recognition Section, Laboratory of Bioorganic Chemistry, National Institute of Diabetes and Digestive and Kidney Diseases, National Institutes of Health, Bethesda, MD 20892, USA.

Abstract:
Molecular modeling has contributed to drug discovery for purinergic GPCRs, including adenosine receptors (ARs) and P2Y receptors (P2YRs). Experimental structures and homology modeling have proven to be useful in understanding and predicting structure activity relationships (SAR) of agonists and antagonists. This review provides an excursus on molecular dynamics (MD) simulations applied to ARs and P2YRs. The binding modes of newly synthesized A1AR- and A3AR-selective nucleoside derivatives, potentially of use against depression and inflammation, respectively, have been predicted to recapitulate their SAR and the species dependence of A3AR affinity. P2Y12R and P2Y1R crystallographic structures, respectively, have provided a detailed understanding of the recognition of anti-inflammatory P2Y14R antagonists and a large group of allosteric and orthosteric antagonists of P2Y1R, an antithrombotic and neuroprotective target. MD of A2AAR (an anticancer and neuroprotective target), A3AR, and P2Y1R has identified microswitches that are putatively involved in receptor activation. The approach pathways of different ligands toward A2AAR and P2Y1R binding sites have also been explored. A1AR, A2AAR, and A3AR were utilizes to study allosteric phenomena, but locating the binding site of structurally diverse allosteric modulators, such as an A3AR enhancer LUF6000, is challenging. Ligand residence time, a predictor of in vivo efficacy, and the structural role of water were investigated through A2AAR MD simulations. Thus, new MD and other modeling algorithms have contributed to purinergic GPCR drug discovery.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 08:21
Monitoring GPCR-β-arrestin1/2 Interactions in Real Time Living Systems to Accelerate Drug Discovery
Published on: June 28, 2019
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:15
G Protein-coupled Receptors
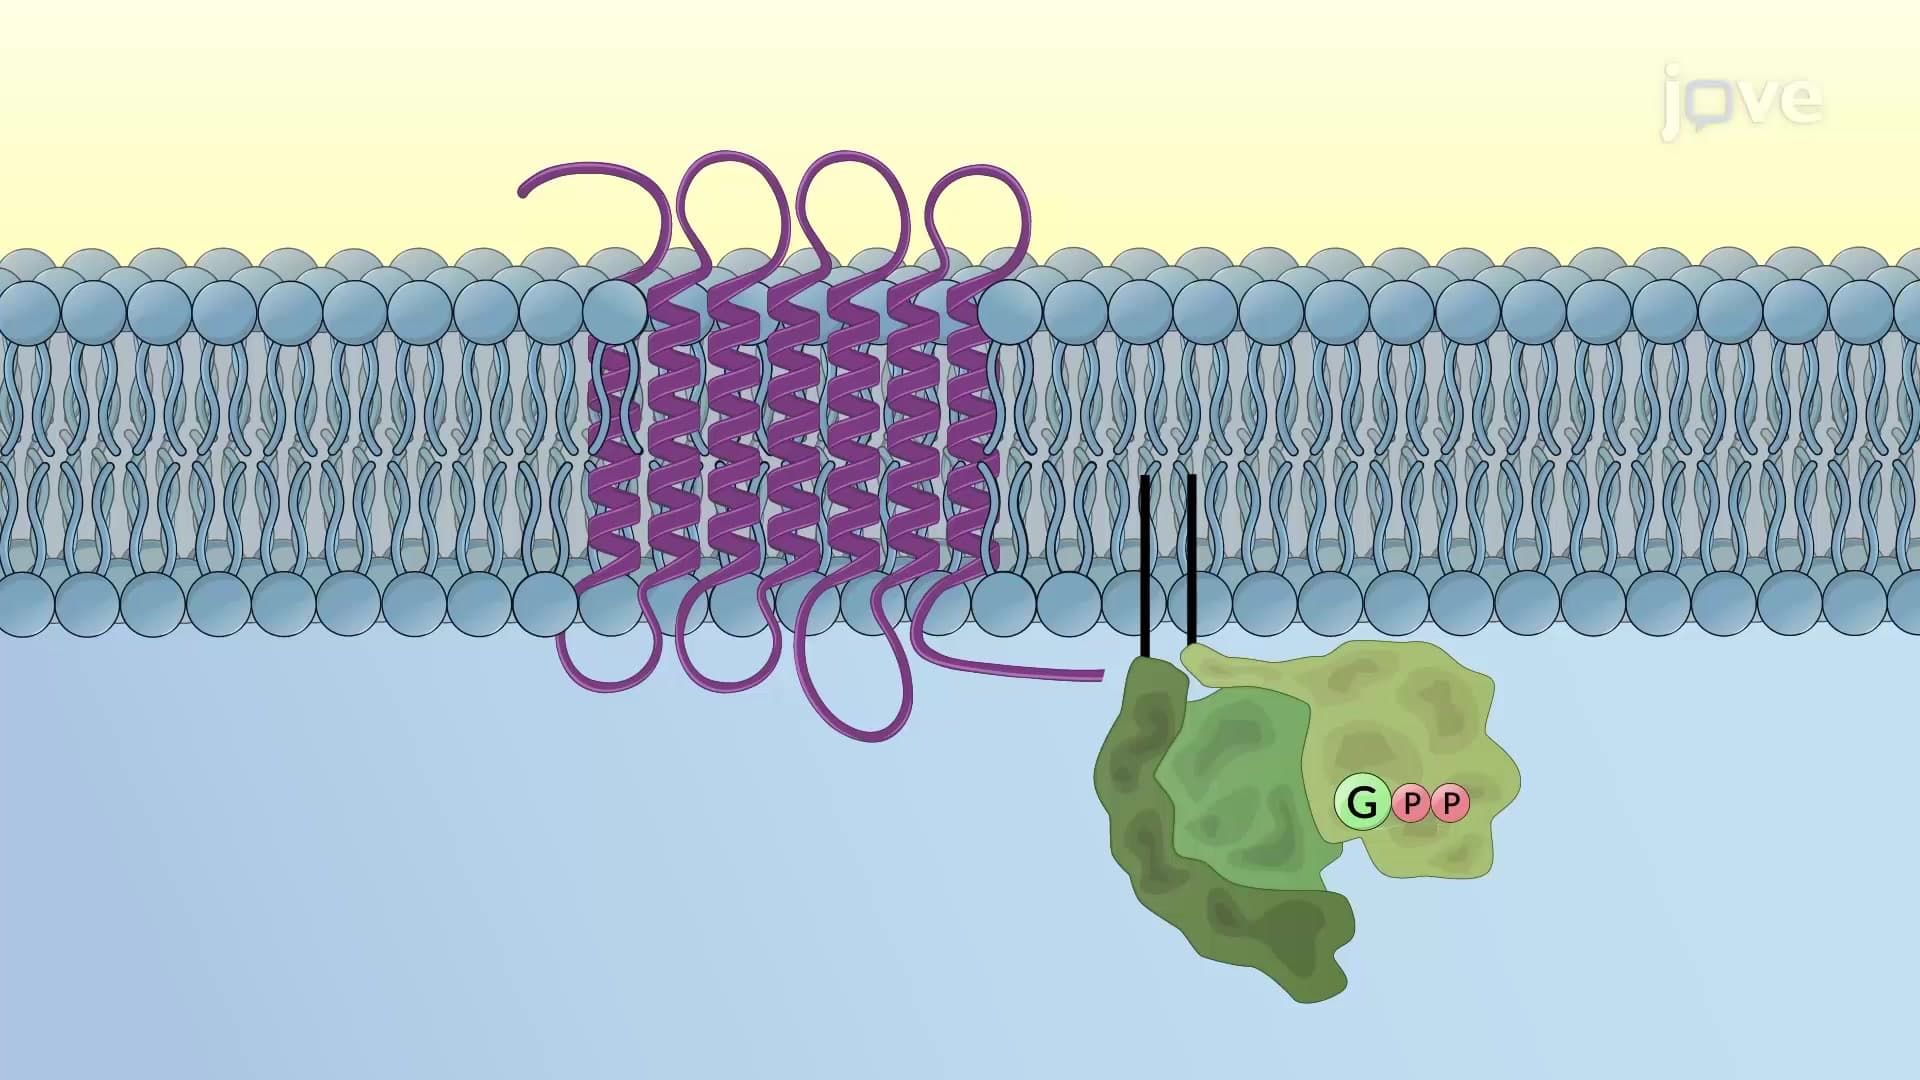
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:09
GPCRs Regulate Adenylyl Cylase Activity
 01:26
Targets for Drug Action: Overview
Receptors are either membrane-spanning or intracellular proteins, which upon binding a ligand, get activated and transmit the signal downstream to elicit a response. Drugs bind receptors, either mimicking the action of endogenous ligands or blocking the receptor activity to bring about a modified response. Nearly 35% of approved drugs target the G...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
 01:27
Transducer Mechanism: Enzyme-Linked Receptors
Major types that are helpful drug targets include: