Related Experiment Video
Updated: Dec 20, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Improved Basis-Set Incompleteness Potentials for Accurate Density-Functional Theory Calculations in Large Systems
A Otero-de-la-Roza1, Gino A DiLabio2,3
1Departamento de Quı́mica Fı́sica y Analítica and MALTA-Consolider Team, Facultad de Quı́mica, Universidad de Oviedo, 33006 Oviedo, Spain.

Abstract:
The accurate calculation of chemical properties using density-functional theory (DFT) requires the use of a nearly complete basis set. In chemical systems involving hundreds to thousands of atoms, the cost of the calculations place practical limitations on the number of basis functions that can be used. Therefore, in most practical applications of DFT to large systems, there exists a basis-set incompleteness error (BSIE). In this article, we present the next iteration of the basis-set incompleteness potentials (BSIPs), one-electron potentials designed to correct for basis-set incompleteness error. The ultimate goal associated with the development of BSIPs is to allow the calculation of molecular properties using DFT with near-complete-basis-set results at a computational cost that is similar to a small basis set calculation. In this work, we develop BSIPs for 10 atoms in the first and second rows (H, B-F, Si-Cl) and 15 common basis sets of the Pople, Dunning, Karlsruhe, and Huzinaga types. Our new BSIPs are constructed to minimize BSIE in the calculation of reaction energies, barrier heights, noncovalent binding energies, and intermolecular distances. The BSIPs were obtained using a training set of 15 944 data points. The fitting approach employed a regularized linear least-squares method with variable selection (the LASSO method), which results in a much better fit to the training data than our previous BSIPs while, at the same time, reducing the computational cost of BSIP development. The proposed BSIPs are tested on various benchmark sets and demonstrate excellent performance in practice. Our new BSIPs are also transferable; i.e., they can be used to correct BSIE in calculations that employ density functionals other than the one used in the BSIP development (B3LYP). Finally, BSIPs can be used in any quantum chemistry program that have implemented effective-core potentials without changes to the software.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I
 01:10
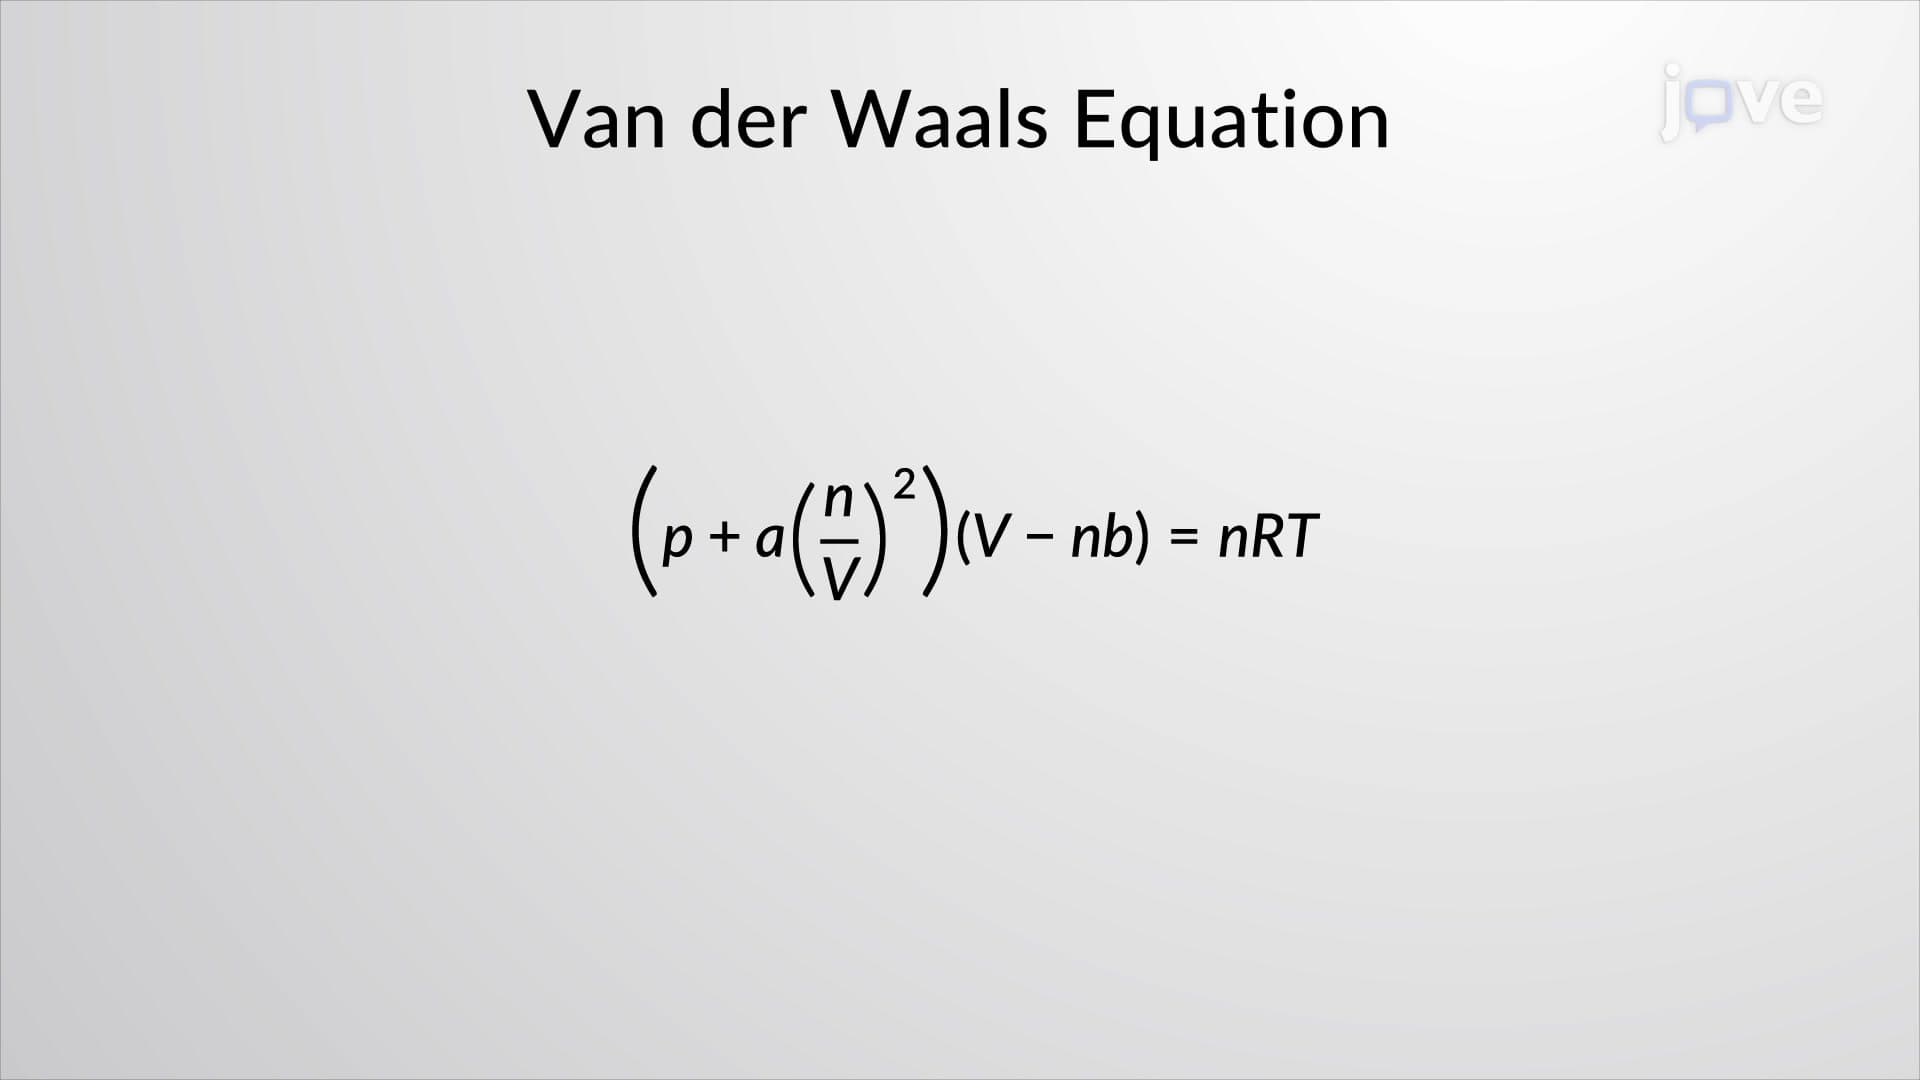
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...