Related Experiment Video
Updated: Dec 20, 2025
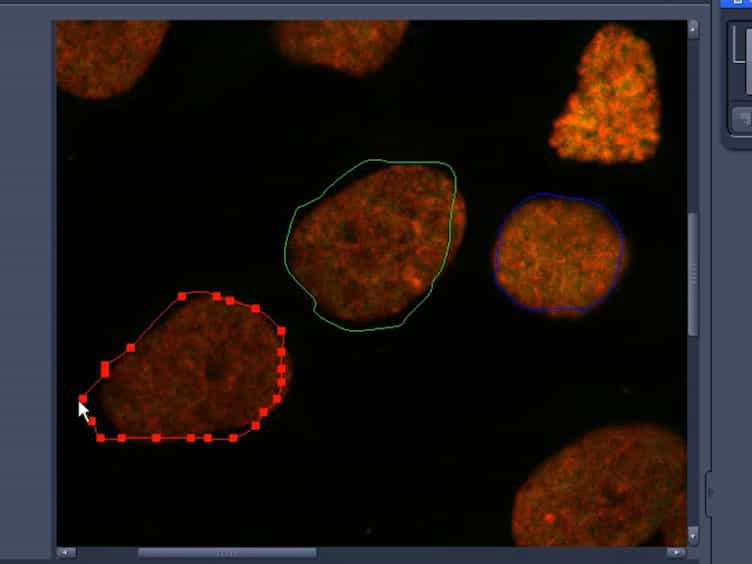

Immunostaining for DNA Modifications: Computational Analysis of Confocal Images
Published on: September 7, 2017
A genome-scale map of DNA methylation turnover identifies site-specific dependencies of DNMT and TET activity
Paul Adrian Ginno1, Dimos Gaidatzis1,2, Angelika Feldmann1,3
1Friedrich Miescher Institute for Biomedical Research, Basel, Switzerland.

Abstract:
DNA methylation is considered a stable epigenetic mark, yet methylation patterns can vary during differentiation and in diseases such as cancer. Local levels of DNA methylation result from opposing enzymatic activities, the rates of which remain largely unknown. Here we developed a theoretical and experimental framework enabling us to infer methylation and demethylation rates at 860,404 CpGs in mouse embryonic stem cells. We find that enzymatic rates can vary as much as two orders of magnitude between CpGs with identical steady-state DNA methylation. Unexpectedly, de novo and maintenance methylation activity is reduced at transcription factor binding sites, while methylation turnover is elevated in transcribed gene bodies. Furthermore, we show that TET activity contributes substantially more than passive demethylation to establishing low methylation levels at distal enhancers. Taken together, our work unveils a genome-scale map of methylation kinetics, revealing highly variable and context-specific activity for the DNA methylation machinery.
Related Concept Videos
 01:37
Epigenetic Regulation
X-chromosome...
 01:46
Epigenetic Regulation