Related Experiment Video
Updated: Dec 19, 2025

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Effect of Protein Flexibility from Coarse-Grained Elastic Network Parameterizations on the Calculation of Free Energy
Hugo A L Filipe1, Margarida I M Esteves1, César A Henriques2
1Coimbra Chemistry Centre, Dept. of Chemistry, University of Coimbra, Rua Larga, Coimbra 3004-535, Portugal.

Abstract:
The characterization of the affinity and binding mechanism of specific molecules to a protein active site is scientifically and industrially relevant for many applications. In principle, this information can be obtained using molecular dynamics (MD) simulations by calculating the free energy profile of the process. However, this is a computationally demanding calculation. Currently, coarse-grained (CG) force fields are very well implemented for MD simulations of biomolecular systems. These computationally efficient force fields are a major advantage to the study of large model systems and/or those requiring long simulation times. The Martini model is currently one of the most popular CG force fields for these systems. For the specific case of protein simulations, to correctly maintain the macromolecular three-dimensional structure, the Martini model needs to include an elastic network (EN). In this work, the effect of protein flexibility, as induced by three EN models compatible with the Martini force field, was tested on the calculation of free energy profiles for protein-ligand binding. The EN models used were ElNeDyn, GoMartini, and GEN. The binding of triolein (TOG) and triacetin (TAG) to a lipase protein (thermomyces lanuginosa lipase-TLL) was used as a case study. The results show that inclusion of greater flexibility in the CG parameterization of proteins is of high importance in the calculation of the free energy profiles of protein-ligand systems. However, care must be taken in order to avoid unjustified large protein deformations. In addition, due to molecular flexibility there may be no absolute need for the center of the ligand to reach the center of the protein-binding site. The calculation of the energy profile to a distance of about 0.5 nm from the active site center can be sufficient to differentiate the affinity of different ligands to a protein.
Related Concept Videos
 02:18
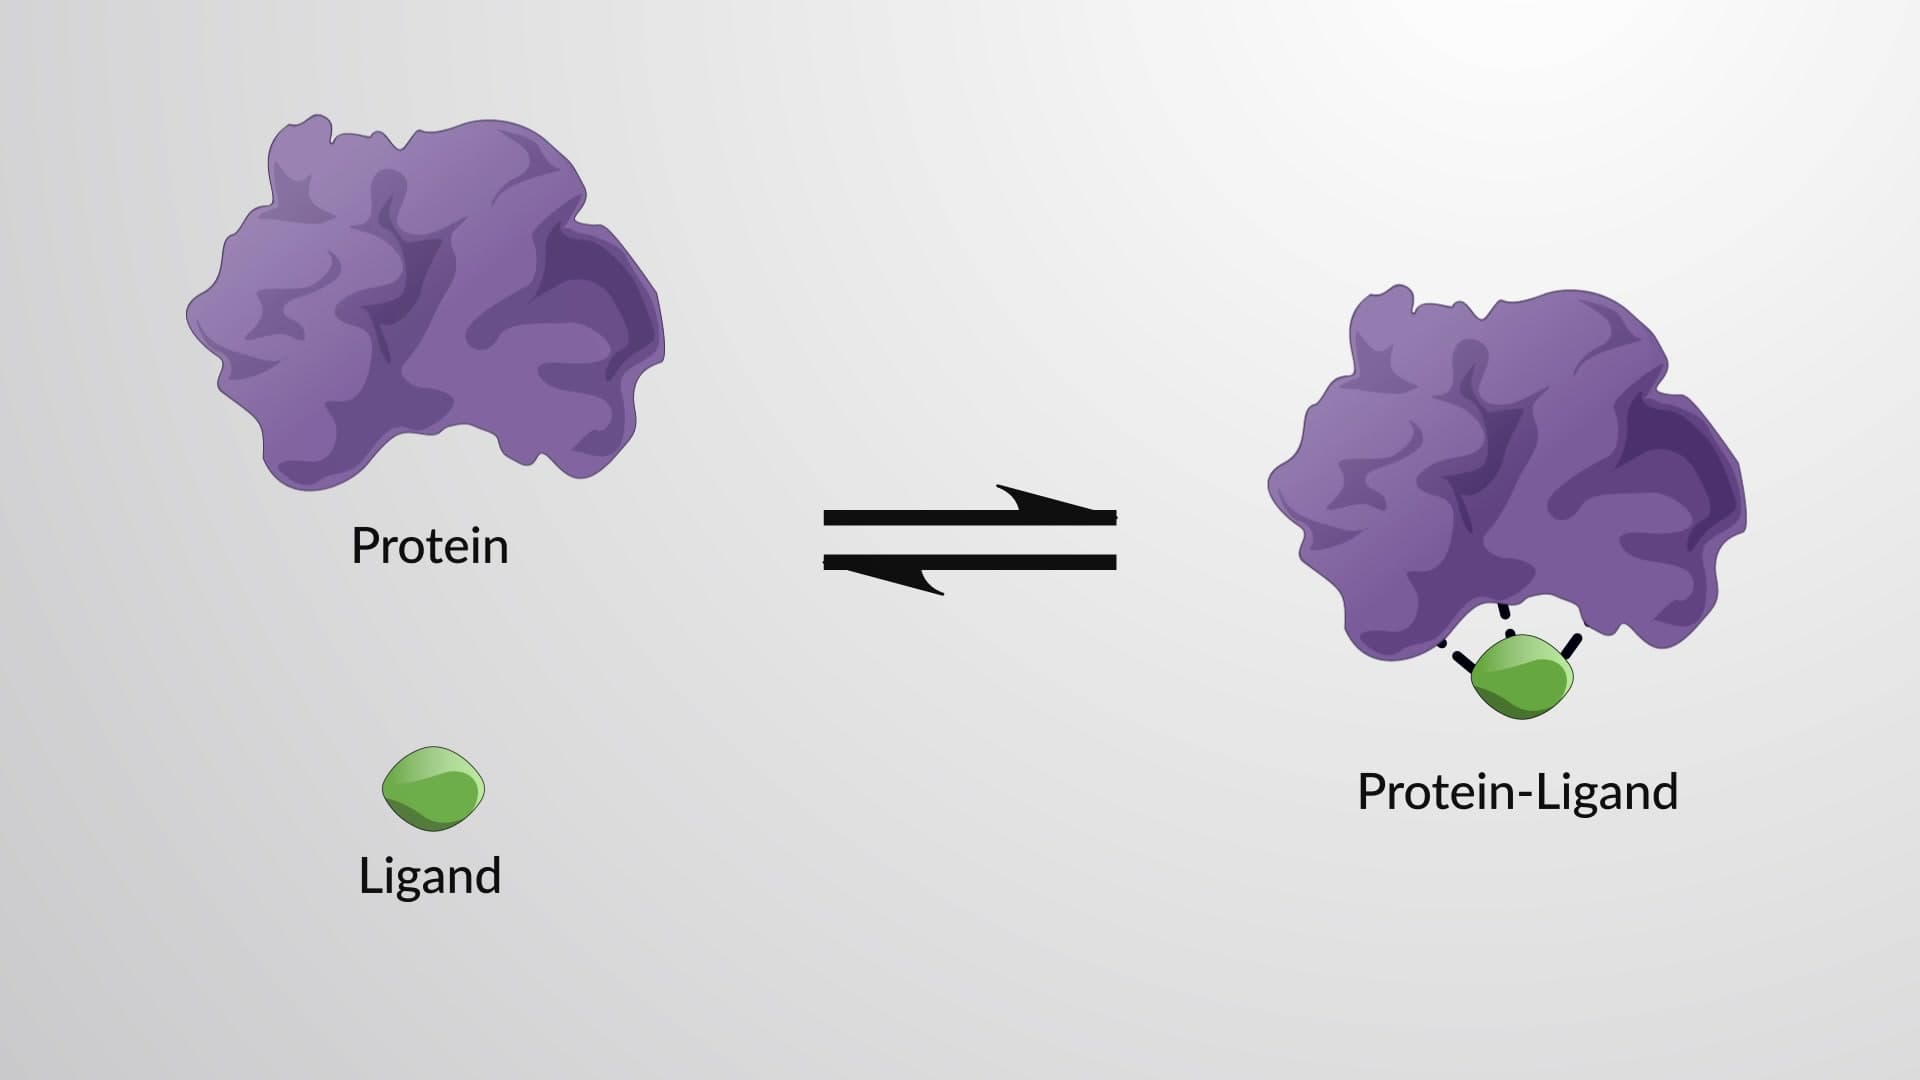
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 00:49
Ligand Binding and Linkage