In silico prediction of potential inhibitors for the main protease of SARS-CoV-2 using molecular docking and dynamics simulation based drug-repurposing
Related Experiment Video
Updated: Dec 17, 2025
Pharmacophore Modeling for Targets with Extensive Ligand Libraries: A Case Study on SARS-CoV-2 Mpro
Published on: September 26, 2025
Yogesh Kumar1, Harvijay Singh2, Chirag N Patel3
1Department of Metabolic & Structural Biology, CSIR-Central Institute of Medicinal & Aromatic Plants, Lucknow 226015, India; Department of Medicine, University Medical Center Hamburg-Eppendorf, 20246 Hamburg, Germany.

Background:
The rapidly enlarging COVID-19 pandemic caused by the novel SARS-corona virus-2 is a global public health emergency of an unprecedented level. Unfortunately no treatment therapy or vaccine is yet available to counter the SARS-CoV-2 infection, which substantiates the need to expand research efforts in this direction. The indispensable function of the main protease in virus replication makes this enzyme a promising target for inhibitors screening and drug discovery to treat novel coronavirus infection. The recently concluded α-ketoamide ligand-bound X-ray crystal structure of SARS-CoV-2 Mpro (PDB ID: 6Y2F) from Zhang et al. has revealed the potential inhibitor binding mechanism and the molecular determinants responsible for substrate binding.
Methods:
For the study, we have targeted the SARS-CoV-2 Mpro for the screening of FDA approved antiviral drugs and carried out molecular docking based virtual screening. Further molecular dynamic simulation studies of the top three selected drugs carried out to investigated for their binding affinity and stability in the SARS-CoV-2 Mpro active site. The phylogenetic analysis was also performed to know the relatedness between the SARS-CoV-2 genomes isolated from different countries.
Results:
The phylogenetic analysis of the SARS-CoV-2 genome reveals that the virus is closely related to the Bat-SL-CoV and does not exhibit any divergence at the genomic level. Molecular docking studies revealed that among the 77 drugs, screened top ten drugs shows good binding affinities, whereas the top three drugs: Lopinavir-Ritonavir, Tipranavir, and Raltegravir were undergone for molecular dynamics simulation studies for their conformational stability in the active site of the SARS-CoV-2 Mpro protein.
Conclusions:
In the present study among the library of FDA approved antiviral drugs, the top three inhibitors Lopinavir-Ritonavir, Tipranavir, and Raltegravir show the best molecular interaction with the main protease of SARS-CoV-2. However, the in-vitro efficacy of the drug molecules screened in this study further needs to be corroborated by carrying out a biochemical and structural investigation.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Related Concept Videos
 02:04
Protein-protein Interfaces
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:16
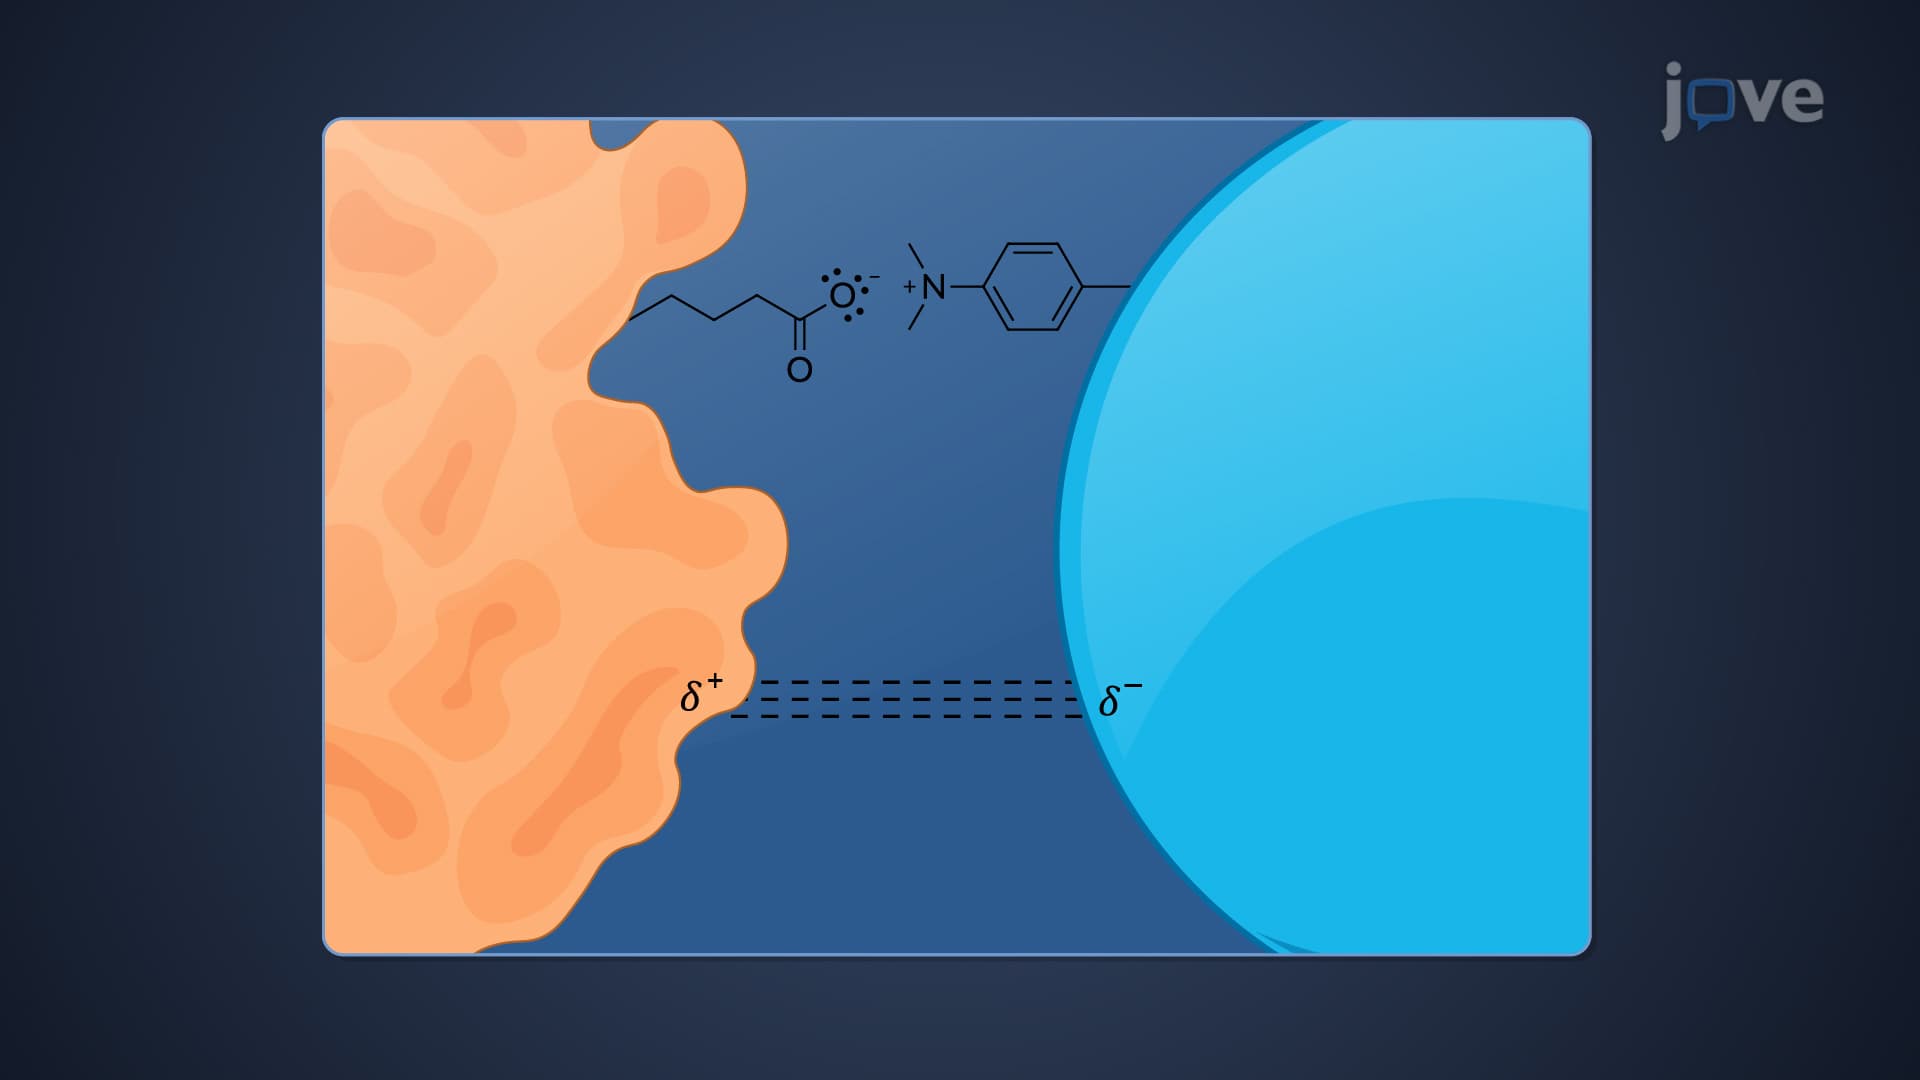
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...