Related Experiment Video
Updated: Dec 17, 2025
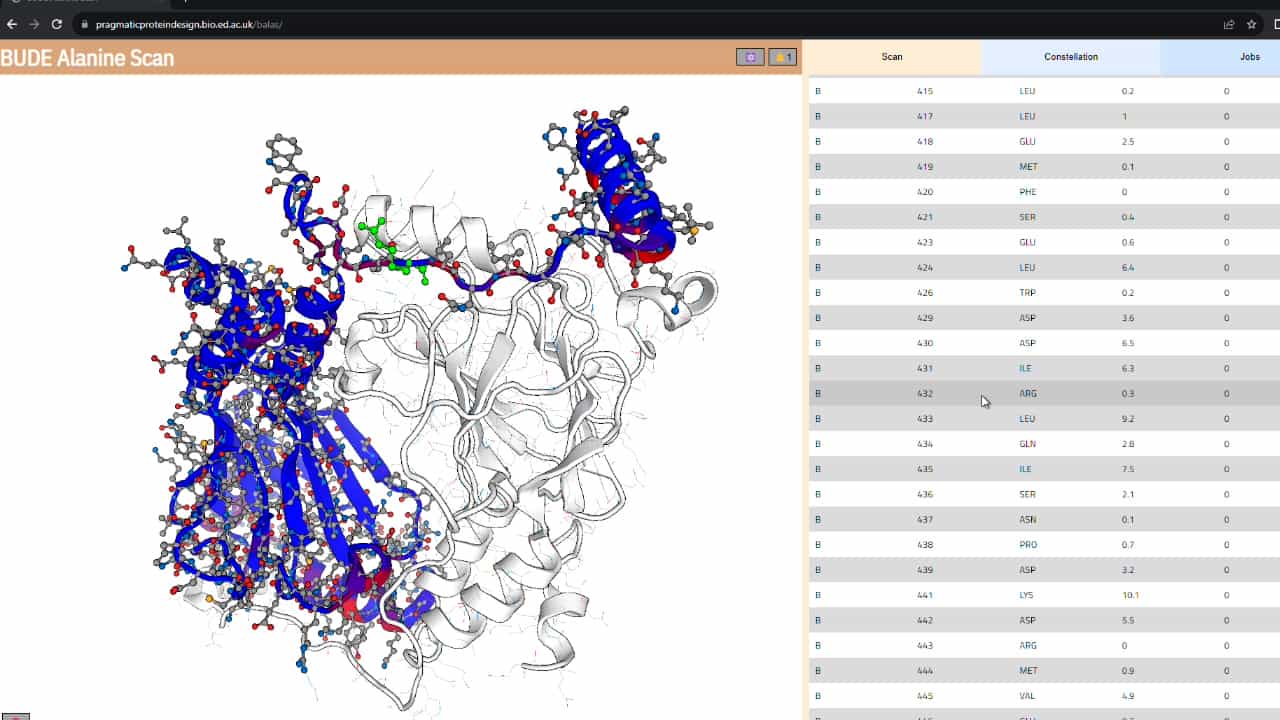
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Computationally predicting binding affinity in protein-ligand complexes: free energy-based simulations and machine
Debby D Wang1, Mengxu Zhu2, Hong Yan3
1School of Medical Instrument and Food Engineering, University of Shanghai for Science and Technology.

Abstract:
Accurately predicting protein-ligand binding affinities can substantially facilitate the drug discovery process, but it remains as a difficult problem. To tackle the challenge, many computational methods have been proposed. Among these methods, free energy-based simulations and machine learning-based scoring functions can potentially provide accurate predictions. In this paper, we review these two classes of methods, following a number of thermodynamic cycles for the free energy-based simulations and a feature-representation taxonomy for the machine learning-based scoring functions. More recent deep learning-based predictions, where a hierarchy of feature representations are generally extracted, are also reviewed. Strengths and weaknesses of the two classes of methods, coupled with future directions for improvements, are comparatively discussed.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
 02:18
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 02:04
Protein-protein Interfaces