Protein Binding Pocket Optimization for Virtual High-Throughput Screening (vHTS) Drug Discovery
Related Experiment Video
Updated: Dec 17, 2025
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
Dimitris Gazgalis1, Mehreen Zaka1,2, Bilal Haider Abbasi2
1Department of Pharmaceutical Sciences, Northeastern University School of Pharmacy, Boston, Massachusetts 02115, United States.

Abstract:
The virtual high-throughput screening (vHTS) approach has been widely used for large database screening to identify potential lead compounds for drug discovery. Due to its high computational demands, docking that allows receptor flexibility has been a challenging problem for virtual screening. Therefore, the selection of protein target conformations is crucial to produce useful vHTS results. Since only a single protein structure is used to screen large databases in most vHTS studies, the main challenge is to reduce false negative rates in selecting compounds for in vitro tests. False negatives are most likely to occur when using apo structures or homology models of protein targets due to the small volume of the binding pocket formed by incorrect side-chain conformations. Even holo protein structures can exhibit high false negative rates due to ligand-induced fit effects, since the shape of the binding pocket highly depends on its bound ligand. To reduce false negative rates and improve success rates for vHTS in drug discovery, we have developed a new Monte Carlo-based approach that optimizes the binding pocket of protein targets. This newly developed Monte Carlo pocket optimization (MCPO) approach was assessed on several datasets showing promising results. The binding pocket optimization approach could be a useful tool for vHTS-based drug discovery, especially in cases when only apo structures or homology models are available.
More Related Videos

Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
01:49Conserved Binding Sites
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:04
Protein-protein Interfaces
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:16
Protein-Drug Binding: Mechanism and Kinetics
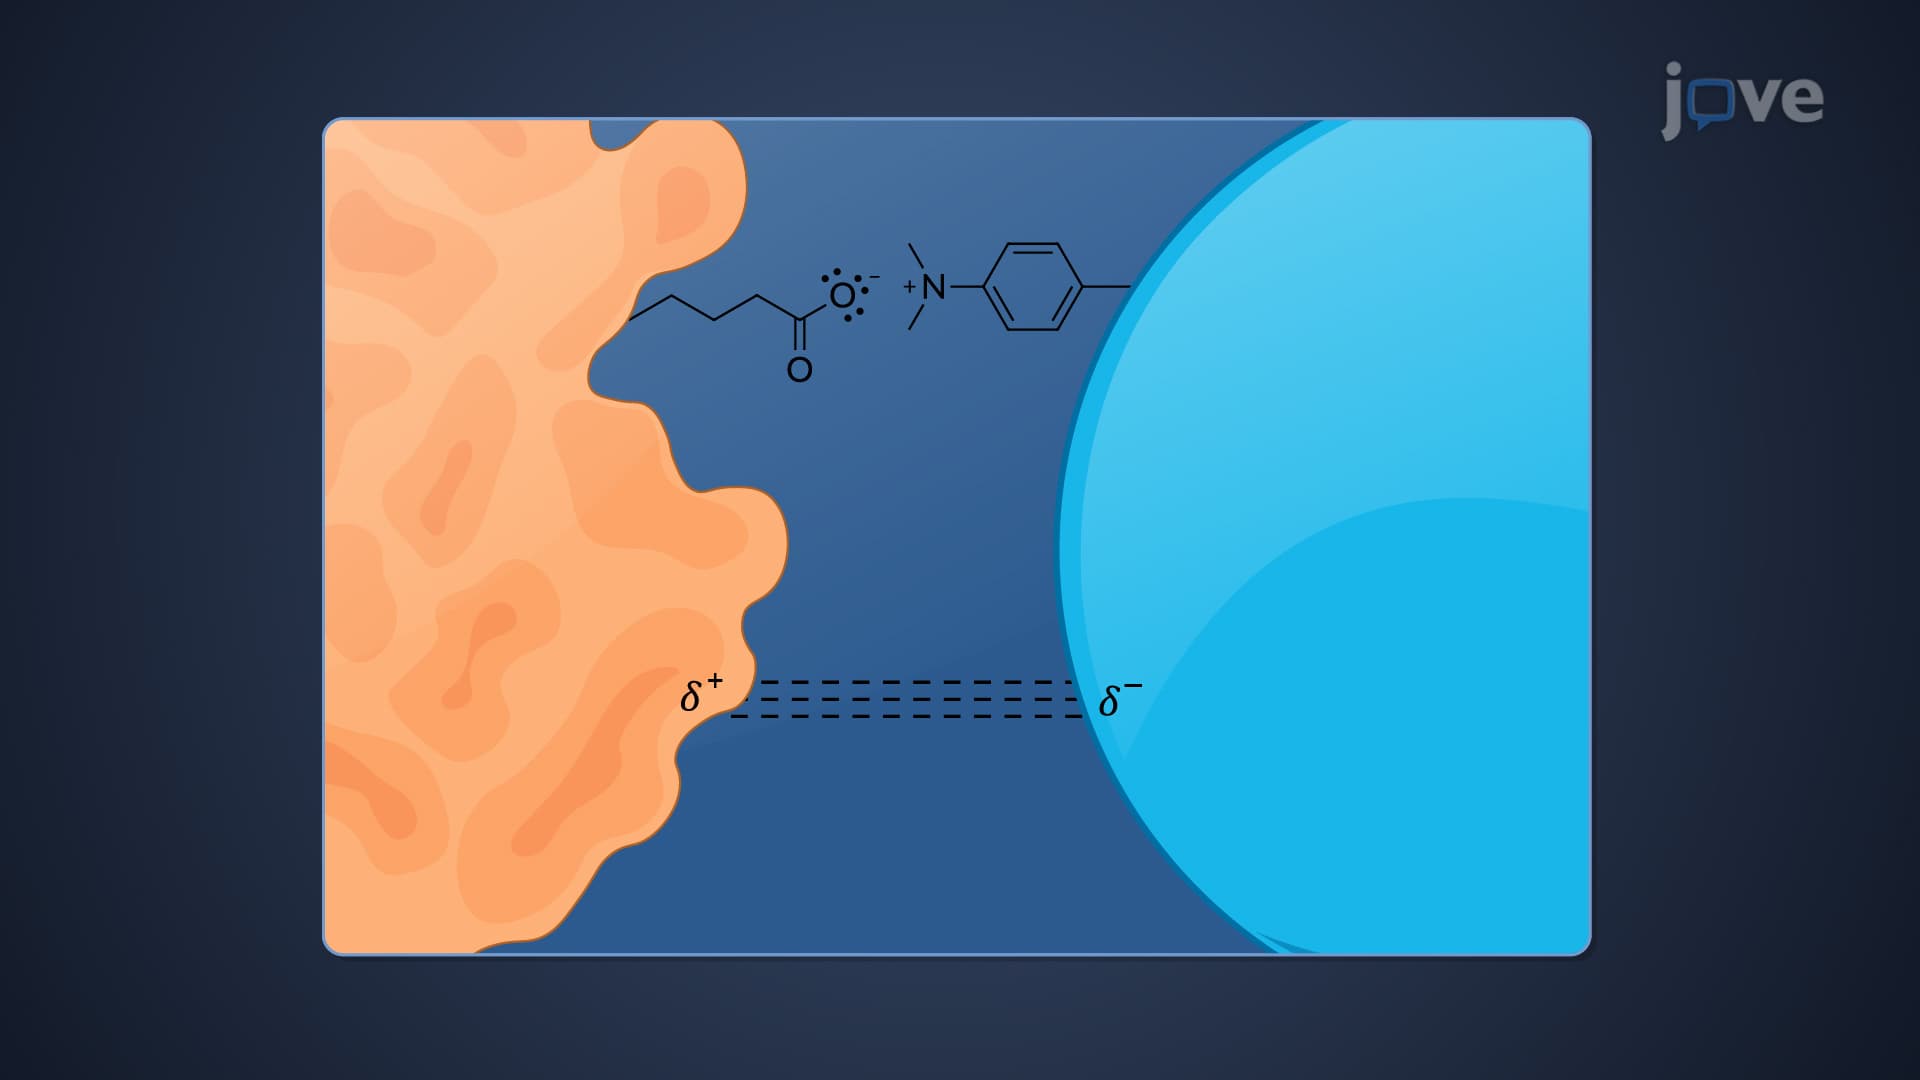
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...