Related Experiment Video
Updated: Dec 15, 2025

Comparing the Affinity of GTPase-binding Proteins using Competition Assays
Published on: October 8, 2015
The molecular basis for immune dysregulation by the hyperactivated E62K mutant of the GTPase RAC2
Megan E Arrington1, Brenda Temple2, Antje Schaefer3
1Department of Chemistry, University of North Carolina, Chapel Hill, North Carolina, USA.

Abstract:
The RAS-related C3 botulinum toxin substrate 2 (RAC2) is a member of the RHO subclass of RAS superfamily GTPases required for proper immune function. An activating mutation in a key switch II region of RAC2 (RAC2E62K) involved in recognizing modulatory factors and effectors has been identified in patients with common variable immune deficiency. To better understand how the mutation dysregulates RAC2 function, we evaluated the structure and stability, guanine nucleotide exchange factor (GEF) and GTPase-activating protein (GAP) activity, and effector binding of RAC2E62K Our findings indicate the E62K mutation does not alter RAC2 structure or stability. However, it does alter GEF specificity, as RAC2E62K is activated by the DOCK GEF, DOCK2, but not by the Dbl homology GEF, TIAM1, both of which activate the parent protein. Our previous data further showed that the E62K mutation impairs GAP activity for RAC2E62K As this disease mutation is also found in RAS GTPases, we assessed GAP-stimulated GTP hydrolysis for KRAS and observed a similar impairment, suggesting that the mutation plays a conserved role in GAP activation. We also investigated whether the E62K mutation alters effector binding, as activated RAC2 binds effectors to transmit signaling through effector pathways. We find that RAC2E62K retains binding to an NADPH oxidase (NOX2) subunit, p67phox, and to the RAC-binding domain of p21-activated kinase, consistent with our earlier findings. Taken together, our findings indicate that the RAC2E62K mutation promotes immune dysfunction by promoting RAC2 hyperactivation, altering GEF specificity, and impairing GAP function yet retaining key effector interactions.
Insights
A mutation in RAS-related C3 botulinum toxin substrate 2 (RAC2) causes immune deficiency by altering guanine nucleotide exchange factor specificity and impairing GTPase-activating protein function, leading to RAC2 hyperactivation.
Area of Science:
- Molecular biology
- Immunology
- Biochemistry
Background:
- RAS-related C3 botulinum toxin substrate 2 (RAC2) is crucial for immune function.
- An activating mutation, RAC2E62K, is linked to common variable immune deficiency.
- Understanding this mutation's impact on RAC2 function is vital.
Purpose of the Study:
- To investigate the structural and functional consequences of the RAC2E62K mutation.
- To assess the mutation's effects on guanine nucleotide exchange factor (GEF) and GTPase-activating protein (GAP) activity.
- To determine the impact on RAC2 effector binding and signaling.
Main Methods:
- Structural and stability assessments of RAC2E62K.
- Evaluation of GEF specificity using DOCK2 and TIAM1.
- Analysis of GAP activity and GTP hydrolysis for RAC2E62K and KRAS.
- Assessment of effector binding to p67phox and p21-activated kinase.
Main Results:
- The E62K mutation does not affect RAC2 structure or stability.
- RAC2E62K exhibits altered GEF specificity, favoring DOCK2 over TIAM1.
- The mutation impairs GAP activity for RAC2 and KRAS, suggesting a conserved role.
- RAC2E62K maintains binding to key effectors like p67phox and p21-activated kinase.
Conclusions:
- The RAC2E62K mutation promotes immune dysfunction through RAC2 hyperactivation.
- Altered GEF specificity and impaired GAP function contribute to disease pathogenesis.
- Despite functional changes, critical effector interactions are preserved.
Related Concept Videos
 01:24
Small GTPases - Ras and Rho
Three regulatory proteins control their activity:
 02:38
The Ras Gene
Ras is a...
 01:21
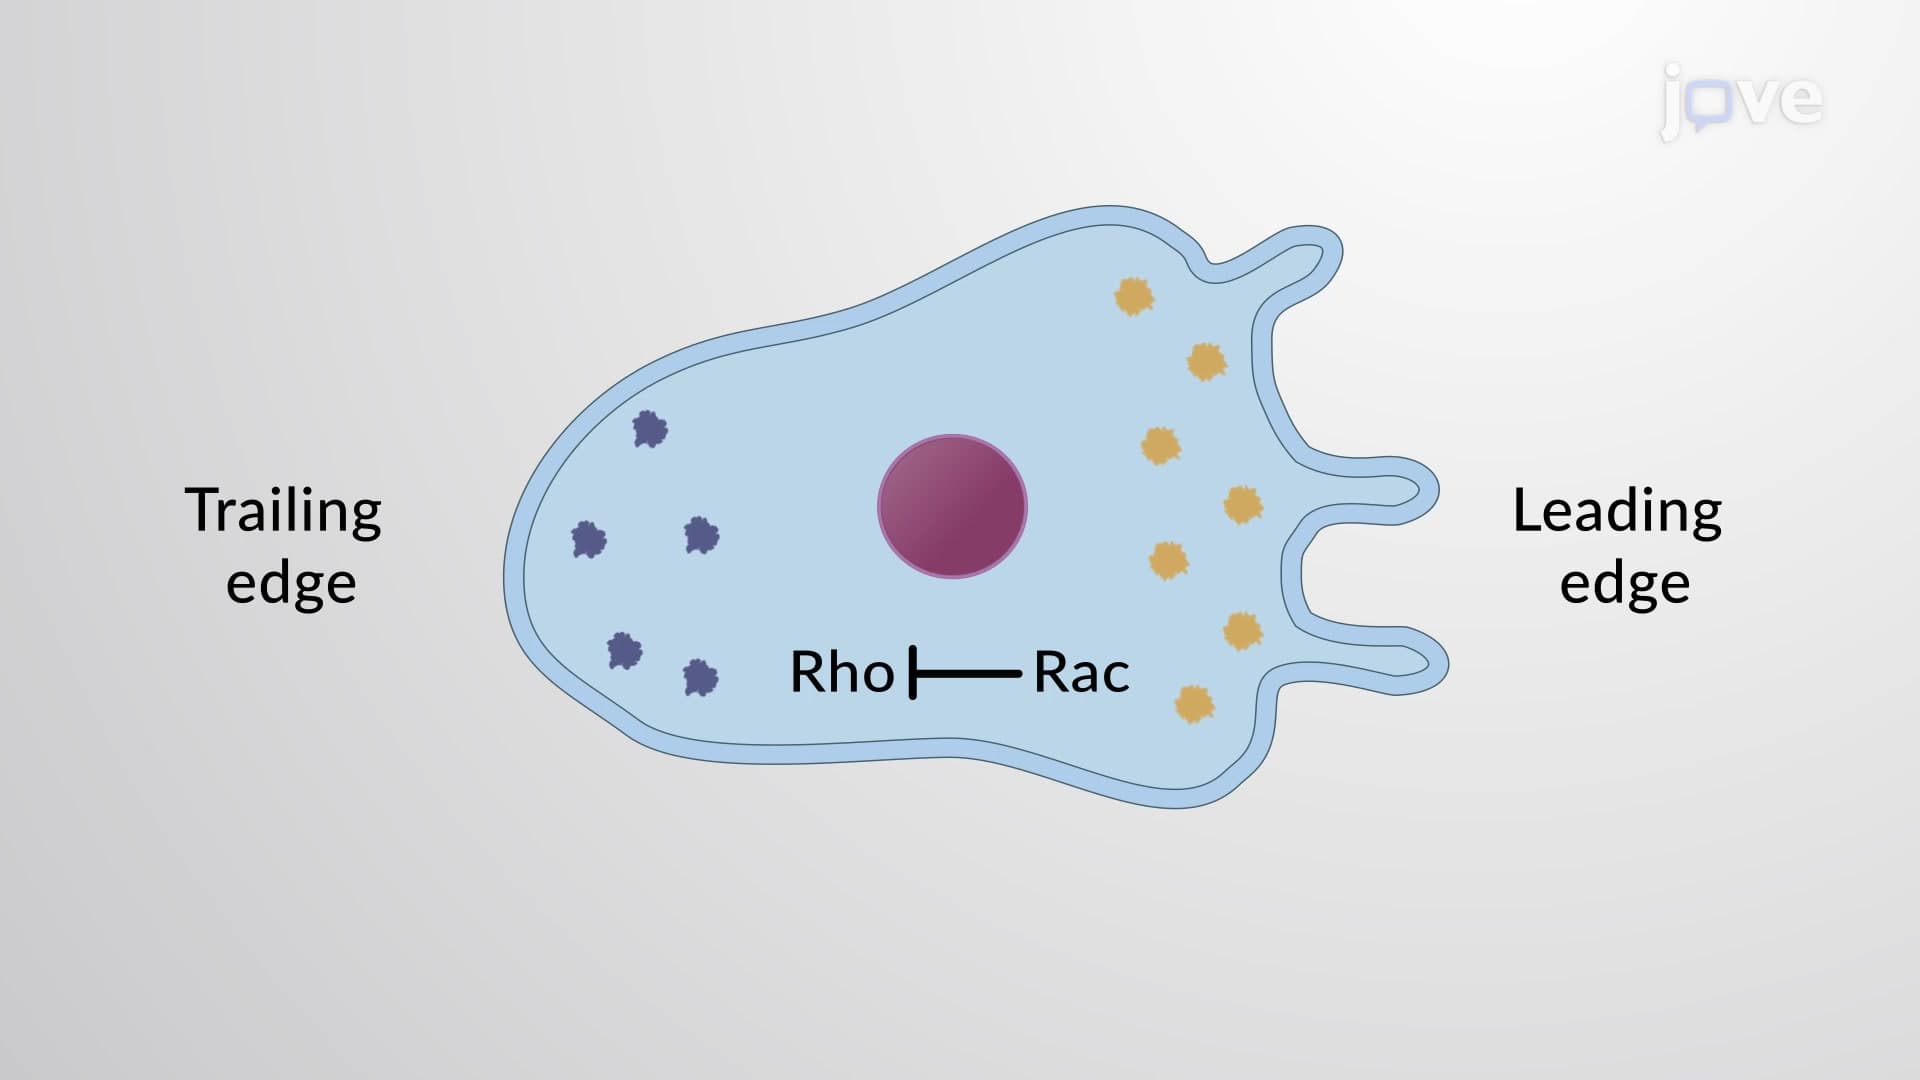
Cell Polarization by Rho Proteins
 01:32
Cytoskeletal Coordination in Cell Migration
 01:22
Activation and Inactivation of G Proteins
 01:07
MAPK Signaling Cascades