Comparison of Normalization Methods for Analysis of TempO-Seq Targeted RNA Sequencing Data
Related Experiment Video
Updated: Dec 15, 2025
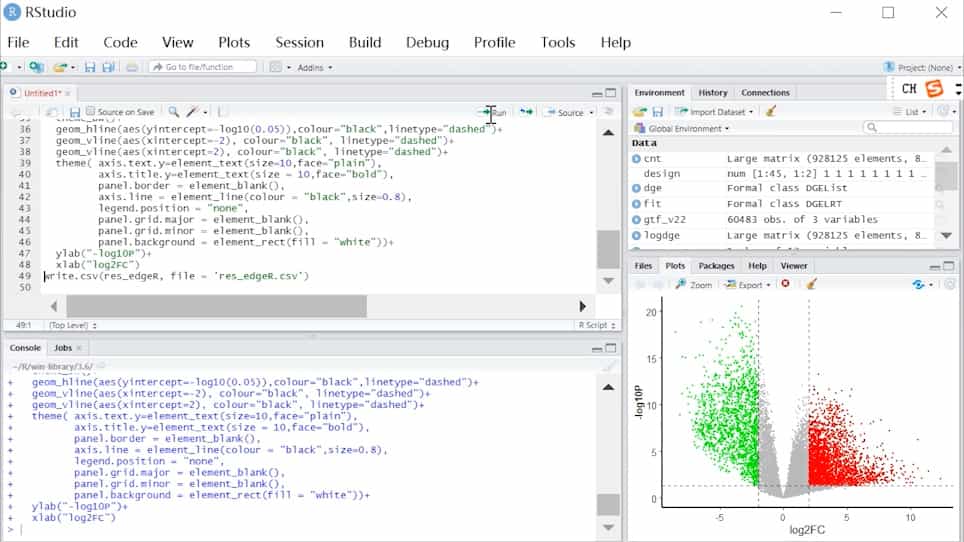
Three Differential Expression Analysis Methods for RNA Sequencing: limma, EdgeR, DESeq2
Published on: September 18, 2021
Pierre R Bushel1,2,3, Stephen S Ferguson3, Sreenivasa C Ramaiahgari3
1Biostatistics and Computational Biology Branch, National Institute of Environmental Health Sciences of National Institutes of Health, Durham, NC, United States.

Abstract:
Analysis of bulk RNA sequencing (RNA-Seq) data is a valuable tool to understand transcription at the genome scale. Targeted sequencing of RNA has emerged as a practical means of assessing the majority of the transcriptomic space with less reliance on large resources for consumables and bioinformatics. TempO-Seq is a templated, multiplexed RNA-Seq platform that interrogates a panel of sentinel genes representative of genome-wide transcription. Nuances of the technology require proper preprocessing of the data. Various methods have been proposed and compared for normalizing bulk RNA-Seq data, but there has been little to no investigation of how the methods perform on TempO-Seq data. We simulated count data into two groups (treated vs. untreated) at seven-fold change (FC) levels (including no change) using control samples from human HepaRG cells run on TempO-Seq and normalized the data using seven normalization methods. Upper Quartile (UQ) performed the best with regard to maintaining FC levels as detected by a limma contrast between treated vs. untreated groups. For all FC levels, specificity of the UQ normalization was greater than 0.84 and sensitivity greater than 0.90 except for the no change and +1.5 levels. Furthermore, K-means clustering of the simulated genes normalized by UQ agreed the most with the FC assignments [adjusted Rand index (ARI) = 0.67]. Despite having an assumption of the majority of genes being unchanged, the DESeq2 scaling factors normalization method performed reasonably well as did simple normalization procedures counts per million (CPM) and total counts (TCs). These results suggest that for two class comparisons of TempO-Seq data, UQ, CPM, TC, or DESeq2 normalization should provide reasonably reliable results at absolute FC levels ≥2.0. These findings will help guide researchers to normalize TempO-Seq gene expression data for more reliable results.
More Related Videos
 12:54
Real-time Analysis of Transcription Factor Binding, Transcription, Translation, and Turnover to Display Global Events During Cellular Activation
Published on: March 7, 2018
 04:58
Author Spotlight: Investigating the Role of Repetitive DNA Misregulation in Cancer Initiation and Immunotherapy Resistance
Published on: December 13, 2024
Related Concept Videos
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 02:24
Ribosome Profiling
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique...
 02:57
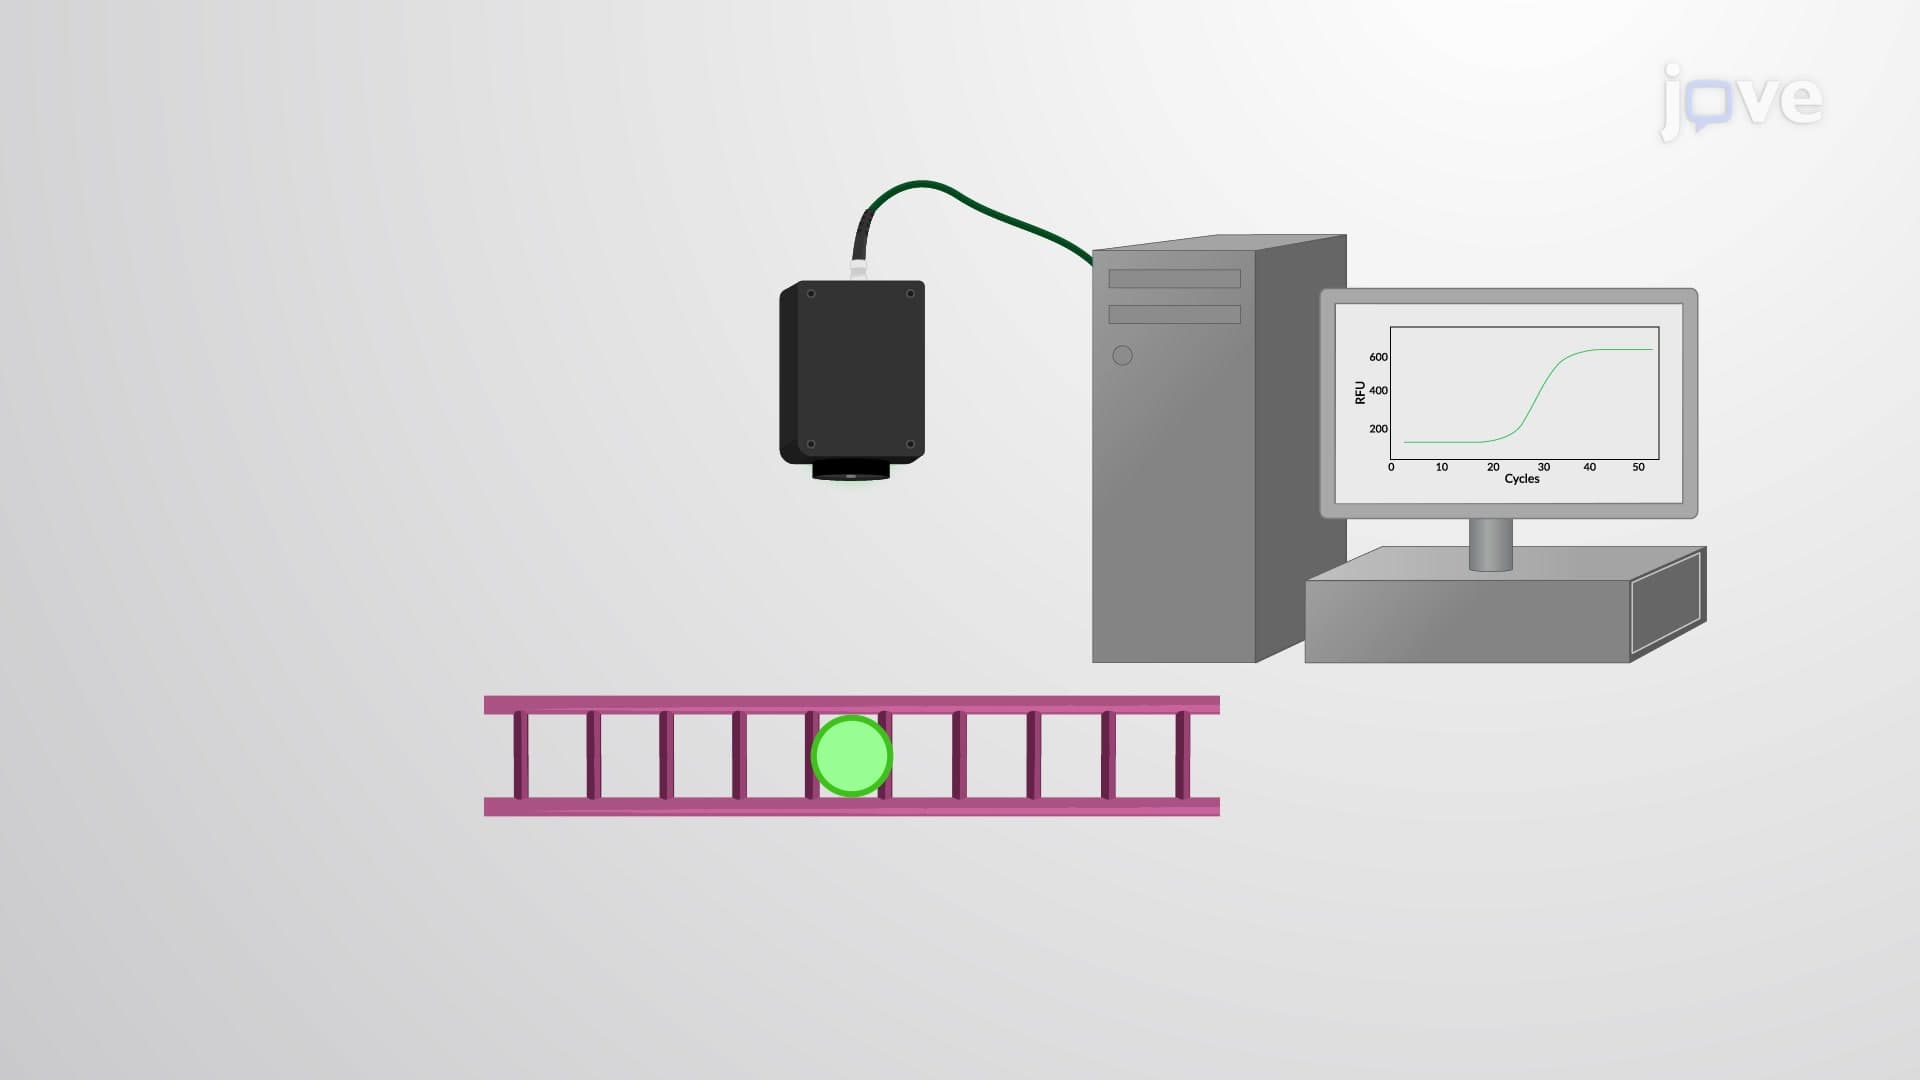
Real Time RT-PCR
The real-time quantification of the number of amplified products is...