Related Experiment Video
Updated: Dec 14, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Hole-hole Tamm-Dancoff-approximated density functional theory: A highly efficient electronic structure method
Christoph Bannwarth1, Jimmy K Yu1, Edward G Hohenstein1
1Department of Chemistry and The PULSE Institute, Stanford University, Stanford, California 94305, USA.

Abstract:
The study of photochemical reaction dynamics requires accurate as well as computationally efficient electronic structure methods for the ground and excited states. While time-dependent density functional theory (TDDFT) is not able to capture static correlation, complete active space self-consistent field methods neglect much of the dynamic correlation. Hence, inexpensive methods that encompass both static and dynamic electron correlation effects are of high interest. Here, we revisit hole-hole Tamm-Dancoff approximated (hh-TDA) density functional theory for this purpose. The hh-TDA method is the hole-hole counterpart to the more established particle-particle TDA (pp-TDA) method, both of which are derived from the particle-particle random phase approximation (pp-RPA). In hh-TDA, the N-electron electronic states are obtained through double annihilations starting from a doubly anionic (N+2 electron) reference state. In this way, hh-TDA treats ground and excited states on equal footing, thus allowing for conical intersections to be correctly described. The treatment of dynamic correlation is introduced through the use of commonly employed density functional approximations to the exchange-correlation potential. We show that hh-TDA is a promising candidate to efficiently treat the photochemistry of organic and biochemical systems that involve several low-lying excited states-particularly those with both low-lying ππ* and nπ* states where inclusion of dynamic correlation is essential to describe the relative energetics. In contrast to the existing literature on pp-TDA and pp-RPA, we employ a functional-dependent choice for the response kernel in pp- and hh-TDA, which closely resembles the response kernels occurring in linear response and collinear spin-flip TDDFT.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:10
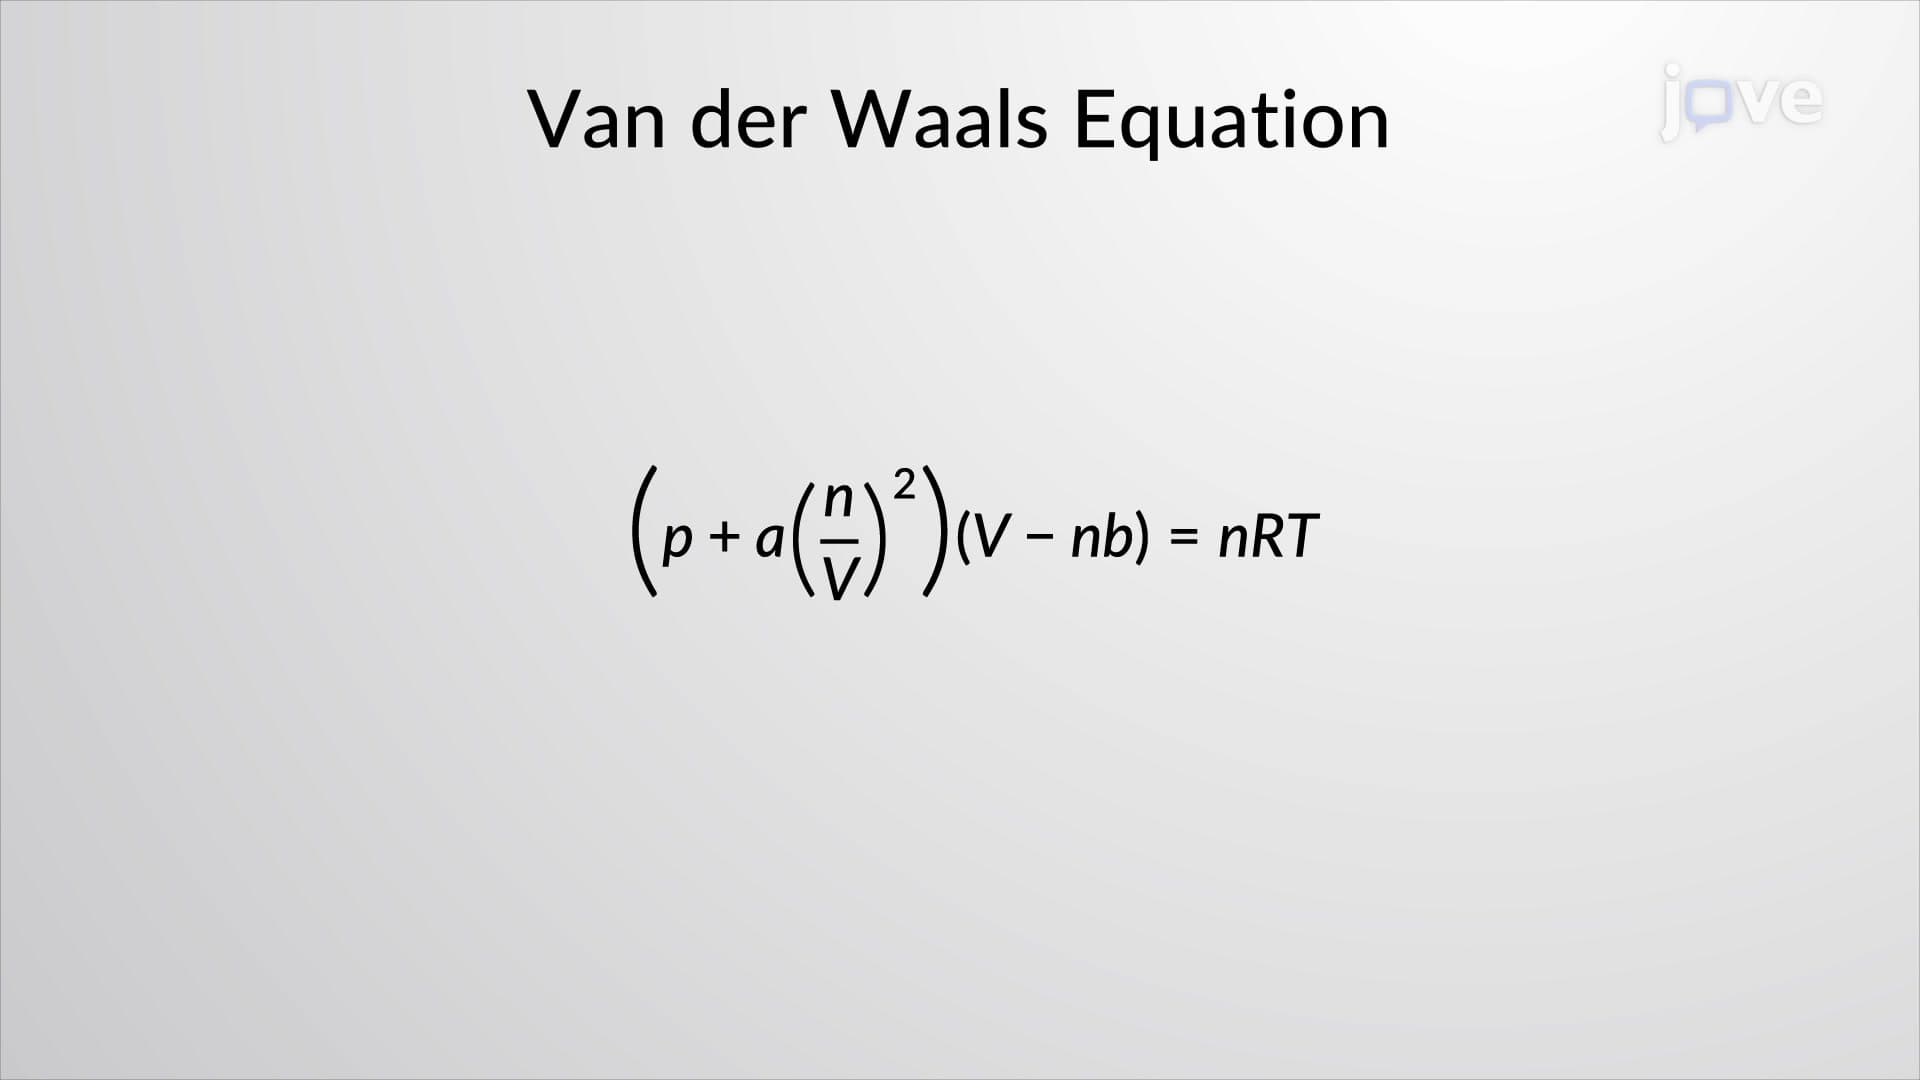
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 02:35
Molecular Orbital Theory I
 02:40
MO Theory and Covalent Bonding
 02:36
Molecular Geometry and Dipole Moments
 02:45
The Quantum-Mechanical Model of an Atom