Related Experiment Video
Updated: Dec 13, 2025


Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
AP-Net: An atomic-pairwise neural network for smooth and transferable interaction potentials
Zachary L Glick1, Derek P Metcalf1, Alexios Koutsoukas2
1Center for Computational Molecular Science and Technology, School of Chemistry and Biochemistry, and School of Computational Science and Engineering, Georgia Institute of Technology, Atlanta, Georgia 30332-0400, USA.

Abstract:
Intermolecular interactions are critical to many chemical phenomena, but their accurate computation using ab initio methods is often limited by computational cost. The recent emergence of machine learning (ML) potentials may be a promising alternative. Useful ML models should not only estimate accurate interaction energies but also predict smooth and asymptotically correct potential energy surfaces. However, existing ML models are not guaranteed to obey these constraints. Indeed, systemic deficiencies are apparent in the predictions of our previous hydrogen-bond model as well as the popular ANI-1X model, which we attribute to the use of an atomic energy partition. As a solution, we propose an alternative atomic-pairwise framework specifically for intermolecular ML potentials, and we introduce AP-Net-a neural network model for interaction energies. The AP-Net model is developed using this physically motivated atomic-pairwise paradigm and also exploits the interpretability of symmetry adapted perturbation theory (SAPT). We show that in contrast to other models, AP-Net produces smooth, physically meaningful intermolecular potentials exhibiting correct asymptotic behavior. Initially trained on only a limited number of mostly hydrogen-bonded dimers, AP-Net makes accurate predictions across the chemically diverse S66x8 dataset, demonstrating significant transferability. On a test set including experimental hydrogen-bonded dimers, AP-Net predicts total interaction energies with a mean absolute error of 0.37 kcal mol-1, reducing errors by a factor of 2-5 across SAPT components from previous neural network potentials. The pairwise interaction energies of the model are physically interpretable, and an investigation of predicted electrostatic energies suggests that the model "learns" the physics of hydrogen-bonded interactions.
More Related Videos
 11:18
Closed-loop Neuro-robotic Experiments to Test Computational Properties of Neuronal Networks
Published on: March 2, 2015
 08:04
Identification and Classification of Position-specific GABAA Receptor Subunit Missense Variants for Their Role In Hippocampal Pyramidal Neurons
Published on: June 6, 2025
Related Concept Videos
 01:23
Propagation of Action Potentials
Neurons (nerve cells) have a resting membrane potential, with a slightly negative charge inside compared to outside. This is maintained by ion channels, such as sodium (Na+) and potassium (K+) channels, which control the flow of ions. When a stimulus, like a touch or a signal from another neuron, triggers the neuron, sodium channels open, allowing sodium ions to...
 01:15
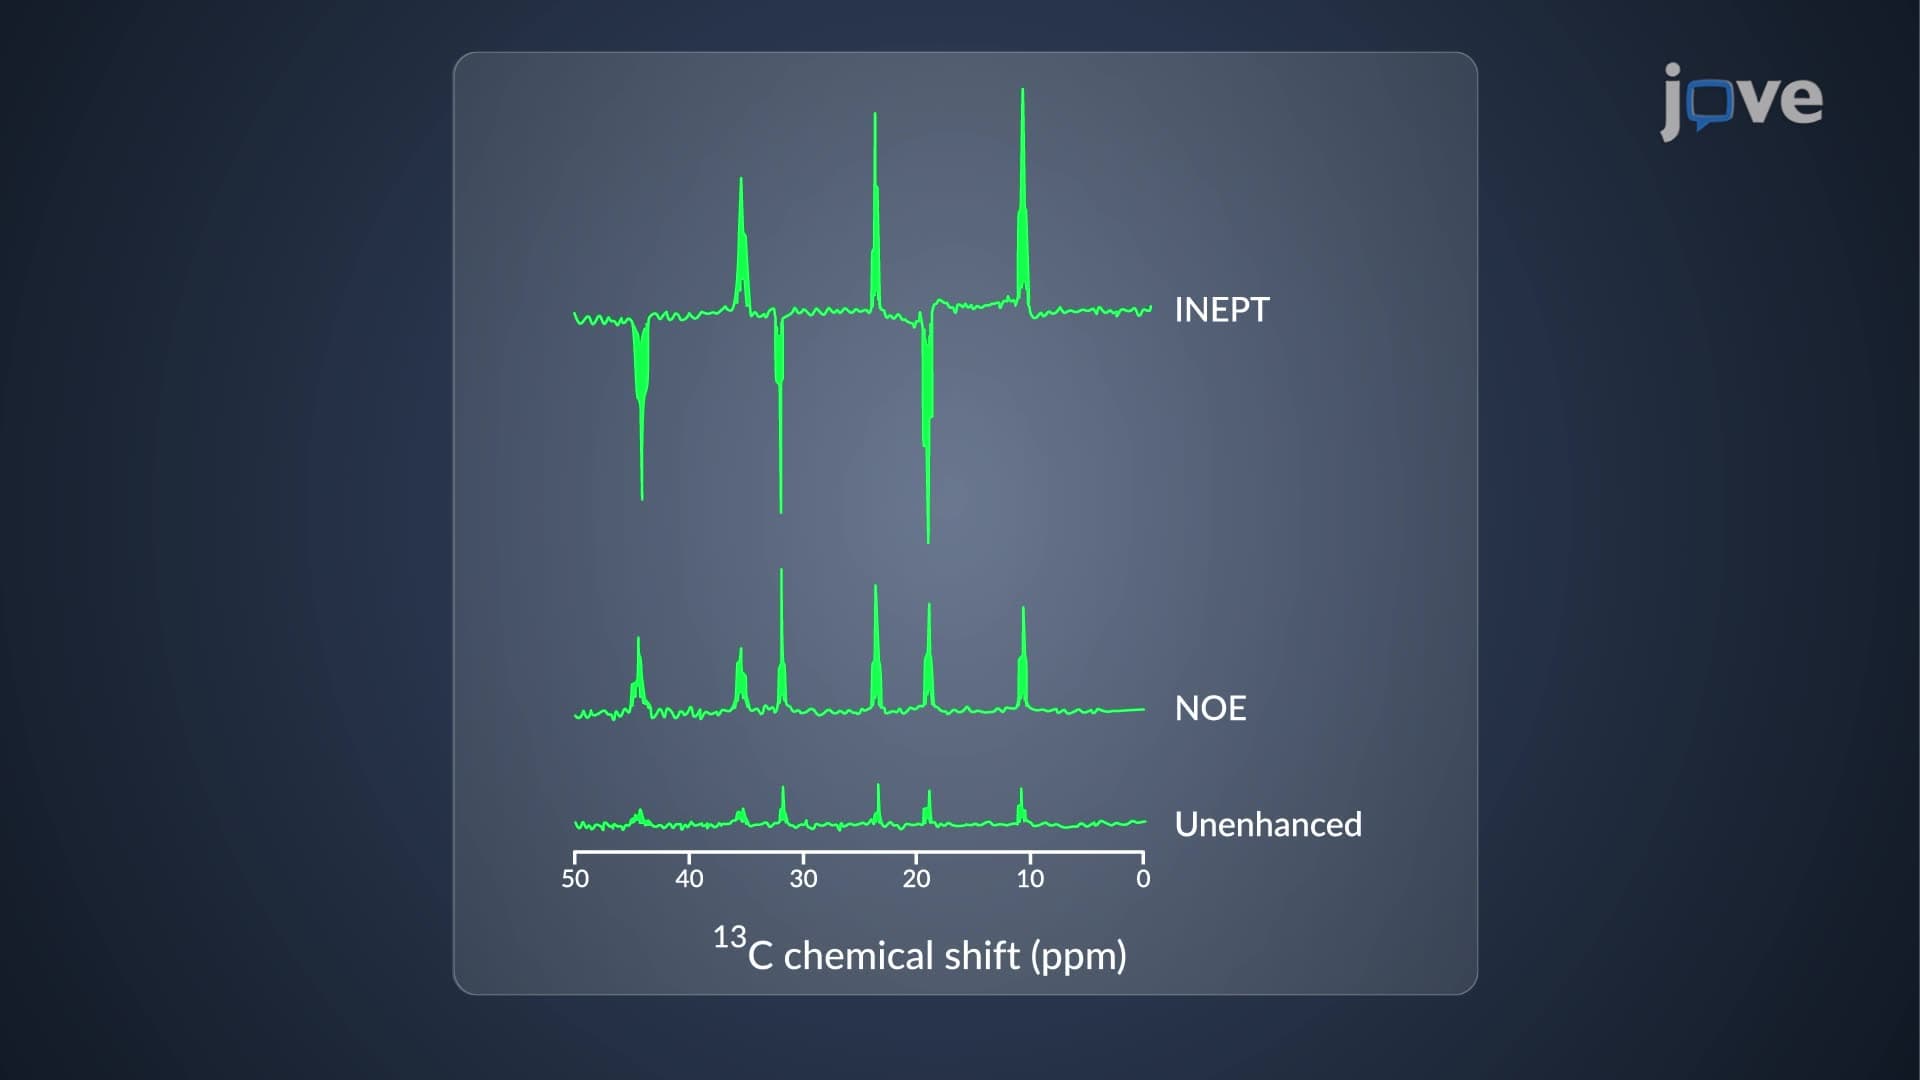
Insensitive Nuclei Enhanced by Polarization Transfer (INEPT)
 01:07
Nuclear Overhauser Enhancement (NOE)
 02:35
Noncovalent Attractions in Biomolecules
02:35Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
 01:28
Integration of Synaptic Events