Related Experiment Video
Updated: Dec 10, 2025

Tissue Triage and Freezing for Models of Skeletal Muscle Disease
Published on: July 15, 2014
TYMP Variants Result in Late-Onset Mitochondrial Myopathy With Altered Muscle Mitochondrial DNA Homeostasis
Dario Ronchi1,2, Leonardo Caporali3, Giulia Francesca Manenti2
1IRCCS Fondazione Ca' Granda Ospedale Maggiore Policlinico, Neurology Unit, Milan, Italy.

Abstract:
Biallelic TYMP variants result in the mitochondrial neurogastrointestinal encephalomyopathy (MNGIE), a juvenile-onset disorder with progressive course and fatal outcome. Milder late-onset (>40 years) form has been rarely described. Gene panel sequencing in a cohort of 60 patients featuring muscle accumulation of mitochondrial DNA (mtDNA) deletions detected TYMP defects in three subjects (5%), two of them with symptom onset in the fifth decade. One of the patients only displayed ptosis and ophthalmoparesis. Biochemical and molecular studies supported the diagnosis. Screening of TYMP is recommended in adult patients with muscle mtDNA instability, even in the absence of cardinal MNGIE features.
Insights
Genetic defects in thymidine phosphorylase (TYMP) cause mitochondrial neurogastrointestinal encephalomyopathy (MNGIE). This study identifies TYMP variants in adult-onset MNGIE patients with muscle mitochondrial DNA instability, even without classic symptoms.
Area of Science:
- Genetics and Molecular Biology
- Neurology
- Mitochondrial Diseases
Background:
- Biallelic thymidine phosphorylase (TYMP) variants cause mitochondrial neurogastrointestinal encephalomyopathy (MNGIE), a severe, juvenile-onset disorder.
- A milder, late-onset form of MNGIE is rarely documented, presenting diagnostic challenges.
Purpose of the Study:
- To investigate the role of TYMP gene defects in adult patients presenting with muscle mitochondrial DNA (mtDNA) instability.
- To identify potential genetic causes for late-onset MNGIE or related phenotypes.
Main Methods:
- Gene panel sequencing was performed on a cohort of 60 patients with muscle mtDNA deletions.
- Biochemical and molecular analyses were conducted to confirm TYMP variant pathogenicity.
Main Results:
- TYMP defects were identified in 5% (3/60) of patients with muscle mtDNA instability.
- Two of these patients exhibited late-onset symptoms (onset >40 years), with one presenting solely with ptosis and ophthalmoparesis.
- Genetic and biochemical findings confirmed TYMP deficiency as the cause of disease in these individuals.
Conclusions:
- TYMP gene screening is crucial for adult patients with unexplained muscle mtDNA instability, even in the absence of typical MNGIE symptoms.
- This highlights the potential for TYMP variants to cause milder, late-onset neurological and ophthalmological manifestations.
More Related Videos
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase COX/SDH Double-labeling Histochemistry
Published on: November 23, 2011
 07:24
Genotyping Single Nucleotide Polymorphisms in the Mitochondrial Genome by Pyrosequencing
Published on: February 10, 2023
Related Concept Videos
 01:19
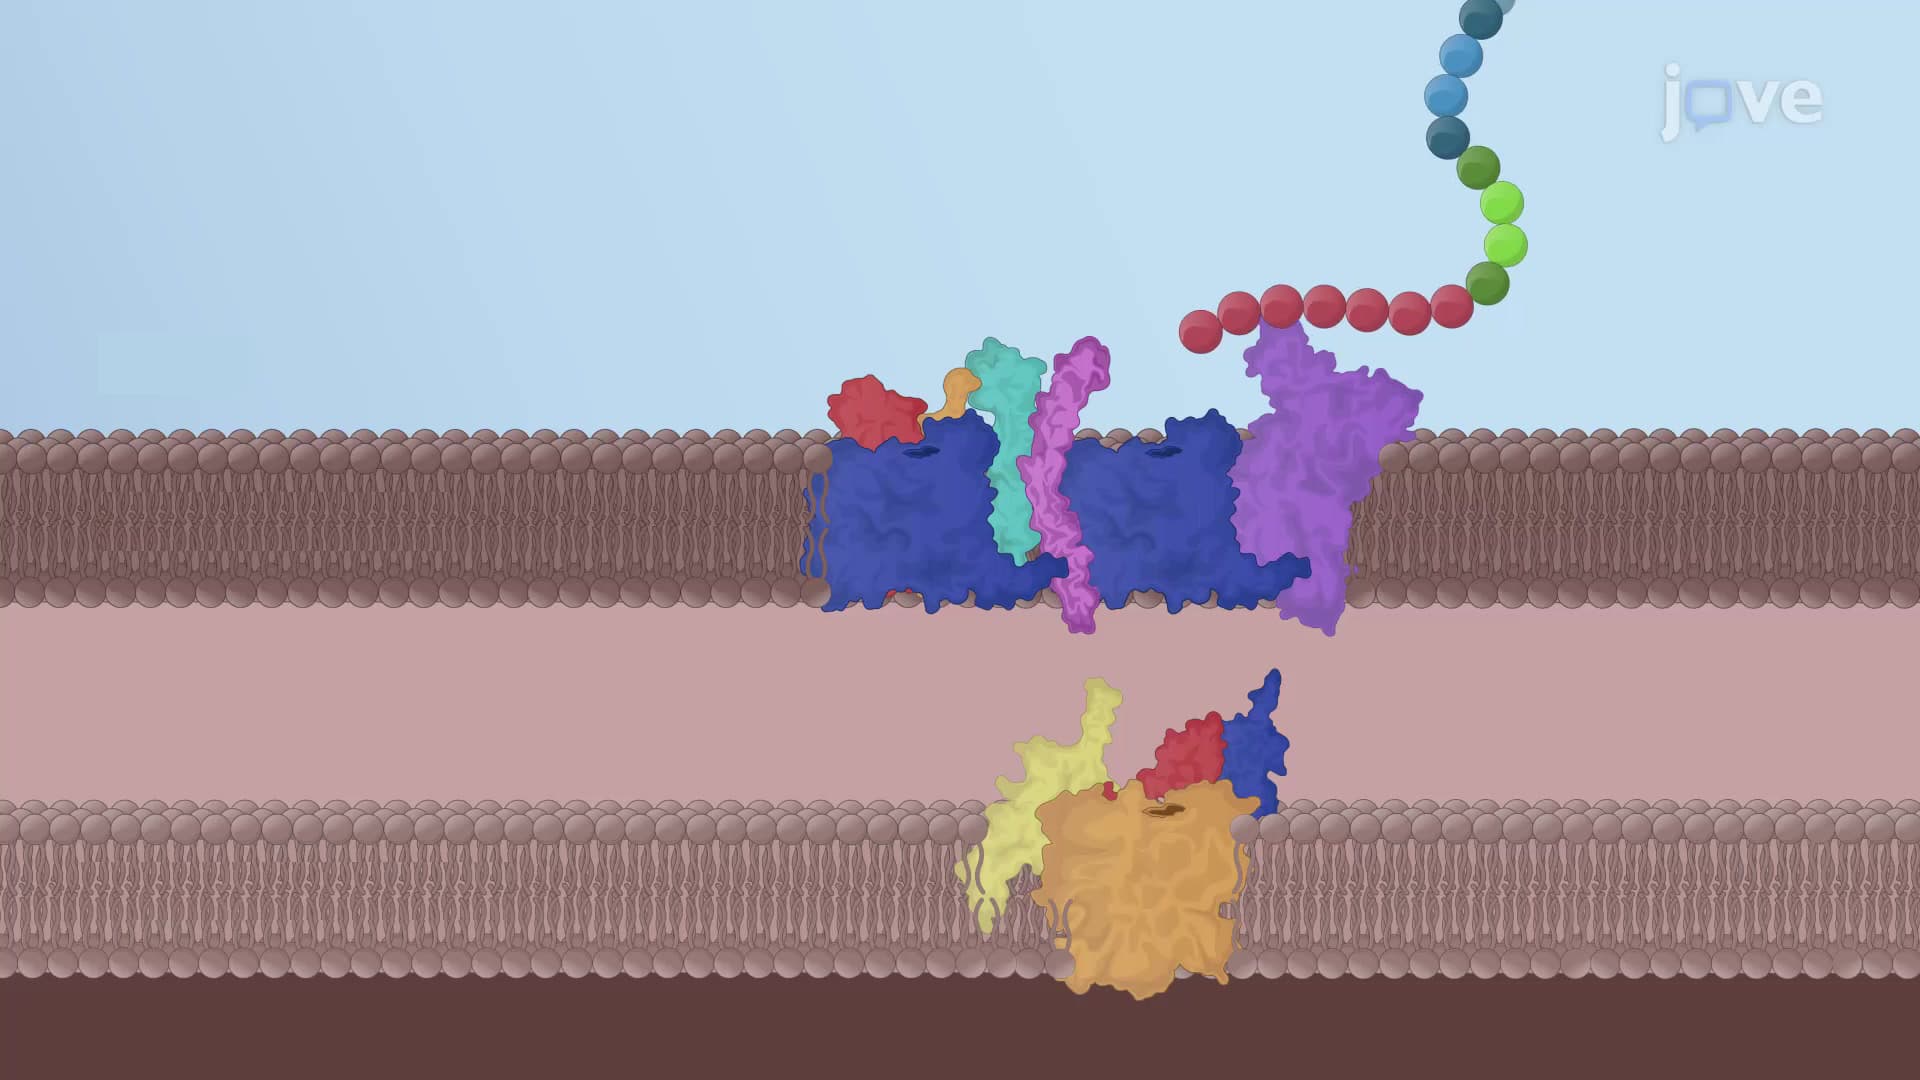
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:48
ATP Synthase: Mechanism
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:37
Energy to Drive Translocation
Generally, polypeptides are unfolded by two distinct...
 02:59
Animal Mitochondrial Genetics