Related Experiment Video
Updated: Dec 10, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
PyFLOSIC: Python-based Fermi-Löwdin orbital self-interaction correction
Sebastian Schwalbe1, Lenz Fiedler1, Jakob Kraus1
1Institute of Theoretical Physics, TU Bergakademie Freiberg, Leipziger Str. 23, D-09599 Freiberg, Germany.

Abstract:
We present pyflosic, an open-source, general-purpose python implementation of the Fermi-Löwdin orbital self-interaction correction (FLO-SIC), which is based on the python simulation of chemistry framework (pyscf) electronic structure and quantum chemistry code. Thanks to pyscf, pyflosic can be used with any kind of Gaussian-type basis set, various kinds of radial and angular quadrature grids, and all exchange-correlation functionals within the local density approximation, generalized-gradient approximation (GGA), and meta-GGA provided in the libxc and xcfun libraries. A central aspect of FLO-SIC is the Fermi-orbital descriptors, which are used to estimate the self-interaction correction. Importantly, they can be initialized automatically within pyflosic; they can also be optimized within pyflosic with an interface to the atomic simulation environment, a python library that provides a variety of powerful gradient-based algorithms for geometry optimization. Although pyflosic has already facilitated applications of FLO-SIC to chemical studies, it offers an excellent starting point for further developments in FLO-SIC approaches, thanks to its use of a high-level programming language and pronounced modularity.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:12
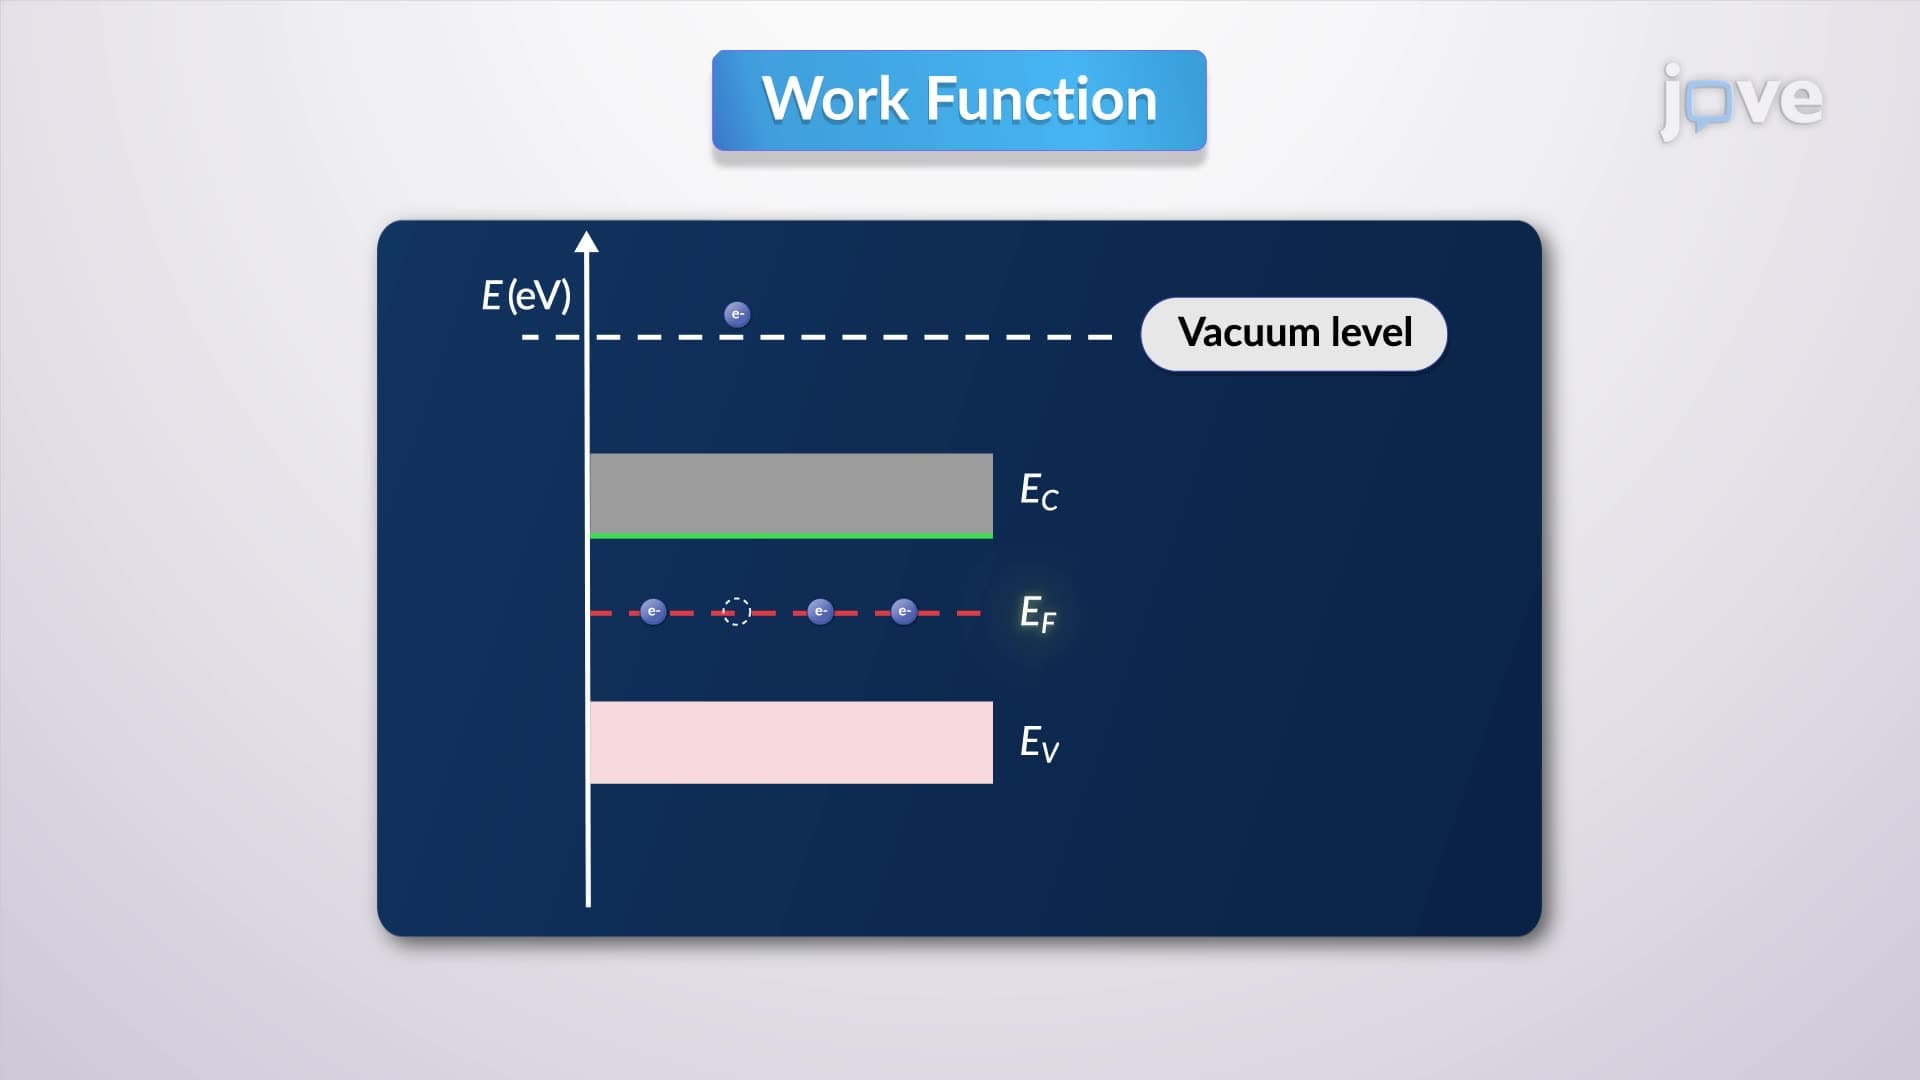
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:44
Atomic Orbitals
 01:18
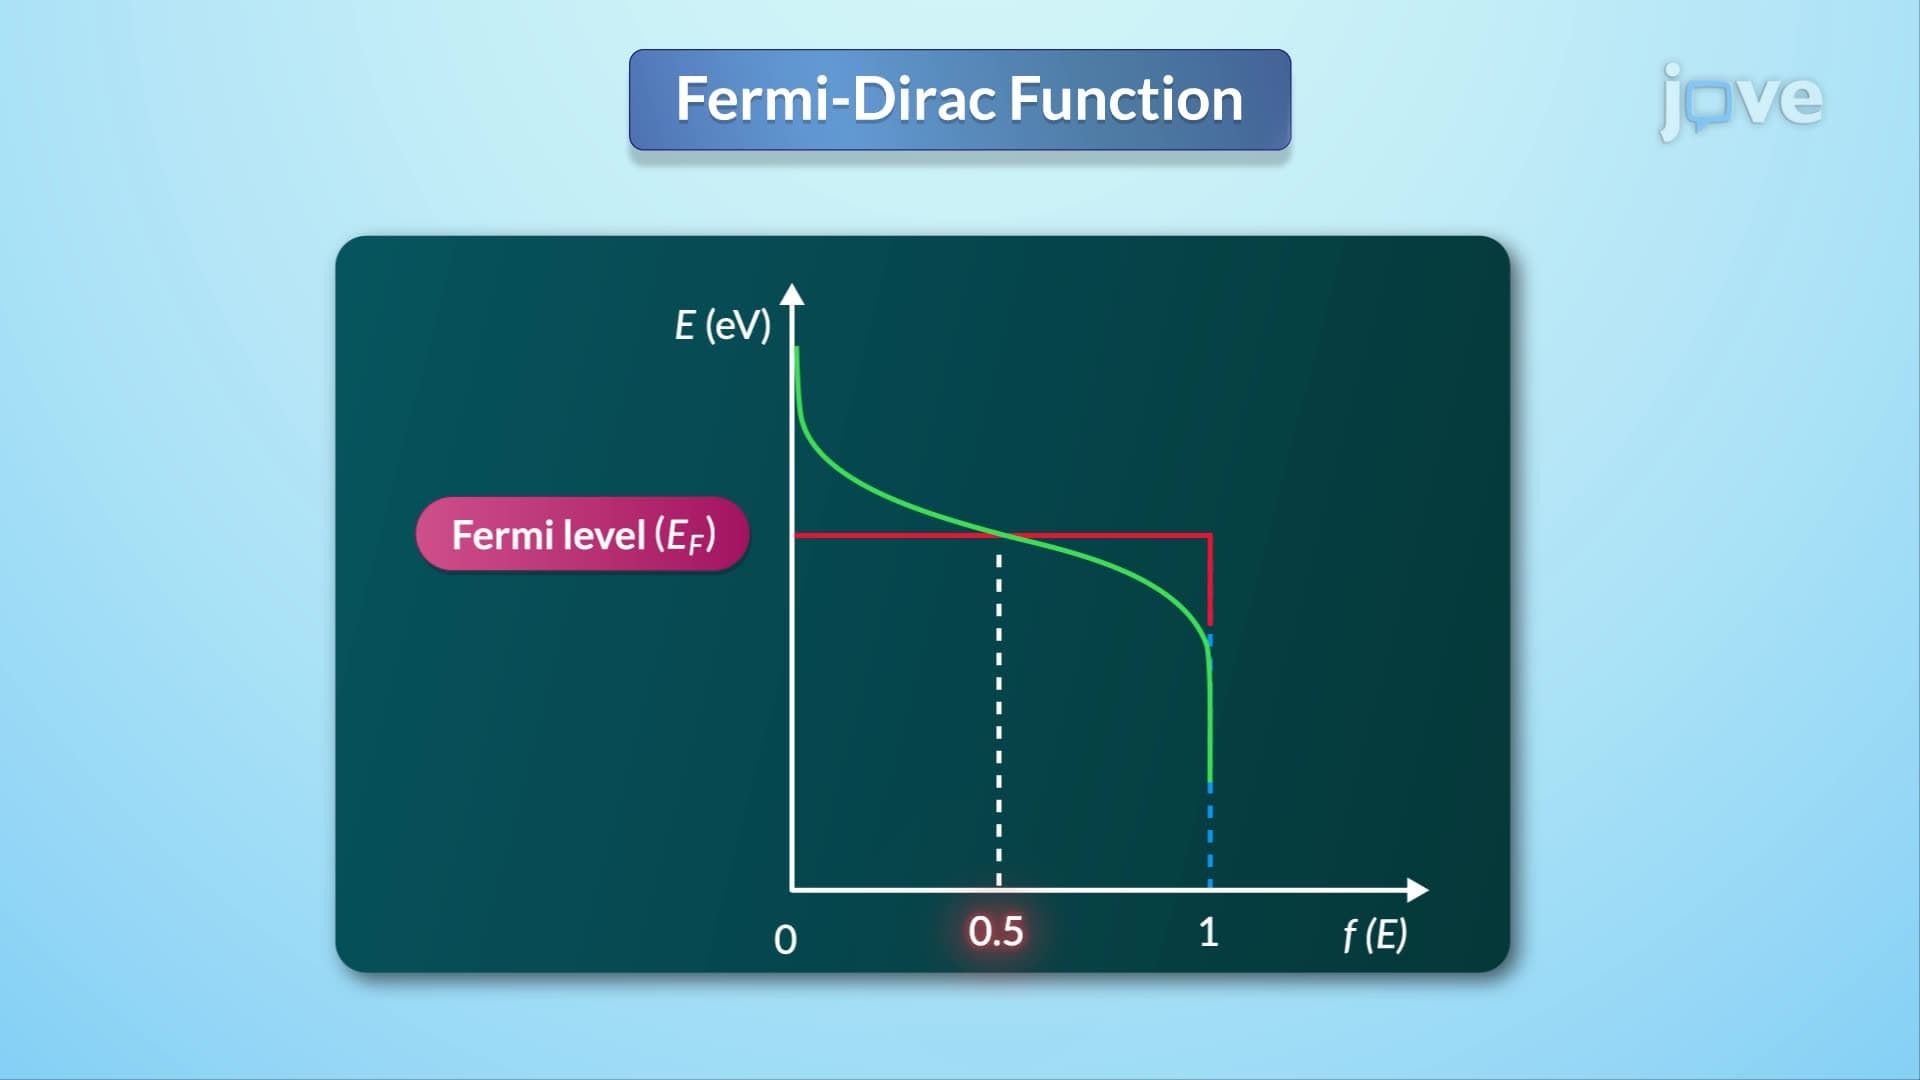
Fermi Level
At absolute zero temperature, electrons fill all energy states up to the Fermi level, leaving upper states empty. As the temperature rises,...
 01:12
Reduced Mass Coordinates: Isolated Two-body Problem
 03:21
The Energies of Atomic Orbitals
 03:51
Molecular Orbital Theory II