Related Experiment Video
Updated: Dec 9, 2025

Pharmacophore Modeling for Targets with Extensive Ligand Libraries: A Case Study on SARS-CoV-2 Mpro
Published on: September 26, 2025
Molecular docking, validation, dynamics simulations, and pharmacokinetic prediction of natural compounds against the
Shivanika C1, Deepak Kumar S2, Venkataraghavan Ragunathan3
1Department of Bio-Engineering, School of Engineering, Vels Institute of Science Technology and Advanced Studies, Chennai, Tamil Nadu, India.

Abstract:
The study aims to evaluate the potency of two hundred natural antiviral phytocompounds against the active site of the Severe Acquired Respiratory Syndrome - Coronavirus - 2 (SARS-CoV-2) Main-Protease (Mpro) using AutoDock 4.2.6. The three- dimensional crystal structure of the Mpro (PDB Id: 6LU7) was retrieved from the Protein Data Bank (PDB), the active site was predicted using MetaPocket 2.0. Food and Drug Administration (FDA) approved viral protease inhibitors were used as standards for comparison of results. The compounds theaflavin-3-3'-digallate, rutin, hypericin, robustaflavone, and (-)-solenolide A with respective binding energy of -12.41 (Ki = 794.96 pM); -11.33 (Ki = 4.98 nM); -11.17 (Ki = 6.54 nM); -10.92 (Ki = 9.85 nM); and -10.82 kcal/mol (Ki = 11.88 nM) were ranked top as Coronavirus Disease - 2019 (COVID-19) Mpro inhibitors. The interacting amino acid residues were visualized using Discovery Studio 3.5 to elucidate the 2-dimensional and 3-dimensional interactions. The study was validated by i) re-docking the N3-peptide inhibitor-Mpro and superimposing them onto co-crystallized complex and ii) docking decoy ligands to Mpro. The ligands that showed low binding energy were further predicted for and pharmacokinetic properties and Lipinski's rule of 5 and the results are tabulated and discussed. Molecular dynamics simulations were performed for 50 ns for those compounds using the Desmond package, Schrödinger to assess the conformational stability and fluctuations of protein-ligand complexes during the simulation. Thus, the natural compounds could act as a lead for the COVID-19 regimen after in-vitro and in- vivo clinical trials.Communicated by Ramaswamy H. Sarma.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Related Concept Videos
 01:16
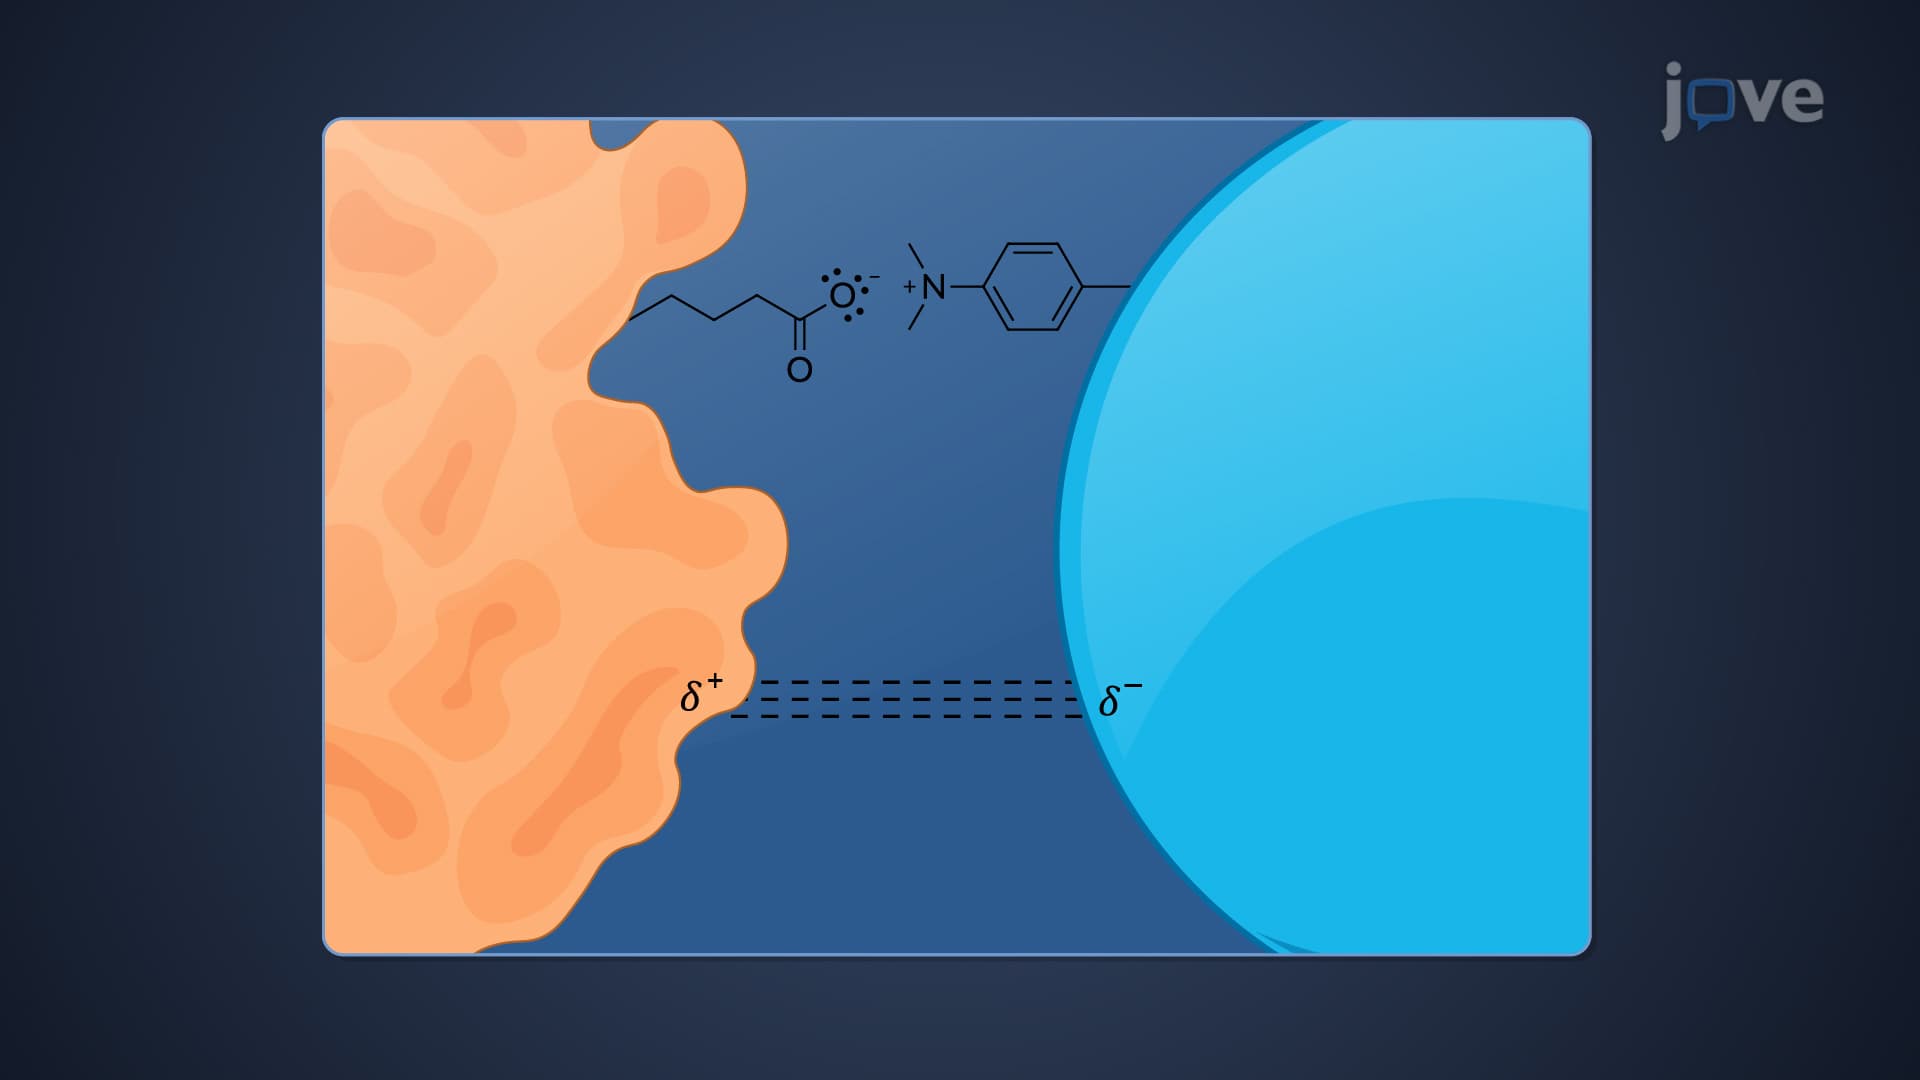
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...