Related Experiment Video
Updated: Dec 9, 2025


Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Efficiency of Homology Modeling Assisted Molecular Docking in G-protein Coupled Receptors
Shome S Bhunia1, Anil K Saxena2
1Global Institute of Pharmaceutical Education and Research, Kashipur, Uttarakhand, India.

Background:
Molecular docking is in regular practice to assess ligand affinity on a target protein crystal structure. In the absence of protein crystal structure, the homology modeling or comparative modeling is the best alternative to elucidate the relationship details between a ligand and protein at the molecular level. The development of accurate homology modeling (HM) and its integration with molecular docking (MD) is essential for successful, rational drug discovery.
Objective:
The G-protein coupled receptors (GPCRs) are attractive therapeutic targets due to their immense role in human pharmacology. The GPCRs are membrane-bound proteins with the complex constitution, and the understanding of their activation and inactivation mechanisms is quite challenging. Over the past decade, there has been a rapid expansion in the number of solved G-protein-coupled receptor (GPCR) crystal structures; however, the majority of the GPCR structures remain unsolved. In this context, HM guided MD has been widely used for structure-based drug design (SBDD) of GPCRs.
Methods:
The focus of this review is on the recent (i) developments on HM supported GPCR drug discovery in the absence of GPCR crystal structures and (ii) application of HM in understanding the ligand interactions at the binding site, virtual screening, determining receptor subtype selectivity and receptor behaviour in comparison with GPCR crystal structures.
Results:
The HM in GPCRs has been extremely challenging due to the scarcity in template structures. In such a scenario, it is difficult to get accurate HM that can facilitate understanding of the ligand-receptor interactions. This problem has been alleviated to some extent by developing refined HM based on incorporating active /inactive ligand information and inducing protein flexibility. In some cases, HM proteins were found to outscore crystal structures.
Conclusion:
The developments in HM have been highly operative to gain insights about the ligand interaction at the binding site and receptor functioning at the molecular level. Thus, HM guided molecular docking may be useful for rational drug discovery for the GPCRs mediated diseases.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
Related Concept Videos
 01:15
G Protein-coupled Receptors
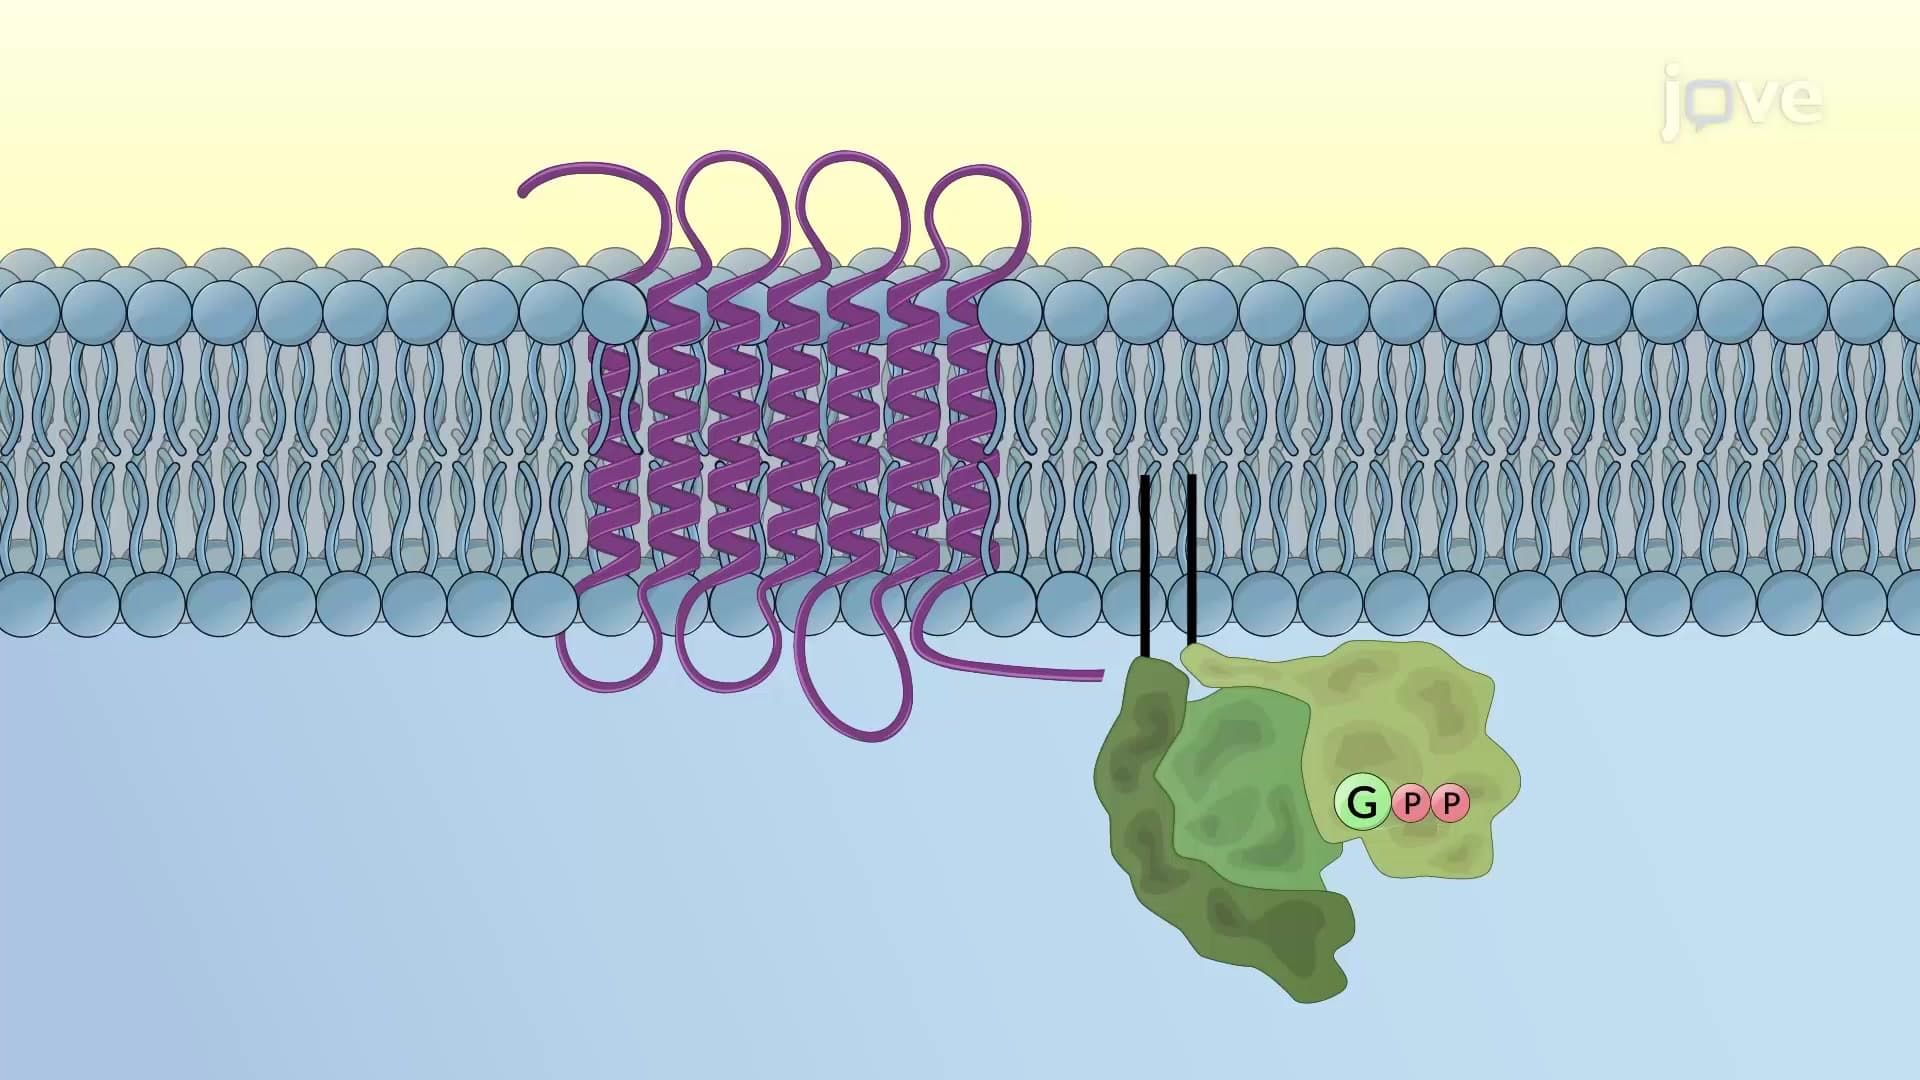
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:30
Transducer Mechanism: G Protein–Coupled Receptors
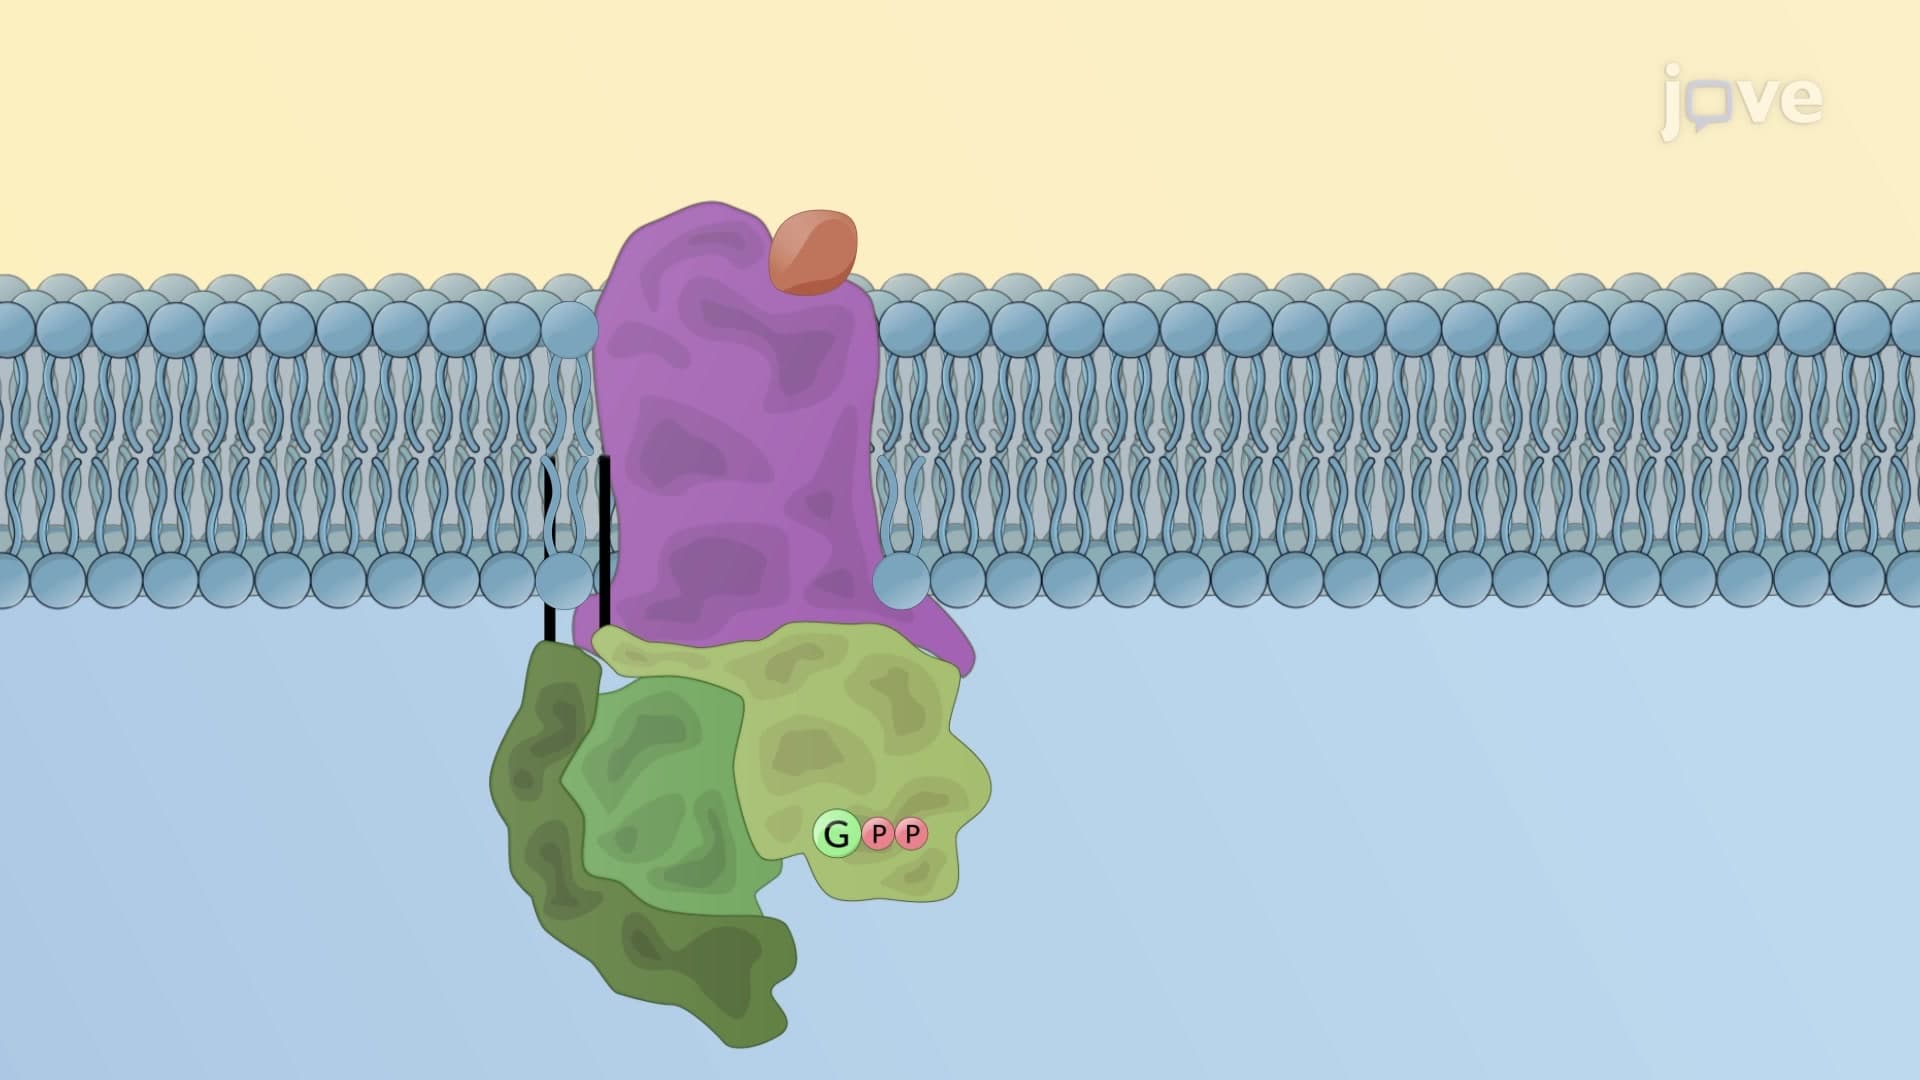
GPCRs are also called heptahelical,...
 01:21
G-protein Coupled Receptors
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces