A machine learning based intramolecular potential for a flexible organic molecule
Related Experiment Video
Updated: Dec 8, 2025

Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Daniel J Cole1, Letif Mones, Gábor Csányi
1School of Natural and Environmental Sciences, Newcastle University, Newcastle upon Tyne NE1 7RU, UK.

Abstract:
Quantum mechanical predictive modelling in chemistry and biology is often hindered by the long time scales and large system sizes required of the computational model. Here, we employ the kernel regression machine learning technique to construct an analytical potential, using the Gaussian Approximation Potential software and framework, that reproduces the quantum mechanical potential energy surface of a small, flexible, drug-like molecule, 3-(benzyloxy)pyridin-2-amine. Challenges linked to the high dimensionality of the configurational space of the molecule are overcome by developing an iterative training protocol and employing a representation that separates short and long range interactions. The analytical model is connected to the MCPRO simulation software, which allows us to perform Monte Carlo simulations of the small molecule bound to two proteins, p38 MAP kinase and leukotriene A4 hydrolase, as well as in water. We demonstrate that our machine learning based intramolecular model is transferable to the condensed phase, and demonstrate that the use of a faithful representation of the quantum mechanical potential energy surface can result in corrections to absolute protein-ligand binding free energies of up to 2 kcal mol-1 in the example studied here.

Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:33
Radical Reactivity: Intramolecular vs Intermolecular
 03:00
Intermolecular vs Intramolecular Forces
 02:36
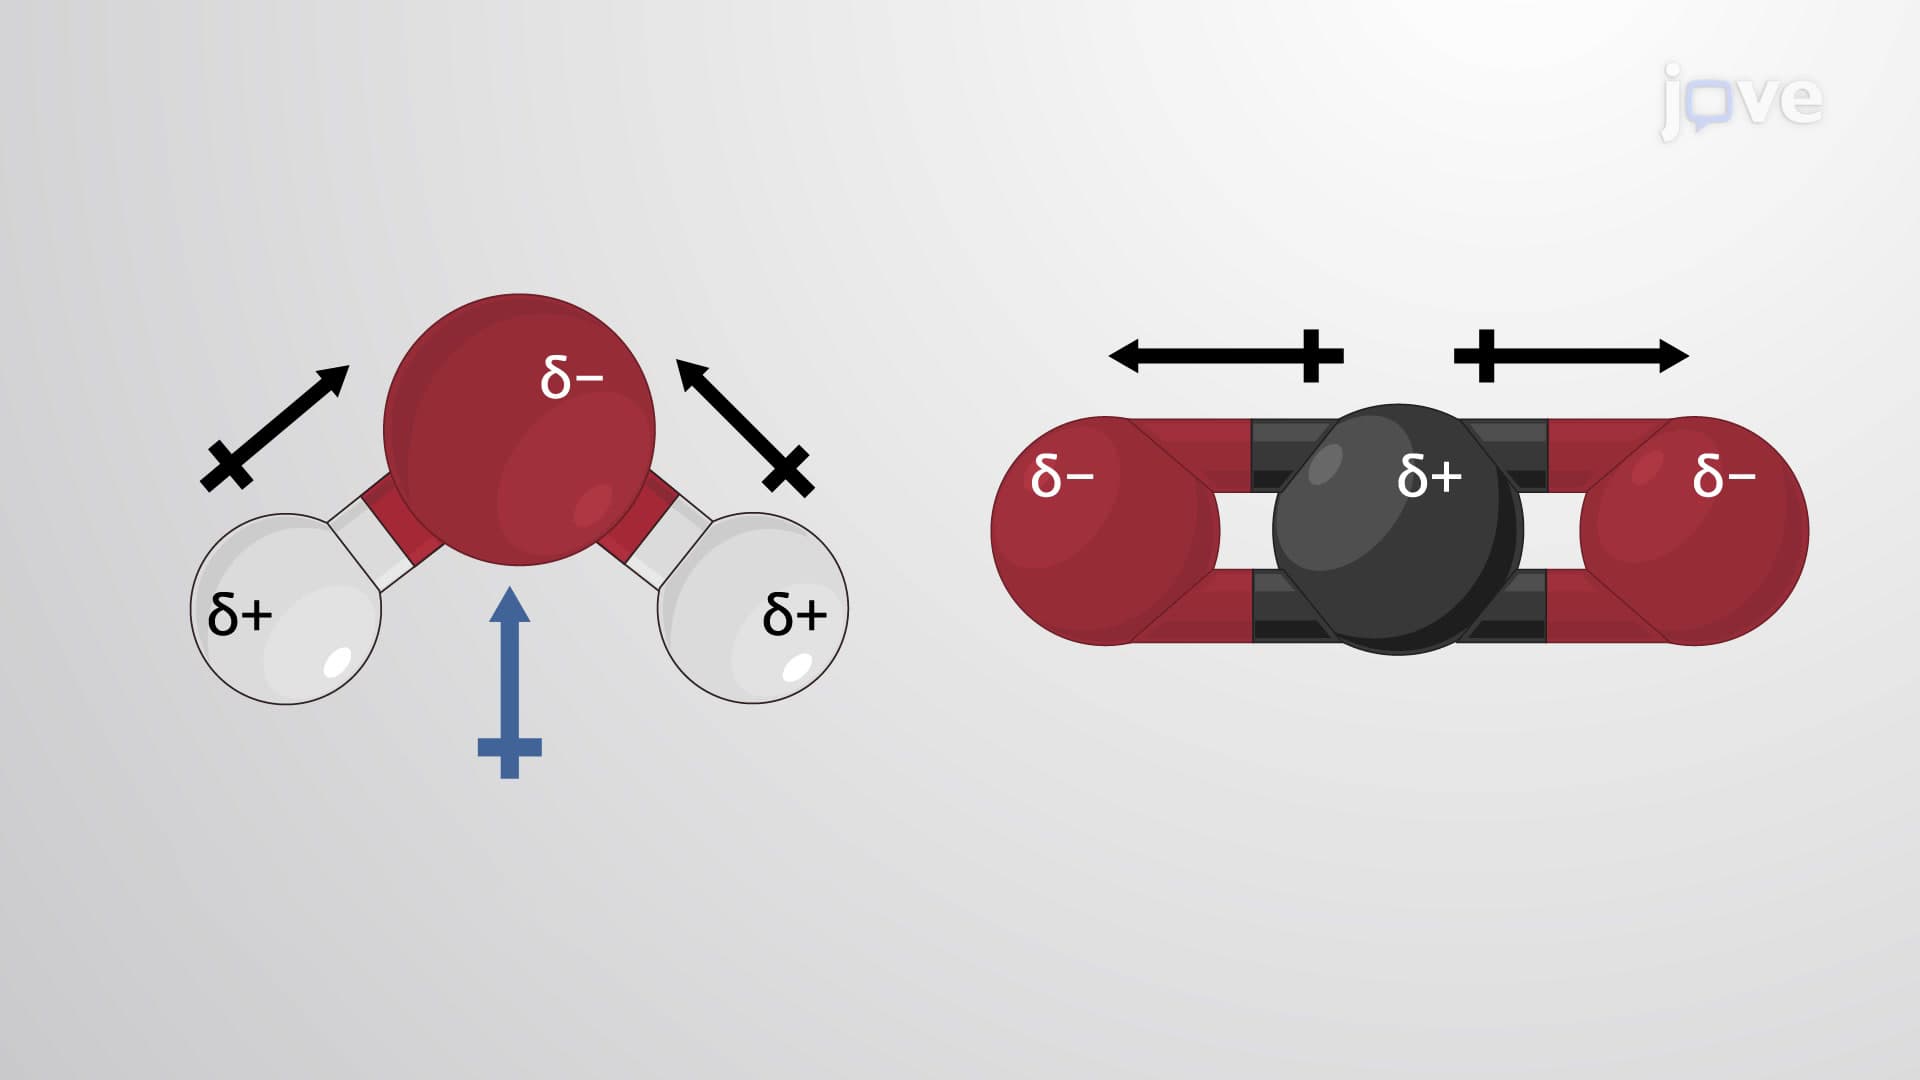
Molecular Geometry and Dipole Moments
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:35
Noncovalent Attractions in Biomolecules