Related Experiment Video
Updated: Dec 8, 2025

Scalable Quantum Integrated Circuits on Superconducting Two-Dimensional Electron Gas Platform
Published on: August 2, 2019
Weak Dimensionality Dependence and Dominant Role of Ionic Fluctuations in the Charge-Density-Wave Transition of
Raffaello Bianco1, Lorenzo Monacelli2,3, Matteo Calandra3,4,5
1Centro de Física de Materiales (CSIC-UPV/EHU), Manuel de Lardizabal pasealekua 5, 20018 Donostia/San Sebastián, Spain.

Abstract:
Contradictory experiments have been reported about the dimensionality effect on the charge-density-wave transition in 2H NbSe_{2}. While scanning tunneling experiments on single layers grown by molecular beam epitaxy measure a charge-density-wave transition temperature in the monolayer similar to the bulk, around 33 K, Raman experiments on exfoliated samples observe a large enhancement of the transition temperature up to 145 K. By employing a nonperturbative approach to deal with anharmonicity, we calculate from first principles the temperature dependence of the phonon spectra both for bulk and monolayer. In both cases, the charge-density-wave transition temperature is estimated as the temperature at which the phonon energy of the mode driving the structural instability vanishes. The obtained transition temperature in the bulk is around 59 K, in rather good agreement with experiments, and it is just slightly increased in the single-layer limit to 73 K, showing the weak dependence of the transition on dimensionality. Environmental factors could motivate the disagreement between the transition temperatures reported by experiments. Our analysis also demonstrates the predominance of ionic fluctuations over electronic ones in the melting of the charge-density-wave order.

Related Concept Videos
 02:54
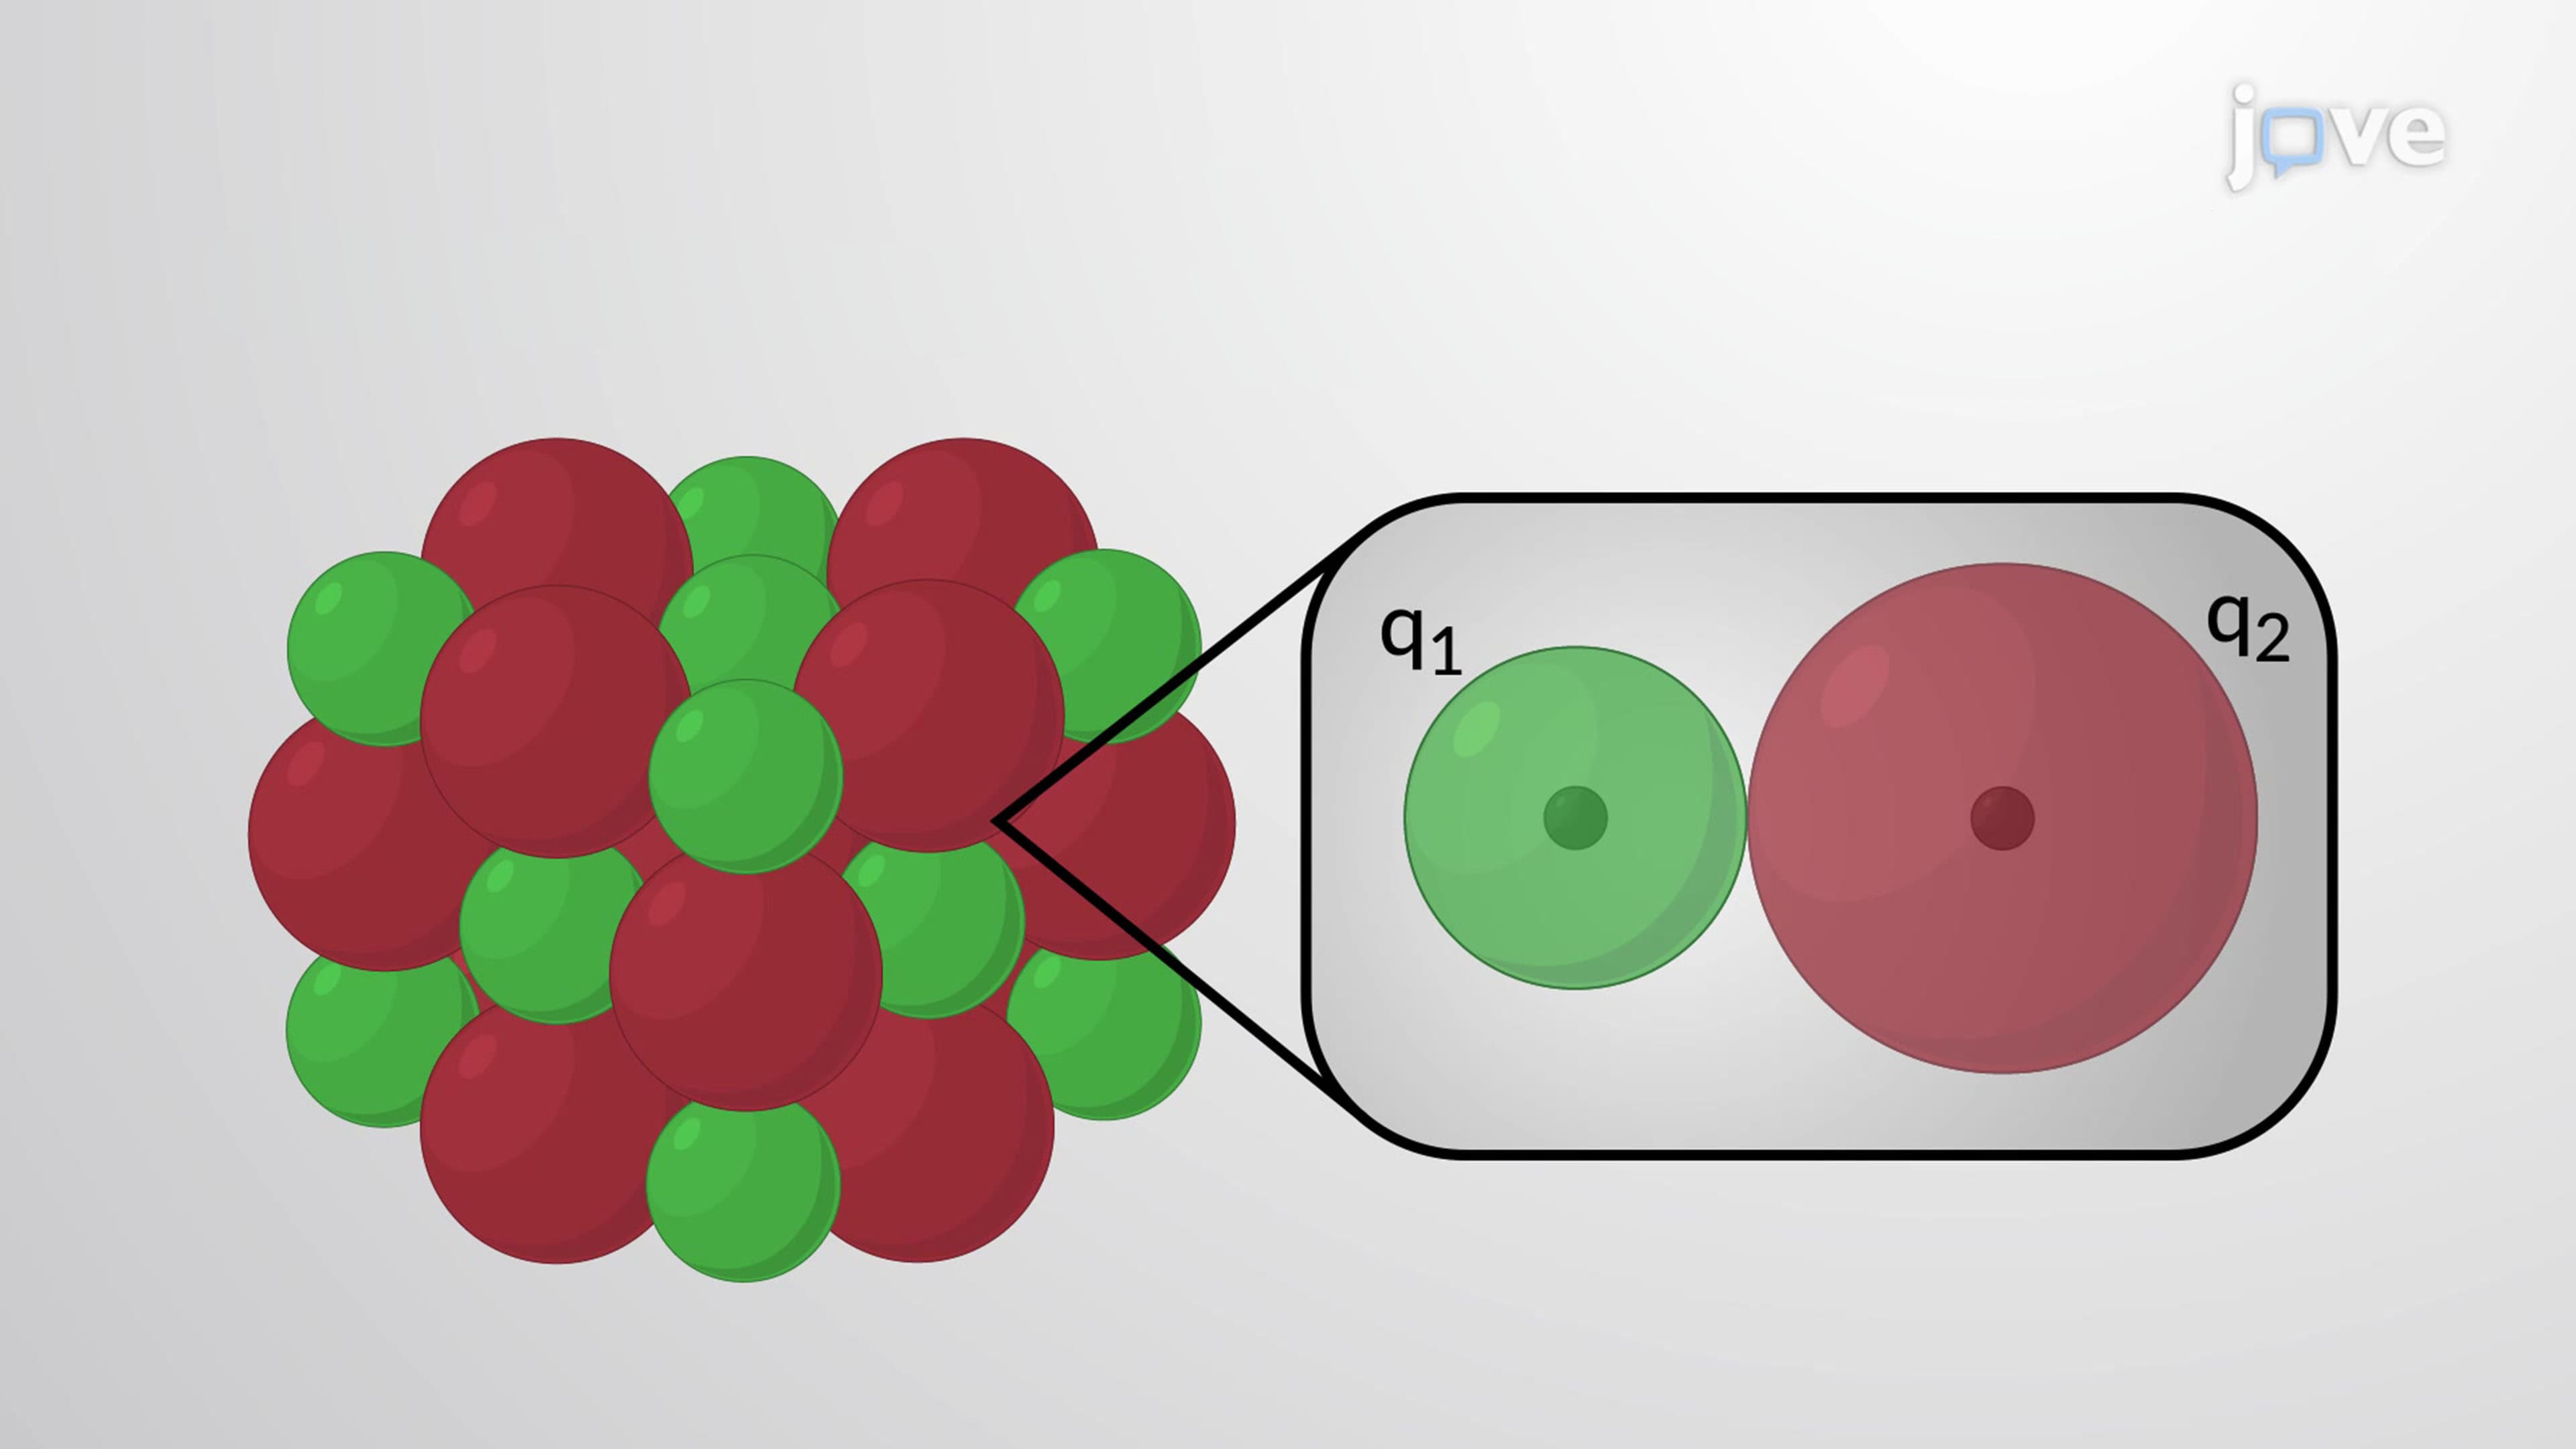
Trends in Lattice Energy: Ion Size and Charge
 02:42
Valence Bond Theory
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 02:32
The de Broglie Wavelength
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...