Related Experiment Video
Updated: Dec 8, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Decoherence-corrected Ehrenfest molecular dynamics on many electronic states
Michael P Esch1, Benjamin G Levine1
1Department of Chemistry, Michigan State University, East Lansing, Michigan 48824, USA.

Abstract:
Decoherence corrections increase the accuracy of mixed quantum-classical nonadiabatic molecular dynamics methods, but they typically require explicit knowledge of the potential energy surfaces of all occupied electronic states. This requirement renders them impractical for applications in which large numbers of electronic states are occupied. The authors recently introduced the collapse to a block (TAB) decoherence correction [M. P. Esch and B. G. Levine, J. Chem. Phys. 152, 234105 (2020)], which incorporates a state-pairwise definition of decoherence time to accurately describe dynamics on more than two electronic states. In this work, TAB is extended by introduction of a scheme for efficiently computing a small number of approximate eigenstates of the electronic Hamiltonian, eliminating the need for explicit knowledge of a large number of potential energy surfaces. This adaptation of TAB for dense manifolds of states (TAB-DMS) is systematically improvable by increasing the number of computed approximate eigenstates. Application to a series of one-dimensional model problems demonstrates that TAB-DMS can be accurate when even a very modest number of approximate eigenstates are computed (four in all models tested here). Comparison of TAB simulations to exact quantum dynamical simulations indicates that TAB is quite accurate so long as the decoherence correction is carefully parameterized.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 01:12
Fermi Level Dynamics
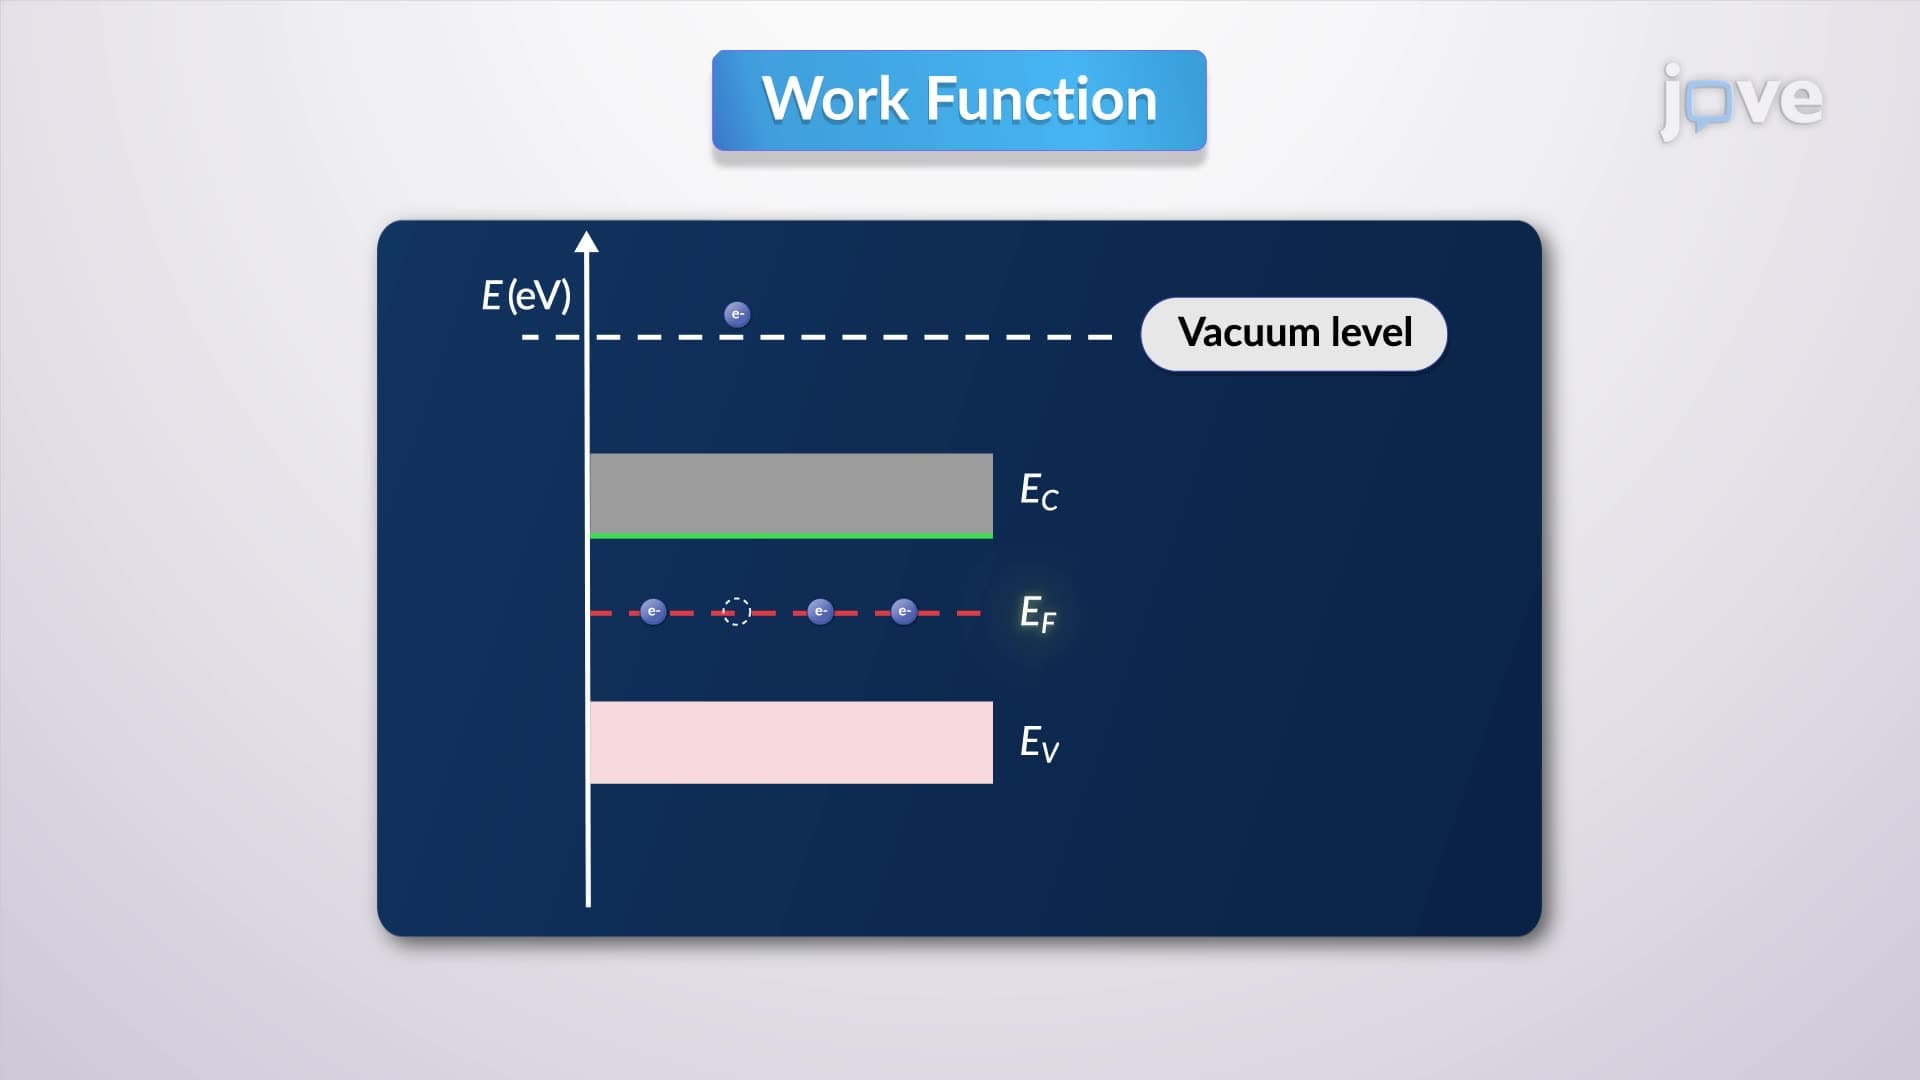
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 03:51
Molecular Orbital Theory II
 03:25
Free Energy Changes for Nonstandard States
 02:35
Molecular Orbital Theory I