Related Experiment Video
Updated: Dec 7, 2025
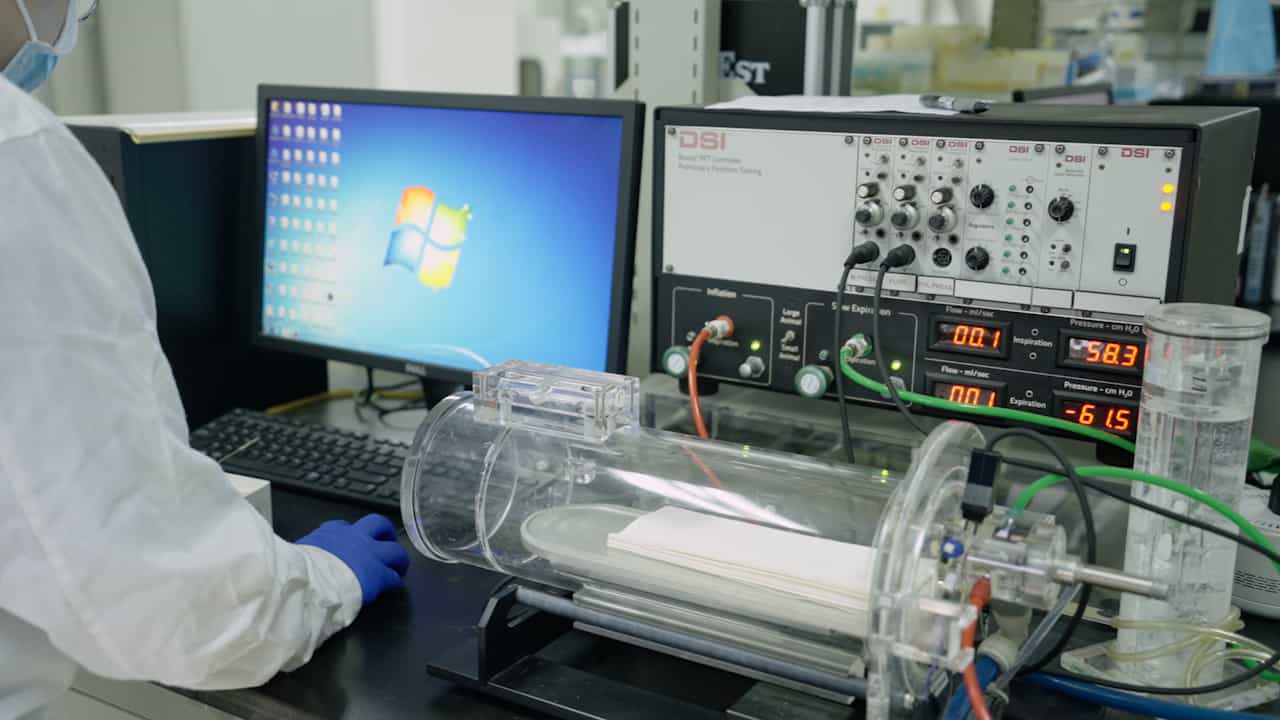

Establishment and Validation of a Rat Model of Pulmonary Arterial Hypertension Associated with Pulmonary Fibrosis
Published on: May 23, 2025
Targeting HIF2α-ARNT hetero-dimerisation as a novel therapeutic strategy for pulmonary arterial hypertension
David Macias1,2, Stephen Moore3,2, Alexi Crosby3
1CRUK Cambridge Centre Early Detection Programme, Dept of Oncology, Hutchison/MRC Research Centre, University of Cambridge, Cambridge, UK.

Abstract:
Pulmonary arterial hypertension (PAH) is a destructive disease of the pulmonary vasculature often leading to right heart failure and death. Current therapeutic intervention strategies only slow disease progression. The role of aberrant hypoxia-inducible factor (HIF)2α stability and function in the initiation and development of pulmonary hypertension (PH) has been an area of intense interest for nearly two decades.Here we determine the effect of a novel HIF2α inhibitor (PT2567) on PH disease initiation and progression, using two pre-clinical models of PH. Haemodynamic measurements were performed, followed by collection of heart, lung and blood for pathological, gene expression and biochemical analysis. Blood outgrowth endothelial cells from idiopathic PAH patients were used to determine the impact of HIF2α-inhibition on endothelial function.Global inhibition of HIF2a reduced pulmonary vascular haemodynamics and pulmonary vascular remodelling in both su5416/hypoxia prevention and intervention models. PT2567 intervention reduced the expression of PH-associated target genes in both lung and cardiac tissues and restored plasma nitrite concentration. Treatment of monocrotaline-exposed rodents with PT2567 reduced the impact on cardiovascular haemodynamics and promoted a survival advantage. In vitro, loss of HIF2α signalling in human pulmonary arterial endothelial cells suppresses target genes associated with inflammation, and PT2567 reduced the hyperproliferative phenotype and overactive arginase activity in blood outgrowth endothelial cells from idiopathic PAH patients. These data suggest that targeting HIF2α hetero-dimerisation with an orally bioavailable compound could offer a new therapeutic approach for PAH. Future studies are required to determine the role of HIF in the heterogeneous PAH population.
Related Concept Videos
 01:18
Treatment for Pulmonary Arterial Hypertension: Endothelin Receptor Antagonists
ETs are synthesized through a complex sequence of enzymatic steps, primarily involving an enzyme referred to as endothelin-converting enzyme...
 01:26
Treatment for Pulmonary Arterial Hypertension: Receptor Tyrosine Kinase Inhibitors and Calcium Channel Blockers
TKIs, such as imatinib (Gleevec), are particularly effective in tackling the growth and mitogenic factors that become upregulated in PAH patients. These factors contribute to the...
 01:23
Treatment for Pulmonary Arterial Hypertension: Prostacyclin Receptor Agonists
These agonists bind to the IPR receptor situated on the plasma membrane of the pulmonary artery smooth muscle cells. This binding triggers a cascade of reactions known as the GS-AC-cAMP-PKA pathway. This pathway results in the relaxation of smooth muscle...
 01:16
Treatment for Pulmonary Arterial Hypertension: Oxygen Therapy for Respiratory Failure
Oxygen therapy is vital in increasing and maintaining blood oxygen levels in PAH patients. As a result, it aids in reducing fatigue,...
 01:30
Pulmonary Hypertension: Classification and Pathogenesis
There are various classifications for PH, each relating to different underlying causes and also...
 01:28
Treatment for Pulmonary Arterial Hypertension: Phosphodiesterase Inhibitors
Among the PDE5 inhibitors, sildenafil (Revatio) stands out as a competitive and selective inhibitor. It operates by elevating cellular levels of cGMP and augmenting signaling through the cGMP-PKG pathway, promoting vasodilation. Upon oral...