Related Experiment Video
Updated: Dec 7, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Theoretical study of the O(3P) + C2H6 reaction based on a new ab initio-based global potential energy surface
J Espinosa-Garcia1, C Rangel, J C Corchado
1Departamento de Química Física and Instituto de Computación Científica Avanzada, Universidad de Extremadura, 06071 Badajoz, Spain. joaquin@unex.es.

Abstract:
A new analytical potential energy surface was developed for the first time for the nine-body O(3P) + C2H6 hydrogen abstraction reaction, named PES-2020, which was fitted to explicitly-correlated high-level electronic structure calculations. This surface simulates the topography of the reactive system, from reactants to products, OH(v,j) + C2H5. The adiabatic energy of reaction, ΔHr(0 K) = -2.33 kcal mol-1, reproduces the experimental evidence, and the barrier height, 10.70 kcal mol-1, agrees with the ab initio calculations used as input. In addition, an intermediate complex in the exit channel is observed, which is stabilized with respect to the products of the reaction. Based on PES-2020 a dynamics study was carried out, where quasi-classical trajectory calculations were performed for collision energies in the range of 7.0-60.0 kcal mol-1, which covers high collision energy regions. The reaction cross section increases with collision energy; the largest fraction of available energy is deposited as translational energy (44-66%), and the scattering distribution evolves from backward to forward with collision energy. These findings reproduce previous theoretical calculations using electronic structure calculations of lower levels. However, where these previous studies failed, viz. in rotational and vibrational OH(v,j) distributions, PES-2020 reproduces practically quantitatively the experimental evidence, i.e., cold vibration and rotation, the rotational distribution peaking at j = 1-3 depending on the collision energy. In sum, this behaviour is typical of gas-phase hydrogen abstraction reactions with direct mechanism and high reaction barrier.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
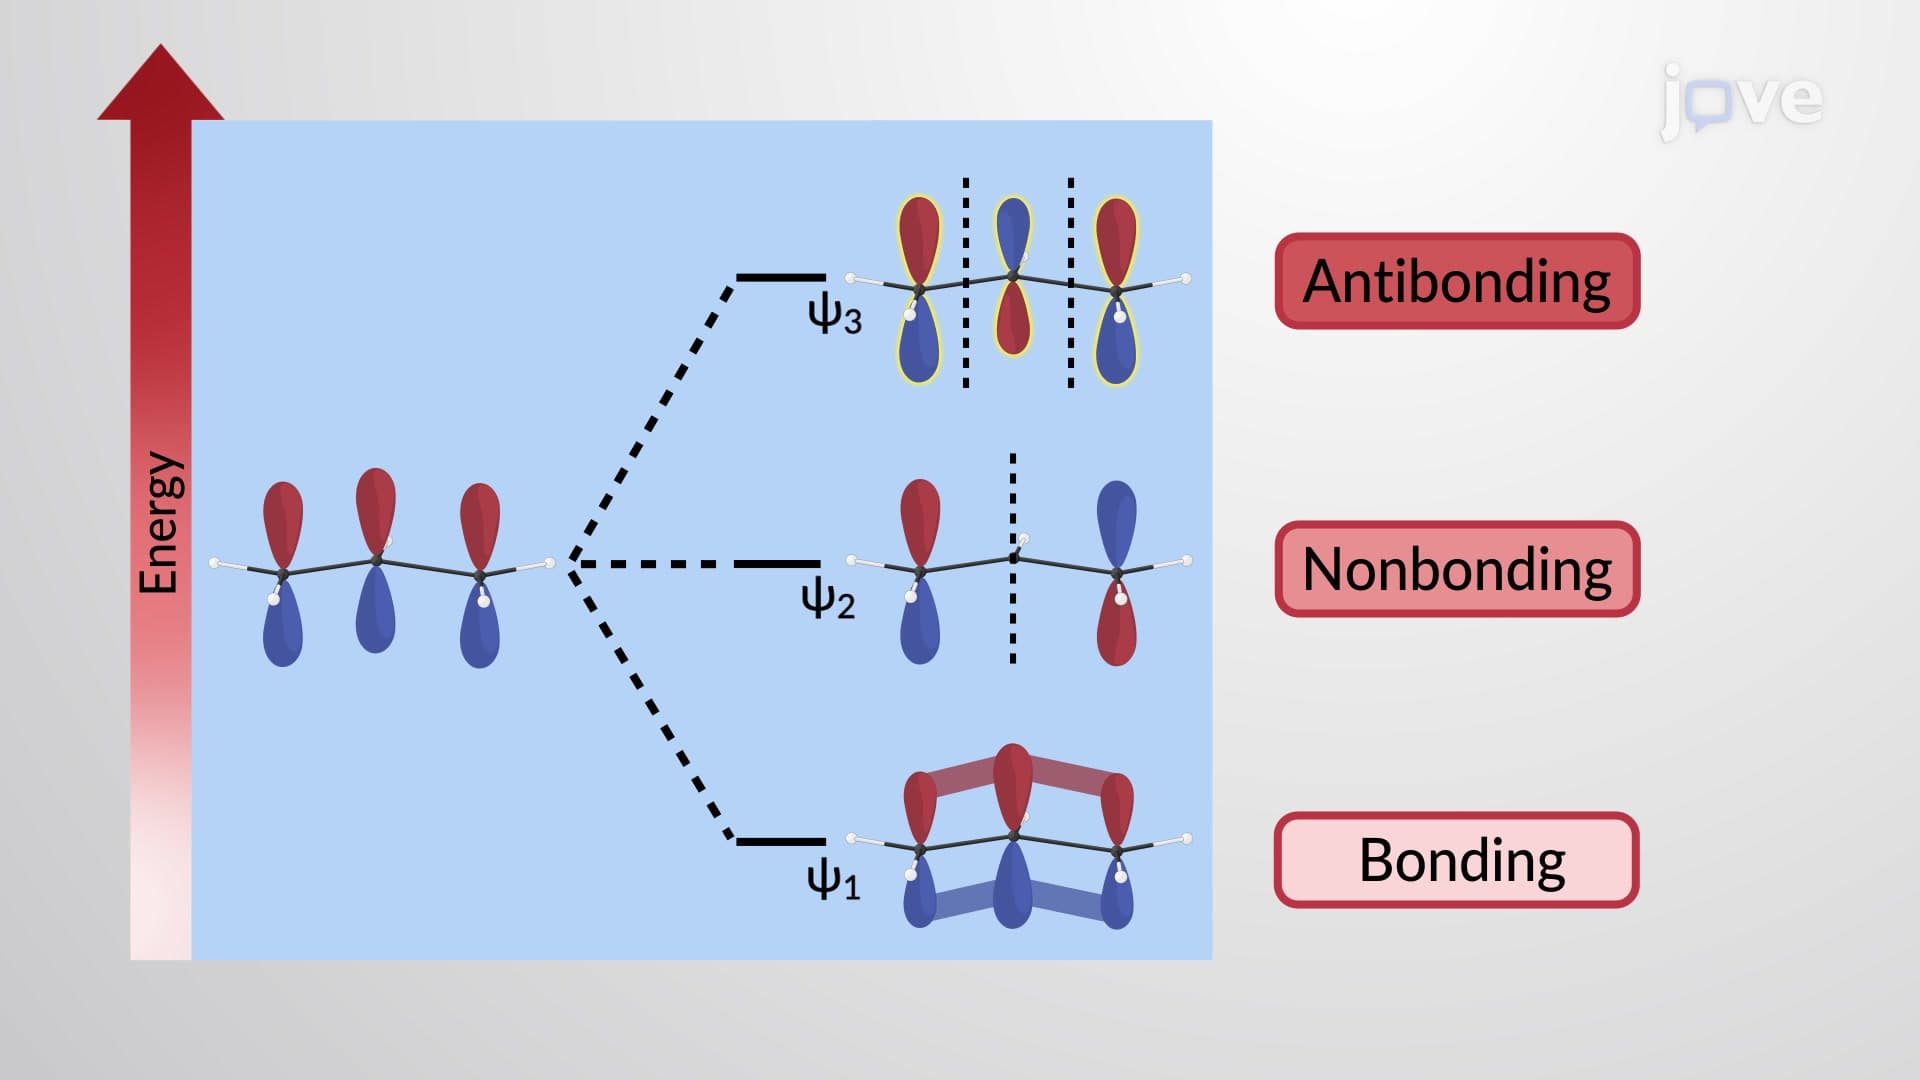 01:18
π Molecular Orbitals of the Allyl Cation and Anion
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...