Related Experiment Video
Updated: Dec 7, 2025
Hyperspectral Imaging as a Tool to Study Optical Anisotropy in Lanthanide-Based Molecular Single Crystals
Published on: April 14, 2020
Crystal structure of rare earth and group III nitride alloys by ab initio calculations
Maciej J Winiarski1, Dorota A Kowalska2
1Institute of Low Temperature and Structure Research, Polish Academy of Sciences, ul. Okólna 2, 50-422, Wrocław, Poland. m.winiarski@int.pan.wroc.pl.

Abstract:
The ground state phases of ternary alloys of rare earth and group III nitride semiconductors have been investigated within the density functional theory. The most energetically favorable crystal phases among possible cubic and hexagonal structures, i.e., the rock salt, zinc blende, wurtzite, and hexagonal BN, were determined. The type of a unit cell and the lattice parameters of the materials are presented as a function of their composition. Furthermore, effects of strain on ground states of group III and rare earth nitride materials are discussed. The findings presented in this work discloses the wurtzite type materials as being stable with relatively low contents of rare earth elements. It is expected that the wurtzite phase will be very persistent only in the La-based systems. Nevertheless, the two-dimensional hexagonal atomic layers are revealed as being a metastable phase for all alloys studied. This finding supports the conclusion of previous experimental reports for Sc-doped GaN systems that the presence of rare earth ions in group III nitride materials leads to flattening of the wurtzite type layers.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 07:24
Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
 03:26
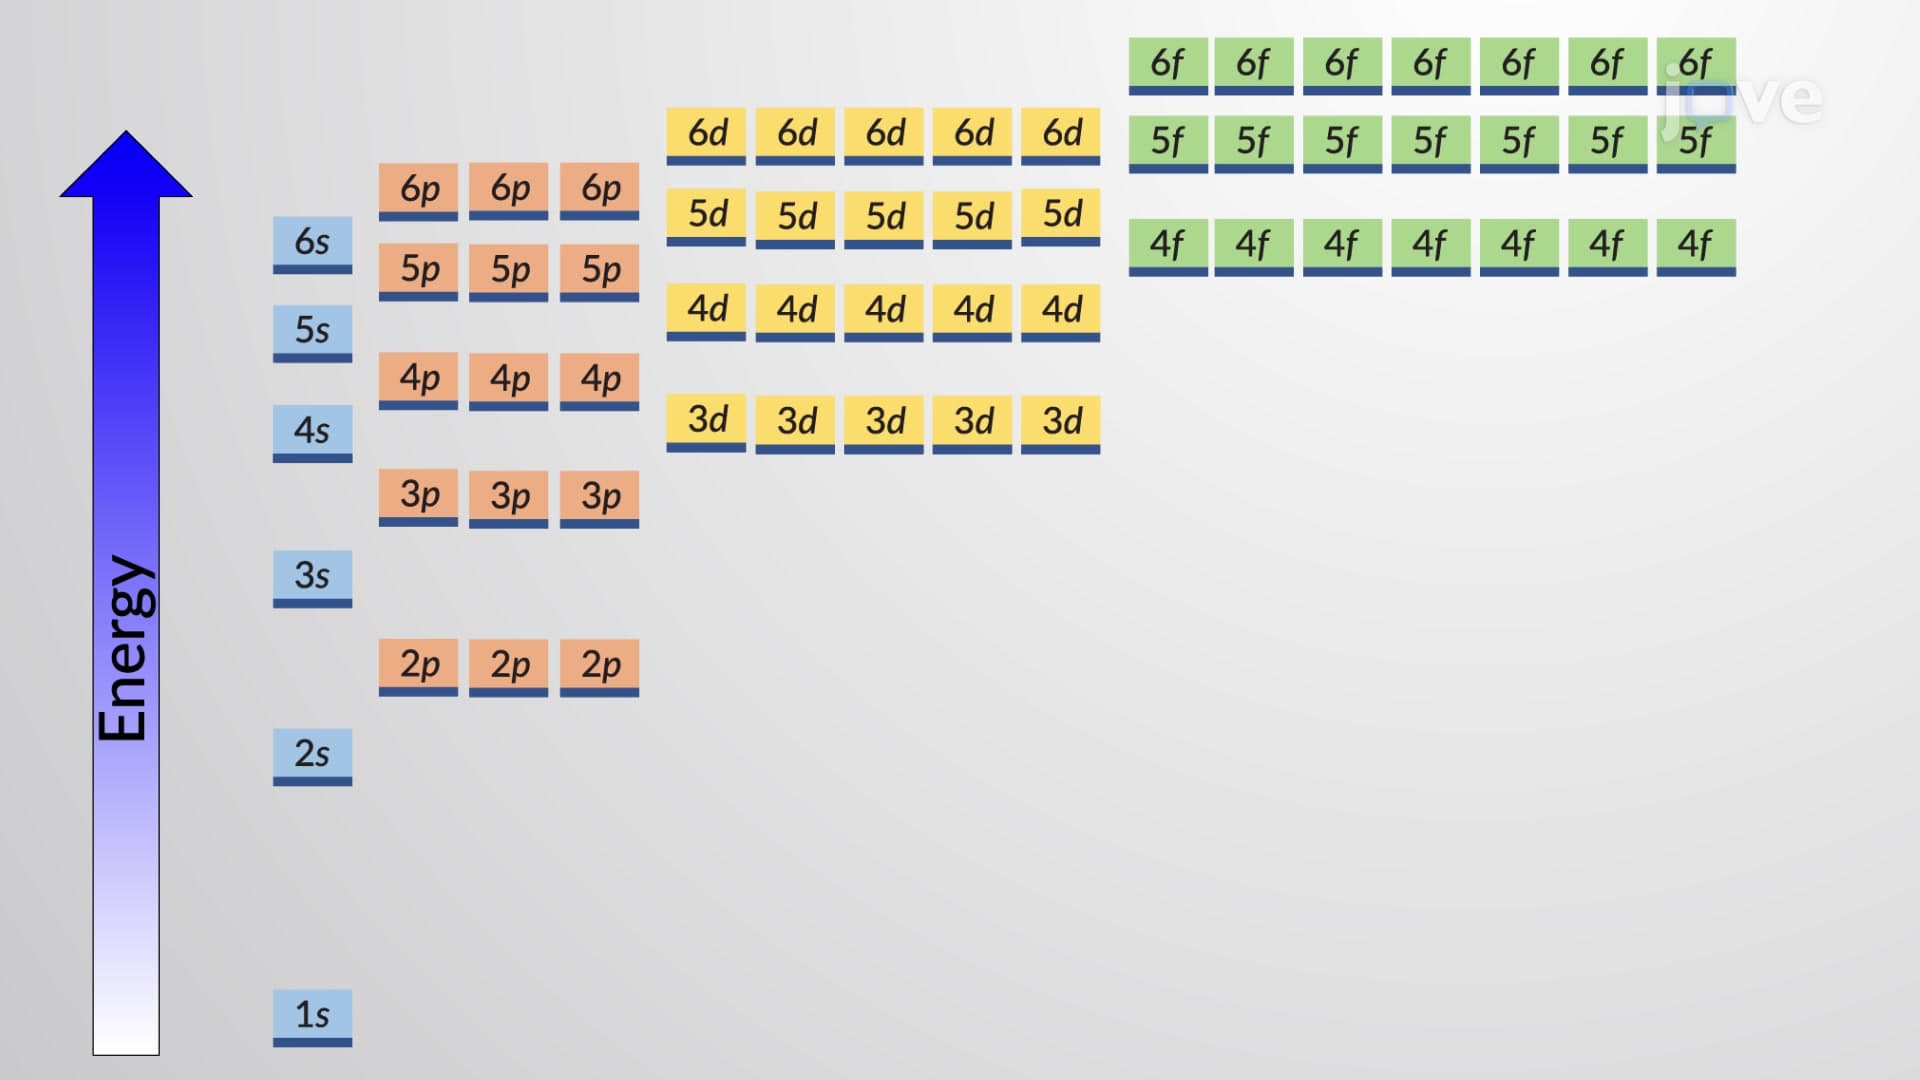
Electron Configuration of Multielectron Atoms
 02:48
Ionic Bonding and Electron Transfer