Related Experiment Video
Updated: Dec 7, 2025

Large Scale Non-targeted Metabolomic Profiling of Serum by Ultra Performance Liquid Chromatography-Mass Spectrometry UPLC-MS
Published on: March 14, 2013
PRiSM: A prototype for exhaustive, restriction-free database searching for mass spectrometry-based identification
Joris Van Houtven1, Kurt Boonen2, Geert Baggerman3
1Flemish Institute for Technological Research (VITO), Boeretang 200, Mol, Belgium.

Rationale:
The current methods for identifying peptides in mass spectral product ion data still struggle to do so for the majority of spectra. Based on the experimental setup and other assumptions, such methods restrict the search space to speed up computations, but at the cost of creating blind spots. The proteomics community would greatly benefit from a method that is capable of covering the entire search space without using any restrictions, thus establishing a baseline for identification.
Methods:
We conceived the "mass pattern paradigm" (MPP) that enables the creation of such an identification method, and we implemented it into a prototype database search engine "PRiSM" (PRotein-Spectrum Matching). We then assessed its operational characteristics by applying it to publicly available high-precision mass spectra of low and high identification difficulty. We used those characteristics to gain theoretical insights into trade-offs between sensitivity and speed when trying to establish a baseline for identification.
Results:
Of 100 low difficulty spectra, PRiSM and SEQUEST agree on 84 identifications (of which 75 are statistically significant). Of 15 of 100 spectra not identified in a previous study (using SEQUEST), 13 are considered reliable after visual inspection and represent 3 proteins (out of 9 in total) not detected previously.
Conclusions:
Despite leaving noise intact, the simple PRiSM prototype can make statistically reliable identifications, while controlling the false discovery rate by fitting a null distribution. It also identifies some spectra previously unidentifiable in an "extremely open" SEQUEST search, paving the way to establishing a baseline for identification in proteomics.
Related Concept Videos
 01:15
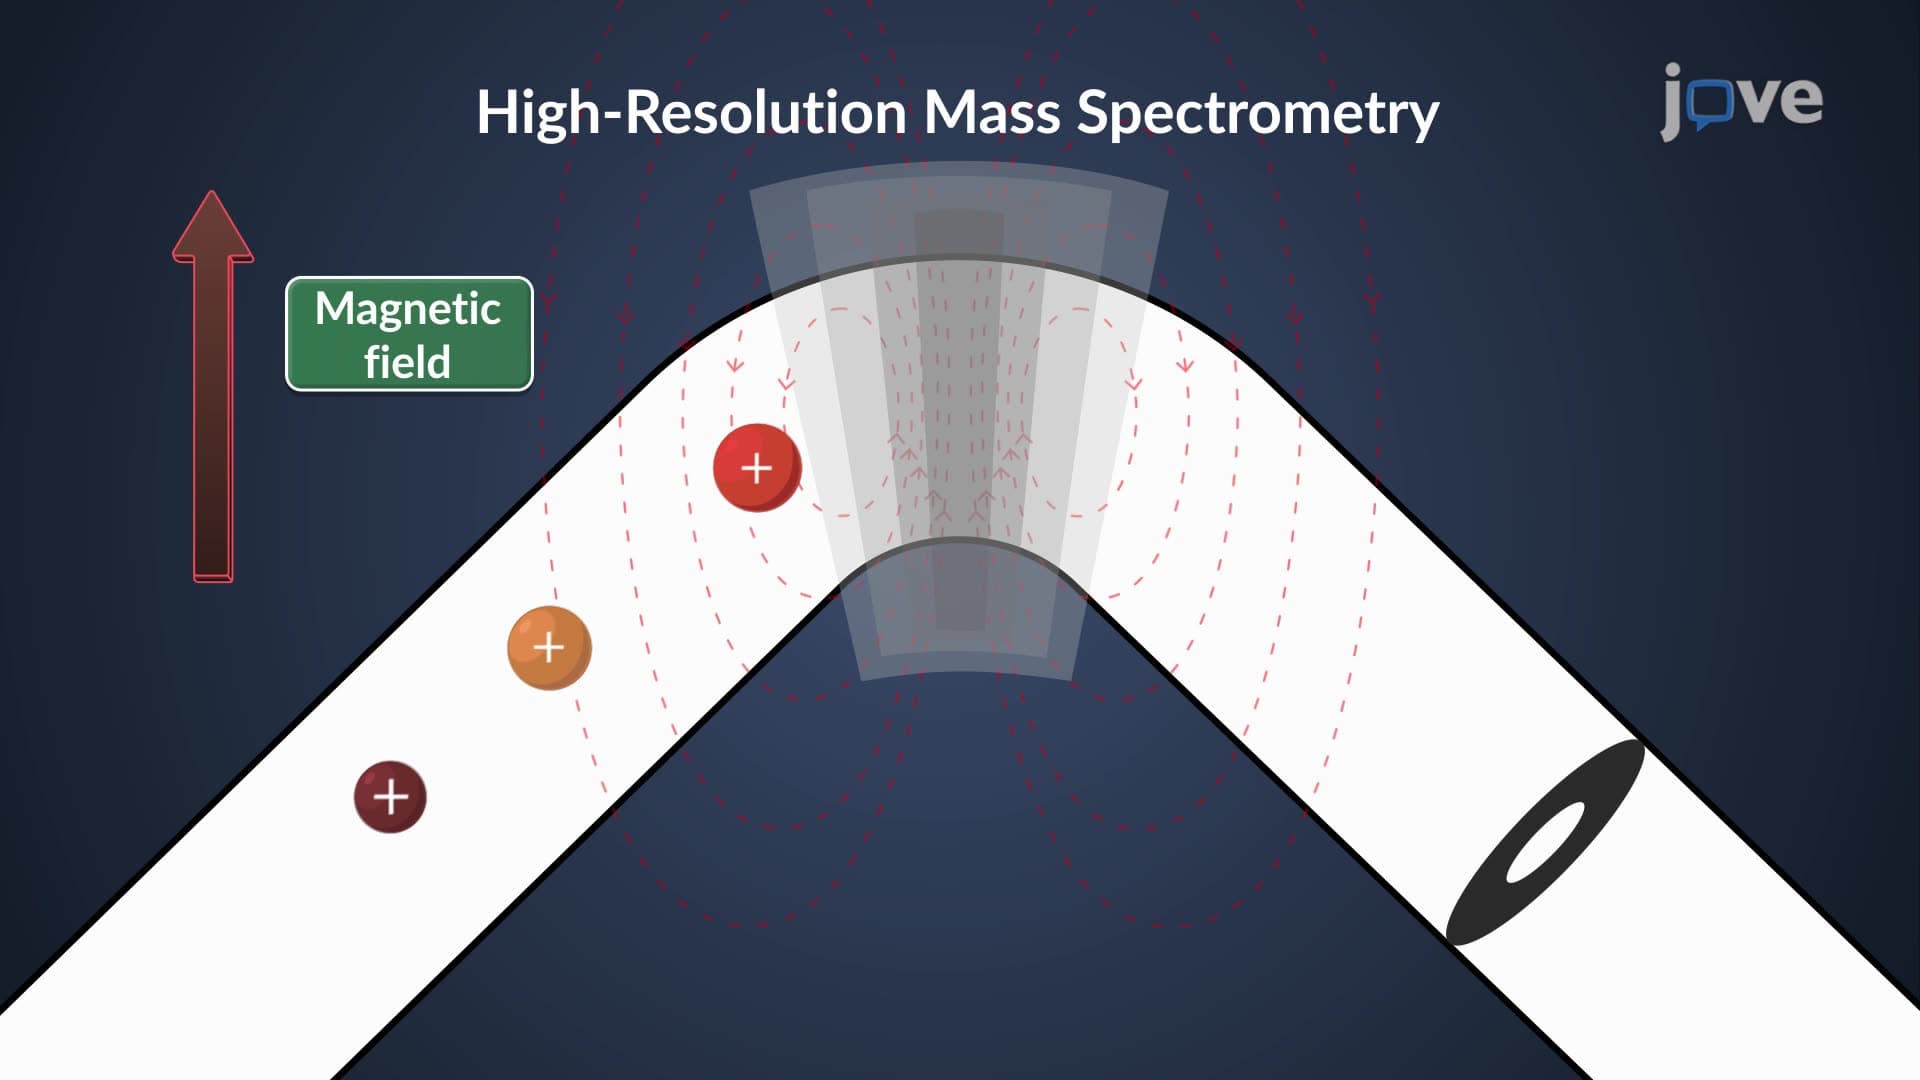
High-Resolution Mass Spectrometry (HRMS)
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:20
Mass Spectrometry: Molecular Fragmentation Overview
One type of fragmentation pattern is the cleavage of a single bond in the molecular ion. The cleavage leads to a radical and a cation. The cleavage can occur at...
 01:13
Mass Analyzers: Overview